TWINBIRD Launches Ultra-Low Temperature Portable Freezer SC-UF25 to Enable a Dry-Ice-Free Cold Chain for Research Samples

TWINBIRD Launches Ultra-Low Temperature Portable Freezer SC-UF25 to Enable a Dry-Ice-Free Cold Chain for Research Samples


TWINBIRD Launches Ultra-Low Temperature Portable Freezer SC-UF25 to Enable a Dry-Ice-Free Cold Chain for Research Samples

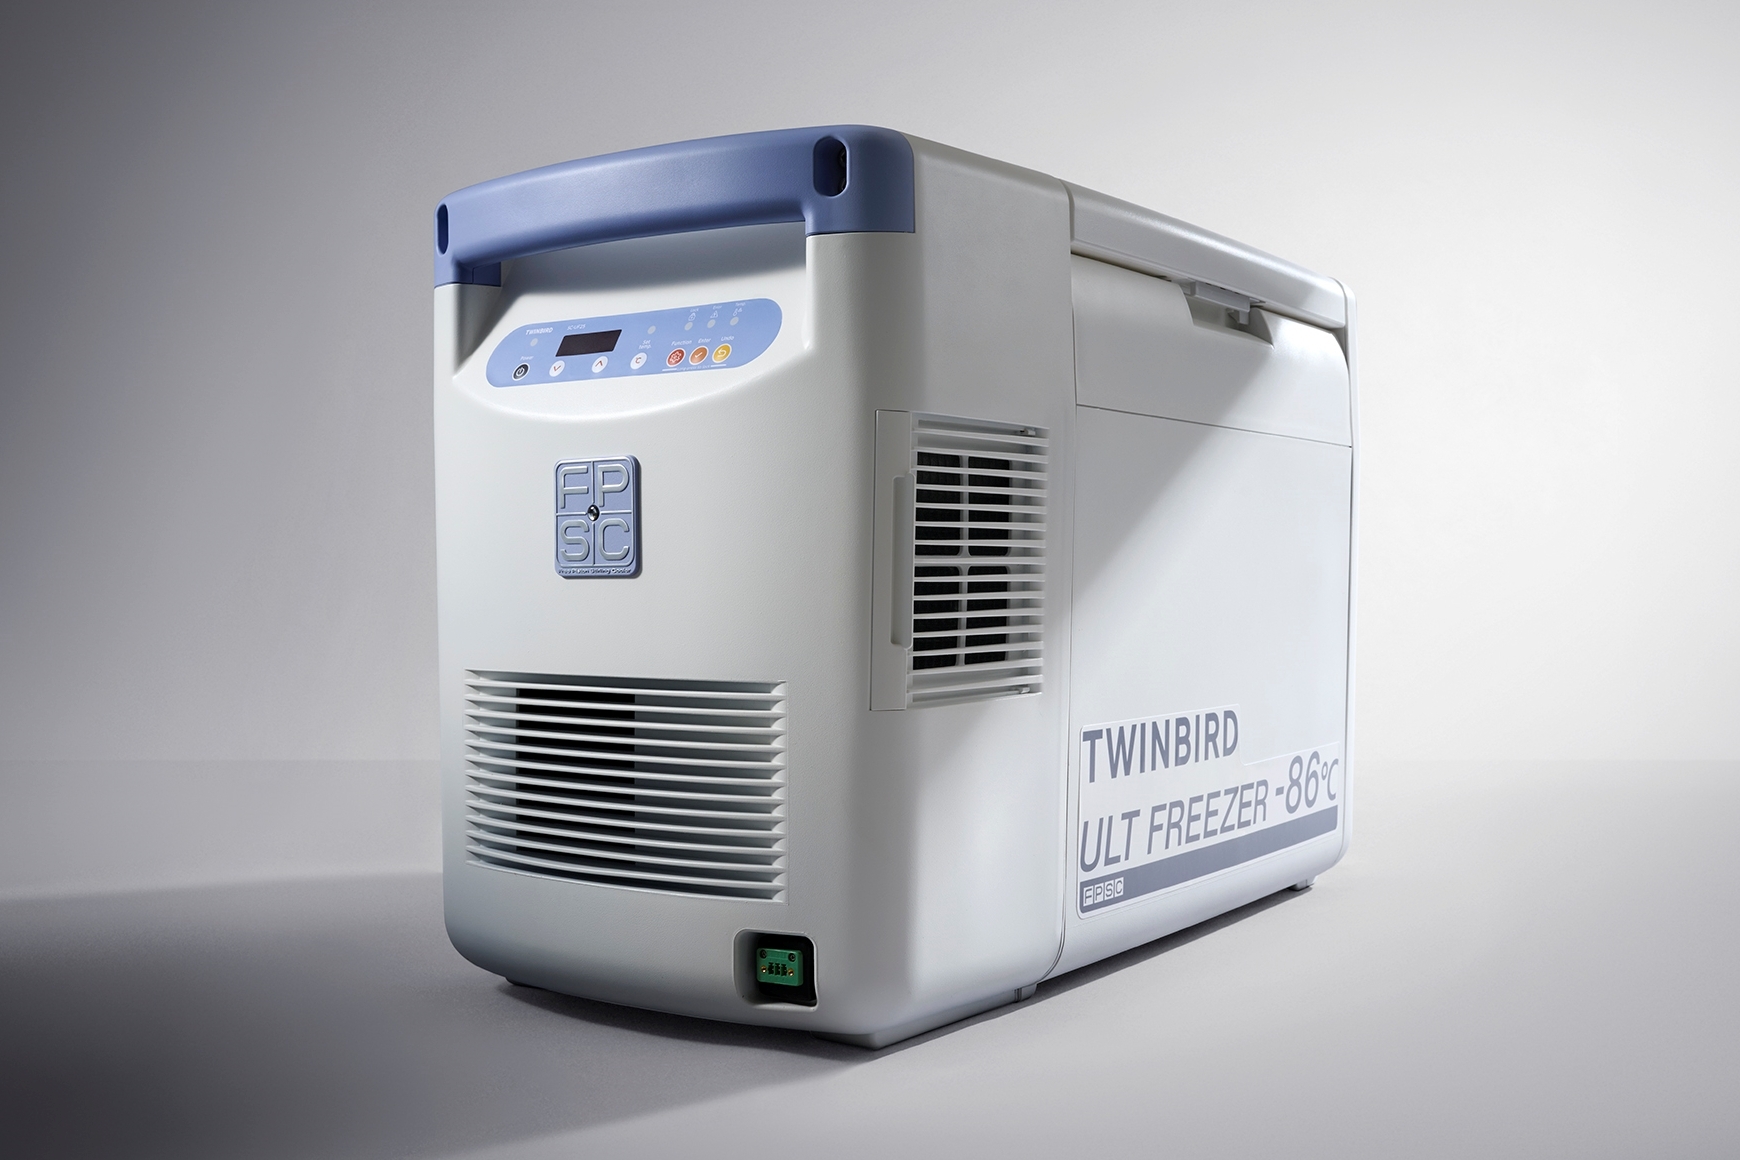
TSUBAME, Japan–(BUSINESS WIRE)–#ColdChain—TWINBIRD Corporation has announced the launch of the SC-UF25 Ultra-Low Temperature Portable Freezer, a compact 25-litre freezer capable of reaching temperatures as low as −86°C. Scheduled for release in November 2026, the new product expands the company’s line-up of solutions powered by its Free-Piston Stirling Cooler (FPSC) technology.


As life-science fields such as regenerative medicine, cell therapy and drug discovery continue to advance, reliable ultra-low-temperature management of research samples is becoming increasingly important. Although storage technologies have evolved, transport and temporary storage often depend on dry ice, posing challenges to temperature stability, operational efficiency and environmental impact. This has increased demand among researchers, laboratories and research organisations for sustainable cold-chain solutions that can maintain ultra-low temperatures from storage through to transport.

To address these challenges, TWINBIRD developed the SC-UF25 using its FPSC technology, which employs helium gas as a refrigerant. The freezer achieves temperatures down to −86°C while remaining compact and lightweight. It supports standard AC power input of 100–240 V, 50/60 Hz, enabling flexible use for stationary storage as well as transport within research facilities and between locations.

Key features include precise temperature control from −86°C to −20°C in 1°C increments, low-vibration and a compact design suitable for laboratories and research institutions where installation space is limited. The product also incorporates precision components manufactured in the renowned Tsubame-Sanjo region of Japan, drawing on advanced engineering expertise developed through decades of manufacturing excellence.

By enabling storage and transport in a −86°C environment, the SC-UF25 helps reduce dependence on dry ice while improving the stability and efficiency of ultra-low temperature cold-chain operations. Through the application of FPSC technology, TWINBIRD aims to support the preservation of high-value research materials and contribute to the advancement of life-science research and operations.

The SC-UF25 will be showcased at MEDICA 2026, one of the world’s largest trade fairs for the medical and life-science sectors, held in Düsseldorf, Germany, from 16 to 19 November 2026, where TWINBIRD will highlight the product’s contribution to sustainable ultra-low temperature cold-chain applications for research, laboratory and life-science use.

Disclaimer: The SC-UF25 is for research, laboratory and life-science cold-chain use only. It is not registered under the EU Medical Device Regulation and is not intended for diagnosis, treatment, clinical decisions, or patient care.

Contacts

SC Business Sales, TWINBIRD Corporation fpsc@twinbird.co.jp

TG Therapeutics Announces Publication of Five-Year Comparative Analysis of BRIUMVI Versus Ocrelizumab and Ofatumumab in Relapsing Multiple Sclerosis

TG Therapeutics Announces Publication of Five-Year Comparative Analysis of BRIUMVI Versus Ocrelizumab and Ofatumumab in Relapsing Multiple Sclerosis


TG Therapeutics Announces Publication of Five-Year Comparative Analysis of BRIUMVI Versus Ocrelizumab and Ofatumumab in Relapsing Multiple Sclerosis

BRIUMVI was associated with a lower risk of 24-week confirmed disability progression and lower annualized relapse rates (ARRs) over five years compared with ocrelizumab and ofatumumab
NEW YORK–(BUSINESS WIRE)–$TGTX–TG Therapeutics, Inc. (NASDAQ: TGTX), today announced the publication of results from a five-year matching-adjusted indirect comparison (MAIC) evaluating long-term relapse activity and disability progression outcomes with BRIUMVI® (ublituximab-xiiy) compared with ocrelizumab and ofatumumab in patients with relapsing multiple sclerosis (RMS). For the analysis, individual patient data from the Phase 3 ULTIMATE I/II pivotal trials for BRIUMVI were reweighted to match published baseline characteristics from the pivotal studies for ocrelizumab and ofatumumab in patients with RMS, revealing favorable long-term outcomes with BRIUMVI compared to the other two comparators. The manuscript, entitled “Long-Term Comparative Outcomes in Relapse Activity and Disability Progression Among Anti-CD20 Therapies in Relapsing Multiple Sclerosis: An Unanchored Matching-Adjusted Indirect Comparison,” was published online in Multiple Sclerosis and Related Disorders.

Michael S. Weiss, Chairman and Chief Executive Officer of TG Therapeutics, stated, “In an era where physicians and patients are fortunate to have multiple anti-CD20 treatment options available for relapsing MS, the lack of comparative data makes choosing the right one challenging. This is particularly important for people living with a chronic disease like MS, where preventing the accumulation of disability over time is an important treatment goal.”

Mr. Weiss continued, “In the absence of head-to-head clinical trials, MAIC provides a valuable statistical approach to evaluating comparative treatment outcomes while accounting for differences in patient populations across studies. We are encouraged that this analysis showed that BRIUMVI was associated with consistently lower relapse activity versus ocrelizumab and a lower risk of confirmed disability progression compared with both ocrelizumab and ofatumumab over five years. We believe these findings add to the growing body of evidence supporting the durable efficacy profile of BRIUMVI and its potential to provide meaningful long-term benefit for people living with MS.”

The manuscript can be accessed at ScienceDirect.com or on the publications page of the TG corporate website at www.tgtherapeutics.com/publications/.

Key Findings from the Publication

Five-year anti-CD20 outcomes from the pivotal studies of BRIUMVI, ocrelizumab, and ofatumumab were compared using an unanchored MAIC (see below for more information about the MAIC). The analysis used individual patient-level data from 545 BRIUMVI (ublituximab)-treated participants in the Phase 3 ULTIMATE I and II trials and their open-label extension and compared outcomes with published aggregate data from the Phase 3 OPERA I/II trials of ocrelizumab and ASCLEPIOS I/II trials of ofatumumab. In the analysis, BRIUMVI patient-level data were re-weighted to align baseline characteristics with the respective comparator trial populations. These adjustments are designed to mitigate the effects of any demographic differences in the study populations on the efficacy results.

The findings included:

  • BRIUMVI was associated with a lower risk of 24-week confirmed disability progression (CDP24) compared with both ocrelizumab and ofatumumab, with a 44% lower relative hazard versus ocrelizumab (HR 0.560; 95% CI: 0.380–0.826) and a 49% lower relative hazard versus ofatumumab (HR 0.509; 95% CI: 0.350–0.741).
  • BRIUMVI was associated with a lower cumulative incidence of CDP24 at 240 weeks, with estimated rates of 9.3% versus 16.1% for ocrelizumab and 9.4% versus 17.9% for ofatumumab.
  • BRIUMVI was associated with more time free from confirmed disability progression over 240 weeks, with an average of 8.1 additional weeks compared with ocrelizumab (95% CI: 2.3–14.1) and 13.1 additional weeks compared with ofatumumab (95% CI: 6.7–19.3).
  • BRIUMVI demonstrated consistently low annualized relapse rates (ARR) over five years of treatment, with MAIC-adjusted ARR declining from 0.118 in Year 1 to 0.032 in Year 5 in the comparison with ocrelizumab, and from 0.116 to 0.030 in the comparison with ofatumumab.
  • These findings are considered exploratory and require confirmation in direct comparative studies.

About the MAIC Analysis

In the absence of randomized head-to-head trials directly comparing anti-CD20 therapies in RMS, investigators conducted an unanchored MAIC to explore comparative outcomes over five years. Individual patient data from the pivotal Phase 3 ULTIMATE I and II studies were reweighted to match published baseline characteristics from the pivotal Phase 3 studies for ocrelizumab (OPERA I/II) and ofatumumab (ASCLEPIOS I/II), including age, sex, baseline Expanded Disability Status Scale (EDSS) score, gadolinium-enhancing lesion prevalence, T2 lesion volume, prior relapse activity and time since diagnosis, where corresponding data were available. Following weighting, baseline characteristics were closely aligned with both comparator populations.

The authors note that the findings should be interpreted within the inherent limitations of an unanchored MAIC. Notably, the methodology cannot adjust for unmeasured differences between trials, and residual confounding cannot be excluded. Accordingly, the analysis is exploratory and the findings require confirmation through direct comparative studies or robust real-world evidence.

The study was funded by Neuraxpharm, TG Therapeutics’ European commercialization partner for BRIUMVI.

ABOUT THE ULTIMATE I & II PHASE 3 TRIALS

ULTIMATE I & II are two randomized, double-blind, double-dummy, parallel group, active comparator-controlled clinical trials of identical design, in patients with RMS treated for 96 weeks. Patients were randomized to receive either BRIUMVI, given as an IV infusion of 150 mg administered in four hours, 450 mg two weeks after the first infusion administered in one hour, and 450 mg every 24 weeks administered in one hour, with oral placebo administered daily; or teriflunomide, the active comparator, given orally as a 14 mg daily dose with IV placebo administered on the same schedule as BRIUMVI. Both studies enrolled patients who had experienced at least one relapse in the previous year, two relapses in the previous two years, or had the presence of a T1 gadolinium (Gd)-enhancing lesion in the previous year. Patients were also required to have an Expanded Disability Status Scale (EDSS) score from 0 to 5.5 at baseline. The ULTIMATE I & II trials enrolled a total of 1,094 patients with RMS across 10 countries. These trials were led by Lawrence Steinman, MD, Zimmermann Professor of Neurology & Neurological Sciences, and Pediatrics at Stanford University. Additional information on these clinical trials can be found at www.clinicaltrials.gov (NCT03277261; NCT03277248).

ABOUT TG THERAPEUTICS

TG Therapeutics is a fully integrated, commercial stage, biotechnology company focused on the acquisition, development and commercialization of novel treatments for B-cell diseases. In addition to a research pipeline, TG Therapeutics has received approval from the U.S. Food and Drug Administration (FDA) for BRIUMVI® (ublituximab-xiiy) to treat adult patients with relapsing forms of multiple sclerosis, including clinically isolated syndrome, relapsing-remitting disease, and active secondary progressive disease, as well as approval from several regulatory agencies outside of the U.S. for BRIUMVI to treat adult patients with RMS who have active disease defined by clinical or imaging features. For more information, visit www.tgtherapeutics.com, and follow us on X (formerly Twitter) @TGTherapeutics and on LinkedIn.

BRIUMVI® is a registered trademark of TG Therapeutics, Inc.

ABOUT BRIUMVI® (ublituximab-xiiy) 150 mg/6 mL Injection for IV

BRIUMVI is a novel monoclonal antibody that targets a unique epitope on CD20-expressing B-cells. Targeting CD20 using monoclonal antibodies has proven to be an important therapeutic approach for the management of autoimmune disorders, such as RMS. BRIUMVI is uniquely designed to lack certain sugar molecules normally expressed on the antibody. Removal of these sugar molecules, a process called glycoengineering, allows for efficient B-cell depletion at low doses.

BRIUMVI is indicated in the U.S. for the treatment of adults with RMS, including clinically isolated syndrome, relapsing-remitting disease, and active secondary progressive disease and in several countries outside of the U.S. for the treatment of adult patients with RMS with active disease defined by clinical or imaging features.

A list of authorized specialty distributors can be found at www.briumvi.com.

IMPORTANT SAFETY INFORMATION

Contraindications: BRIUMVI is contraindicated in patients with:

  • Active Hepatitis B Virus infection
  • A history of life-threatening infusion reaction to BRIUMVI

WARNINGS AND PRECAUTIONS

Infusion Reactions: BRIUMVI can cause infusion reactions, which can include pyrexia, chills, headache, influenza-like illness, tachycardia, nausea, throat irritation, erythema, and an anaphylactic reaction. In MS clinical trials, the incidence of infusion reactions in BRIUMVI-treated patients who received infusion reaction-limiting premedication prior to each infusion was 48%, with the highest incidence within 24 hours of the first infusion. 0.6% of BRIUMVI-treated patients experienced infusion reactions that were serious, some requiring hospitalization.

Observe treated patients for infusion reactions during the infusion and for at least one hour after the completion of the first two infusions unless infusion reaction and/or hypersensitivity has been observed in association with the current or any prior infusion. Inform patients that infusion reactions can occur up to 24 hours after the infusion. Administer the recommended pre-medication to reduce the frequency and severity of infusion reactions. If life-threatening, stop the infusion immediately, permanently discontinue BRIUMVI, and administer appropriate supportive treatment. Less severe infusion reactions may involve temporarily stopping the infusion, reducing the infusion rate, and/or administering symptomatic treatment.

Infections: Serious, life-threatening or fatal, bacterial and viral infections have been reported in BRIUMVI-treated patients. In MS clinical trials, the overall rate of infections in BRIUMVI-treated patients was 56% compared to 54% in teriflunomide-treated patients. The rate of serious infections was 5% compared to 3% respectively. There were 3 infection-related deaths in BRIUMVI-treated patients. The most common infections in BRIUMVI-treated patients included upper respiratory tract infection (45%) and urinary tract infection (10%). Delay BRIUMVI administration in patients with an active infection until the infection is resolved.

Consider the potential for increased immunosuppressive effects when initiating BRIUMVI after immunosuppressive therapy or initiating an immunosuppressive therapy after BRIUMVI.

Hepatitis B Virus (HBV) Reactivation: HBV reactivation occurred in an MS patient treated with BRIUMVI in clinical trials. Fulminant hepatitis, hepatic failure, and death caused by HBV reactivation have occurred in patients treated with anti-CD20 antibodies. Perform HBV screening in all patients before initiation of treatment with BRIUMVI. Do not start treatment with BRIUMVI in patients with active HBV confirmed by positive results for HB surface antigen (HBsAg) and anti-HB tests. For patients who are negative for HBsAg and positive for HB core antibody [HBcAb+] or are carriers of HBV [HBsAg+], consult a liver disease expert before starting and during treatment.

Progressive Multifocal Leukoencephalopathy (PML): PML is an opportunistic viral infection of the brain caused by the JC virus (JCV) that typically only occurs in patients who are immunocompromised, and that usually leads to death or severe disability. JCV infection resulting in PML has been observed in patients treated with anti-CD20 antibodies, including BRIUMVI, and other MS therapies.

If PML is suspected, withhold BRIUMVI and perform an appropriate diagnostic evaluation. Typical symptoms associated with PML are diverse, progress over days to weeks, and include progressive weakness on one side of the body or clumsiness of limbs, disturbance of vision, and changes in thinking, memory, and orientation leading to confusion and personality changes.

MRI findings may be apparent before clinical signs or symptoms; monitoring for signs consistent with PML may be useful. Further investigate suspicious findings to allow for an early diagnosis of PML, if present. Following discontinuation of another MS medication associated with PML, lower PML-related mortality and morbidity have been reported in patients who were initially asymptomatic at diagnosis compared to patients who had characteristic clinical signs and symptoms at diagnosis.

If PML is confirmed, treatment with BRIUMVI should be discontinued.

Vaccinations: Administer all immunizations according to immunization guidelines: for live or live-attenuated vaccines, at least 4 weeks and, whenever possible, at least 2 weeks prior to initiation of BRIUMVI for non-live vaccines. BRIUMVI may interfere with the effectiveness of non-live vaccines. The safety of immunization with live or live-attenuated vaccines during or following administration of BRIUMVI has not been studied. Vaccination with live virus vaccines is not recommended during treatment and until B-cell repletion.

Vaccination of Infants Born to Mothers Treated with BRIUMVI During Pregnancy: In infants of mothers exposed to BRIUMVI during pregnancy, assess B-cell counts prior to administration of live or live-attenuated vaccines as measured by CD19+ B-cells. Depletion of B-cells in these infants may increase the risks from live or live-attenuated vaccines. Inactivated or non-live vaccines may be administered prior to B-cell recovery. Assessment of vaccine immune responses, including consultation with a qualified specialist, should be considered to determine whether a protective immune response was mounted.

Fetal Risk: Based on data from animal studies, BRIUMVI may cause fetal harm when administered to a pregnant woman. Transient peripheral B-cell depletion and lymphocytopenia have been reported in infants born to mothers exposed to other anti-CD20 B-cell depleting antibodies during pregnancy. Advise females of reproductive potential to use effective contraception during BRIUMVI treatment and for 6 months after the last dose.

Reduction in Immunoglobulins: As expected with any B-cell depleting therapy, decreased immunoglobulin levels were observed. Decrease in immunoglobulin M (IgM) was reported in 0.6% of BRIUMVI-treated patients compared to none of the patients treated with teriflunomide in RMS clinical trials. Monitor the levels of quantitative serum immunoglobulins during treatment, especially in patients with opportunistic or recurrent infections, and after discontinuation of therapy, until B-cell repletion. Consider discontinuing BRIUMVI therapy if a patient with low immunoglobulins develops a serious opportunistic infection or recurrent infections, or if prolonged hypogammaglobulinemia requires treatment with intravenous immunoglobulins.

Liver Injury: Clinically significant liver injury, without findings of viral hepatitis, has been reported in the postmarketing setting in patients treated with anti-CD20 B-cell depleting therapies approved for the treatment of MS, including BRIUMVI. Signs of liver injury, including markedly elevated serum hepatic enzymes with elevated total bilirubin, have occurred from weeks to months after administration.

Patients treated with BRIUMVI found to have an alanine aminotransaminase (ALT) or aspartate aminotransferase (AST) greater than 3x the upper limit of normal (ULN) with serum total bilirubin greater than 2x ULN are potentially at risk for severe drug-induced liver injury.

Obtain liver function tests prior to initiating treatment with BRIUMVI, and monitor for signs and symptoms of any hepatic injury during treatment. Measure serum aminotransferases, alkaline phosphatase, and bilirubin levels promptly in patients who report symptoms that may indicate liver injury, including new or worsening fatigue, anorexia, nausea, vomiting, right upper abdominal discomfort, dark urine, or jaundice. If liver injury is present and an alternative etiology is not identified, discontinue BRIUMVI.

Most Common Adverse Reactions: The most common adverse reactions in RMS trials (incidence of at least 10%) were infusion reactions and upper respiratory tract infections.

Physicians, pharmacists, or other healthcare professionals with questions about BRIUMVI should visit www.briumvi.com.

ABOUT BRIUMVI PATIENT SUPPORT in the U.S.

BRIUMVI Patient Support is a flexible program designed by TG Therapeutics to support U.S. patients through their treatment journey in a way that works best for them. More information about the BRIUMVI Patient Support program can be accessed at www.briumvipatientsupport.com.

ABOUT MULTIPLE SCLEROSIS

Relapsing multiple sclerosis (RMS) is a chronic demyelinating disease of the central nervous system (CNS) and includes people with relapsing-remitting multiple sclerosis (RRMS) and people with secondary progressive multiple sclerosis (SPMS) who continue to experience relapses. RRMS is the most common form of multiple sclerosis (MS) and is characterized by episodes of new or worsening signs or symptoms (relapses) followed by periods of recovery. It is estimated that nearly 1 million people are living with MS in the United States and approximately 85% are initially diagnosed with RRMS.1,2 The majority of people who are diagnosed with RRMS will eventually transition to SPMS, in which they experience steadily worsening disability over time. Worldwide, more than 2.3 million people have a diagnosis of MS.1

Cautionary Statement

This press release contains forward-looking statements that involve a number of risks and uncertainties. For those statements, we claim the protection of the safe harbor for forward-looking statements contained in the Private Securities Litigation Reform Act of 1995.

Any forward-looking statements in this press release are based on management’s current expectations and beliefs and are subject to a number of risks, uncertainties and important factors that may cause actual events or results to differ materially from those expressed or implied by any forward-looking statements contained in this press release. In addition to the risk factors identified from time to time in our reports filed with the U.S. Securities and Exchange Commission (SEC), factors that could cause our actual results to differ materially include the below.

Such forward-looking statements include, but are not limited to, statements regarding the potential significance and implications of the published results; the potential for BRIUMVI (ublituximab-xiiy) to provide meaningful long-term benefit for people living with relapsing multiple sclerosis (RMS); the potential for the published findings to support the efficacy profile, differentiation, utilization, acceptance or commercial opportunity for BRIUMVI; and expectations regarding the continued commercialization, healthcare professional and patient acceptance, and use of BRIUMVI in the United States and jurisdictions outside the United States.

Additional factors that could cause actual results to differ materially include: the inherent limitations of matching-adjusted indirect comparisons and other cross-trial analyses; the risk that the results of the analysis may not be confirmed in other clinical studies or real-world evidence; the risk that differences in trial design, patient populations, treatment exposure, follow-up, unmeasured or unavailable prognostic factors, treatment effect modifiers, or other sources of residual confounding may affect the interpretation of the results; the risk that the results may not be indicative of outcomes in broader patient populations or clinical practice; and the uncertainties generally inherent in research and development and the interpretation of clinical and comparative effectiveness data.

Any forward-looking statements set forth in this press release speak only as of the date of this press release. We do not undertake to update any of these forward-looking statements to reflect events or circumstances that occur after the date hereof. This press release and prior releases are available at www.tgtherapeutics.com. The information found on our website is not incorporated by reference into this press release and is included for reference purposes only.

1.

MS Prevalence. National Multiple Sclerosis Society website. https://www.nationalmssociety.org/About-the-Society/MS-Prevalence. Accessed October 26, 2020.

2.

Multiple Sclerosis International Federation, 2013 via Datamonitor p. 236.


Contacts

Investor Relations:
Email: ir@tgtxinc.com
Telephone: 1.877.575.TGTX (8489), Option 4

Media Relations:
Email: media@tgtxinc.com
Telephone: 1.877.575.TGTX (8489), Option 6
Fujifilm Launches Synapse Pathology v3.0, Advancing the Digital Pathology Experience With Expanded AI Capabilities

Fujifilm Launches Synapse Pathology v3.0, Advancing the Digital Pathology Experience With Expanded AI Capabilities


Fujifilm Launches Synapse Pathology v3.0, Advancing the Digital Pathology Experience With Expanded AI Capabilities

Major software release introduces more than 50 new features and functions, AI orchestration capabilities, and Fujifilm’s pathology-specific foundation model, Synapse Pathology AI Foundry

LEXINGTON, Mass.–(BUSINESS WIRE)–FUJIFILM Healthcare Americas Corporation, a leading provider of diagnostic and enterprise imaging solutions, today announced the launch of Synapse® Pathology v3.0. Featuring more than 50 new features and functions, Synapse Pathology v3.0 is designed to enhance the user experience while expanding the platform’s artificial intelligence (AI) capabilities to support the evolving needs of pathologists and laboratories. The new release will be showcased at Pathology Visions 2026 in Fujifilm booth #603, taking place October 17–20 in San Diego, California.


“Digital pathology is evolving rapidly, and our customers are looking for solutions that not only help improve the experience of working in a digital environment today, but also prepare them for the AI-driven workflows of tomorrow,” said Dr. Mark Lloyd, vice president of digital pathology, FUJIFILM Healthcare Americas Corporation. “Synapse Pathology v3.0 represents a major step forward for our platform. We listened closely to our customers and translated their feedback into meaningful enhancements across navigation, measurement, classification and organization, while simultaneously creating new opportunities to bring AI directly into the pathology workflow.”

New enhancements focused on the user experience

User experience is at the center of Synapse Pathology v3.0. Fujifilm incorporated customer feedback to introduce dozens of new and enhanced capabilities designed to help pathologists navigate, evaluate, measure, classify and organize digital pathology cases more efficiently. New enhancements include Smart Worklist sorting, new navigation devices, slide tagging, new measurement tools and enhancements to collaboration tools. The release builds upon Synapse Pathology’s existing capabilities while providing laboratories with an increasingly flexible platform designed to support digital workflows as their needs evolve.

Among the most significant advancements in v3.0 is expanded support for AI. Fujifilm has enhanced its existing proprietary algorithms, including TissueSync® and TissueDirect®. Synapse Pathology v3.0 also introduces new capabilities designed to simplify access to third-party AI applications, including Ibex’s AI-powered solutions for malignancy detection in prostate, breast and gastric specimens.

Creating a connected ecosystem for pathology AI

Synapse Pathology v3.0 incorporates Synapse AI Orchestrator, Fujifilm’s fully automated distribution hub designed to connect Synapse Pathology with Fujifilm’s growing ecosystem of third-party AI partners. Synapse AI Orchestrator is designed to streamline the exchange of information between the Synapse Pathology platform and compatible AI applications, creating an infrastructure where laboratories can incorporate specialized algorithms into their digital pathology workflows.

Rather than requiring pathologists to navigate separate systems and applications, Synapse AI Orchestrator is designed to provide a scalable foundation for integrating AI capabilities into the broader digital pathology environment.

Introducing Synapse Pathology AI Foundry, Fujifilm’s pathology foundation model (RUO)*

At Pathology Visions, Fujifilm will also introduce Synapse Pathology AI Foundry, the company’s large-scale, pathology-specific AI foundation model trained using millions of pathology images.

Unlike AI models developed for a single, narrowly defined application, foundation models are designed to provide a broad technological base that can support multiple downstream applications. Synapse Pathology AI Foundry will serve as Synapse Pathology’s foundation for a range of current and future pathology AI capabilities.

During Pathology Visions, Fujifilm will showcase the world’s first foundation model for screening in pathology. The model is designed to identify and distinguish normal, inflammatory, atypical and malignant conditions in gastrointestinal specimens.

The demonstration will offer attendees a look at how large-scale, pathology-specific AI models could support the future development of tools designed to assist pathologists with increasingly complex diagnostic workflows. Fujifilm will also present a scientific poster at Pathology Visions detailing the research and development behind Synapse Pathology AI Foundry. The presentation, “From Diagnostic to Prognostic Risk Scores in GI Cancers: Using AI in H&E WSIs for Clinical Insights” will take place on Friday, October 16 at 5:00 p.m. at display #9 during Session #1 at the reception.

Exploring the future of pathology AI at Pathology Visions

In addition to demonstrating Synapse Pathology v3.0, AI Orchestrator and Synapse Pathology AI Foundry, Fujifilm will welcome experts to Fujifilm’s booth to discuss emerging approaches to AI and data science in pathology:

  • On October 17 from 2:45–3:30 p.m., Michael Wan, Ph.D., Research Scientist at the Institute for Experiential AI, will present “Seeing the Forest and the Trees: Leveraging Hierarchical Vision AI for Whole Slide Image Analysis.”
  • Later that day, from 5:35–6:05 p.m., Saeed Amal, Ph.D., Visiting Professor at Harvard Medical School and Founder of the Amal Lab for Precision Medicine, will discuss “Developing Innovative Agentic AI for Digital Pathology.”
  • On October 18 from 11:30 a.m.–12:30 p.m., Gene Tunik, Ph.D., Professor and Senior Associate Dean of Research & Innovation at Bouvé College of Health Sciences, will present “Data Acquisition for Pathology LLMs.”

These sessions will explore emerging approaches to whole slide image analysis, agentic AI and the development of large language models for pathology, complementing Fujifilm’s demonstrations of its latest digital pathology technologies.

To learn more about Synapse Pathology, click here. To schedule a demo of Synapse Pathology at PathVisions 2026, click here.

Fujifilm’s pathology foundation, Synapse Pathology AI Foundry, is for research use only.*

About Fujifilm

FUJIFILM Healthcare Americas Corporation is a comprehensive healthcare company that has an extensive range of technology and expertise in the detection, diagnosis, and treatment of diseases. Fujifilm’s innovative portfolio includes solutions spanning diagnostic imaging, enterprise imaging, endoscopic imaging, surgical imaging, and in-vitro diagnostics. The Non-Destructive Testing group delivers radiography solutions to ensure high accuracy inspection of transportation infrastructure, and assets within aerospace, and oil and gas industries.

The company is headquartered in Lexington, Massachusetts. For more information on healthcare offerings, please visit healthcaresolutions-us.fujifilm.com, and for NDT portfolio, please visit https://www.fujifilm.com/us/en/business/industrial-materials/non-destructive-testing.

FUJIFILM Holdings Corporation, headquartered in Tokyo, leverages its depth of knowledge and proprietary core technologies to deliver innovative products and services across the globe through the four key business segments of healthcare, electronics, business innovation, and imaging with over 70,000 employees. Guided and united by our Group Purpose of “giving our world more smiles,” we address social challenges and create a positive impact on society through our products, services, and business operations. Under its medium-term management plan, VISION2030, which ends in FY2030, we aspire to continue our evolution into a company that creates value and smiles for various stakeholders as a collection of global leading businesses and achieve a global revenue of 4 trillion yen. For more information, please visit: https://holdings.fujifilm.com/en.

FUJIFILM-US-Healthcare

Contacts

Media Contact
Marissa Confredo

FUJIFILM Holdings America Corporation

914-343-4761

Marissa.confredo@fujifilm.com

Bristol Myers Squibb Announces Phase 3 ALOFT-IPF Trial Evaluating Admilparant in Adults with Idiopathic Pulmonary Fibrosis Met Primary Endpoint

Bristol Myers Squibb Announces Phase 3 ALOFT-IPF Trial Evaluating Admilparant in Adults with Idiopathic Pulmonary Fibrosis Met Primary Endpoint


Bristol Myers Squibb Announces Phase 3 ALOFT-IPF Trial Evaluating Admilparant in Adults with Idiopathic Pulmonary Fibrosis Met Primary Endpoint

In the trial, admilparant at the 60 mg dose demonstrated a statistically significant benefit in absolute change in forced vital capacity (FVC) from baseline to Week 52 compared with placebo

Admilparant demonstrated a favorable safety and tolerability profile

The totality of the findings supports the potential of admilparant as a clinically meaningful first-in-class treatment

PRINCETON, N.J.–(BUSINESS WIRE)–$BMY #ALOFT—Bristol Myers Squibb (NYSE: BMY) today announced topline results from the pivotal Phase 3 ALOFT-IPF trial evaluating the efficacy, safety and tolerability of admilparant, a potential first-in-class, orally administered, potent and selective lysophosphatidic acid receptor 1 (LPA1) antagonist, in adults with idiopathic pulmonary fibrosis (IPF). The trial evaluated two dose levels of admilparant, 60 mg and 120 mg, twice daily for ≥52 weeks against placebo (1:1:1).


The trial met its primary endpoint at the 60 mg dose, with admilparant demonstrating a statistically significant improvement compared to placebo in absolute change in forced vital capacity (FVC) from baseline at Week 52. At the 120 mg dose, admilparant slowed the decline of FVC but was not statistically significant.

Notably, admilparant demonstrated a well-tolerated safety profile across both doses in the ALOFT-IPF trial. In both admilparant arms of the trial, discontinuations due to adverse events were low and similar to placebo.

“Taken together, these topline data reinforce the profile of admilparant as a promising, clinically meaningful investigational therapy for IPF – one that reflects the innovation of targeting the LPA1 pathway as a first-in-class approach,” said Cristian Massacesi, MD, executive vice president, chief medical officer and head of development, Bristol Myers Squibb. “We also look forward to sharing the topline results from the ALOFT-PPF trial early next year, as we continue working to improve outcomes for patients with pulmonary fibrosis.”

“IPF remains one of the most complex and challenging lung diseases to treat, with unpredictable progression, irreversible scarring and a profound impact on patients’ daily lives,” said Fernando Martinez, MD, chair of the ALOFT program clinical steering committee and the Joseph D. Early Chair in Biomedical Research, interim chair and professor of medicine at UMass Chan Medical School. “These admilparant results represent an important advancement as there remains a need for therapies that are both effective and tolerable for patients living with IPF.”

The company will complete a comprehensive evaluation of the Phase 3 ALOFT-IPF trial data and engage with health authorities to support planned regulatory submissions, with detailed results to be presented at a future medical meeting.

Admilparant is also being studied in the fully enrolled Phase 3 ALOFT-PPF trial in progressive pulmonary fibrosis (PPF), with topline results anticipated in early 2027.

Bristol Myers Squibb thanks the patients and investigators who participated in the ALOFT-IPF clinical trial.

About Pulmonary Fibrosis

Pulmonary fibrosis is a chronic, life-threatening interstitial lung disease (ILD) that occurs when lung tissue becomes irreversibly damaged and scarred, impacting how the lungs function. Idiopathic pulmonary fibrosis (IPF) is the most common type of progressive fibrosing ILD. As an idiopathic disease, there is no identifiable cause, and approximately 1.4 million people live with IPF globally. Progressive pulmonary fibrosis (PPF) describes patients with non-idiopathic forms of ILD. These include connective tissue disease-related lung diseases, such as rheumatoid arthritis-associated ILD and scleroderma-associated ILD, and other types, such as environmental lung fibrosis, which is caused by exposure to dust or chemicals.

Many people with pulmonary fibrosis are physically impaired, experience a progressive decline in lung function, have difficulty performing simple daily activities due to breathlessness and chronic cough and require continuous supplemental oxygen to ease the burden of normal breathing.

Pulmonary fibrosis is a fatal disease with a median survival time of three to five years following diagnosis and a five-year survival rate of approximately 45%. Innovation in treatment has been limited, with very few new therapies approved in more than 10 years. Currently approved treatments can slow the decline of lung function; however, there remains a need for additional safe and tolerable therapies which allow patients to manage their disease while they continue to perform daily activities.

About the Phase 3 ALOFT Trial Program

ALOFT-IPF (IM027-068; NCT06003426) and ALOFT-PPF (IM027-1015; NCT06025578) are Phase 3, multicenter, randomized, double-blind, placebo-controlled trials. ALOFT-IPF enrolled approximately 1,255 adults (≥40 years) with idiopathic pulmonary fibrosis (IPF), while ALOFT-PPF enrolled approximately 1,057 adults (≥21 years) with progressive pulmonary fibrosis (PPF), with or without stable background antifibrotic therapy. The trials include both a 60 mg and 120 mg dose arm (twice daily for ≥52 weeks) against placebo (1:1:1), assessing the efficacy, safety and tolerability of admilparant.

The primary endpoint for the registrational portions of the trials is absolute change in forced vital capacity (FVC) from baseline to Week 52.

About Admilparant (BMS-986278)

Admilparant is a potential first-in-class, orally administered, potent, selective lysophosphatidic acid receptor 1 (LPA1) antagonist currently being evaluated as a novel antifibrogenic treatment for patients with idiopathic pulmonary fibrosis (IPF) and progressive pulmonary fibrosis (PPF). Increased LPA levels and activation of LPA1 are involved in the pathogenesis of pulmonary fibrosis.

In a global Phase 2 trial with parallel IPF and PPF cohorts, 26 weeks of treatment with admilparant 60 mg twice daily resulted in a 62% relative reduction in the rate of lung function decline versus placebo in patients with IPF and a 69% relative reduction in patients with PPF. Treatment effect was consistent with or without background therapy and admilparant was well tolerated in both cohorts, with rates of adverse events and treatment discontinuation comparable to placebo.

Bristol Myers Squibb’s research approach to IPF begins not at the point where fibrosis is visible, but at its origin, where epithelial cell injury and dysfunctional repair first emerge. Admilparant targets three key drivers of fibrogenesis: faulty epithelial cell repair, inflammation/vascular leakage and fibroblast activation and expansion.

Bristol Myers Squibb: Pioneering Paths Forward in Immunology to Transform Patients’ Lives

Bristol Myers Squibb is inspired by a single vision — transforming patients’ lives through science. Combining our patient-centric approach and deep understanding of the immune system that spans over 20 years of experience, we continue to pursue bold science to deliver life-changing medicines that elevate new standards of care across rheumatology, dermatology and pulmonology. Through our sequential immunotherapy research framework, which aims to address the root cause of disease by controlling inflammation, resetting the immune system and promoting immune homeostasis with the goal of achieving transformational efficacy, we strive to bring forward tailored approaches, treatments and combinations that may lead to durable remissions, improved quality of life and functional cures.

About Bristol Myers Squibb: Transforming Patients’ Lives Through Science

At Bristol Myers Squibb, our mission is to discover, develop and deliver innovative medicines that help patients prevail over serious diseases. We are pursuing bold science to define what’s possible for the future of medicine and the patients we serve. For more information, visit us at BMS.com and follow us on LinkedIn, X, YouTube, Facebook and Instagram.

Cautionary Statement Regarding Forward-Looking Statements

This press release contains “forward-looking statements” within the meaning of the Private Securities Litigation Reform Act of 1995 regarding, among other things, the research, development and commercialization of pharmaceutical products. All statements that are not statements of historical facts are, or may be deemed to be, forward-looking statements. Such forward-looking statements are based on current expectations and projections about our future financial results, goals, plans and objectives and involve inherent risks, assumptions and uncertainties, including internal or external factors that could delay, divert or change any of them in the next several years, that are difficult to predict, may be beyond our control and could cause our future financial results, goals, plans and objectives to differ materially from those expressed in, or implied by, the statements. These risks, assumptions, uncertainties and other factors include, among others, that future study results may not be consistent with the results to date and may be unfavorable, that admilparant may not receive regulatory approval for the indications described in this release, any marketing approvals, if granted, may have significant limitations on their use, and, if approved, whether admilparant for the indications described in this release will be commercially successful. No forward-looking statement can be guaranteed. Forward-looking statements in this press release should be evaluated together with the many risks and uncertainties that affect Bristol Myers Squibb’s business and market, particularly those identified in the cautionary statement and risk factors discussion in Bristol Myers Squibb’s Annual Report on Form 10-K for the year ended December 31, 2025, as updated by our subsequent Quarterly Reports on Form 10-Q, Current Reports on Form 8-K and other filings with the Securities and Exchange Commission. The forward-looking statements included in this document are made only as of the date of this document and except as otherwise required by applicable law, Bristol Myers Squibb undertakes no obligation to publicly update or revise any forward-looking statement, whether as a result of new information, future events, changed circumstances or otherwise.

corporatefinancial-news

Contacts

Bristol Myers Squibb

Media Inquiries:
media@bms.com

Investors:
investor.relations@bms.com

Bristol Myers Squibb Announces Positive Topline Results from the Interim Analysis of Phase 3 ACTION-1 for RYZ101, RayzeBio’s Radiopharmaceutical Therapy, in Patients with GEP-NETs

Bristol Myers Squibb Announces Positive Topline Results from the Interim Analysis of Phase 3 ACTION-1 for RYZ101, RayzeBio’s Radiopharmaceutical Therapy, in Patients with GEP-NETs


Bristol Myers Squibb Announces Positive Topline Results from the Interim Analysis of Phase 3 ACTION-1 for RYZ101, RayzeBio’s Radiopharmaceutical Therapy, in Patients with GEP-NETs

The study met the primary endpoint with a statistically significant and clinically meaningful improvement in progression free survival (PFS)

PRINCETON, N.J.–(BUSINESS WIRE)–$BMY #ACTION—Bristol Myers Squibb (NYSE: BMY) today announced positive topline results from the planned interim analysis of the ACTION-1 trial (NCT05477576) conducted by RayzeBio, Inc., a Bristol Myers Squibb company. The study met the primary endpoint with a statistically significant and clinically meaningful improvement in progression-free survival by blinded independent central review, and safety results consistent with the known RYZ101 safety profile. ACTION-1 is a randomized Phase 3 clinical trial evaluating RYZ101 (225Ac-DOTATATE) versus investigator’s choice of standard of care (high-dose or high-frequency long-acting somatostatin analogues, everolimus, or sunitinib) in patients with somatostatin receptor-positive (SSTR+) gastroenteropancreatic neuroendocrine tumors (GEP-NETs) who have progressed following Lutetium-177 (Lu177)-based therapy.


“The improvement in progression-free survival is a defining moment for RYZ101 and for the patients we built this program around — people with advanced GEP-NETs whose cancer has progressed after Lu177-based therapy and who have few remaining options,” said Amreen Husain, MD, chief medical officer of RayzeBio. “This is the kind of impact we set out to deliver with actinium-based radiopharmaceuticals, and it is an important milestone supporting the pipeline we are building across our RPT platform. These are the first results from a Phase 3 trial of an alpha-isotope based targeted RPT (225Ac) in any indication, and we are moving with urgency to bring these data to the FDA and, ultimately, provide a novel option for patients.”

The FDA granted RYZ101 Orphan Drug Designation for the treatment of GEP-NETs in August 2026 based on the significant unmet need facing patients with this disease. That designation, combined with the results of this interim analysis, supports RayzeBio’s continued development efforts and plans to engage with the FDA to discuss next steps toward a regulatory submission. Detailed results are expected to be presented at a future medical meeting.

“Lu177-based therapy is widely used in GEP-NETs, but unfortunately the vast majority of patients will eventually progress. Determining the optimal treatment after Lu177 has been an important question for my patients, and until now there has been no randomized Phase 3 evidence to guide it,” said Simron Singh, MD, medical oncologist in the Odette Cancer Centre, Sunnybrook Health Sciences Centre, Professor of Medicine at the University of Toronto, and principal investigator of the ACTION-1 study. “ACTION-1 provides the first Phase 3 results for an alpha-based targeted RPT, and a clinically meaningful improvement in progression-free survival is a crucial development in improving the care of patients with GEP-NETs.”

Individuals currently enrolled will continue to receive care consistent with the study protocol, and participants will be followed for overall survival.

About Gastroenteropancreatic Neuroendocrine Tumors (GEP-NETs)

GEP-NETs are rare tumors, with an estimated incidence of approximately 18,000 patients diagnosed annually in the United States and a prevalence of fewer than 200,000 patients. Somatostatin receptor type 2 (SSTR2) is expressed in 80–90% of GEP-NET tumors, and metastatic disease is present at diagnosis in the vast majority of cases. Lu177-based radiopharmaceutical therapy has shown significant clinical benefit and is approved for adult patients with SSTR-positive GEP-NETs, but most patients invariably progress following treatment, with few effective treatment options available.

About RYZ101

RYZ101 is an investigational, targeted radiopharmaceutical therapy comprised of DOTATATE bound to Actinium-225 (225Ac), a highly potent alpha-emitting radioisotope. 225Ac emits a high-energy, short-range alpha particle that can cause double-stranded DNA damage in cancer cells, potentially resulting in higher chance of cancer cell death. In a Phase 1b trial of 17 previously Lu177-treated patients with disease progression, RYZ101 was well tolerated, with no treatment-related serious adverse events or dose discontinuations due to any adverse event. RYZ101 is also being evaluated in earlier-stage trials in hormone receptor-positive (HR+) metastatic breast cancer.

About RayzeBio

RayzeBio, Inc., a Bristol Myers Squibb company, develops precision radiopharmaceuticals designed to open new pathways for patients living with cancer. Since its founding in 2020, RayzeBio has built a broad, highly differentiated pipeline supported by capabilities spanning discovery, development, isotope production, and manufacturing across its sites. The company’s lead program, RYZ101, is in Phase 3 development for gastroenteropancreatic neuroendocrine tumors (GEP-NETs), alongside programs targeting GPC3, ACP3, next-generation SSTR2-directed therapies, and a number of undisclosed targets. Together, these programs reflect RayzeBio’s ambition to advance the next generation of radiopharmaceutical therapy in service of unmet need for patients. Headquartered in San Diego, California, RayzeBio pairs a nimble, innovative culture with the scale and resources of Bristol Myers Squibb. For more information, visit rayzebio.com.

About Bristol Myers Squibb: Transforming Patients’ Lives Through Science

At Bristol Myers Squibb, our mission is to discover, develop and deliver innovative medicines that help patients prevail over serious diseases. We are pursuing bold science to define what’s possible for the future of medicine and the patients we serve. For more information, visit us at BMS.com and follow us on LinkedIn, X, YouTube, Facebook and Instagram.

Cautionary Statement Regarding Forward-Looking Statements

This press release contains “forward-looking statements” within the meaning of the Private Securities Litigation Reform Act of 1995 regarding, among other things, the research, development and commercialization of pharmaceutical products. All statements that are not statements of historical facts are, or may be deemed to be, forward-looking statements. Such forward-looking statements are based on current expectations and projections about our future financial results, goals, plans and objectives and involve inherent risks, assumptions and uncertainties, including internal or external factors that could delay, divert or change any of them in the next several years, that are difficult to predict, may be beyond our control and could cause our future financial results, goals, plans and objectives to differ materially from those expressed in, or implied by, the statements. These risks, assumptions, uncertainties and other factors include, among others, that future study results may not be consistent with the results to date, and that RYZ101 (225Ac-DOTATATE) may not receive regulatory approval for the indication described in this release in the currently anticipated timeline or at all, any marketing approvals, if granted, may have significant limitations on their use, and, if approved, whether RYZ101 for such indication will be commercially successful. No forward-looking statement can be guaranteed. Forward-looking statements in this press release should be evaluated together with the many risks and uncertainties that affect Bristol Myers Squibb’s business and market, particularly those identified in the cautionary statement and risk factors discussion in Bristol Myers Squibb’s Annual Report on Form 10-K for the year ended December 31, 2025, as updated by our subsequent Quarterly Reports on Form 10-Q, Current Reports on Form 8-K and other filings with the Securities and Exchange Commission. The forward-looking statements included in this document are made only as of the date of this document and except as otherwise required by applicable law, Bristol Myers Squibb undertakes no obligation to publicly update or revise any forward-looking statement, whether as a result of new information, future events, changed circumstances or otherwise.

corporatefinancial-news

Contacts

Bristol Myers Squibb

Media Inquiries:
media@bms.com

Investors:
investor.relations@bms.com

Bausch + Lomb and Bausch Health Announce Appointment of Thomas J. Appio as Interim Bausch + Lomb CEO, Expect to Raise Full-Year 2026 Guidance

Bausch + Lomb and Bausch Health Announce Appointment of Thomas J. Appio as Interim Bausch + Lomb CEO, Expect to Raise Full-Year 2026 Guidance


Bausch + Lomb and Bausch Health Announce Appointment of Thomas J. Appio as Interim Bausch + Lomb CEO, Expect to Raise Full-Year 2026 Guidance

VAUGHAN, Ontario–(BUSINESS WIRE)–Bausch + Lomb Corporation (NYSE/TSX: BLCO), a leading global eye health company, and its parent company, Bausch Health Companies Inc. (NYSE/TSX: BHC), today announced that Bausch + Lomb Board of Directors Chairman Thomas J. Appio has been appointed interim chief executive officer, effective immediately. Appio will continue to serve as CEO of Bausch Health while assuming leadership of Bausch + Lomb.


Appio was appointed chairman of the Bausch + Lomb Board of Directors in August 2026. The Board believes Appio’s deep knowledge of the business, global leadership experience and longstanding connection to Bausch + Lomb position him well to build on the company’s momentum and lead its next phase of growth.

“The opportunity to lead Bausch + Lomb is both a privilege and a responsibility that I embrace with great enthusiasm,” said Appio. “Our company is built on the strength of our people, the quality of our portfolio and a relentless commitment to serving customers and patients around the world. With a strong foundation, we are well positioned to build on our momentum and deliver sustainable growth.”

Appio has served as CEO of Bausch Health since 2022. Under his leadership, Bausch Health (excluding Bausch + Lomb) has delivered 13 consecutive quarters of year-over-year growth in revenue and adjusted EBITDA (non-GAAP).1 He held several leadership roles within Bausch + Lomb prior to the company’s initial public offering in 2022. Appio joined Bausch + Lomb in 2010 as managing director, China, and in 2016 assumed the role of president, Bausch + Lomb International.

Brent Saunders, who has served as Bausch + Lomb CEO since 2023, and the Board of Directors mutually agreed that Saunders will depart the company.

“When I returned to Bausch + Lomb, I believed this company had tremendous potential,” said Saunders. “Today, it is stronger, more focused and better positioned for the future, with a clearer strategy, a more robust pipeline and significant opportunities ahead. The progress we’ve made reflects the talent, passion and commitment of colleagues across the organization, and I’m proud of everything we’ve accomplished together.”

Preliminary Update on 2026 Guidance2

Based on preliminary third-quarter results, Bausch + Lomb and Bausch Health currently expect to raise full-year 2026 adjusted EBITDA excluding Acquired IPR&D1 guidance.3 The companies are currently in a blackout period and will provide additional details when they report third-quarter results on Oct. 28.

1

This is a non-GAAP measure or a non-GAAP ratio. For further information on non-GAAP measures and non-GAAP ratios, please refer to the “Non-GAAP Information” section of this news release.

2

The preliminary update on guidance in this news release speaks only as of the date given, October 12, 2026, and will not be further updated or affirmed unless and until each company publicly announces updated or affirmed guidance. Distribution or reference of this news release following October 12, 2026, does not constitute the company reaffirming guidance. There is no assurance that Bausch + Lomb or Bausch Health will achieve its forecasted results within the relevant period or otherwise. Actual results may differ materially from these estimates as a result of actual quarter-end results, the completion of normal quarter-end accounting procedures and adjustments, including the execution of our internal control over financial reporting, the completion of the preparation and management’s review of our financial statements for the relevant period and the subsequent occurrence or identification of events prior to the release of our financial results for the relevant period. See the “Forward-looking Statements” section for further information.

3

The increase in anticipated full-year adjusted EBITDA (excluding Acquired IPR&D) is a result of continued strong business performance in the third quarter of 2026 and anticipated stronger business performance for the remainder of 2026.

About Bausch + Lomb

Our mission is simple – we help people see better to live better, all over the world. For nearly two centuries we’ve evolved with the changing needs of patients and customers, and our commitment to innovation and improving the standard of care in eye health has never been stronger. From contact lenses to prescription products, over-the-counter options, surgical devices and more, we’re turning bold ideas into better outcomes through passion, perseverance and purpose. Learn more at www.bausch.com and connect with us on Facebook, Instagram, LinkedIn, X and YouTube.

About Bausch Health

Bausch Health Companies Inc. (NYSE:BHC)(TSX:BHC) is a global, diversified pharmaceutical company enriching lives through our relentless drive to deliver better health care outcomes. We develop, manufacture and market a range of products primarily in gastroenterology, hepatology, neuroscience, dermatology, dentistry, aesthetics, international pharmaceuticals and eye health, through our controlling interest in Bausch + Lomb Corporation. Our ambition is to be a globally integrated healthcare company, trusted and valued by patients, HCPs, employees and investors. For more information about Bausch Health, visit www.bauschhealth.com and connect with us on LinkedIn.

Forward-looking Statements

This news release may contain forward-looking information and statements within the meaning of applicable securities laws (collectively, “forward-looking statements”). Forward-looking statements may generally be identified by the use of the words “anticipates,” “hopes,” “seeks,” “intends,” “expects,” “believes,” “plans,” “projects,” “predicts,” “forecasts,” “should,” “could,” “would,” “may,” “might,” “will,” “strive,” “estimates,” “potential,” “target,” “guidance,” “outlook,” “continue,” “pending” or “proposed” and positive and negative variations or similar expressions and phrases and statements that certain actions, events or results may, could, should or will be achieved, received or taken, or will occur or result. Forward-looking statements include statements regarding the companies’ full-year 2026 guidance (including expected increases to such guidance) and Bausch + Lomb’s ability to deliver sustainable growth and create lasting value for shareholders. These statements are based upon the current expectations and beliefs of management and are provided for the purpose of providing additional information about such expectations and beliefs, and readers are cautioned that these statements may not be appropriate for other purposes. The expected increase to full-year 2026 guidance is based on certain material factors and assumptions, including preliminary, unaudited third-quarter 2026 results that remain subject to completion of quarter-end closing and review procedures, stronger business performance for the remainder of 2026, prevailing foreign exchange rates and other assumptions. Forward-looking statements are subject to certain risks and uncertainties that could cause actual results to differ materially from those described in the forward-looking statements. These risks and uncertainties include, but are not limited to, the risks and uncertainties discussed in Bausch + Lomb’s and Bausch Health’s filings with the U.S. Securities and Exchange Commission (the “SEC”) and the Canadian Securities Administrators (the “CSA”), which factors are incorporated herein by reference. Readers are cautioned not to place undue reliance on any of these forward-looking statements. These forward-looking statements speak only as of the date hereof. Neither Bausch + Lomb nor Bausch Health undertake any obligation to update any of these forward-looking statements to reflect events or circumstances after the date of this news release or to reflect actual outcomes, unless required by law.

Non-GAAP Information

To supplement the financial measures prepared in accordance with U.S. generally accepted accounting principles (GAAP), Bausch + Lomb and Bausch Health use certain non-GAAP financial measures and ratios. Management of each company use these non-GAAP measures and ratios as key metrics in the evaluation of each company’s performance and the consolidated financial results and, in part, in the determination of cash bonuses for its executive officers. Bausch + Lomb and Bausch Health believe these non-GAAP measures and ratios are useful to investors in their assessment of each company’s operating performance and the valuation of each company. In addition, these non-GAAP measures and ratios address questions the companies routinely receives from analysts and investors, and in order to assure that all investors have access to similar data, the companies have determined that it is appropriate to make this data available to all investors. These measures and ratios do not have any standardized meaning under GAAP and other companies may use similarly titled non-GAAP financial measures and ratios that are calculated differently from the way we calculate such measures and ratios. Accordingly, our non-GAAP financial measures and ratios may not be comparable to similar non-GAAP measures and ratios of other companies. We caution investors not to place undue reliance on such non-GAAP measures and ratios, but instead to consider them with the most directly comparable GAAP measures and ratios. Non-GAAP financial measures and ratios have limitations as analytical tools and should not be considered in isolation. They should be considered as a supplement to, not a substitute for, or superior to, the corresponding measures calculated in accordance with GAAP.

Bausch + Lomb does not provide a reconciliation of forward-looking Adjusted EBITDA excluding Acquired IPR&D (non-GAAP)2 to GAAP net income (loss) attributable to Bausch + Lomb Corporation, and Bausch Health does not provide a reconciliation of forward-looking Adjusted EBITDA (non-GAAP)2 to GAAP net income (loss), in each case due to the inherent difficulty in forecasting and quantifying certain amounts that are necessary for such reconciliations. These amounts may be material and, therefore, could result in the projected GAAP measure or ratio being materially different or less than the projected non-GAAP measure or ratio.

Specific Non-GAAP Measures

Adjusted EBITDA excluding Acquired IPR&D

EBITDA (non-GAAP) is Net income (loss) attributable to Bausch + Lomb Corporation (its most directly comparable U.S. GAAP financial measure) adjusted for interest, income taxes, depreciation and amortization. Adjusted EBITDA (non-GAAP) is EBITDA (non-GAAP) further adjusted for the items described below. Management believes that Adjusted EBITDA (non-GAAP), along with the GAAP measures used by management, most appropriately reflect how the company measures the business internally and sets operational goals and incentives. In particular, the company believes that Adjusted EBITDA (non-GAAP) focuses management on the company’s underlying operational results and business performance. As a result, the company uses Adjusted EBITDA (non-GAAP) both to assess the actual financial performance of the company and to forecast future results as part of its guidance. Management believes Adjusted EBITDA (non-GAAP) is a useful measure to evaluate current performance. Adjusted EBITDA (non-GAAP) is intended to show our unleveraged, pre-tax operating results and therefore reflects our financial performance based on operational factors. In addition, cash bonuses for the company’s executive officers and other key employees are based, in part, on the achievement of certain Adjusted EBITDA (non-GAAP) targets.

Adjusted EBITDA (non-GAAP) is Net income (loss) attributable to Bausch + Lomb Corporation (its most directly comparable U.S. GAAP financial measure) adjusted for interest expense, net, (benefit from) provision for income taxes, depreciation and amortization and further adjusted for the following items:

  • Asset impairments: The company has excluded the impact of impairments of finite-lived and indefinite-lived intangible assets as such amounts are inconsistent in amount and frequency and are significantly impacted by the timing and/or size of acquisitions and divestitures. The company believes that the adjustments of these items correlate with the sustainability of the company’s operating performance. Although the company excludes impairments of intangible assets from measuring the performance of the company and its business, the company believes that it is important for investors to understand that intangible assets contribute to revenue generation.
  • Restructuring, integration and transformation costs: The company has incurred restructuring costs as it implemented certain strategies, which involved, among other things, improvements to its infrastructure and operations, internal reorganizations and impacts from the divestiture of assets and businesses. With regard to infrastructure and operational improvements which the company has taken to improve efficiencies in the businesses and facilities, these tend to be costs intended to right size the business or organization that fluctuate significantly between periods in amount, size and timing, depending on the improvement project, reorganization or transaction. Additionally, with the completion of the Bausch + Lomb IPO, as the company prepares for post-separation operations, the company is launching certain transformation initiatives that will result in certain changes to and investment in its organizational structure and operations. These transformation initiatives arise outside of the ordinary course of continuing operations and, as is the case with the company’s restructuring efforts, costs associated with these transformation initiatives are expected to fluctuate between periods in amount, size and timing. These out-of-the-ordinary-course charges include third-party advisory costs, as well as certain compensation-related costs. Investors should understand that the outcome of these transformation initiatives may result in future restructuring actions and certain of these charges could recur. The company believes that the adjustments of these items provide supplemental information with regard to the sustainability of the company’s operating performance, allow for a comparison of the financial results to historical operations and forward-looking guidance and, as a result, provide useful supplemental information to investors.
  • Acquisition-related costs and adjustments excluding amortization of intangible assets: The company has excluded the impact of acquisition-related costs and fair value inventory step-up resulting from acquisitions as the amounts and frequency of such costs and adjustments are not consistent and are significantly impacted by the timing and size of its acquisitions. In addition, the company excludes the impact of acquisition-related contingent consideration non-cash adjustments due to the inherent uncertainty and volatility associated with such amounts based on changes in assumptions with respect to fair value estimates, and the amount and frequency of such adjustments are not consistent and are significantly impacted by the timing and size of the company’s acquisitions, as well as the nature of the agreed-upon consideration.
  • Share-based compensation: The company excludes costs relating to share-based compensation. The company believes that the exclusion of share-based compensation expense assists investors in the comparisons of operating results to peer companies. Share-based compensation expense can vary significantly based on the timing, size and nature of awards granted.
  • Separation costs and separation-related costs: The company has excluded certain costs incurred in connection with activities taken to: (i) separate the Bausch + Lomb business from the remainder of BHC and (ii) register the Bausch + Lomb business as an independent publicly traded entity. Separation costs are incremental costs directly related to effectuating the separation of the Bausch + Lomb business from the remainder of BHC and include, but are not limited to, legal, audit and advisory fees, talent acquisition costs and costs associated with establishing a new Board of Directors and Audit Committee. Separation-related costs are incremental costs indirectly related to the separation of the Bausch + Lomb business from the remainder of BHC and include, but are not limited to, IT infrastructure and software licensing costs, rebranding costs and costs associated with facility relocation and/or modification. As these costs arise from events outside of the ordinary course of continuing operations, the company believes that the adjustments of these items provide supplemental information with regard to the sustainability of the company’s operating performance, allow for a comparison of the financial results to historical operations and forward-looking guidance and, as a result, provide useful supplemental information to investors.
  • Loss on extinguishment of debt: The company has excluded loss on extinguishment of debt as this represents a loss from refinancing our existing debt and is not a reflection of our operations for the period. Further, the amount and frequency of such amounts are not consistent and are significantly impacted by the timing and size of debt financing transactions and other factors in the debt market that are not within management’s control.
  • Other Non-GAAP adjustments: The company also excludes certain other amounts, including IT infrastructure investment, litigation and other matters, gain/(loss) on sales of assets and certain other amounts that are the result of other, non-comparable events to measure operating performance if and when present in the periods presented. These events arise outside of the ordinary course of continuing operations. Given the unique nature of the matters relating to these costs, the company believes these items are not routine operating expenses. For example, legal settlements and judgments vary significantly, in their nature, size and frequency, and, due to this volatility, the company believes the costs associated with legal settlements and judgments are not routine operating expenses. The company excluded these costs as this event is outside of the ordinary course of continuing operations and is infrequent in nature. The company believes that the exclusion of such out-of-the-ordinary-course amounts provides supplemental information to assist in the comparison of the financial results of the company from period to period and, therefore, provides useful supplemental information to investors. However, investors should understand that many of these costs could recur and that companies in our industry often face litigation.

Adjusted EBITDA excluding Acquired In-Process Research and Development (IPR&D) (non-GAAP) is Adjusted EBITDA (non-GAAP) further adjusted to exclude Acquired IPR&D. The IPR&D expenditures represent costs directly resulting from business development transactions and not through the normal course of business. The company believes that the exclusion of such out-of-the-ordinary-course amounts provides supplemental information to assist in the comparison of the financial results of the company from period to period and, therefore, provides useful supplemental information to investors in assessing our performance. However, investors should understand that the company may enter into additional business development transactions in the future and, as a result, such Acquired IPR&D may recur in the future.

Adjusted EBITDA Margin (non-GAAP) and Adjusted EBITDA Margin excluding Bausch + Lomb (non-GAAP)

Adjusted EBITDA Margin (non-GAAP) is defined as Adjusted EBITDA (non-GAAP) (as previously defined) divided by Revenue. The most directly comparable U.S. GAAP financial measure is Net income (loss) divided by Revenue. Adjusted EBITDA Margin excluding Bausch + Lomb (non-GAAP) is defined as Adjusted EBITDA excluding Bausch + Lomb (non-GAAP) divided by Revenue excluding revenue attributable to Bausch + Lomb. Adjusted EBITDA excluding Bausch + Lomb (non-GAAP) is Adjusted EBITDA (non-GAAP) adjusted to remove Adjusted EBITDA attributable to Bausch + Lomb (non-GAAP). Adjusted EBITDA Margin excluding Bausch + Lomb (non-GAAP) is not intended to represent continuing or discontinued operations under GAAP and is provided as a supplemental management metric. This measure may not be indicative of the results of operations that would have been reported had Bausch + Lomb been accounted for as a discontinued operation. Management believes Adjusted EBITDA Margin (non-GAAP) and Adjusted EBITDA Margin excluding Bausch + Lomb (non-GAAP) provide investors with useful supplemental information regarding the Company’s operating profitability and operational efficiency.

©2026 Bausch + Lomb.

Contacts

Bausch + Lomb Media Contact:

T.J. Crawford

tj.crawford@bausch.com
(908) 705-2851

Bausch Health Media Contact:
Katie Savastano

corporate.communications@bauschhealth.com
(908) 569-3692

Bausch + Lomb Investor Contact:
George Gadkowski

george.gadkowski@bausch.com
(877) 354-3705 (toll free)

(908) 927-0735

Bausch Health Investor Contact:
Garen Sarafian

ir@bauschhealth.com
(877) 281-6642 (toll free)


Final MINT Study Results Strengthen Real-World Evidence for Mogamulizumab (Poteligeo®) in Patients With Mycosis Fungoides and Sézary Syndrome

Final MINT Study Results Strengthen Real-World Evidence for Mogamulizumab (Poteligeo®) in Patients With Mycosis Fungoides and Sézary Syndrome


Final MINT Study Results Strengthen Real-World Evidence for Mogamulizumab (Poteligeo®) in Patients With Mycosis Fungoides and Sézary Syndrome

  • Final analysis of 109 patients from 18 German centres demonstrates durable treatment outcomes in routine clinical practice1
  • Median time-to-next-treatment (TTNT), the study’s primary endpoint, was 14.9 months, alongside a 62.4% best overall response rate, 20.2-month overall median progression-free survival and improved quality of life from baseline reported in routine care1
  • Exploratory analyses show meaningful responses in clinically relevant subgroups, including large cell transformation, folliculotropic mycosis fungoides and mogamulizumab-associated rash status, supporting individualised treatment for patients with mycosis fungoides and Sézary syndrome1

GALASHIELS, Scotland & MARLOW, England–(BUSINESS WIRE)–Kyowa Kirin International plc, a wholly owned subsidiary of Kyowa Kirin Co., Ltd. (TSE:4151, Kyowa Kirin), today announced final results from the German MINT study evaluating mogamulizumab in patients with mycosis fungoides (MF) and Sézary Syndrome (SS). Presented at the European Organisation for Research and Treatment of Cancer Cutaneous Lymphoma Tumour Group (EORTC-CLTG) Annual Meeting 2026 in Berlin, Germany, the findings represent one of the largest real-world evaluations of mogamulizumab in Germany and add to the growing body of evidence supporting its use in routine clinical practice.1-3


Conducted as a company-sponsored, multicentric, non-interventional, combined retro-prospective real-world study across 18 centres throughout Germany, MINT brought together cutaneous T‑cell lymphoma (CTCL) specialists to evaluate treatment outcomes with mogamulizumab in a real-world setting, with time-to-next-treatment (TTNT) as the study’s primary endpoint.1 The study captured a broad and heterogeneous patient population, including clinically relevant subgroups such as patients with large cell transformation (LCT) and folliculotropic mycosis fungoides (fMF), and stratified by disease stage and mogamulizumab-associated rash (MAR) status, providing insights that complement evidence generated through controlled clinical studies.1,4

MF and SS are forms of CTCL, a type of non-Hodgkin’s lymphoma that manifests on the skin,5 though the disease can also involve the blood, lymph nodes and visceral organs.6 Symptoms can include persistent, itchy and scaly rashes, changes in skin colour and patches, plaques and/or tumours.7 These visible and often difficult-to-hide symptoms can make everyday activities harder and significantly affect patients’ quality of life, including their work, relationships and social lives,8 and for some patients the disease can progress to affect long-term survival.9

“For patients living with mycosis fungoides and Sézary syndrome, understanding how treatments perform outside the controlled setting of clinical trials is critically important. The final MINT results demonstrate meaningful clinical outcomes across a broad patient population treated in routine practice and highlight the value of collaborative real-world research in helping clinicians make informed treatment decisions,” said Professor Chalid Assaf, Director of the Department of Dermatology and Venereology, Helios Klinikum Krefeld, Health and Medical University Düsseldorf-Krefeld, Germany.

The final analysis included 109 patients, comprising 56 patients with MF and 53 patients with SS, with a median follow-up of 25.2 months.1 The majority of patients had advanced disease when they started treatment, with 83.5% at stage IIB or above.1 The study demonstrated durable treatment outcomes, reporting a median TTNT of 14.9 months, a best overall response rate of 62.4%, and an overall median progression-free survival of 20.2 months.1 Median overall survival was not estimable at the time of analysis due to the limited number of deaths observed, despite up to 71 months of follow-up in some cases.1

Looking more closely at treatment response in clinically relevant patient subgroups, exploratory analyses reported best overall response rates of 54.5% in patients with LCT (n=11; 95% CI: 23.4–83.3) and 50.0% in patients with fMF (n=12; 95% CI: 21.1–78.9). Additional exploratory analyses by disease stage showed best overall response rates of 50% in patients with early-stage disease (n=18; 95% CI: 26.0–74.0) and 64.8% in patients with advanced stage disease (n=91; 95% CI: 54.1–74.6).10 Disease control rate was also reported in patients with MAR (n=47), at 95.7%.4

TTNT is an important measure in real-world clinical practice as it reflects both disease control and the duration of clinical benefit before a patient requires an alternative treatment approach. The study reported a median TTNT of 12.0 months in patients with MF and 20.2 months in patients with SS.4 Patients also experienced improvements in quality of life, with median Skindex-29 scores improving by 38.5% from 49.1 at treatment initiation (n=21) to 30.2 at Month 12 (n=7).1

“MINT provides valuable real-world evidence from Germany on the use of mogamulizumab across the heterogeneous population of patients with mycosis fungoides and Sézary syndrome, including clinically relevant subgroups that are often underrepresented in clinical trials. By evaluating time-to-next-treatment as a clinically meaningful outcome, the study offers practical insights that can support CTCL specialists in making informed, individualised treatment decisions,” said Dr. Azeem Danish, Medical Affairs Director, DACH, Kyowa Kirin.

The investigators concluded that, in a real-world setting, mogamulizumab demonstrated a safety profile comparable to that observed in the pivotal Phase 3 MAVORIC study, while providing clinical benefits in treatment response and quality of life for patients with MF and SS.1,5 Grade 3 or higher treatment-related adverse events were reported in 38.5% of patients, most commonly drug eruption and lymphopenia.1 Serious treatment-related adverse events leading to discontinuation occurred in 22.0% of patients.1

Kyowa Kirin has supported the generation of real-world evidence in CTCL through collaborations with investigators and treatment centres across Europe. The MINT study contributes to a growing evidence base exploring outcomes in patients treated with mogamulizumab in routine clinical practice1-3 and reflects a shared commitment between investigators and industry to advancing understanding of rare cancers and improving care for patients living with CTCL.

About the MINT Study

MINT (Mogamulizumab In a Non-interventional study to evaluate the safety and outcome in mycosis fungoides and Sézary syndrome Treatment) is a German multicentric, non-interventional, retro-prospective real-world study evaluating the effectiveness and tolerability of mogamulizumab in patients with mycosis fungoides (MF) and Sézary Syndrome (SS). Data were collected from patient medical records across 18 study sites in Germany. The primary endpoint was time-to-next-treatment, with secondary endpoints including response rate, progression-free survival, overall survival, quality of life and tolerability.1

About Kyowa Kirin

Kyowa Kirin aims to discover novel medicines with life-changing value. As a global specialty pharmaceutical company, we have invested in drug discovery and biotechnology innovation for more than 75 years and are currently working to engineer the next generation of antibodies and cell and gene therapies with the potential to help patients affected by severe and rare diseases. A shared commitment to our values, to sustainable growth, and to making people smile unites us across our three regions: JAPAC, North America, and EMEA.

You can learn more about the business of Kyowa Kirin at: https://kyowakirininternational.com/.

About mycosis fungoides (MF) and Sézary Syndrome (SS)

MF and SS are two subtypes of cutaneous T‑cell lymphoma (CTCL), which is itself a rare form of non-Hodgkin’s lymphoma that presents and persists in the skin.5 CTCL is treatable, but is not generally considered to be curable, and there has been a clear unmet need for novel treatment options. As well as the obvious impact of symptoms upon patients, there can be significant erosions to quality of life for those caring for an individual living with CTCL.11

MF and SS are characterised by localisation of cancerous white blood cells called T lymphocytes (T cells), to the skin.9 These cancerous T cells consistently express a protein called CCR4, which enables them to move from the blood to the skin.12-14 When these cancerous T cells move to the skin, this results in the visible early skin symptoms of red patches or plaques which can resemble psoriasis or eczema in the early stages of the disease.7 Later, for some patients, skin involvement may evolve to include tumours or reddening of the majority of the skin’s surface (erythroderma).7

MF—the most common CTCL subtype—accounts for approximately 60% of all CTCLs15 and is typically indolent, characterised by skin symptoms including patches or plaques, skin redness and tumours. SS is much rarer, accounting for around 5% of CTCLs,6 and is more aggressive,7 with high levels of blood involvement.6 It can cause severe itching, erythroderma, intense scaling of the skin and hair loss.7 CTCL can take, on average, between 2 and 7 years for individuals to receive a confirmed diagnosis.16

About POTELIGEO® (mogamulizumab)

Mogamulizumab is a first-in-class humanised monoclonal antibody (mAb) directed against CC-chemokine receptor 4 (CCR4), a protein consistently expressed on cancerous cells seen in both mycosis fungoides and Sézary syndrome.12-14 Once mogamulizumab binds to CCR4, it increases attraction of immune cells from the immune system to destroy the cancerous cells.17

About POTELIGEO® (mogamulizumab) in Plain Language

Mogamulizumab is a monoclonal antibody designed to recognise and attach to CCR4, a protein commonly found on certain cancerous cells in mycosis fungoides and Sézary syndrome. By targeting these cells, it helps the body’s immune system identify and destroy them.

REFERENCES

1 Assaf C DG, Schlaak M, Booken N, Oymanns M, Klespe K-C, et al. Final results from the German “Mogamulizumab In a Non-interventional study to evaluate the safety and outcome in mycosis fungoides and Sézary syndrome Treatment” (MINT) study [Abstract]. Presented at: European Organisation for Research and Treatment of Cancer – Cutaneous Lymphoma Tumour Group (EORTC-CLTG) Annual Meeting; 2026 Oct 9–10; Berlin, Germany.

2 Broccoli A, Argnani L, Pileri A, Mazzoni C, Sordi B, Pimpinelli N, et al. Quality and duration of responses with mogamulizumab in cutaneous T-cell lymphomas: Insights into long-lasting outcomes. Br J Haematol. 2025;207(6):2578-82.

3 Bozonnat A, Beylot-Barry M, Dereure O, D’Incan M, Quereux G, Guenova E, et al. Real-life efficacy of immunotherapy for Sézary syndrome: a multicenter observational cohort study. eClinicalMedicine. 2024;73(102679).

4 Assaf C DG, Schlaak M, Booken N, Oymanns M, Klespe K-C, et al. Final results from the German “Mogamulizumab In a Non-interventional study to evaluate the safety and outcome in mycosis fungoides and Sézary syndrome Treatment” (MINT) study [Oral presentation]. Presented at: European Organisation for Research and Treatment of Cancer – Cutaneous Lymphoma Tumour Group (EORTC-CLTG) Annual Meeting; 2026 Oct 9–10; Berlin, Germany.

5 Kim YH, Bagot M, Pinter-Brown L, Rook AH, Porcu P, Horwitz SM, et al. Mogamulizumab versus vorinostat in previously treated cutaneous T-cell lymphoma (MAVORIC): an international, open-label, randomised, controlled phase 3 trial. Lancet Oncol. 2018;19(9):1192-204.

6 Trautinger F, Eder J, Assaf C, Bagot M, Cozzio A, Dummer R, et al. European Organisation for Research and Treatment of Cancer consensus recommendations for the treatment of mycosis fungoides/Sézary syndrome – Update 2017. Eur J Cancer. 2017;77:57-74.

7 Lymphoma Coalition. Cutaneous lymphoma – a patient’s guide. Available at: https://lymphomacoalition.org/wp-content/uploads/Cutaneous_lymphoma_-_patients_guide_-.pdf Last accessed: October 2026.

8 Martín-Carrasco P, Morillo-Andújar M, Sendín-Martín M, Conejo-Mir J. Quality of Life in Patients with Cutaneous T-Cell Lymphoma. Actas Dermo-Sifiliográficas. 2019;110(6):511-3.

9 Scarisbrick JJ, Prince HM, Vermeer MH, Quaglino P, Horwitz S, Porcu P, et al. Cutaneous Lymphoma International Consortium Study of Outcome in Advanced Stages of Mycosis Fungoides and Sézary Syndrome: Effect of Specific Prognostic Markers on Survival and Development of a Prognostic Model. J Clin Oncol. 2015;33(32):3766-73.

10 Kyowa Kirin. Data on file. 2026.

11 Williams K, Gibson A, McNamara L, Jones T, Lloyd A. Health state utilities associated with caring for an individual with cutaneous T-cell lymphoma (CTCL). J Med Econ. 2020;23:1142-50.

12 Ferenczi K, Fuhlbrigge RC, Pinkus JL, Pinkus GS, Kupper TS. Increased CCR4 Expression in Cutaneous T Cell Lymphoma. Journal of Investigative Dermatology. 2002;119(6):1405-10.

13 Yoshie O, Fujisawa R, Nakayama T, Harasawa H, Tago H, Izawa D, et al. Frequent expression of CCR4 in Adult T-cell leukemia and human T-cell leukemia virus type 1-transformed T cells. Blood. 2002;99:1505-11.

14 Ishida T, Utsunomiya A, Iida S, Inagaki H, Takatsuka Y, Kusumoto S, et al. Clinical significance of CCR4 expression in adult T-cell leukemia/lymphoma: its close association with skin involvement and unfavorable outcome. Clin Cancer Res. 2003;9(10 Pt 1):3625-34.

15 Willemze R, Cerroni L, Kempf W, Berti E, Facchetti F, Swerdlow SH, et al. The 2018 update of the WHO-EORTC classification for primary cutaneous lymphomas. Blood. 2019;133(16):1703-14.

16 Cutaneous Lymphoma Foundation. A patient’s guide to understanding cutaneous lymphoma [Internet]. 2nd ed. Cutaneous Lymphoma Foundation; 2018. Why it takes time to get a diagnosis; p. 21. Available from: https://flipbook.clfoundation.org/A_Patients_Guide/files/assets/basic-html/page29.html Last accessed: October 2026.

17 Duvic M, Evans M, Wang C. Mogamulizumab for the treatment of cutaneous T-cell lymphoma: recent advances and clinical potential. Ther Adv Hematol. 2016;7(3):171-4.

Contacts

James Read

james.read@kyowakirin.com
+44 (0)7917 922566

Or email: comms@kyowakirin.com

Mirum Pharmaceuticals to Host Investor Call to Share Topline Results from the Phase 3 EXPAND Study of LIVMARLI® (maralixibat) in Ultra-Rare Cholestatic Liver Diseases

Mirum Pharmaceuticals to Host Investor Call to Share Topline Results from the Phase 3 EXPAND Study of LIVMARLI® (maralixibat) in Ultra-Rare Cholestatic Liver Diseases


Mirum Pharmaceuticals to Host Investor Call to Share Topline Results from the Phase 3 EXPAND Study of LIVMARLI® (maralixibat) in Ultra-Rare Cholestatic Liver Diseases

FOSTER CITY, Calif.–(BUSINESS WIRE)–Mirum Pharmaceuticals, Inc. (Nasdaq: MIRM), a leading rare disease company, today announced that it will host an investor call on Monday, October 12, 2026 at 8:30 a.m. ET/5:30 a.m. PT to share topline results from the Phase 3 EXPAND study evaluating LIVMARLI® (maralixibat) for the treatment of cholestatic pruritus in patients aged 6 months or older with ultra-rare cholestatic liver diseases, including biliary atresia.

Conference Call Details:


US/Toll-Free: +1 833 461 5787
International: +1 585 542 9983
Access Code: 486697901

You may also access the call via webcast by visiting the Investors section of Mirum’s corporate website. The archived webcast will be available for replay.

About Mirum Pharmaceuticals

Mirum Pharmaceuticals (NASDAQ: MIRM) is a leading rare disease company with a global footprint of approved products and a broad pipeline of investigational medicines. Purpose-built to bring forward breakthrough medicines for people with overlooked conditions, Mirum focuses on rare liver and rare genetic diseases, where it has built deep expertise and strong connections to patient communities. The company’s commercial portfolio includes LIVMARLI® (maralixibat) for Alagille syndrome (ALGS) and progressive familial intrahepatic cholestasis (PFIC), ATEBRIOZ™ (zilurgisertib) for fibrodysplasia ossificans progressiva (FOP), CHOLBAM® (cholic acid) for bile-acid synthesis disorders and CTEXLI® (chenodiol) for cerebrotendinous xanthomatosis (CTX).

Mirum’s clinical-stage pipeline includes volixibat, an IBAT inhibitor in late-stage development for primary sclerosing cholangitis (PSC) and primary biliary cholangitis (PBC), brelovitug, a fully human monoclonal antibody in late-stage development for chronic hepatitis delta virus (HDV) and MRM-3379, a PDE4D inhibitor being evaluated for Fragile X syndrome (FXS).

Mirum’s success is driven by a team dedicated to advancing high impact medicines through strategic development, disciplined execution and purposeful collaboration across the rare disease ecosystem. Learn more at www.mirumpharma.com and follow Mirum on Facebook, LinkedIn, Instagram and X.

Contacts

Investor Contact:
Andrew McKibben
ir@mirumpharma.com

Media Contact:
Meredith Kiernan
media@mirumpharma.com

Merck Presents One-Year Results from the Pivotal Phase 2b/3 BRUNELLO Study Evaluating Remigromig, a Tri-specific Agonist of the Wingless-related Integration Site (Wnt) Pathway, in Adults with Diabetic Macular Edema at AAO 2026

Merck Presents One-Year Results from the Pivotal Phase 2b/3 BRUNELLO Study Evaluating Remigromig, a Tri-specific Agonist of the Wingless-related Integration Site (Wnt) Pathway, in Adults with Diabetic Macular Edema at AAO 2026


Merck Presents One-Year Results from the Pivotal Phase 2b/3 BRUNELLO Study Evaluating Remigromig, a Tri-specific Agonist of the Wingless-related Integration Site (Wnt) Pathway, in Adults with Diabetic Macular Edema at AAO 2026

Remigromig demonstrated non-inferiority in mean change from baseline in best corrected visual acuity versus ranibizumab at one year in patients with diabetic macular edema (DME)

Remigromig is the first biologic with a novel mechanism of action to demonstrate non-inferior visual acuity compared with anti-VEGF therapy in a pivotal DME trial

Despite available therapies, up to 40% of people with DME have persistent disease after six months, driving need for new approaches

RAHWAY, N.J.–(BUSINESS WIRE)–$MRK #MRK–Merck (NYSE: MRK), known as MSD outside of the United States and Canada, today announced the first presentation of one-year results from the pivotal Phase 2b/3 BRUNELLO trial evaluating remigromig (MK-3000, formerly EYE103), an investigational, potentially first-in-class tetravalent, tri-specific antibody designed to activate the Wingless-related integration site (Wnt) pathway, for the treatment of diabetic macular edema (DME). Results were presented today at the American Academy of Ophthalmology (AAO) 2026 Annual Meeting in New Orleans: BRUNELLO – 1-Year Pivotal Trial Results of Remigromig (MK-3000, formerly EYE103) for the Treatment of Diabetic Macular Edema; D. D’Amico.

In the BRUNELLO study, both remigromig dose arms met the primary endpoint independently, demonstrating non-inferiority in mean change from baseline in best corrected visual acuity (BCVA) at one year compared with monthly 0.5mg ranibizumab. Mean BCVA gains at Year 1 were +9.1 letters (95% CI, 8.1-10.1; p=0.0129, multiplicity-adjusted) with remigromig 0.5 mg and +8.7 letters (95% CI, 7.7-9.8; p=0.0222, multiplicity-adjusted) with remigromig 0.8 mg, compared with +11.8 letters with ranibizumab (95% CI, 10.7-12.8). Differences between treatment groups were within the prespecified non-inferiority margin and were not considered clinically meaningful. No secondary endpoints demonstrated superiority to ranibizumab.

While both doses of remigromig were generally well tolerated, adverse events related to proliferative diabetic retinopathy (PDR) occurred more frequently with remigromig than with ranibizumab (6.7% and 6.1% with remigromig 0.5 mg and 0.8 mg, respectively, versus 0.9%), inclusive of new-onset PDR.1 In addition, higher rates of treatment discontinuations due to adverse events were observed in the remigromig treatment arms compared with ranibizumab (4.9% and 4.5% with remigromig 0.5 mg and 0.8 mg, respectively, versus 0.9%). Additional analyses are underway to further characterize these findings. PDR rates may reflect natural disease progression in DME patients since remigromig focuses on repair and maintenance of the blood-retinal barrier rather than directly suppressing new vessel growth. PDR cases were generally manageable with standard-of-care treatment.

“Although anti-VEGF therapies remain the foundation of treatment for diabetic macular edema, up to two thirds of patients fail to achieve clinically meaningful vision gains after one year, underscoring the need to explore new biological pathways,” said Donald J. D’Amico, M.D., chair of Ophthalmology at Weill Cornell Medicine, Ophthalmologist-in-Chief, New York-Presbyterian Hospital and presenting author of the BRUNELLO study. “These findings ‒ which demonstrate for the first time that a biologic targeting the Wnt pathway can achieve visual acuity comparable to an established anti-VEGF therapy ‒ support continued evaluation of this novel approach as we look forward to additional Phase 3 results from the BAROLO study.”

“Despite important advances in care, up to 40% of patients with diabetic macular edema have persistent disease activity after six months of treatment, meaning that they do not respond or only partially respond to therapy, underscoring the need for new therapeutic approaches,” said Dr. David Guyer, founder, chief executive officer and president of EyeBio, a wholly owned subsidiary of Merck. “The BRUNELLO study provides important evidence that activating the Wnt pathway, which is involved in the repair and maintenance of the blood-retinal barrier, may offer a potential new way to help patients with DME and expand treatment options for retinal specialists and their patients.”

“The BRUNELLO results represent an important milestone for people living with diabetic macular edema, a leading cause of vision loss among people living with diabetes, which impacts nearly 1.6 million people in the U.S.,” said Dr. Dean Y. Li, president, Merck Research Laboratories. “As the first new mechanism of action in more than 20 years to demonstrate non-inferior visual acuity compared with standard-of-care anti-VEGF therapy in a pivotal DME study, remigromig has the potential to expand treatment options for retinal specialists and patients. We thank the patients, caregivers and investigators whose participation made this study possible.”

The BRUNELLO trial data further strengthens Merck’s advancing ophthalmology pipeline, which is aimed at promoting retinal recovery by addressing specific retinal diseases associated with vascular leakage and neovascularization, including DME, neovascular (wet) age-related macular degeneration (NVAMD) and macular edema secondary to retinal vein occlusion (RVO).

Results from BRUNELLO will be discussed with regulatory authorities. Remigromig is also being evaluated in the ongoing pivotal Phase 2b/3 BAROLO study (NCT06957080) in patients with DME and in a Phase 2 proof-of-concept study (SUPER TUSCAN) in patients with NVAMD and RVO (NCT07205887).

The company is also developing MK-8748 (also known as Tiespectus, EYE201), a novel investigational bispecific antibody with a dual mechanism that directly activates the Tie2 pathway and inhibits VEGF with the goal of stabilizing retinal and choroidal blood vessels and reducing fluid accumulation in the macula. MK-8748 is currently being studied in two pivotal Phase 2b/3 trials for the treatment of NVAMD, TORRONTES (NCT07496567) and MALBEC (NCT07440225), and in two pivotal Phase 3 studies for the treatment of DME, SANGIOVESE (NCT07705555) and SYRAH (NCT07705607).

About the BRUNELLO trial

BRUNELLO is a randomized, double masked pivotal Phase 2b/3 trial (NCT06571045) evaluating the safety and efficacy of two dose levels (0.5 mg and 0.8 mg) of intravitreal (IVT) remigromig (MK-3000, formerly EYE103) versus active control 0.5mg ranibizumab in adults with DME. The trial enrolled 984 participants who were randomized 1:1:1 to receive low and high dose regimens of remigromig or ranibizumab every four weeks for the first year. In the second year, the frequency of treatment for participants will shift based on a personalized treatment interval (PTI) algorithm. The primary endpoint is mean change in best-corrected visual acuity (BCVA) from baseline to week 52 in the study eye of the participants, using standardized Early Treatment of Diabetic Retinopathy Study (ETDRS) vision testing. The key secondary endpoints at week 52 are mean change from baseline in BCVA (superiority of remigromig vs. ranibizumab), and mean change from baseline in Central Subfield Thickness (CST) (superiority of remigromig vs. ranibizumab).

About Remigromig

Remigromig (MK-3000, formerly EYE103) is an investigational, potentially first-in-class tetravalent, tri-specific antibody designed to activate the Wingless-related integration site (Wnt) pathway, which is involved in the repair and maintenance of the blood-retinal-barrier. Remigromig is being studied in patients with certain retinal diseases including diabetic macular edema (DME) and neovascular age-related macular degeneration (NVAMD). Remigromig is administered by intravitreal injection.

About diabetic macular edema

Diabetic macular edema (DME) is a serious retinal condition and a leading cause of vision loss for people with diabetes. An estimated 1.6 million people are living with DME in the United States. DME occurs when damaged blood vessels leak into the retina, resulting in swelling in the macula, the central region of the retina crucial for precise vision necessary for everyday activities. The prevalence of DME is anticipated to rise with the increasing incidence of diabetes. While currently available treatments have helped to improve outcomes, a significant number of patients, up to 40%, do not respond or only partially respond to therapy, underscoring the need for new treatment approaches.

About Merck

At Merck, known as MSD outside of the United States and Canada, we are unified around our purpose: We use the power of leading-edge science to save and improve lives around the world. For more than 130 years, we have brought hope to humanity through the development of important medicines and vaccines. We aspire to be the premier research-intensive biopharmaceutical company in the world – and today, we are at the forefront of research to deliver innovative health solutions that advance the prevention and treatment of diseases in people and animals. We foster a diverse and inclusive global workforce and operate responsibly every day to enable a safe, sustainable and healthy future for all people and communities. For more information, visit www.merck.com and connect with us on X (formerly Twitter), Facebook, Instagram, YouTube and LinkedIn.

Forward-Looking Statement of Merck & Co., Inc., Rahway, N.J., USA

This news release of Merck & Co., Inc., Rahway, N.J., USA (the “company”) includes “forward-looking statements” within the meaning of the safe harbor provisions of the U.S. Private Securities Litigation Reform Act of 1995. These statements are based upon the current beliefs and expectations of the company’s management and are subject to significant risks and uncertainties. There can be no guarantees with respect to pipeline candidates that the candidates will receive the necessary regulatory approvals or that they will prove to be commercially successful. If underlying assumptions prove inaccurate or risks or uncertainties materialize, actual results may differ materially from those set forth in the forward-looking statements.

Risks and uncertainties include, but are not limited to, general industry conditions and competition; general economic factors, including interest rate and currency exchange rate fluctuations; the impact of pharmaceutical industry regulation and health care legislation in the United States and internationally; global trends toward health care cost containment; technological advances, new products and patents attained by competitors; challenges inherent in new product development, including obtaining regulatory approval; the company’s ability to accurately predict future market conditions; manufacturing difficulties or delays; financial instability of international economies and sovereign risk; dependence on the effectiveness of the company’s patents and other protections for innovative products; and the exposure to litigation, including patent litigation, and/or regulatory actions.

The company undertakes no obligation to publicly update any forward-looking statement, whether as a result of new information, future events or otherwise. Additional factors that could cause results to differ materially from those described in the forward-looking statements can be found in the company’s Annual Report on Form 10-K for the year ended December 31, 2025 and the company’s other filings with the Securities and Exchange Commission (SEC) available at the SEC’s Internet site (http://www.sec.gov/).

1 PTs indicative of PDR: diabetic retinopathy (if LLT ‘proliferative diabetic retinopathy’), vitreous hemorrhage, iris neovascularization, retinal neovascularization, glaucoma (if LLT ‘neovascular glaucoma’), neovascularization

Contacts

Media Contacts:

Elizabeth Sell

elizabeth.sell@merck.com

Shannon Hatch

Shannon.hatch@merck.com

Investor Contacts:

Peter Dannenbaum

(732) 594-1579

Hayley Kasko

(732) 594-4237

Genentech Presents New Phase III One-Year Data for Vamikibart in Uveitic Macular Edema (UME), a Serious Cause of Vision Loss

Genentech Presents New Phase III One-Year Data for Vamikibart in Uveitic Macular Edema (UME), a Serious Cause of Vision Loss


Genentech Presents New Phase III One-Year Data for Vamikibart in Uveitic Macular Edema (UME), a Serious Cause of Vision Loss

  • In the MEERKAT and SANDCAT studies, vamikibart demonstrated sustained improvements in vision and reductions in macular thickness in patients with UME at 52 weeks
  • UME is the leading cause of moderate and severe vision loss in patients with uveitis, posing a significant risk of blindness if left untreated
  • The FDA has accepted Genentech’s Biologics License Application (BLA), with an approval decision expected by July 2027 


SOUTH SAN FRANCISCO, Calif.–(BUSINESS WIRE)–$RHHBY–Genentech, a member of the Roche Group (SIX: RO, ROP; OTCQX: RHHBY), announced today new one-year data from the Phase III MEERKAT and SANDCAT studies evaluating investigational vamikibart compared with a sham procedure that mimics intravitreal (IVT) injections in adults with uveitic macular edema (UME). Across both studies, data continue to support the potential for rapid and sustained improvements in vision and reductions in macular thickness (swelling in the back of the eye due to retinal fluid) with vamikibart treatment. The data were presented at the American Academy of Ophthalmology 2026 Annual Meeting (AAO 2026) in New Orleans, LA.

Additionally, the U.S. Food and Drug Administration (FDA) has accepted Genentech’s Biologics License Application (BLA) for vamikibart for the treatment of UME. The filing acceptance is based on the results from the MEERKAT and SANDCAT studies. The FDA is expected to make a decision on approval by July 2027. If approved, vamikibart would be the first non-steroid targeted treatment for UME, with potential to establish a new standard of care. Regulatory submissions have also been filed and accepted in the European Union, China and Japan.

“These one-year results strengthen vamikibart’s clinical profile, confirming that initial visual and anatomical improvements are maintained over 52 weeks,” said Levi Garraway, M.D., Ph.D., chief medical officer and head of Global Product Development. “The acceptance of our application by the FDA and other regulatory bodies around the world brings us closer to offering a transformative, non-steroid therapy for people living with this sight-threatening condition.”

“Currently, UME is commonly treated with off-label therapies or ocular steroids, the latter carrying significant long-term side effects like glaucoma and cataract formation,” said Nisha Acharya, M.D., M.S., Distinguished Professor of Ophthalmology at UCSF and Director of the Uveitis and Ocular Inflammatory Disease Service. “Vamikibart is the first non-steroid targeted therapy to demonstrate meaningful and sustained visual and anatomical improvements in people with UME, offering a potential new treatment option that directly targets the inflammation driving this sight-threatening condition.”

The 16-week primary analysis results were presented at the AAO meeting in October 2025. In both MEERKAT and SANDCAT at 52 weeks, a numerically higher proportion of vamikibart-treated patients achieved vision gains compared to sham treatment on the primary endpoint. Key secondary endpoints showed this benefit was maintained, with sustained improvements in average change from baseline in best corrected visual acuity (BCVA) and average change from baseline in central subfield thickness (CST), a key measure of macular edema. Vamikibart demonstrated durable clinical benefit with a low treatment burden: approximately two-thirds of eligible patients required no retreatment after 16 weeks, while the majority of those who did required only one additional injection. Vamikibart was well tolerated, maintaining a favorable safety profile, with a low incidence of treatment-related ocular adverse events (AEs) and intraocular inflammation (IOI) events.

UME is the leading cause of moderate-to-severe vision loss in individuals with uveitis, predominantly affecting working-age adults. Approximately 20% of UME patients progress to sustained blindness within three years of diagnosis. Standard-of-care treatments, primarily high-dose or local corticosteroids, carry substantial long-term risks, including secondary cataracts and elevated intraocular pressure leading to glaucoma. As a targeted anti-IL-6 monoclonal antibody, vamikibart is being developed to address these clinical unmet needs and deliver sustained visual preservation.

About the MEERKAT and SANDCAT studies

MEERKAT (NCT05642312) and SANDCAT (NCT05642325) are identical Phase III, global, parallel, multicenter, randomized, double-masked, sham comparator-controlled, 52-week trials of intravitreal (IVT) vamikibart in uveitic macular edema (UME). In both trials, patients were randomized and received treatment every four weeks with either 0.25 mg vamikibart, 1 mg vamikibart or sham IVT injection, for up to 16 weeks, followed by PRN treatment through week 52. The primary endpoint of both Phase III trials was the proportion of participants with a 15 letter or more improvement from baseline in best corrected visual acuity (BCVA) at week 16. Key secondary endpoints included the average change from baseline in BCVA and CST at week 16. The safety of vamikibart was assessed through adverse events (AEs) such as treatment related ocular AEs, intraocular inflammation (IOI) and retinal occlusive vasculitis. The studies included participants with and without prior IVT treatment history and included patients with history of raised IOP and glaucoma.

About uveitic macular edema (UME)

UME is characterized by the buildup of fluid in the macula due to uveitis, an inflammatory eye condition. Although rare compared to other eye diseases, UME has a disproportionate impact on vision loss and blindness globally. It is the leading cause of moderate and severe vision loss in people with uveitis, and the most frequent sight-threatening complication in uveitis. Uveitis accounts for 10% to 20% of blindness in the United States and Europe, and up to 25% of blindness in the developing world. UME has a significant negative impact on people’s quality of life, including physical and mental health, social functioning, and visual function for day-to-day activities such as driving and reading. Steroids, the current standard of care for UME, are associated with significant serious side effects such as increased pressure in the eye, glaucoma and cataracts, and have recognized efficacy limitations.

About Vamikibart

Vamikibart is an investigational monoclonal antibody that has been specifically engineered for IVT administration. It targets interleukin-6 (IL-6), a key cytokine in the inflammatory pathway in UME. In the Phase I DOVETAIL study, vamikibart provided rapid vision improvements and resolution of macular edema in people with UME. Vamikibart was also well tolerated, with no treatment-related serious adverse events reported. Based on the promising Phase I DOVETAIL data, Genentech initiated the two identical Phase III vamikibart studies MEERKAT and SANDCAT. Vamikibart is being investigated in retinal diseases with recognized inflammatory pathways. Vamikibart has orphan drug designation in the United States and European Union.

About Genentech in Ophthalmology

Genentech is researching and developing new treatments for people living with a range of eye diseases that cause significant visual impairment and blindness, including wet age-related macular degeneration (AMD), diabetic macular edema (DME), diabetic retinopathy (DR), geographic atrophy (GA) and other retinal diseases, including rare and inherited conditions.

About Genentech

Founded 50 years ago, Genentech is a leading biotechnology company that discovers, develops, manufactures and commercializes medicines to treat patients with serious and life-threatening medical conditions. The company, a member of the Roche Group, has headquarters in South San Francisco, California. For additional information about the company, please visit http://www.gene.com.

Contacts

Media Contact:

Nicole Burkart

(650) 467-6800

Advocacy Contact:

Meg Harrison

(617) 694-7060

Investor Contacts:

Loren Kalm

(650) 225-3217

Bruno Eschli

+41 61 687 5284