AngioDynamics Announces Inducement Grants Under Nasdaq Listing Rule 5635(c)(4)

AngioDynamics Announces Inducement Grants Under Nasdaq Listing Rule 5635(c)(4)


AngioDynamics Announces Inducement Grants Under Nasdaq Listing Rule 5635(c)(4)

LATHAM, N.Y.–(BUSINESS WIRE)–AngioDynamics, Inc. (NASDAQ: ANGO), a leading and transformative medical technology company focused on restoring healthy blood flow in the body’s vascular system, expanding cancer treatment options and improving patient quality of life, today announced the grant of inducement awards to Eric Honroth in connection with his appointment as Chief Executive Officer of the Company effective as of November 2, 2026.

On November 2, 2026 (the “Grant Date”), as an inducement material to Mr. Honroth entering into employment with the Company in accordance with Nasdaq Listing Rule 5635(c)(4) and pursuant to approval by the independent Compensation Committee of the Board of Directors of the Company (the “Committee”), the Company granted to Mr. Honroth (i) time-based restricted stock units with respect to 62,928 shares of Common Stock of the Company (the “First Sign-On Award”), (ii) performance-based restricted stock units with respect to 62,928 shares of Common Stock of the Company at the target level of performance (the “Second Sign-On Award”) and (iii) time-based restricted stock units with respect to 38,265 shares of Common Stock of the Company (the “Third Sign-On Award”).

The First Sign-On Award will vest in equal annual installments over four years, subject to Mr. Honroth’s continued service through each applicable vesting date.

The Second Sign-On Award will vest, if at all, based on a combination of the Company’s revenue, adjusted EBITDA, and relative total shareholder return over a three-year performance period, subject to Mr. Honroth’s continued service through the date of the Committee’s certification as to whether or not the threshold performance metrics have been met. The number of shares that may be earned under the PSUs ranges from 0% of target to 240% of target.

The Third Sign-On Award will vest in two equal installments on November 2, 2027 and November 2, 2028, subject to Mr. Honroth’s continued service through each applicable vesting date.

About AngioDynamics, Inc.

AngioDynamics is a leading and transformative medical technology company focused on restoring healthy blood flow in the body’s vascular system, expanding cancer treatment options and improving patient quality of life.

The Company’s innovative technologies and devices are chosen by talented physicians in fast-growing healthcare markets to treat unmet patient needs. For more information, visit www.angiodynamics.com.

Safe Harbor

This release contains forward-looking statements within the meaning of the Private Securities Litigation Reform Act of 1995. All statements regarding AngioDynamics’ expected future financial position, results of operations, cash flows, business strategy, budgets, projected costs, capital expenditures, products, competitive positions, growth opportunities, plans and objectives of management for future operations, as well as statements that include the words such as “expects,” “reaffirms,” “intends,” “anticipates,” “plans,” “believes,” “seeks,” “estimates,” “projects,” “optimistic,” or variations of such words and similar expressions, are forward-looking statements. These forward-looking statements are not guarantees of future performance and are subject to risks and uncertainties. Investors are cautioned that actual events or results may differ materially from AngioDynamics’ expectations, expressed or implied. Factors that may affect the actual results achieved by AngioDynamics include, without limitation, the scale and scope of the COVID-19 global pandemic, the ability of AngioDynamics to develop its existing and new products, technological advances and patents attained by competitors, infringement of AngioDynamics’ technology or assertions that AngioDynamics’ technology infringes the technology of third parties, the ability of AngioDynamics to effectively compete against competitors that have substantially greater resources, future actions by the FDA or other regulatory agencies, domestic and foreign healthcare reforms and government regulations, results of pending or future clinical trials, overall economic conditions (including inflation, tariffs, labor shortages and supply chain challenges including the cost and availability of raw materials), the results of on-going litigation, challenges with respect to third-party distributors or joint venture partners or collaborators, the results of sales efforts, the effects of product recalls and product liability claims, changes in key personnel, the ability of AngioDynamics to execute on strategic initiatives, the effects of economic, credit and capital market conditions, general market conditions, market acceptance, foreign currency exchange rate fluctuations, the effects on pricing from group purchasing organizations and competition, the ability of AngioDynamics to obtain regulatory clearances or approval of its products, or to integrate acquired businesses, as well as the risk factors listed from time to time in AngioDynamics’ SEC filings, including but not limited to its Annual Report on Form 10-K for the year ended May 31, 2026. AngioDynamics does not assume any obligation to publicly update or revise any forward-looking statements for any reason.

AngioDynamics and the AngioDynamics logo are trademarks and/or registered trademarks of AngioDynamics, Inc., an affiliate or subsidiary. All other trademarks are property of their respective owners.

Contacts

Investor Contact:
Stephen Trowbridge

Executive Vice President & CFO

518-795-1408

strowbridge@angiodynamics.com

Media Contact:
Saleem Cheeks

Vice President, Communications

518-795-1174

scheeks@angiodynamics.com

bioAffinity Technologies Reports Third Quarter 2026 Results and Continued Momentum in CyPath® Lung Commercialization

bioAffinity Technologies Reports Third Quarter 2026 Results and Continued Momentum in CyPath® Lung Commercialization


bioAffinity Technologies Reports Third Quarter 2026 Results and Continued Momentum in CyPath® Lung Commercialization

CyPath® Lung test unit sales double year to date over 2025, with solid gains in physician utilization and recurring orders; testing revenue increases 79% year-to-date

Company advances precision medicine pipeline with new asthma and COPD research, collaboration with Pictor and continued development of topical skin cancer therapeutics

SAN ANTONIO–(BUSINESS WIRE)–$BIAF #BIAF—bioAffinity Technologies, Inc. (Nasdaq: BIAF; BIAFW), a biotechnology company developing noninvasive healthcare solutions for the early detection and monitoring of lung disease and topically delivered therapeutics for skin cancer, today reported financial results and business highlights for the third quarter (Q3) ended September 30, 2026.


Q3 2026 Highlights and Recent Events

  • CyPath® Lung test unit sales increased 100% through Q3 compared to the first nine months of 2025.
  • CyPath Lung testing revenue for the first nine months of 2026 increased 79% to approximately $1.1 million, compared to the same period in 2025.
  • In July 2026, the Company surpassed the total number of CyPath Lung test reports delivered to physicians during 2025.
  • The number of ordering physicians increased by 89% in the third quarter of 2026 compared to the third quarter of 2025, continuing to expand clinical utilization of CyPath Lung. Physicians already ordering CyPath Lung tests increased their volume by 48% year over year.
  • The Company expanded its government healthcare distribution network and commercial reach through an agreement with AvMEDICAL, a Service-Disabled Veteran-Owned Small Business and established distributor of medical-surgical and laboratory products and services to government agencies, including the Department of Veterans Affairs (VA).
  • The ongoing longitudinal clinical trial designed to evaluate the clinical performance of CyPath Lung continued to advance. As of September 30, 2026, 11 clinical sites were active, including nine VA and military medical centers that had begun patient enrollment. The John P. Murtha Cancer Center Research Program at the Uniformed Services University of the Health Sciences is providing support and funding for the trial.
  • In July, the Company published a comprehensive clinical review and white paper authored by Chief Medical Officer Gordon H. Downie, MD, PhD, presenting a practical framework for incorporating CyPath Lung into pulmonology and oncology practices. Dr. Downie presented the framework to physicians in October at the annual meeting of the American College of Chest Physicians (CHEST 2026).
  • The Company advanced CyPath Lung for potential use in surveillance of lung cancer survivors as a complement to imaging to monitor patients for cancer recurrence and new primary lung cancers.
  • The Company presented positive research results at CHEST 2026 relating to its development of noninvasive tests to characterize lung inflammation and investigate biologic drug targets for asthma and COPD therapies. Further development and validation are underway to determine whether this approach can identify patients who may benefit from specific biologic therapies or support monitoring of lung inflammation over time.
  • The Company announced a collaboration with Pictor®, Inc., a targeted proteomic platform company, to support development and potential commercialization of bioAffinity Technologies’ next-generation diagnostic tests designed to provide a more complete picture of lung inflammation in patients with asthma and COPD.
  • The Company continued research into potential topically delivered therapeutics for squamous and basal cell skin cancers. Preliminary research findings have demonstrated that self-delivering, stabilized small interfering RNAs (siRNAs) selectively kill squamous and basal carcinoma cells in vitro while sparing noncancerous cutaneous cells.
  • In September, bioAffinity announced the grant of a Hong Kong patent covering CyPath Lung’s proprietary flow cytometry + AI platform for predicting the likelihood of lung cancer by characterizing cell populations in sputum. In October, the Company received a Notice of Allowance from the Japan Patent Office indicating its intention to grant a patent for the method of predicting the likelihood of lung cancer used by the CyPath Lung test.
  • The Company raised an aggregate of $8.0 million in gross proceeds through a private placement offering in August 2026 and a registered direct offering and concurrent private placement in October 2026 for working capital and general corporate purposes, including support for the ongoing commercialization of CyPath Lung.

Management Commentary

“Our third-quarter results reflect CyPath Lung’s continued momentum in addressing the critical need for early lung cancer detection, driven by both new physicians ordering the test and increased use by existing customers. We believe this growth reflects increasing physician engagement and recognition of the role CyPath Lung can play in managing care for patients at risk for lung cancer. We are also advancing its potential use in surveillance following curative-intent treatment, where CyPath Lung could complement imaging in monitoring for recurrence and new cancers,” said Maria Zannes, President of CEO of bioAffinity Technologies. “Peer-to-peer education and growing clinical experience with the test remain important drivers of adoption, while our longitudinal study is designed to generate robust, real-world data across multiple sites to further evaluate how CyPath Lung may support risk assessment and clinical decision-making. Together, these efforts advance our objective of establishing CyPath Lung as a standard of care for evaluating patients at risk for lung cancer.

“We continue to build the foundation for long-term growth. Beyond lung cancer, we are advancing our platform to take advantage of precision medicine opportunities in asthma and COPD, where sputum-based analysis may help characterize lung inflammation and help physicians better match individual patients to available treatments, including Tezspire, Dupixent and Nucala. The tests we are developing are designed to meet a critical need for precision diagnostics in the multi-billion-dollar asthma and COPD therapeutic markets.

“Our patent portfolio now includes 22 awarded U.S. and foreign patents and 36 pending applications. The recent Hong Kong patent and the Japanese patent allowance, once awarded, are important additions to our patent portfolio. Our distribution agreement with AvMEDICAL is intended to expand our reach into government healthcare markets, including the VA, where the substantial burden of lung cancer underscores the need for more effective ways to evaluate pulmonary nodules and improve patient outcomes.

Ms. Zannes concluded, “Our priorities remain disciplined execution, expanding commercial adoption and advancing our clinical and research programs. We are also pursuing topical therapeutic approaches for squamous and basal cell skin cancers based on promising findings involving the CD320 and LRP2 receptors. We believe our expertise in flow cytometry, automated data analysis and siRNA technology provides a foundation for developing differentiated products across these markets. Together, these programs demonstrate our commitment to translating scientific innovation into meaningful new opportunities for patients and the Company.”

Third Quarter 2026 Financial Results

Total revenue for the quarter ended September 30, 2026, was approximately $1.1 million, compared with approximately $1.4 million for the same period in 2025. The decrease is due primarily to a decrease in average reimbursement per CyPath Lung test and a slight decline in total cases completed at Precision Pathology Laboratory Services (PPLS) compared to the same period in 2025, in part due to internal business changes at three PPLS research clients. Multiple accounts previously serviced under these clients have since returned to PPLS. The Company is working to restore additional accounts and improve reimbursement levels.

Net loss for the quarter ended September 30, 2026, was approximately $5.3 million, compared with approximately $5.1 million for the third quarter of 2025. The increase in net loss is primarily due to the increase in loss from operations of approximately $3.5 million from approximately $2.3 million in the prior-year quarter, partially offset by a $1.2 million loss from the change in fair value of warrants, compared with a $2.7 million loss in the prior-year quarter.

Cash and cash equivalents were approximately $2.6 million as of September 30, 2026, compared with approximately $6.4 million at December 31, 2025. Subsequent to quarter-end, the Company completed a financing that generated approximately $4.0 million in gross proceeds.

Operating expenses for the third quarter of 2026 were approximately $4.6 million, compared with approximately $3.7 million in the third quarter of 2025.

  • Direct costs and expenses were approximately $1.1 million, compared with $942,000 in the prior-year period. The increase primarily reflected additional personnel to process CyPath Lung commercial tests and tests associated with the longitudinal clinical study.
  • Research and development expenses increased 18% year over year to approximately $389,000, primarily due to laboratory supplies and costs associated with relocating laboratory facilities in June 2026.
  • Clinical development expenses increased 75% to approximately $251,000, compared with approximately $143,000 in the third quarter of 2025. The increase primarily reflected the initiation of clinical trial sites and patient enrollment in the longitudinal clinical study.
  • Selling, general and administrative expenses increased 28% to approximately $2.8 million, compared with approximately $2.2 million in the prior-year period. The increase primarily reflected additional sales and marketing personnel and support services to expand commercialization of CyPath Lung.

About CyPath Lung

CyPath Lung by bioAffinity Technologies is a noninvasive test designed to improve the early detection of lung cancer in patients at high risk for the disease. CyPath Lung analyzes the lung microenvironment by using advanced flow cytometry and proprietary artificial intelligence (AI) to identify cell populations in patient sputum that indicate malignancy. The test incorporates a novel fluorescent porphyrin that is preferentially taken up by cancer and cancer-related cells. In a published clinical trial of high-risk patients, CyPath Lung demonstrated 92% sensitivity, 87% specificity, 88% accuracy and 99% negative predictive value (NPV) in detecting lung cancer in patients at high risk for the disease who had small indeterminate lung nodules less than 20 millimeters. The high NPV gives physicians greater confidence that a negative result is truly negative, potentially sparing patients from unnecessary invasive and costly procedures. CyPath Lung is marketed as a Laboratory Developed Test (LDT) and is not intended for use as a sole diagnostic tool and should be considered alongside other clinical findings. For more information, visit www.cypathlung.com.

About bioAffinity Technologies, Inc.

bioAffinity Technologies, Inc. addresses the need for noninvasive diagnosis of early-stage cancer and other diseases of the lung and broad-spectrum cancer treatments. The Company’s first product, CyPath Lung, is a noninvasive test that has shown high sensitivity, specificity and accuracy for the detection of early-stage lung cancer. CyPath Lung is marketed as a Laboratory Developed Test (LDT) by Precision Pathology Laboratory Services, a subsidiary of bioAffinity Technologies. LDTs are overseen under the Clinical Laboratory Improvement Amendments (CLIA), which are administered by the Centers for Medicare & Medicaid Services. For more information, visit www.bioaffinitytech.com.

Forward-Looking Statements

Certain statements in this press release constitute “forward-looking statements” within the meaning of the federal securities laws. Words such as “may,” “might,” “will,” “should,” “believe,” “expect,” “anticipate,” “estimate,” “continue,” “predict,” “forecast,” “project,” “plan,” “intend” and similar expressions may identify forward-looking statements, although not all forward-looking statements contain these words. Actual results could differ materially from those expressed or implied because of risks and uncertainties, including the Company’s ability to expand the commercial adoption of CyPath Lung and capitalize on government healthcare market opportunities; the timing, cost, enrollment and outcome of clinical studies; the possibility that preliminary, in vitro or study-specific findings will not be replicated or translate into safe and effective therapies or commercial products for asthma, COPD or skin cancer; the need for further validation of new applications, including surveillance; the extent and timing of physician adoption, repeat utilization and customer retention; payer coverage, reimbursement rates and collections; and the Company’s ability to obtain, maintain and enforce intellectual property rights. Additional risks are discussed in the Company’s Annual Report on Form 10-K for the year ended December 31, 2025, and subsequent SEC filings, including its reports on Forms 10-Q and 8-K. Forward-looking statements reflect expectations and assumptions only as of the date of this release. Readers should not place undue reliance on them. The Company undertakes no obligation to update these statements, except as required by applicable law.

bioAffinity Technologies, Inc.

Condensed Consolidated Balance Sheets


September 30,

2026


December 31,

2025


(unaudited)


ASSETS


Current assets:


Cash and cash equivalents


$

2,550,034


$

6,449,782


Accounts and other receivables, net


742,144


541,962


Inventory


108,754


53,548


Prepaid expenses and other current assets


427,230


519,916


Total current assets


3,828,162


7,565,208


Non-current assets:


Property and equipment, net


362,359


265,593


Operating lease right-of-use asset, net


730,572


334,289


Finance lease right-of-use asset, net


538,326


661,575


Goodwill


1,404,486


1,404,486


Intangible assets, net


673,056


716,806


Other assets


16,710


12,815


Total assets


$

7,553,671


$

10,960,772


LIABILITIES AND STOCKHOLDERS’ EQUITY


Current liabilities:


Accounts payable


$

949,018


$

761,901


Accrued expenses


1,143,227


1,717,989


Unearned revenue


48,874


42,405


Operating lease liability, current portion


168,534


139,220


Finance lease liability, current portion


78,926


139,490


Notes payable, current portion


144,715


105,161


Total current liabilities


2,533,294


2,906,166


Non-current liabilities:


Operating lease liability, net of current portion


596,086


202,878


Finance lease liability, net of current portion


475,772


532,759


Notes payable, net of current portion


33,958


41,313


Total liabilities


3,639,110


3,683,116


Stockholders’ equity:


Preferred Stock, par value $0.001 per share; 20,000,000 shares authorized; 150 shares and 700 shares issued and outstanding at September 30, 2026, and December 31, 2025, respectively


1


1


Common Stock, par value $0.007 per share; 350,000,000 shares authorized; 1,166,625 and 299,826 issued and outstanding at September 30, 2026, and December 31, 2025, respectively


8,166


2,097


Additional paid-in capital


84,734,173


75,829,622


Accumulated deficit


(80,827,779

)


(68,554,064

)


Total stockholders’ equity


3,914,561


7,277,656


Total liabilities and stockholders’ equity


$

7,553,671


$

10,960,772


bioAffinity Technologies, Inc.

Unaudited Condensed Consolidated Statements of Operations


Three Months Ended

September 30,

Nine Months Ended

September 30,


2026

2025

2026

2025


Net revenue

$

1,070,280


$

1,446,066


$

3,932,386


$

4,569,146


Operating expenses:


Direct costs and expenses


1,114,176


942,211


3,130,649


3,326,673


Research and development


389,122


330,589


1,100,404


1,009,347


Clinical development


251,043


143,311


1,060,968


410,943


Selling, general and administrative


2,759,769


2,209,441


8,859,961


6,876,551


Depreciation and amortization


66,842


113,360


242,916


381,177


Total operating expenses


4,580,952


3,738,912


14,394,898


12,004,691


Loss from operations


(3,510,672

)


(2,292,846

)


(10,462,512

)


(7,435,545

)


Other income (expense):


Interest income


4,734


1,088


18,118


3,655


Interest expense


(10,713

)


(5,358

)


(37,123

)


(31,303

)

Other income


3,144


513


6,516


38,568


Other expense


(13,072

)


(3,839

)


(48,270

)


(16,828

)

Offering costs allocated to warrant liability


(532,209

)


—


(532,209

)


(479,696


Loss on warrant liabilities


(1,218,235

)


(2,747,460

)


(1,218,235

)


(3,810,278

)


Total other income (expense), net


(1,766,351

)


(2,755,056

)


(1,811,203

)


(4,295,882

)


Net loss before provision for income tax expense


(5,277,023

)


(5,047,902

)


(12,273,715

)


(11,731,427

)


Income tax expense


—


3,182


—


40,861


Net loss

$

(5,277,023

)

$

(5,051,084

)

$

(12,273,715

)

$

(11,772,288

)


Net loss per common share, basic and diluted

$

(5.89

)

$

(71.05

)

$

(23.72

)

$

(219.47

)


Weighted average common shares outstanding


896,271


71,090


517,490


53,640


Contacts

bioAffinity Technologies
Julie Anne Overton

Director of Communications

investors@bioaffinitytech.com

Kardigan Completes Enrollment in Phase 2b KATALYST-AV Trial and Announces New Evidence Supporting Patient Selection and Endpoints for the Evaluation of Ataciguat in Aortic Stenosis

Kardigan Completes Enrollment in Phase 2b KATALYST-AV Trial and Announces New Evidence Supporting Patient Selection and Endpoints for the Evaluation of Ataciguat in Aortic Stenosis


Kardigan Completes Enrollment in Phase 2b KATALYST-AV Trial and Announces New Evidence Supporting Patient Selection and Endpoints for the Evaluation of Ataciguat in Aortic Stenosis

Primary endpoint of aortic valve calcium in KATALYST-AV to be assessed at Week 48 alongside aortic valve area and measures of cardiac structure and function; topline data expected 2H 2027

Novel real-world data help identify key markers of risk, rate of progression of disease and worsening outcomes in aortic stenosis, informing patient selection criteria and potential endpoints for Phase 3 registrational trial

Company to host an investor webcast while at the American Heart Association Scientific Sessions on Saturday, November 7, 2026, at 8:00 AM ET

SOUTH SAN FRANCISCO, Calif. & PRINCETON, N.J.–(BUSINESS WIRE)–$KARD #Kardigan–Kardigan®, Inc. (Nasdaq: KARD), a precision cardiovascular company, today announced enrollment completion in the Phase 2b KATALYST-AV trial of ataciguat in people with aortic stenosis (AS). The Company also shared insights from a pre-planned analysis of the trial’s baseline data, as well as new evidence on disease progression from one of the largest real-world analyses conducted in approximately 45,000 people with AS. These analyses support the primary endpoint assessment of aortic valve calcium (AVC) at Week 48, alongside measures of cardiac structure and function, and an internal interim analysis at Week 24.


“Completing enrollment in KATALYST-AV is a foundational milestone for our program and an important one for people with aortic stenosis,” said Tassos Gianakakos, co-founder, chief executive officer and board chair of Kardigan. “As with our other clinical programs, KATALYST-AV was specifically designed to enable us to learn and adapt as evidence accumulates. The real-world data give us a clearer picture of which AS patients progress more quickly to worsening outcomes and the markers associated with that progression, directly informing how we design our upcoming registrational trial. In addition, our analysis of aortic valve calcium in the KATALYST-AV baseline data is consistent with the variability observed in the real-world data, which indicates more time is needed to assess a robust reduction in calcium with ataciguat versus placebo. Together, these findings reinforce our development strategy addressing both valve calcification and ventricular dysfunction with ataciguat and strengthen our confidence in its potential to become the first therapy to slow the progression of aortic stenosis.”

KATALYST-AV Baseline Data

A pre-planned analysis of the KATALYST-AV baseline characteristics showed that AVC varied widely across the enrolled population. Baseline data from the first 139 participants (out of 156 total enrolled) in the KATALYST-AV intent-to-treat population are consistent with the real-world findings (reported below) of markers and predictors of rapid progression, including 30% with left ventricular (LV) hypertrophy, 21% with elevated filling pressures, 14% with low flow and 26% with left atrial enlargement. Baseline AVC ranged from 320 to 1,983 Agatston units, with substantial overlap in participants with or without ventricular dysfunction.

Because calcium accrues at each patient’s own rate across this wide range, the Company amended the KATALYST-AV protocol to allow more time to assess the change in AVC with the treatment of ataciguat versus placebo in people with aortic stenosis. An interim analysis at Week 24 will be reviewed internally to evaluate rates of progression and directional mechanistic trends of ataciguat on the valve and the ventricle to further inform the design of the Phase 3 registrational trial.

Real-World Analysis of Aortic Stenosis Progression1

The Company analyzed disease progression in a retrospective, de-identified dataset of approximately 45,000 people with mild-to-moderate AS from multiple U.S. academic medical centers. The real-world analysis showed that AS progresses through both the valve and the myocardium, two areas potentially targeted by the mechanism of ataciguat. Notably, AS patients with signs of cardiac damage on routine echocardiography were more likely to progress rapidly and experience worse outcomes. This analysis was designed to identify at-risk patients for the upcoming Phase 3 registrational trial for ataciguat.

In the analysis, baseline aortic valve area (AVA) was the strongest predictor of progression. Approximately 27% of patients progressed rapidly (defined as a decline in AVA of at least 0.1 cm²/year), across both mild and moderate AS. Patients with signs of cardiac damage on routine echocardiography (LV hypertrophy, enlarged left atrium, low flow, or impaired relaxation) were more likely to progress rapidly, and rapid progression predicted higher rates of valve replacement, hospitalization and worsening heart failure.

AVC, an established predictor of adverse outcomes and a mechanistic marker of effect for ataciguat, varied widely in an exploratory analysis of the real-world data. Unlike AVA, AVC did not distinguish between rapid and normal progressors, indicating that a single baseline calcium measurement potentially reflects risk but not the pace of disease progression.

“These data add to the growing body of evidence that mild-to-moderate aortic stenosis is a continuum, with valvular disease and cardiac remodeling evolving at different rates across patients,” said Philippe Pibarot, DVM, PhD, FAHA, head of cardiology research, Université Laval. “Assessing changes in valve calcium over a longer period, alongside measures of disease progression, will provide a more complete picture of how changes in the valve relate to the course of the disease and outcomes that matter to patients.”

Additional details from the baseline data from KATALYST-AV and the real-world analysis will be presented during the Company’s webcast on Saturday, November 7.

KATALYST-AV Development Timeline

The primary endpoint in KATALYST-AV of AVC will now be assessed at Week 48, together with additional analyses evaluating change in AVA, LV mass index, stroke volume index, and E/e′. This allows for the effect of ataciguat on valve calcification to be read against the clinical course of disease. At Week 48, KATALYST-AV is powered to detect the same treatment effect assumed in the original design. At Week 24, the Company will conduct an internal interim analysis to assess the rates of progression and directional mechanistic trends of ataciguat on the valve and the ventricle. Together with the Week 48 results, these analyses will further inform the design and operational conduct of the upcoming Phase 3 registrational trial, including inclusion/exclusion criteria, endpoint selection, sample size, and follow-up duration.

The protocol amendment to the ongoing KATALYST-AV trial was made under blinded conditions without review of unblinded efficacy data and was not prompted by any safety finding. Kardigan expects to provide an update on its Phase 3 development plans in the first half of 2027 and to report Week 48 topline data in the second half of 2027.

Investor Webcast

The Company will host an investor webcast while at the American Heart Association Scientific Sessions on Saturday, November 7, 2026, at 8:00 AM ET, to provide more information on the real-world data of aortic stenosis disease progression and KATALYST-AV baseline data. Registration for the webcast is available on the Investors & Media section of the Kardigan website at https://ir.kardigan.bio/. Following the event, a replay of the presentations will be available on the website for a limited time.

About Ataciguat

Ataciguat is a once-daily, oral soluble guanylate cyclase activator being developed as a potential first-in-class treatment for AS and an alternative to the current “watchful waiting” approach and progression to valve replacement that is today’s standard of care. Ataciguat has the potential to revolutionize the management of AS as the first pharmacological therapy. Kardigan acquired rights to ataciguat, an investigational drug developed by, and in-licensed from, Sanofi and Mayo Clinic.

About Kardigan

Kardigan is a clinical-stage precision cardiovascular company developing medicines that target the root cause of specific cardiovascular diseases where no approved treatments exist. Led by a proven and experienced management team, Kardigan is reimagining cardiovascular drug discovery and development through an integrated approach that unites deep cardiovascular biology, real-world patient data, and advanced analytics to enable more precise, efficient, and informed development of novel therapies. Learn more about Kardigan and its mission at Kardigan.bio.

Cautionary Note Regarding Forward-Looking Statements

This press release contains forward-looking statements within the meaning of the Private Securities Litigation Reform Act of 1995. These forward-looking statements include express and implied statements regarding: Kardigan’s business, technology and strategy; the potential safety, efficacy and therapeutic benefits of its product candidates, including ataciguat’s potential effects on aortic valve calcification, the progression of aortic stenosis and myocardial structure and function; Kardigan’s plans and expectations concerning the design, conduct, timing, endpoints and results of its current and future clinical trials, including the ongoing KATALYST-AV Phase 2b clinical trial; the anticipated use of baseline, real-world and observational data to inform future research into aortic stenosis and future clinical development; and Kardigan’s expectations regarding its ability to advance product candidates into, and successfully complete, clinical trials and its commercialization efforts for any of its future approved products, if any.

The words “anticipate,” “believe,” “could,” “expect,” “intend,” “may,” “might,” “plan,” “potential,” “should,” “will,” “would” and similar expressions are intended to identify forward-looking statements, although not all forward-looking statements contain these identifying words.

Forward-looking statements are based on Kardigan’s current expectations and assumptions and are subject to risks and uncertainties that could cause actual results to differ materially from those expressed or implied by such statements. These risks and uncertainties include, among others: risks relating to Kardigan’s research and development activities; Kardigan’s ability to execute on its strategy, including obtaining the requisite regulatory approvals on the expected timeline, if at all; uncertainties relating to preclinical and clinical development activities; Kardigan’s dependence on third parties to conduct clinical trials, manufacture its product candidates and develop and commercialize its product candidates, if approved; Kardigan’s ability to attract, integrate and retain key personnel; risks related to Kardigan’s financial condition and need for substantial additional funds in order to complete development activities and commercialize a product candidate, if approved; risks related to regulatory developments and approval processes of the U.S. Food and Drug Administration and comparable foreign regulatory authorities; risks related to establishing and maintaining Kardigan’s intellectual property protections; and risks related to the competitive landscape for Kardigan’s product candidates; as well as additional risks described under the heading “Risk Factors” in Kardigan’s most recent Quarterly Report on Form 10-Q and in Kardigan’s subsequent filings with the Securities and Exchange Commission.

Forward-looking statements are not guarantees of future performance, and undue reliance should not be placed on them. Forward-looking statements speak only as of the date made and should not be relied upon as representing Kardigan’s views as of any subsequent date. Except as required by applicable law, Kardigan undertakes no obligation to update any forward-looking statement as a result of new information, future events or otherwise.

____________________________

1 The real-world analysis was conducted using the nference nSights platform containing de-identified real-world data from the nference Federated Healthcare Network.


Contacts

Media:
Michael Bachner

Executive Director, Corporate Communications

+1 (609) 664-7308

mbachner@kardigan.bio

Investors:
Clayton Robertson

Head of Investor Relations

investors@kardigan.bio

Alpha Cognition to Participate in Upcoming Investor Events

Alpha Cognition to Participate in Upcoming Investor Events


Alpha Cognition to Participate in Upcoming Investor Events

VANCOUVER, British Columbia & GRAPEVINE, Texas–(BUSINESS WIRE)–Alpha Cognition Inc. (Nasdaq: ACOG) (“Alpha Cognition”, or the “Company”), a biopharmaceutical company developing and commercializing novel therapeutics for debilitating neurodegenerative disorders, today announced that Michael McFadden, Chief Executive Officer, and Lauren D’Angelo, Chief Operating Officer, will participate in the upcoming investor events:


Craig-Hallum Alpha Select Conference

Format: 1×1 meetings

Date: Tuesday, November 17, 2026

Stonegate CEO Series

Format: Fireside chat
Date/Time: Thursday, December 10, 2026 at 12pm EST

For further information regarding these events, please reach out to your institutional representative.

About Alpha Cognition Inc.

Alpha Cognition Inc. is a commercial stage, biopharmaceutical company dedicated to developing treatments for patients with neurodegenerative diseases, such as Alzheimer’s disease and Cognitive Impairment with mild Traumatic Brain Injury (“mTBI”), for which there are limited or no approved treatment options.

ZUNVEYL is a patented drug approved as a new generation acetylcholinesterase inhibitor for the treatment of Alzheimer’s disease, with expected minimal gastrointestinal side effects. ZUNVEYL’s active metabolite is differentiated from donepezil and rivastigmine in that it binds neuronal nicotinic receptors, most notably the alpha-7 subtype, which is known to have a positive effect on cognition. ALPHA-1062 is also being developed in combination with memantine to treat moderate to severe Alzheimer’s dementia, and as an intranasal formulation for Cognitive Impairment with mTBI.

Contacts

For further information:

Investor Relations

LifeSci Advisors

PJ Kelleher

pkelleher@lifesciadvisors.com

African Medicines Agency Marks First Year of Operations with New Platforms to Strengthen Regulation and Support Manufacturing

African Medicines Agency

/ Key word(s): Miscellaneous

African Medicines Agency Marks First Year of Operations with New Platforms to Strengthen Regulation and Support Manufacturing

06.11.2026 / 20:15 CET/CEST

The issuer is solely responsible for the content of this announcement.


African Medicines Agency Marks First Year of Operations with New Platforms to Strengthen Regulation and Support Manufacturing

Inaugural AMA Week brings together regulators, manufacturers and partners to advance access to safe, quality-assured and effective medical products

KIGALI, Rwanda, November 6, 2026/APO Group/ — African Medicines Agency (AMA) (www.au-AMA.africa) has marked its first year of operations with the inauguration of its Technical Committees and the launch of two hubs to strengthen regulatory capacity and support medical-product innovation and manufacturing across Africa.

These milestones formed part of the inaugural AMA Week, which was held under the theme, Building Africa’s Regulatory Future Together: From National Strength to Continental Impact, from 25 to 30 October 2026 in Kigali, Rwanda. The gathering brought together National Regulatory Authorities (NRAs), Regional Economic Communities (RECs), policymakers, manufacturers, investors, researchers and global health partners to review progress and advance priorities for the Agency’s next phase of work.

AMA became operational in October 2025. Its mandate is to coordinate and strengthen Africa’s regulatory systems, helping countries draw on shared scientific expertise and reduce duplication while maintaining rigorous standards of quality, safety and efficacy.

Fragmented regulatory systems can leave gaps in oversight and delay access to medical products. Manufacturers seeking to supply several African markets may face different requirements, procedures and timelines. AMA works alongside national authorities to harmonise requirements, support joint assessments and promote reliance on trusted scientific evaluations. National authorities retain responsibility for marketing authorisation.

“Africa’s strength lies in the expertise of its national authorities. By sharing that expertise and relying on each other’s work, we can bring quality medical products to patients faster across the continent,” said H.E. Dr Delese Mimi Darko, Director-General of AMA.

Strengthening scientific expertise and regulatory capacity

On 26 October, AMA inaugurated seven Technical Committees, bringing together African experts to provide independent scientific advice across key areas of medical-product regulation. Their work will support the Agency’s collaboration with national and regional regulatory institutions.

“The Technical Committees have been entrusted with a highly important mission, which is to help build a regulatory system that is efficient, coherent, transparent and at the service of the population. Their members bring skills, experience and integrity that will be used to protect the health of Africa.” said Dr. Yossounon Chabi, Chair, AMA Governing Board.

On the third day of AMA Week, AMA launched its Capacity Strengthening Hub and Regulatory Innovation and Manufacturing Readiness Hub, both based at its headquarters in Kigali.

The Capacity Strengthening Hub will support the development of regulatory expertise across national authorities and regional bodies. The Regulatory Innovation and Manufacturing Readiness Hub will provide a platform for early engagement between regulators, researchers and manufacturers, helping developers understand regulatory expectations and generate the evidence needed to bring quality-assured products to market.

Connecting regulation, manufacturing and patient access

AMA Week opened with Manufacturing and Innovation Day on 25 October. Discussions brought manufacturers, investors, regulators, academia and patient representatives together to examine how predictable regulatory pathways, workforce development and technology transfer can support sustainable local production.

The Week also included the 11th African Medicines Regulators Conference, co-convened with the World Health Organization on 29–30 October. The conference provided a forum for African regulators and partners to advance cooperation, reliance and trust across regulatory systems.

Together, these engagements reinforced the connection between strong national authorities, effective continental coordination and a manufacturing environment capable of responding to Africa’s health needs.

As AMA enters its second year of operations, its priorities include advancing the work of its Technical Committees, implementing the two hubs’ programmes, strengthening safety monitoring and deepening collaboration with national authorities and regional bodies.

Patients remain the ultimate measure of this work: stronger protection against substandard and falsified products, better safety monitoring and more timely access to quality-assured treatments.

Further information about AMA’s work is available at https://au-AMA.africa/.

Distributed by APO Group on behalf of African Medicines Agency.

Download Images (1): https://apo-opa.co/3V4i4i2

Download Images (2): https://apo-opa.co/3WvdNob

Download Images (3): https://apo-opa.co/4z5zFnJ

Media contact:
Communications Unit
African Medicines Agency
Email: AMAComms@au-ama.africa / bizimunguj@au-ama.africa
Tel: (+250) 78 824 1588

About the African Medicines Agency:
The African Medicines Agency (AMA) is a specialised agency of the African Union, headquartered in Kigali, Rwanda. Its objective is to strengthen the capacity of States Parties and Regional Economic Communities to regulate medical products and improve access to quality, safe and effective medical products across Africa.

About AMA Week:
The inaugural AMA Week brought together African regulators, policymakers, manufacturers, researchers and partners to advance the Agency’s mandate, strengthen regulatory cooperation and support access to quality-assured medical products.


06.11.2026 CET/CEST Dissemination of a Corporate News, transmitted by EQS News – a service of EQS Group.
The issuer is solely responsible for the content of this announcement.

The EQS Distribution Services include Regulatory Announcements, Financial/Corporate News and Press Releases.


Language: English
Company: African Medicines Agency
Germany
EQS News ID: 2430376

 
End of News EQS News Service

2430376  06.11.2026 CET/CEST

BIOTECanada: Federal Budget Builds on Momentum for Canada’s Life Sciences Sector

BIOTECanada: Federal Budget Builds on Momentum for Canada’s Life Sciences Sector


BIOTECanada: Federal Budget Builds on Momentum for Canada’s Life Sciences Sector

OTTAWA, Ontario–(BUSINESS WIRE)–BIOTECanada welcomes measures in Budget 2026 that can help strengthen Canada’s life sciences sector, including expanded support for research and development, new investment incentives and additional funding for strategic sectors, including biomanufacturing.


“Expanding SR&ED eligibility to include certain regulatory approval and intellectual property costs is an important step for biotechnology companies,” said Wendy Zatylny, President and CEO of BIOTECanada. “These are significant costs companies face as they move toward commercialization, and recognizing them can help advance new treatments for patients while supporting the growth of Canadian companies.”

The budget proposes expanded SR&ED tax support, expected to provide $930 million over five years, to cover certain regulatory and intellectual property costs. It also introduces the proposed Canada Growth Incentive to encourage investment in Canadian companies along with an additional $1 billion over six years for the existing Strategic Response Fund, which identifies biomanufacturing as an area that will receive support. Alongside initial intellectual property and commercialization reforms, these measures align with several priorities BIOTECanada outlined in its 2026 Federal Budget Submission.

These new measures build on momentum from earlier announcements in 2026 including announced SR&ED expansion, the Life Sciences Venture Capital Catalyst Initiative funding, and the federal government’s Cabinet Directive on Regulatory Competitiveness and Red Tape Reduction.

The budget also acknowledges the 39 recommendations of the Pharmaceutical and Life Sciences Sector Task Force, of which BIOTECanada was a participant. With the government continuing to assess these recommendations, BIOTECanada is encouraging a clear and expedited path forward for implementation, including a coordinated Life Sciences Industrial Strategy.

“Budget 2026 represents important progress, but there is much more work to do,” added Zatylny. “The Budget recognizes the importance of timely access to critical vaccines and life-saving medicines for Canadian patients. Now we need to build on that recognition with concrete action on the recommendations of the task force, including investment and regulatory modernization, so more patients can benefit from new treatments and more Canadian companies can grow and compete here at home. Many other economies around the world are actively supporting their life sciences sectors. With each passing day, Canada risks falling further behind and losing out on important opportunities for investment and growth.”

BIOTECanada looks forward to continuing its work with the federal government as these measures move forward.

BIOTECanada is the national industry association voice. With over 200 member companies, the Association’s membership is representative of the broader Canadian biotech ecosystem which is national in scope and comprised of pre-commercial biotech companies, global pharma and biotech companies, investors, incubator and accelerator organizations, and academia. In the context of global population growth and corresponding climate change, food sustainability, and human health needs, the biotechnology industry plays a vital role in developing solutions to help address the resulting human health and environmental challenges. Visit: biotech.ca

Contacts

Nina Lewis

Communications Manager

Nina.Lewis@biotech.ca

First U.S. Convintro™ Herbicide Delivery Launches a New Weed-Control Option for Corn and Soybean Growers

First U.S. Convintro™ Herbicide Delivery Launches a New Weed-Control Option for Corn and Soybean Growers


First U.S. Convintro™ Herbicide Delivery Launches a New Weed-Control Option for Corn and Soybean Growers

New Group 12 pre-emergent herbicide brings a new active ingredient to U.S. corn and soybean growers confronting resistant waterhemp and Palmer amaranth.


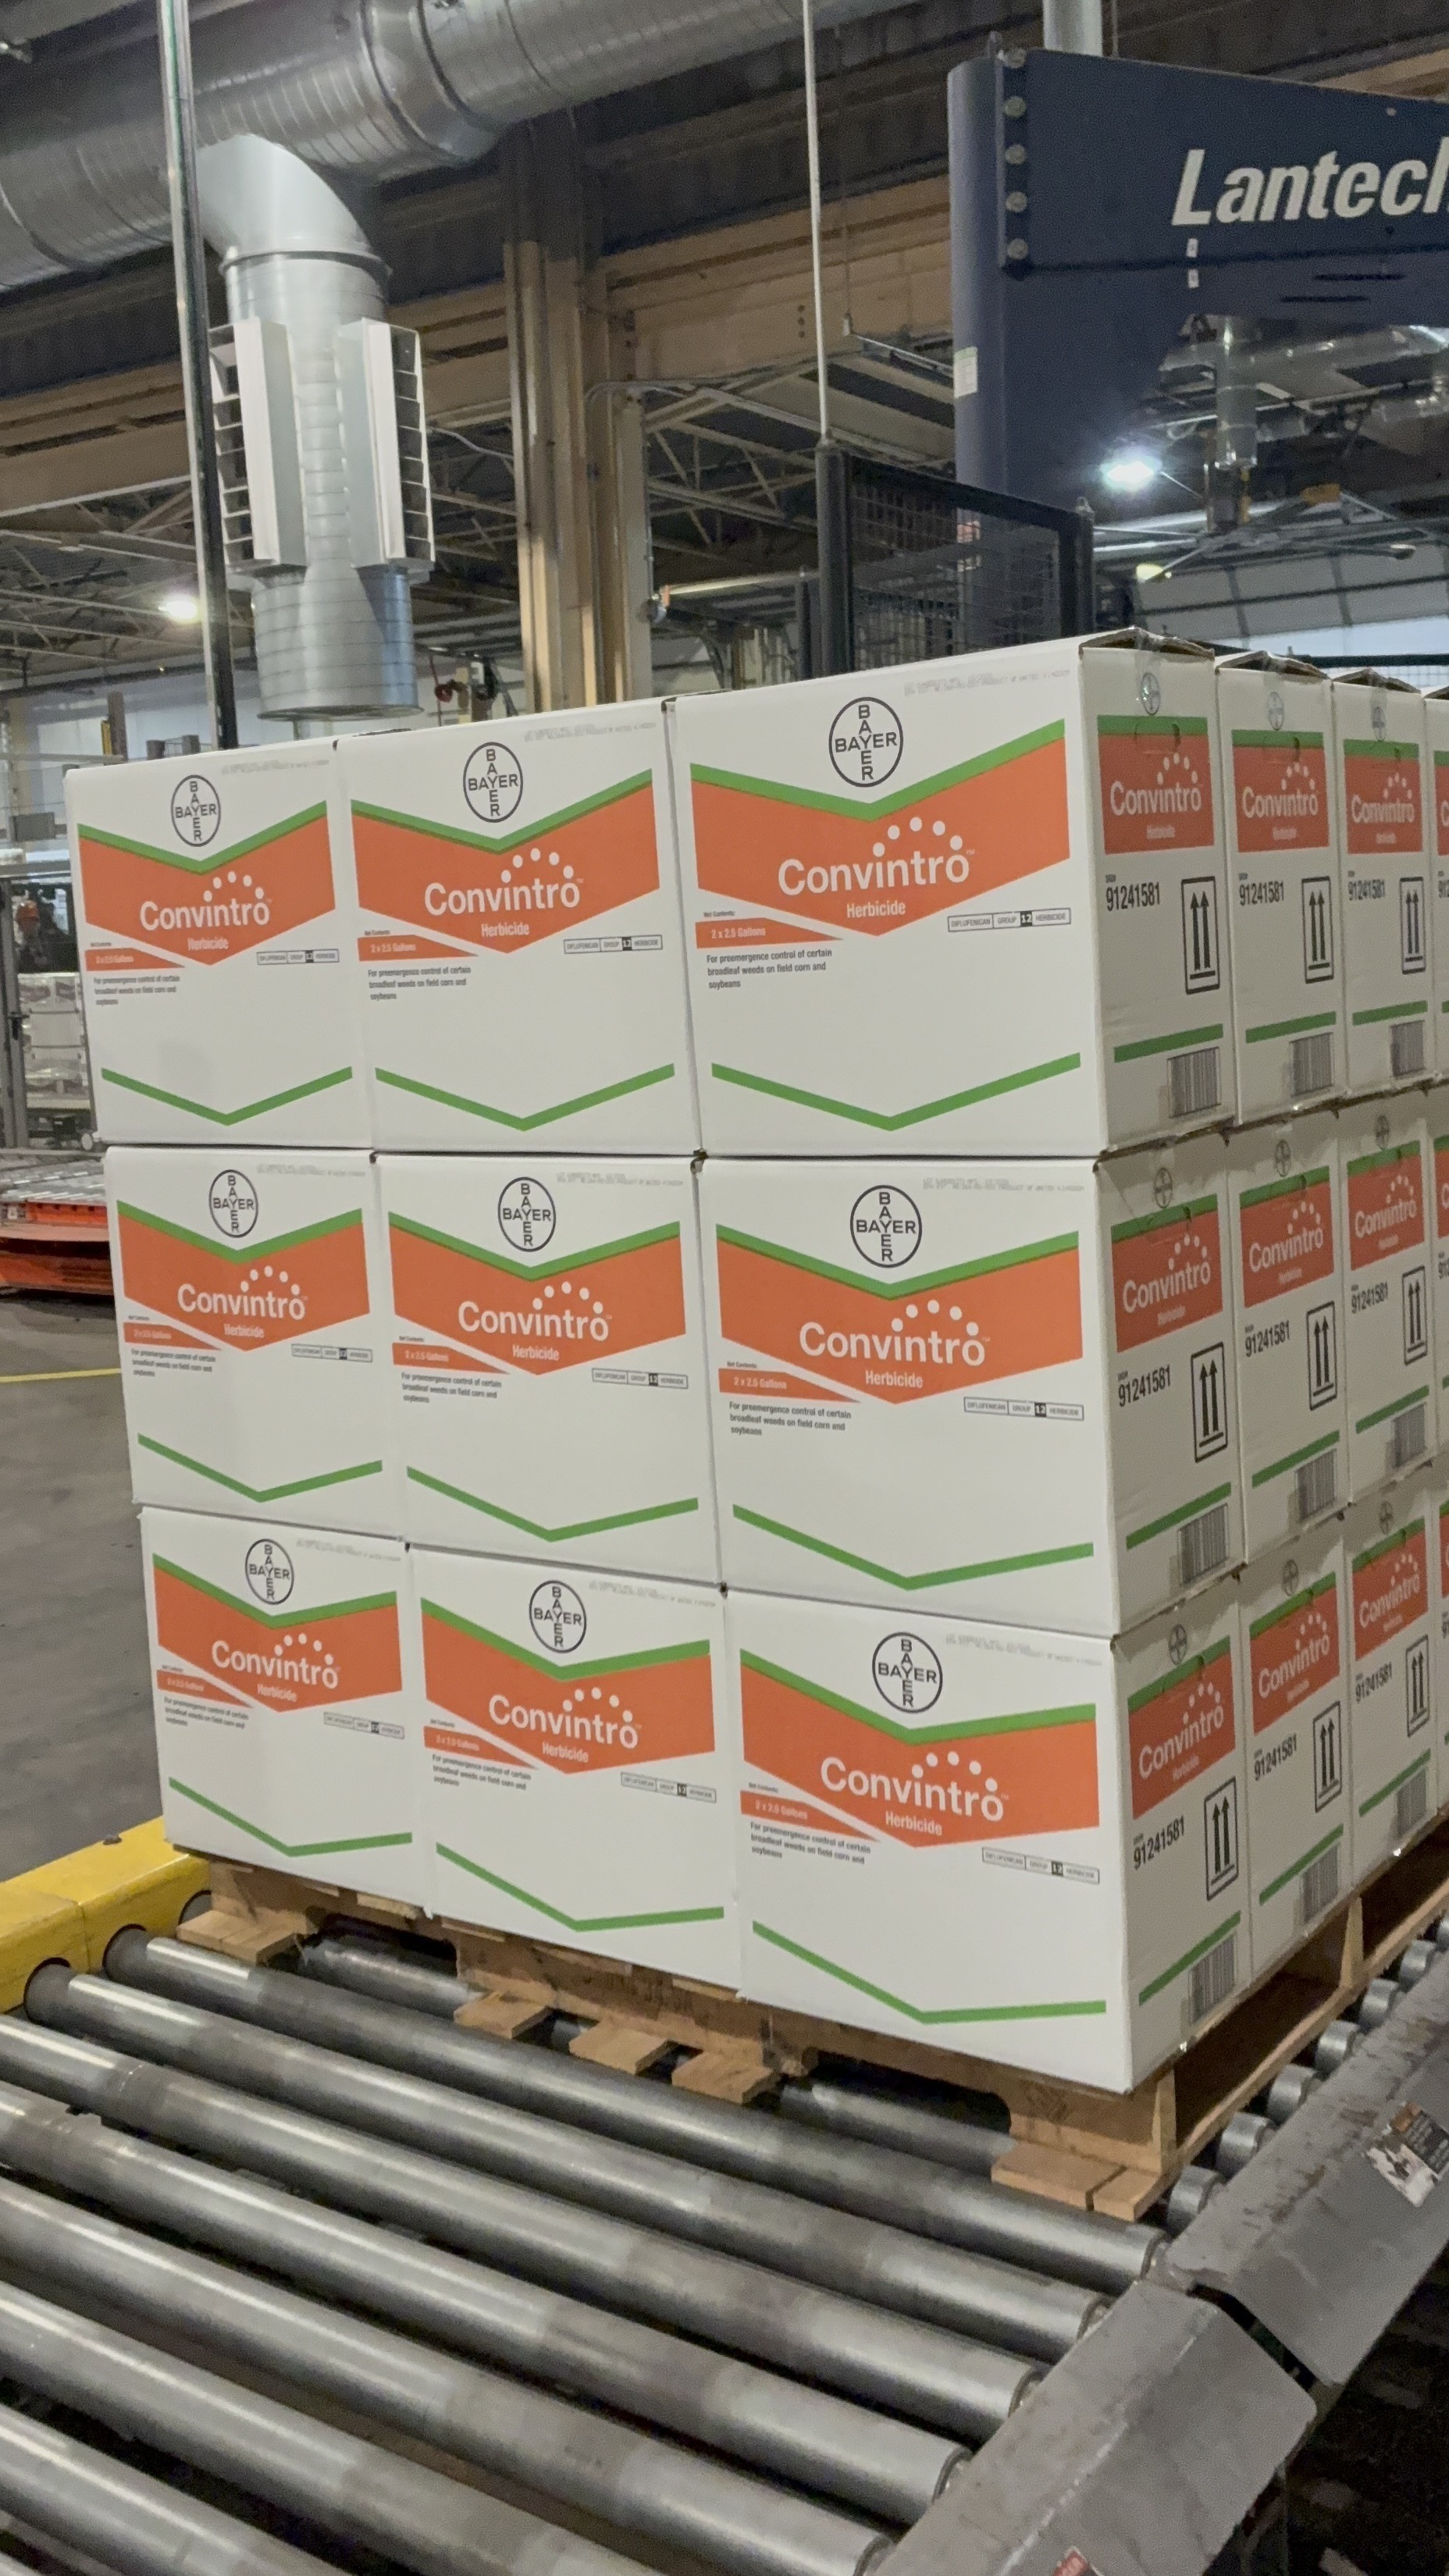
RUTHVEN, Iowa–(BUSINESS WIRE)–Bayer today marked the first U.S. delivery of Convintro™ herbicide, transported from its Kansas City manufacturing facility to New Century FS in Ruthven, Iowa. The milestone moves a new pre-emergent weed-control option closer to corn and soybean growers for the 2027 season.

Convintro is designed to help manage resistant waterhemp and Palmer amaranth – two of agriculture’s most destructive super-weeds, with populations now showing resistance to up to nine different herbicide sites of action.1

Convintro brings Group 12 chemistry to U.S. corn and soybean production. Its active ingredient, diflufenican, inhibits carotenoid biosynthesis and adds a different site of action to integrated weed management programs.

“There’s more than herbicide on this truck – there’s a new chapter in the fight against resistant weeds,” said Kenin Eubank, Bayer U.S. Crop Protection Business Lead. “Helping growers tackle their toughest challenges is what drives us forward, and we’re excited to bring them another tool that addresses an urgent grower need while strengthening their broader weed-management toolbox.”

How Herbicide-Resistant Waterhemp and Palmer Amaranth Threaten Crop Yields

These resilient weeds can spread rapidly, produce large seedbanks and reduce yield potential, making a single weed escape costly for years.

  • Waterhemp: Reported average yield reduction of 43% over three years when uncontrolled.2
  • Palmer amaranth: Soybean yield potential can be reduced up to 79% when there are 2.5 plants per foot of row.3
  • Seed Production: Without competition, each female waterhemp and Palmer amaranth plant can produce up to 1 million seeds.4
  • Seed Dispersal: Seeds from these species can be spread by harvesting and tillage equipment, wildlife, flooding, manure or compost, and gravity. Strong winds can move seeds about 1,000 feet and pollen about 1,600 feet.5

Convintro Key Benefits and Product Highlights

Palmer amaranth and waterhemp can grow up to three inches per day and become difficult to control above 3 to 4 inches. Convintro provides an early-season foundation that helps limit resistant weeds before they gain an advantage.

  • Weed Management: Targeted activity to help control early-season, pre-emergent Palmer amaranth and waterhemp in corn and soybeans.
  • New Active Ingredient: Diflufenican is a Group 12 PDS inhibitor and a new active ingredient in the United States. It can be an important part of integrated weed management programs.
  • Tank Mix Flexibility: A flexible partner for many groups of residual herbicides, helping growers customize programs for field-specific weed spectrum and resistance patterns.
  • Extended Residual: Added residual activity during the critical early season, helping reduce pressure on post-emergence programs.

On the Way for the 2027 season

With EPA federal approval secured, Convintro is well on the way to providing another tool for weed management in corn and soybean production. Visit http://www.cropscience.bayer.us/ for comprehensive strategies, expert insights, to find your local Bayer representative, and to stay updated on state approval status in your region. For more information on Convintro, visit Convintro.us.

About Bayer

Bayer is a global enterprise with core competencies in the life science fields of health care and nutrition. In line with its mission, “Health for all, Hunger for none,” the company’s products and services are designed to help people and the planet thrive by supporting efforts to master the major challenges presented by a growing and aging global population. Bayer is committed to driving sustainable development and generating a positive impact with its businesses. At the same time, the Group aims to increase its earning power and create value through innovation and growth. The Bayer brand stands for trust, reliability and quality throughout the world. In fiscal 2025, the Group employed around 88,000 people and had sales of 45.6 billion euros. R&D expenses amounted to 5.8 billion euros. For more information, go to www.bayer.com.

Sources

1.

International Survey of Herbicide Resistant Weeds. 2024. Species pages: Amaranthus palmeri; Amaranthus tuberculatus. https://www.weedscience.org/Pages/Species.aspx

2.

Hager, A.G., Wax, L.M., and Stoller, E.W. 2009. Common waterhemp (Amaranthus rudis) interference in soybean. Weed Science. 50(5): 607-610.

3.

I Will Take Action. 2024. Palmer amaranth (pigweed). https://iwilltakeaction.com/weeds/identification/palmer-amaranth-pigweed/

4.

Schwartz, L.M., Norsworthy, J.K., Young, B.G., Bradley, K.W., Kruger, G.R., Davis, V.M., Steckel, L.E., and Walsh, M.J. 2016. Tall waterhemp (Amaranthus tuberculatus) and Palmer amaranth (Amaranthus palmeri) seed production and retention at soybean maturity. Weed Technology 30:284-290.

5.

Legleiter, T. and Johnson, B. 2013. Palmer amaranth biology, identification, and management. WS-51. Purdue Weed Science. Purdue Extension. Purdue University. https://www.extension.purdue.edu/extmedia/WS/WS-51-W.pdf

ALWAYS READ AND FOLLOW PESTICIDE LABEL DIRECTIONS.

The distribution, sale, or use of an unregistered pesticide is a violation of federal and/or state law and is strictly prohibited. Check with your state department(s) of agriculture, local dealer or representative for the product registration status in your state. Not all products are registered for use in all states and may be subject to use restrictions. Bayer, Bayer Cross and Convintro™ are trademarks of Bayer Group. For additional product information call toll-free 1-866-99-BAYER (1-866-992-2937) or visit our website at www.BayerCropScience.us. Bayer CropScience LLC, 800 North Lindbergh Boulevard, St. Louis, MO 63167. ©2026 Bayer Group. All rights reserved.

Contacts

Contact for media inquiries:
Kyel Richard (573) 356-0241
Email: kyel.richard@bayer.com

New Data Show Twice-Yearly Lenacapavir Achieved Nearly Twice the One-Year Persistence of Daily Oral F/TDF in HIV Prevention

New Data Show Twice-Yearly Lenacapavir Achieved Nearly Twice the One-Year Persistence of Daily Oral F/TDF in HIV Prevention


New Data Show Twice-Yearly Lenacapavir Achieved Nearly Twice the One-Year Persistence of Daily Oral F/TDF in HIV Prevention

– Zero HIV Acquisitions Occurred Among Participants Receiving Twice-Yearly Lenacapavir During the 52-Week Study Period –

– PURPOSE 5 Study Findings Highlight the Potential of Long-Acting HIV Prevention Option to Address Persistence Challenges with Up to a Full Year of Protection After Second Dose –

FOSTER CITY, Calif.–(BUSINESS WIRE)–#GILD–Gilead Sciences, Inc. (Nasdaq: GILD) today announced results from the PURPOSE 5 trial showing significantly higher persistence—defined as consistent and continuous on-time use—among people starting twice-yearly injectable lenacapavir for HIV prevention as pre-exposure prophylaxis (PrEP), compared with those starting daily oral emtricitabine/tenofovir disoproxil fumarate (F/TDF). The trial included people in the United Kingdom (UK) and France who would benefit from PrEP. These data will be presented at HIV Glasgow 2026.
Key Highlights:

  • Twice-yearly lenacapavir achieved significantly higher one-year PrEP persistence than daily oral F/TDF, with 85.6% of participants receiving twice-yearly lenacapavir remaining persistent through Week 52 compared with 46.3% for participants taking once-daily F/TDF (p<0.0001).
  • PURPOSE 5 is the first randomized study specifically designed to compare persistence between a long-acting HIV prevention option and daily oral PrEP among individuals not currently taking PrEP.
  • Zero HIV acquisitions occurred among participants receiving twice-yearly lenacapavir during the 52-week study period.
  • Results highlight the potential of twice-yearly lenacapavir to address persistence challenges that can limit the effectiveness of HIV prevention efforts.

Despite substantial progress in HIV prevention, approximately 1.2 million people acquired HIV globally in 2025, according to UNAIDS, underscoring the need for prevention options that better support long-term use and protection. Persistence on PrEP remains one of the most important determinants of real-world prevention effectiveness. Daily oral F/TDF is currently the most commonly used oral PrEP medication globally.

“While starting on PrEP is important, persisting on PrEP is critical to preventing HIV acquisition,” said Professor Chloe Orkin, MBE, Clinical Professor of Infection and Inequities, Queen Mary University of London. “In PURPOSE 5, people in France and the United Kingdom receiving twice-yearly lenacapavir injections were much more likely to keep taking PrEP effectively compared with those receiving daily PrEP tablets. These findings suggest long-acting injections for PrEP may help address a very important barrier to sustained HIV prevention for people who need it. It is notable that the study was able to reach a high proportion of new PrEP users.”

“This study was designed specifically to evaluate one of the most important challenges in HIV prevention today: helping people remain on PrEP for as long as they need protection,” said Jared Baeten, MD, PhD, Senior Vice President, Clinical Development, Virology Therapeutic Area Head, Gilead Sciences. “These results demonstrate that twice-yearly lenacapavir has the potential to help close the gap between starting PrEP and staying on PrEP which can have an important impact on our efforts to end the HIV epidemic in Europe and around the world.”

PURPOSE 5 Results

PURPOSE 5 is an ongoing Phase 2, randomized, open-label study evaluating PrEP persistence among individuals with HIV prevention needs in the United Kingdom and France who were not taking PrEP at the time of enrollment. Participants were randomized 2:1 to receive either twice-yearly lenacapavir or daily oral F/TDF and were followed for 52 weeks.

A total of 267 participants enrolled in the study, including cisgender men (72.3%), cisgender women (7.1%) and gender-diverse individuals (20.6%). 37.8% were PrEP-naïve at baseline.

At Week 52:

  • 85.6% (154/180) of participants receiving twice-yearly lenacapavir remained persistent on PrEP
  • 46.3% (37/80) of participants receiving daily oral F/TDF remained persistent on PrEP
  • The difference of 39.3 (95% CI: 26.66% – 51.41%) was statistically significant (p<0.0001)

Zero HIV acquisitions occurred among participants receiving lenacapavir. One HIV acquisition was reported in the F/TDF group; drug-level analysis at the time of diagnosis was consistent with fewer than two doses of F/TDF per week. Participant retention across both groups remained high throughout the study, with 89.4% of participants in the lenacapavir group and 83.9% of participants in the F/TDF group attending the Week 52 study visit.

For the purpose of this study Gilead used strict protocol definitions of persistence. For lenacapavir persistence was defined as receipt of on-time LEN injection at Day 1/baseline and week 26 (+/- 2 weeks) and on-time follow-up visit at week 52. For F/TDF persistence was defined as dried blood spot (DBS) tenofovir diphosphate (TFV-DP) concentrations consistent with ≥ 4 doses/week (≥ 700 fmol/punch) at Weeks 13, 26, 39, and 52.

Safety

The safety profile observed in PURPOSE 5 was generally consistent with previous studies of lenacapavir for HIV prevention. Serious adverse events were uncommon and occurred in 3.9% of participants receiving lenacapavir and 3.4% of participants receiving F/TDF. Investigators determined that no serious adverse events were related to study treatment. No participants discontinued treatment due to adverse events with lenacapavir. The most common treatment-related events were mild or moderate injection-site reactions (ISRs) associated with subcutaneous lenacapavir administration, and included nodules (67.8%), pain (60.0%), induration (20.6%), and erythema (17.8%).

Addressing a Critical HIV Prevention Challenge

Although awareness and availability of PrEP have increased, persistence remains a challenge across many settings. Daily oral PrEP is highly effective when taken as prescribed, but real-world studies have consistently shown that many individuals discontinue PrEP within the first year of use. By reducing dosing frequency to one dose every six months, twice-yearly lenacapavir has the potential to offer a discreet and convenient HIV prevention option that may better align with the needs and preferences of some individuals more vulnerable to HIV acquisition.

About PURPOSE 5

PURPOSE 5 is the first randomized study designed specifically to compare persistence between a long-acting injectable HIV prevention option and daily oral PrEP among individuals who would benefit from PrEP and were not currently taking oral PrEP. The Phase 2 study conducted in the United Kingdom and France evaluates whether twice-yearly lenacapavir can improve persistence compared with daily oral F/TDF over 52 weeks.

PURPOSE 5 is part of Gilead’s broader PURPOSE clinical development program evaluating lenacapavir across diverse populations and settings with the goal of expanding HIV prevention choice through long-acting prevention options.

About the PURPOSE Program

Gilead’s landmark PURPOSE program is the most comprehensive and diverse HIV prevention trial program ever conducted. The program comprises five HIV prevention trials around the world that are focused on innovation in science, trial design, community engagement, and health equity.

The PURPOSE 1 and PURPOSE 2 trials evaluated the safety and efficacy of twice-yearly injectable lenacapavir to reduce the chance of getting HIV. The ongoing Phase 2 and 3 program, consisting of PURPOSE 1-5 studies, is assessing the potential of lenacapavir to help a diverse range of people around the world who could benefit from PrEP.

More information about the PURPOSE program, including individual trial descriptions, populations and locations, can be found at www.purposestudies.com.

About Lenacapavir

Twice-yearly lenacapavir is approved in multiple countries as pre-exposure prophylaxis (PrEP) to reduce the risk of sexually acquired HIV in adults and adolescents who are at risk of HIV acquisition. Lenacapavir is also approved in multiple countries for the treatment of multi-drug-resistant HIV in adults, in combination with other antiretrovirals.

The multi-stage mechanism of action of lenacapavir is distinguishable from other currently approved classes of antiviral agents. While most antivirals act on just one stage of viral replication, lenacapavir is designed to inhibit HIV at multiple stages of its lifecycle and has no known cross resistance exhibited in vitro to other existing drug classes.

Lenacapavir is being evaluated as a long-acting option in multiple ongoing and planned early and late-stage clinical studies in Gilead’s HIV prevention and treatment research program, including a once-yearly injection for HIV prevention. Lenacapavir is being developed as a foundation for potential future HIV therapies with the goal of offering both long-acting oral and injectable options with several dosing frequencies, in combination or as a mono agent, that help address individual needs and preferences of people and communities affected by HIV. The journal Science named lenacapavir its 2024 “Breakthrough of the Year.”

Indication for Yeytuo® (lenacapavir)

Yeytuo injection is indicated in combination with safer sex practices for pre-exposure prophylaxis (PrEP) to reduce the risk of sexually acquired HIV-1 infection in adults and adolescents with increased HIV-1 acquisition risk weighing at least 35kg.

Indication for Yeztugo® (lenacapavir)

Yeztugo is indicated for pre-exposure prophylaxis (PrEP) to reduce the risk of sexually acquired HIV-1 in adults and adolescents (≥35kg) who are at risk for HIV-1 acquisition. Individuals must have a negative HIV-1 test prior to initiating Yeztugo.

Important Safety Information for Yeztugo

BOXED WARNING: RISK OF DRUG RESISTANCE WITH USE OF YEZTUGO IN UNDIAGNOSED HIV-1 INFECTION

  • Individuals must be tested for HIV-1 infection prior to initiating Yeztugo, and with each subsequent injection of Yeztugo, using a test approved or cleared by the FDA for the diagnosis of acute or primary HIV-1 infection. Drug-resistant HIV-1 variants have been identified with use of Yeztugo by individuals with undiagnosed HIV-1 infection. Do not initiate Yeztugo unless negative infection status is confirmed. Individuals who acquire HIV-1 while receiving Yeztugo must transition to a complete HIV-1 treatment regimen.

Contraindications

  • Yeztugo is contraindicated in individuals with unknown or positive HIV-1 status.

Warnings and precautions

  • Comprehensive risk management:

    • Use Yeztugo to reduce the risk of HIV-1 acquisition as part of a comprehensive prevention strategy including adherence to the administration schedule and safer sex practices, including condoms, to reduce the risk of sexually transmitted infections (STIs).
    • HIV-1 acquisition risk includes behavioral, biological, or epidemiologic factors including, but not limited to, condomless sex, past or present STIs, self-identified HIV risk, having sexual partners of unknown HIV-1 viremic status, or sexual activity in a high-prevalence area or network. Counsel individuals on the use of other prevention methods to help reduce their risk.
    • Use Yeztugo only in individuals confirmed to be HIV-1 negative. Evaluate for current or recent signs or symptoms consistent with HIV-1 infection. Confirm HIV-1 negative status prior to initiating, prior to each subsequent injection, and as clinically appropriate.
  • Potential risk of resistance:

    • There is a potential risk of developing resistance to Yeztugo if an individual acquires HIV-1 before or when receiving Yeztugo, or following discontinuation. HIV-1 resistance substitutions may emerge in individuals with undiagnosed HIV-1 infection taking only Yeztugo, because Yeztugo alone is not a complete regimen for HIV-1 treatment.
    • To minimize this risk, it is essential to test before each injection and additionally as clinically appropriate. Individuals confirmed to have HIV-1 must immediately begin a complete HIV-1 treatment regimen.
    • Alternative forms of PrEP should be considered after discontinuation of Yeztugo for those who are at continuing risk of HIV-1 acquisition and should be initiated within 28 weeks of the last Yeztugo injection.
  • Long-acting properties and potential associated risks:

    • Residual concentrations of Yeztugo may remain in systemic circulation for up to 12 months or longer after the last injection.
    • Select individuals who agree to the required injection dosing schedule because nonadherence or missed doses could lead to HIV-1 acquisition and development of resistance.
  • Serious injection site reactions: Improper administration (intradermal injection) has been associated with serious injection site reactions, including necrosis and ulcer. Only administer Yeztugo subcutaneously.

Adverse reactions

  • Most common adverse reactions (≥5%) in Yeztugo clinical trials were injection site reactions, headache, and nausea.

Drug interactions

  • Strong or moderate CYP3A inducers may significantly decrease Yeztugo concentrations. Dosage modifications are recommended when initiating these inducers.
  • It is not recommended to use Yeztugo with combined P-gp, UGT1A1, and strong CYP3A inhibitors.
  • Coadministration of Yeztugo with sensitive substrates of CYP3A or P-gp may increase their concentrations and result in the increased risk of their adverse events. Yeztugo may increase the exposure of drugs primarily metabolized by CYP3A initiated within 9 months after the last injection of Yeztugo.

Dosage and administration

  • HIV screening: Test for HIV-1 infection prior to initiating, prior to each subsequent injection, and as clinically appropriate using an approved or cleared test for the diagnosis of acute or primary HIV-1 infection.
  • Dosage: Initiation dosing (injections and tablets) followed by once-every-6-months continuation injection dosing. Tablets may be taken with or without food.

    • Initiation: Day 1: 927 mg by subcutaneous injection (2 x 1.5-mL injections) and 600 mg orally (2 x 300-mg tablets). Day 2: 600 mg orally.
    • Continuation: 927 mg by subcutaneous injection every 6 months (26 weeks) from date of last injection ±2 weeks.
  • Anticipated delayed injections: If scheduled 6-month injection is anticipated to be delayed by more than 2 weeks, Yeztugo tablets may be taken on an interim basis (for up to 6 months) until injections resume. Dosage is 300 mg orally (1 x 300-mg tablet) once every 7 days. Resume continuation injections within 7 days of the last oral dose.
  • Missed injections: If more than 28 weeks have elapsed since the last injection and Yeztugo tablets have not been taken, restart with initiation dosing if clinically appropriate.
  • Dosage modifications of Yeztugo are recommended when initiating with strong or moderate CYP3A inducers. Consult the full Prescribing Information for recommendations.

About Gilead HIV

For more than 35 years, Gilead has been a leading innovator in the field of HIV, driving advances in treatment, prevention and cure research. Gilead researchers have developed 14 HIV medications, including the first single-tablet regimen to treat HIV, the first antiretroviral for pre-exposure prophylaxis (PrEP) to help reduce new HIV infections, and the first long-acting injectable HIV treatment medication administered twice-yearly. Our advances in medical research have helped to transform HIV into a treatable, preventable, chronic condition for millions of people.

Gilead is committed to continued scientific innovation to provide solutions for the evolving needs of people affected by HIV around the world. Through partnerships, collaborations and charitable giving, the company also aims to improve education, expand access and address barriers to care, with the goal of ending the HIV epidemic worldwide. Gilead has been repeatedly recognized as one of the top two leading philanthropic funders of HIV-related programs in a report released by Funders Concerned About AIDS.

Discover more about Gilead’s unique collaborations worldwide and the work to help end the HIV epidemic.

About Gilead Sciences

Gilead Sciences, Inc. is a biopharmaceutical company that has pursued and achieved breakthroughs in medicine for more than three decades, with the goal of creating a healthier world for all people. The company is committed to advancing innovative medicines to prevent and treat life-threatening diseases, including HIV, viral hepatitis, COVID-19, cancer and inflammation. In 2025, Gilead announced a $32 billion investment to further strengthen its U.S. footprint to power the next era of discovery, job creation and public health preparedness – while continuing to invest globally to ensure patients everywhere benefit from its scientific innovation. Gilead operates in more than 35 countries worldwide, with headquarters in Foster City, Calif.

Forward-Looking Statements

This press release includes forward-looking statements within the meaning of the Private Securities Litigation Reform Act of 1995 that are subject to risks, uncertainties and other factors, including Gilead’s ability to initiate, progress and complete clinical trials in the anticipated timelines or at all, and the possibility of unfavorable results from ongoing and additional clinical trials, including those involving lenacapavir (such as PURPOSE 5); uncertainties relating to regulatory applications and related filing and approval timelines, including regulatory applications for lenacapavir for PrEP, and the risk that any regulatory approvals, if granted, may be subject to significant limitations on use or subject to withdrawal or other adverse actions by the applicable regulatory authority; the possibility that Gilead may make a strategic decision to discontinue development of lenacapavir for indications currently under evaluation and, as a result, lenacapavir may never be successfully commercialized for such indications; and any assumptions underlying any of the foregoing. These and other risks, uncertainties and factors are described in detail in Gilead’s Quarterly Report on Form 10-Q for the quarter ended September 30, 2026, as filed with the U.S. Securities and Exchange Commission. These risks, uncertainties and other factors could cause actual results to differ materially from those referred to in the forward-looking statements. All statements other than statements of historical fact are statements that could be deemed forward-looking statements. The reader is cautioned that any such forward-looking statements are not guarantees of future performance and involve risks and uncertainties, and is cautioned not to place undue reliance on these forward-looking statements. All forward-looking statements are based on information currently available to Gilead, and Gilead assumes no obligation and disclaims any intent to update any such forward-looking statements.

U.S. full Prescribing Information for Truvada (F/TDF) and Yeztugo, including Boxed Warning, are available at www.gilead.com.

Gilead and the Gilead logo, Truvada, Yeztugo and Yeytugo are registered trademarks of Gilead Sciences, Inc., or its related companies.

For more information about Gilead, please visit the company’s website at www.gilead.com, follow Gilead on X/Twitter (@Gilead Sciences) and LinkedIn (@Gilead-Sciences).

Contacts

Priscilla White, Media

public_affairs@gilead.com

Jacquie Ross, Investors

investor_relations@gilead.com
American College of Rheumatology and OpenEvidence Collaborate to Expand Access to Trusted Patient Education Resources

American College of Rheumatology and OpenEvidence Collaborate to Expand Access to Trusted Patient Education Resources


American College of Rheumatology and OpenEvidence Collaborate to Expand Access to Trusted Patient Education Resources

Collaboration brings English- and Spanish-language disease and treatment fact sheets to clinicians caring for people with rheumatic conditions


MIAMI & ORLANDO, Fla.–(BUSINESS WIRE)–#ClinicalAI–The American College of Rheumatology (ACR), the professional organization representing more than 10,600 rheumatology professionals worldwide, and OpenEvidence, the most widely used medical AI platform among U.S. physicians, today announced a collaboration at ACR Convergence 2026 to make ACR’s patient education resources available within OpenEvidence, helping rheumatologists, as well as primary care physicians and other specialists managing rheumatic conditions, share authoritative, evidence-based information with patients and caregivers at the point of care.

The collaboration brings ACR’s disease, condition, and treatment fact sheets, in both English and Spanish, into OpenEvidence, where they surface automatically when clinically relevant, alongside peer-reviewed literature, with a direct link back to the source. Clinicians can use them to connect patients and caregivers with accessible information that reinforces conversations with their healthcare teams.

Many rheumatic diseases are chronic and complex, and patients and caregivers often manage them for years. More than 53 million U.S. adults have been diagnosed with arthritis, and many people with rheumatic conditions are cared for by primary care physicians and other specialists as well as by rheumatologists. Clear, consistent educational materials help patients and caregivers understand a diagnosis, follow a treatment plan, and know what to ask at the next appointment.

“Patients and caregivers need information they can understand, trust, and return to after an appointment,” said William Harvey, MD, MSc, FACR, president of the American College of Rheumatology. “Our collaboration with OpenEvidence helps clinicians connect people with resources that reinforce important conversations about rheumatic diseases and their treatments. The goal is to help patients feel better informed and more prepared to participate in their care.”

ACR’s patient fact sheets explain rheumatic conditions in accessible language, covering causes, common signs and symptoms, diagnosis, and treatment. They also identify topics patients and caregivers can discuss with healthcare providers, helping them prepare questions about a diagnosis or treatment plan.

The collection includes disease-specific information about conditions such as gout and psoriatic arthritis, alongside treatment information on topics such as nonsteroidal anti-inflammatory drugs (NSAIDs) and biologic medications. By bringing disease and treatment education together, the resources help clinicians address questions that arise throughout a patient’s care.

“Rheumatic diseases are lifelong for many patients, and much of how they fare depends on how well they understand their condition and their treatment between visits,” said Vishal Patel, MD, Rheumatologist and VP, Clinical Partnerships & AI Education. “The ACR sets the standard for rheumatology care, and its patient fact sheets carry that same authority. Bringing them into OpenEvidence means any clinician caring for a patient with a rheumatic disease, wherever they practice, can share trusted information in English or Spanish at the point of care.”

Through this collaboration, ACR and OpenEvidence aim to strengthen the connection between evidence-based information, patient education, and informed participation in care.

About the American College of Rheumatology

Founded in 1934, the American College of Rheumatology (ACR) is a not-for-profit, professional association committed to advancing the specialty of rheumatology that serves over 10,600 physicians, health professionals, researchers and scientists worldwide. In doing so, the ACR offers education, research, advocacy and practice management support to help its members continue their innovative work and provide quality patient care. Rheumatology professionals are experts in the diagnosis, management and treatment of more than 100 different types of arthritis and rheumatic diseases.

About OpenEvidence

OpenEvidence is the most popular AI for doctors in the United States. According to independent studies, more American physicians use OpenEvidence than all other AI platforms combined. OpenEvidence is trusted by millions of verified U.S. healthcare professionals making high-stakes clinical decisions at the point of care, with evidence that is sourced, cited, and grounded in peer-reviewed medical literature. Founded with the mission to organize and expand the world’s collective medical knowledge, OpenEvidence is now used daily, on average, by the majority of physicians in the United States. OpenEvidence is free for all verified U.S. clinicians. Learn more at openevidence.com.

Contacts

OpenEvidence: media@openevidence.com

American College of Rheumatology: Teri Arnold, tarnold@rheumatology.org / 757-272-7002

DeciBio Launches Market Intelligence Portal With DeciChat, an AI Assistant Powered by DeciBio’s Proprietary Precision Medicine Research

DeciBio Launches Market Intelligence Portal With DeciChat, an AI Assistant Powered by DeciBio’s Proprietary Precision Medicine Research


DeciBio Launches Market Intelligence Portal With DeciChat, an AI Assistant Powered by DeciBio’s Proprietary Precision Medicine Research

DeciChat synthesizes thousands of data points across primary data sources to deliver deep insights into precision medicine markets instantly.


LOS ANGELES–(BUSINESS WIRE)–#ai–DeciBio Consulting (“DeciBio”), a precision medicine-focused strategy consulting and market intelligence firm, launched the DeciBio Market Intelligence Portal. The portal brings DeciBio’s Market Reports, DxBooks lab survey databases, Oncology Genomics Trackers, and Transcript Database into one subscription platform. It also introduces DeciChat, an AI assistant that performs cross-product querying and synthesis. Life science tools, diagnostics, and biopharma teams can ask a question and receive an answer drawn from across DeciBio’s research, with citations to the source data.

“Precision medicine is moving faster than traditional market research can track. And decisions that once took months of analysis now need to be made in weeks,” said Rebecca Burnham, PhD, Head of DeciBio’s Market Reports Division. “The Market Intelligence Portal and DeciChat give our clients direct access to the primary research that drives those decisions, in a form they can act on right away.”

DeciChat: One Question, Every Source

DeciChat helps teams get evidence-backed answers to the market questions behind their most important decisions, from sizing an opportunity to prioritizing markets or evaluating partners. Its AI agents search proprietary data from DeciBio’s precision medicine experts, then synthesize the findings into a single response. Each answer links back to the underlying report, dataset, or transcript, so teams can verify the numbers and insights and build on them in their own analyses.

“General-purpose AI tools are only as good as what they can find, and most precision medicine market data isn’t public,” said Andrew Stiegler, PhD, Head of DeciBio’s DxBooks Division. “DeciChat’s agents work directly on our private survey data, forecasts, and interview transcripts. Automated pipelines mean every new DxBook release, tracker update, and transcript is searchable as soon as the research is completed.”

What’s Included in the Market Intelligence Portal

The Market Intelligence Portal integrates DeciChat and all of DeciBio’s proprietary data products, including:

  • Global Market Reports: Comprehensive market assessments with market sizing, segmentation, and 5-year forecasts, plus competitive analysis and assessments of key players, technologies, drivers, and moderators. Report topics span life science tools and diagnostics, including Digital & Computational Pathology, Oncology Liquid Biopsy, Next-Generation Sequencing, Spatial Biology, Single Cell Analysis, Proteomics Tools, Clinical Oncology Biomarker Testing, Blood-Based Testing in Neurodegenerative Diseases, Bioprocessing, Cell Therapy R&D and Manufacturing, Pharma Real-World Data & Evidence, and more.
  • Snapshot Reports: Qualitative deep dives on emerging sub-segments, such as Pharma CDx Development & Commercialization, Biopharma’s Use of AI in Drug Discovery, and Precision Medicine Opportunities in Women’s Health.
  • DxBooks: Databases built from annual surveys of clinical labs:

    • NGS DxBook: Panel testing, sample types, volumes, and reimbursement across 275 US and EU labs.
    • Infectious Disease DxBook: Testing settings, assays, and vendor selection for 30 analytes across 1,000 US clinical labs.
    • Digital Pathology DxBook: Scanner and IMS usage, algorithm integration, and vendor satisfaction across 400 US and EU labs.
  • Oncology Genomics Trackers: Interactive dashboards tracking oncology genomic testing across 100 US labs (quarterly) and EU labs (semiannually), characterizing the use of genomic profiling, MRD / recurrence monitoring, and treatment response monitoring.
  • Transcript Database: 450+ searchable expert interviews with stakeholders across large and small biopharma, academic medical centers, community hospitals, reference labs, CROs, and diagnostics companies.

New editions, data releases, and transcripts are added to the portal as they are published.

Availability

The DeciBio Market Intelligence Portal is available now through enterprise subscriptions to the full DeciBio Knowledgebase, custom packages built around an organization’s priority topics, or individual product subscriptions, with DeciChat available on any plan. Existing DxBooks and Oncology Genomics Trackers subscribers will be transitioned to the portal at no extra cost. The DeciBio Market Intelligence Portal is available now. Existing DxBook and Oncology Genomics Trackers subscribers will be moved to the Portal at no extra cost. To request a demo, visit www.decibio.com/intelligence-portal or contact rebecca.burnham@decibio.com.

About DeciBio

DeciBio Consulting is a precision medicine-focused strategy consulting and market intelligence firm whose mission is to accelerate the adoption of precision medicine. DeciBio helps leadership teams across therapeutics, diagnostics, research tools, and advanced manufacturing make strategy decisions grounded in scientific, technical, and execution reality. Its data division develops market intelligence software, databases, and reports covering life science research tools and diagnostics. DeciBio serves clients ranging from startups to Fortune 500 companies from offices in Los Angeles, New York City, Oakland, and London. Learn more at decibio.com.

Contacts

Media Contact
Rebecca Burnham

rebecca.burnham@decibio.com
310.451.4510