World-First Achievement: SUSMED’s Blockchain-Based Clinical Trial System Supports Successful Marketing Authorization for Pharmaceutical Product

World-First Achievement: SUSMED’s Blockchain-Based Clinical Trial System Supports Successful Marketing Authorization for Pharmaceutical Product


World-First Achievement: SUSMED’s Blockchain-Based Clinical Trial System Supports Successful Marketing Authorization for Pharmaceutical Product

– Pioneering Solutions for Drug Lag and Loss –

TOKYO–(BUSINESS WIRE)–SUSMED, Inc. (TOKYO:4263) announced that “SUSMED SourceDataSync®” (“SUSMED SDS”), its proprietary blockchain-based clinical trial system, has contributed to obtaining marketing authorization for a pharmaceutical product1,2,3.

This marks the world’s first pharmaceutical approval based on the results of an industry-sponsored clinical trial utilizing blockchain technology4.


1: https://newsroom.viatris.com/2026-09-16-Viatris-Announces-Approval-of-WAKIX-R-Pitolisant-in-Japan,-a-First-in-Class-Treatment-for-Narcolepsy-and-Excessive-Daytime-Sleepiness-Associated-With-Obstructive-Sleep-Apnea-Syndrome

2 :“Aculys Pharma Commences Phase 3 Clinical Trial of Pitolisant in Japan Utilizing SUSMED’s Blockchain-Based Clinical Trial System”

https://susmed.co.jp/wp-content/uploads/2022/11/aw5ser6xc7tv8y9uboni870.pdf

3: “Aculys Pharma Commences Phase 3 Clinical Trial of Pitolisant in Japan for Excessive Daytime Sleepiness Associated with Obstructive Sleep Apnea, Utilizing SUSMED’s Blockchain-Based Clinical Trial System”

https://susmed.co.jp/wp-content/uploads/2023/01/roeadxfgfuihoi34w5e6r8789tfyc.pdf

4: Based on SUSMED’s internal research of medical databases, clinical trial registries, and research tools, including PubMed (https://pubmed.ncbi.nlm.nih.gov/), ClinicalTrials.gov (https://www.clinicaltrials.gov/) managed by the U.S. National Library of Medicine at the National Institutes of Health (NIH), and the EU Clinical Trials Register (https://www.clinicaltrialsregister.eu/ctr-search/search) managed by the European Medicines Agency (EMA).

Challenges in Pharmaceutical Development

Global research and development (R&D) expenditure for pharmaceuticals continues to rise, increasing at an annual rate of approximately 8.5% over the past decade5. Key drivers of the rise in R&D expenditure include advancement of R&D and the increasing complexity of protocols. In Japan, both labor-intensive and over-engineering practices intended to comply with the Good Clinical Practice(GCP) ordinance have been widely highlighted as major contributing factors. It also results in the fact that the number of clinical trial sites managed per Clinical Research Associate (CRA) in Japan is only one-third of that in the United States6. Consequently, rising development costs lead to serious drug lag and drug loss, which in turn is posing significant challenges to social and economic securities.

In recent years, the updated international guideline for Good Clinical Practice, ICH-E6(R3), outlined the Quality by Design (QbD) approach. The guideline states that the sponsor should avoid unnecessary complexity and not place unnecessary burden on participants and investigators. Implementing an appropriate system to manage quality throughout all stages of the trial process is also required.

5: Congressional Budget Office: “Research and Development in the Pharmaceutical Industry”, April 2021

6: “The Day Japan Is Excluded from Global Clinical Trials: Mindset and Behavioral Reform to Avoid the Worst-Case Scenario” (PhRMA / EFPIA Japan Joint Seminar at the 18th Conference on CRCs and Clinical Trials 2018 in Toyama).

Our Solution and Initiatives

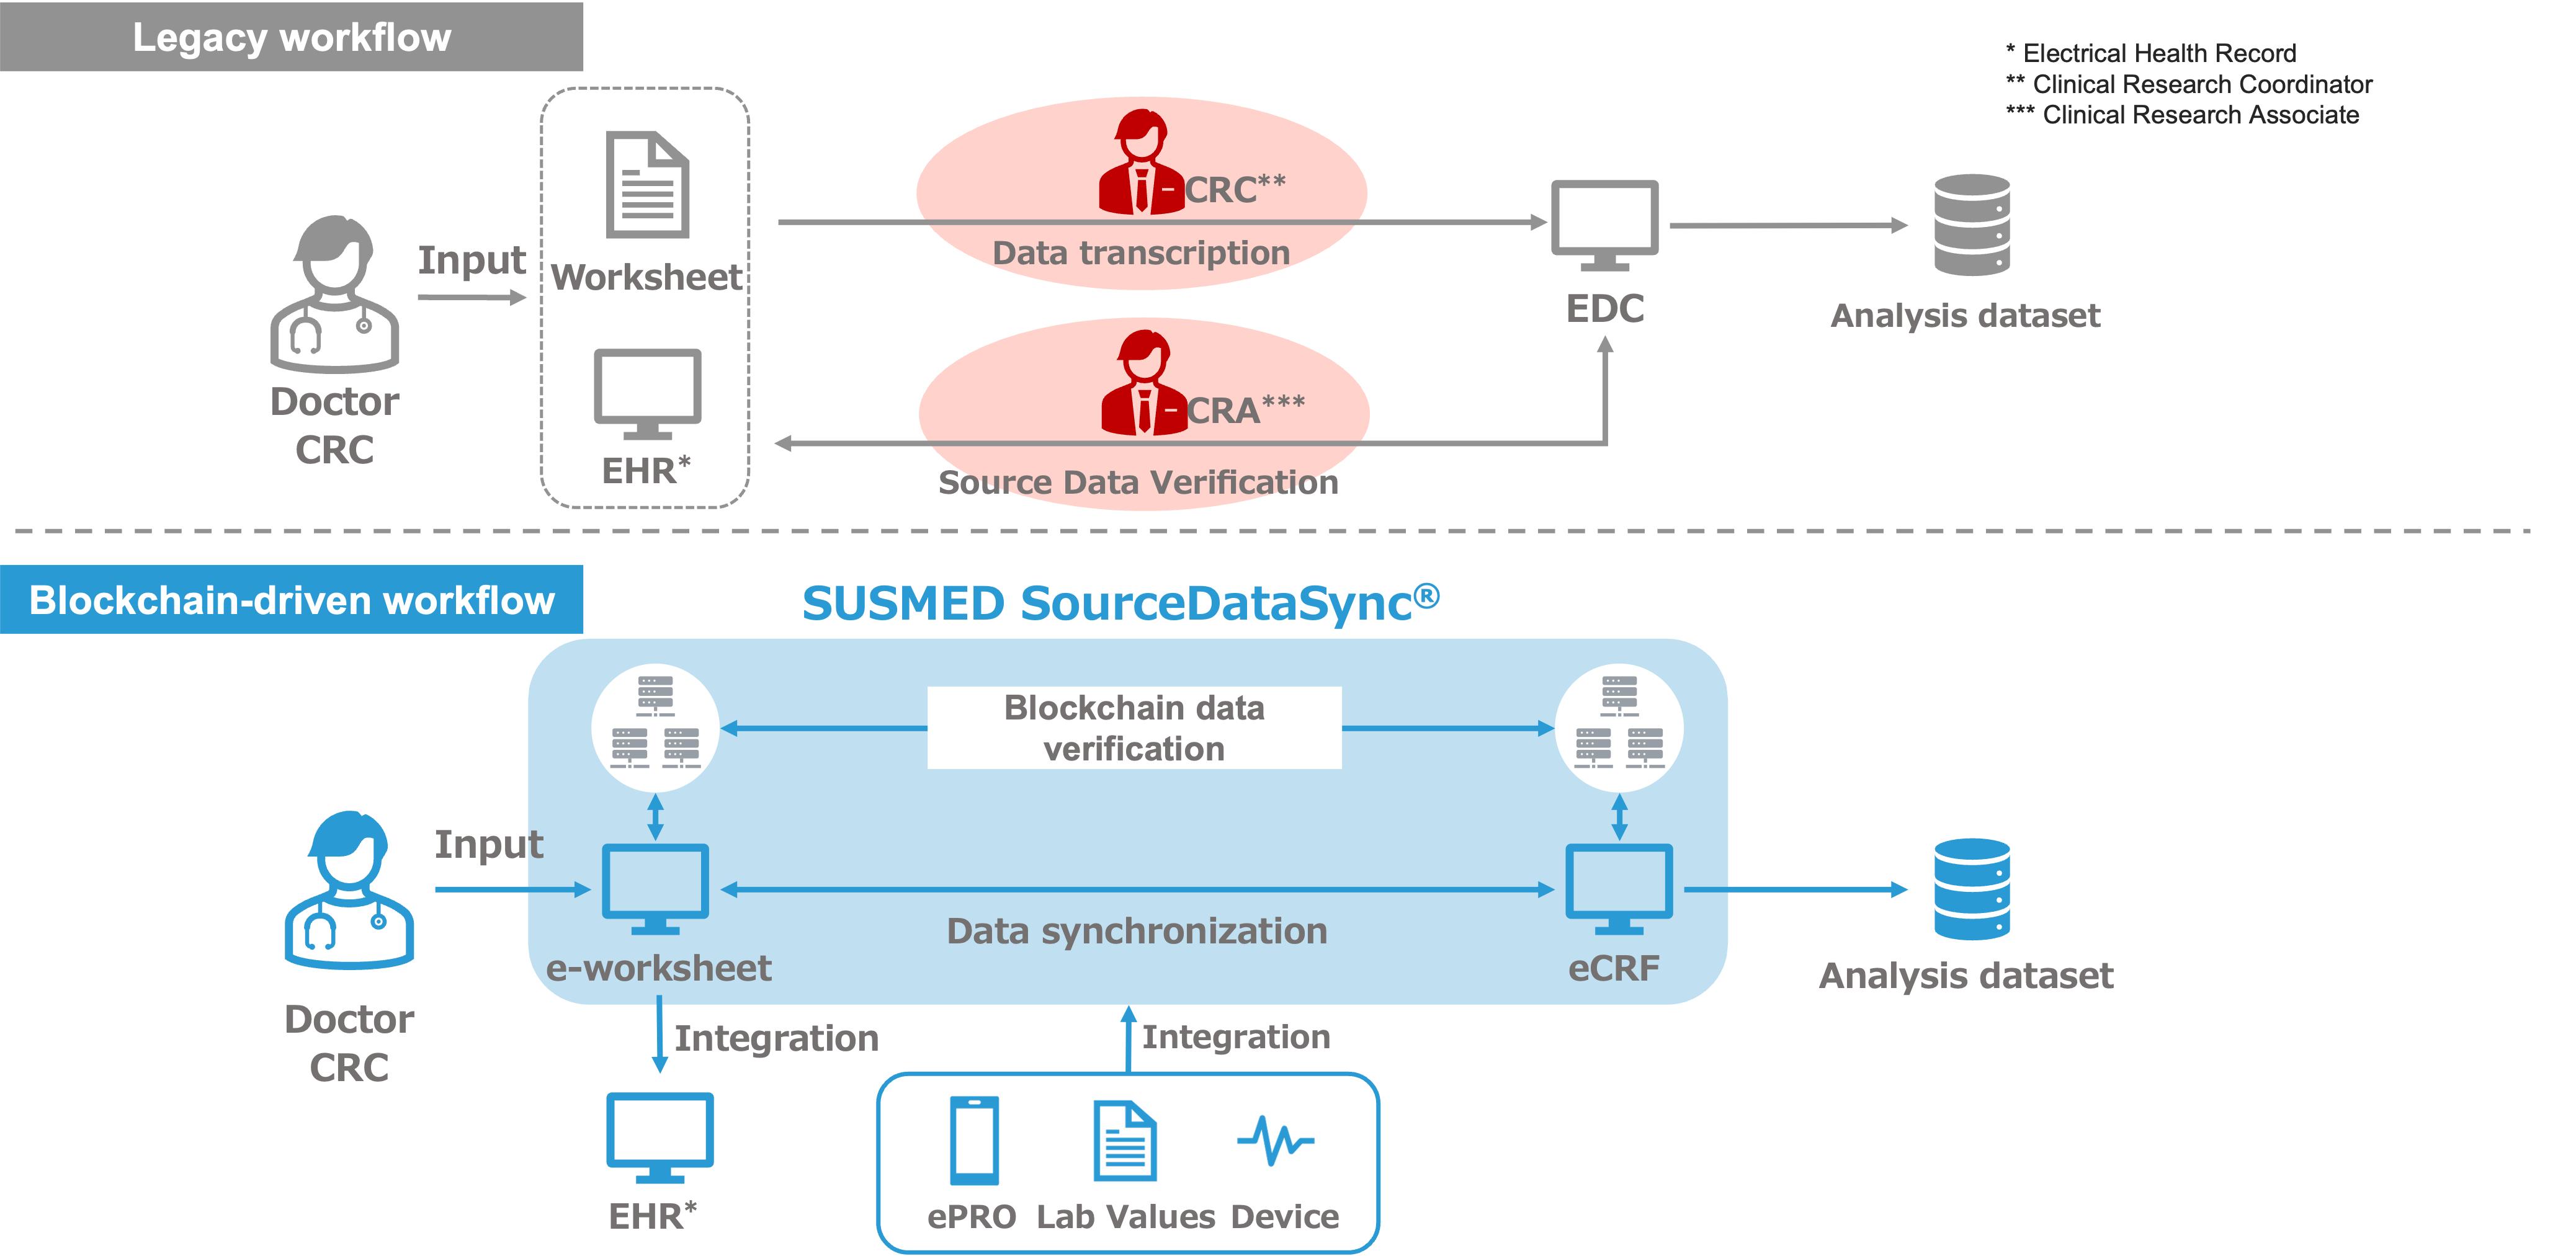
SUSMED SDS is a clinical trial management system that utilizes blockchain technology to securely syncronize medical data captured at medical institutions with Case Report Form (CRF) data. This enables a significant reduction in both data entry workload at medical sites and regulatory-mandated monitoring tasks. The verification results in the Regulatory Sandbox system were published in a peer-reviewed international medical journal in June 2020 (Hirano et al., 2020, JMIR). Based on these findings, SUSMED submitted an application under the “System of Inquiry on Regulatory Gray-Zone Areas” pursuant to the Industrial Competitiveness Enhancement Act and, on December 4, 2020, the Ministry of Health, Labour and Welfare (MHLW) issued an official notification7 confirming that substituting monitoring tasks with blockchain technology is compliant with Japan’s GCP ordinance.

Furthermore, a comparative evaluation between conventional clinical trial workflows and the SUSMED SDS workflow, conducted under the Research and Development Promotion Network Project supported by the Japan Agency for Medical Research and Development (AMED), demonstrated that monitoring workload decreased by 45.9%, medical institution workload decreased by 32.0%, and data queries for verification and correction decreased by 46.7%8. These findings indicate that, while substantially reducing overall operational overhead, the use of blockchain technology for clinical trial data management contributes significantly to streamlining processes, improving productivity in alignment with Quality by Design (QbD) principles.

7: Ministry of Health, Labour and Welfare (MHLW), “Publication of Responses to Inquiries Regarding Regulatory Confirmation” (December 4, 2020).

https://www.mhlw.go.jp/content/000702071.pdf

8: AMED R&D Database,“Development of an efficient and reliable monitoring method using blockchain technology.”

https://amedfind.amed.go.jp/amed/theme-list

Additionally, GAMP 5 Second Edition, the GxP guidance published by the International Society for Pharmaceutical Engineering (ISPE) in 2022, highlighted blockchain technology as a key solution for enhancing data integrity, which signals growing global expectations for the expanded use of blockchain across the clinical trial ecosystem.

Achievements and Future Outlook

The regulatory approval of a pharmaceutical product based on a clinical trial utilizing blockchain technology marks the world-first achievement, driving expectations for the broader implementation of this technology to accelerate the establishment of a modern clinical trial ecosystem.

Comment from Taro Ueno, President and CEO of SUSMED, Inc.:

“I, as a clinician, believe that the issue of Drug Loss, where pharmaceuticals used for patient care overseas remain unavailable in Japan, is a critical challenge.

In 2020, through our participation in the Regulatory Sandbox system and System of Inquiry on Regulatory Gray-Zone Areas under the Industrial Competitiveness Enhancement Act, we demonstrated that utilizing blockchain technology enables efficient compliance with the GCP ordinance.

I am deeply moved that marketing approval has now been granted for a pharmaceutical product based on this deregulation, and that is opening the door to delivering new treatment options to patients. I extend my heartiest gratitude to all stakeholders whose dedicated efforts made this milestone possible.

SUSMED remains committed to contributing to the optimal development of new therapies through the advancement of establishment of an innovative clinical trial ecosystem.”

In addition, Hirohiko Nakahara, Secretary-General, Secretariat of Intellectual Property Strategy Headquarters, Cabinet Office (former Counsellor, Economic Revitalization Bureau, Cabinet Secretariat), and Kazumi Nishikawa, Deputy Director-General, Minister’s Secretariat, Ministry of Economy, Trade and Industry (former Director, Healthcare Industries Division, Ministry of Economy, Trade and Industry), sent us the following comments:

Comment from Mr. Hirohiko Nakagawa, Secretary-General, Secretariat of Intellectual Property Strategy Headquarters, Cabinet Office:

“I would like to offer my sincere congratulations on the marketing approval granted by the Ministry of Health, Labour and Welfare for a pharmaceutical product validated through blockchain-based clinical trial system. It is profoundly meaningful that this approval was achieved based on the results obtained through the Regulatory Sandbox system. We look forward to seeing innovative technologies and solutions continue to contribute to resolving societal challenges in the future.

Comment from Mr. Kazumi Nishikawa, Deputy Director-General, Minister’s Secretariat, Ministry of Economy, Trade and Industry:

“I am delighted to see SUSMED’s initiative progressing appropriately as an effort leading the improvement of healthcare quality through the application of technology, as well as a concrete successful example of utilizing the Regulatory Sandbox system. I look forward to their further endeavors toward achieving a ‘Well Aging Society’ in the future.”

This world-first achievement, wherein regulatory marketing approval was granted for a pharmaceutical validated through a clinical trial leveraging blockchain technology following deregulation, represents a major step forward in addressing the challenges such as drug lag and drug loss. We have built a solid track record of reducing both costs and operational workload in clinical development through the utilization of SUSMED SDS, and we look forward to expanding its implementation among a broader range of clinical trials of pharmaceutical companies. Furthermore, by contributing to the establishment of a clinical trial ecosystem anchored in QbD principles, we aim to enhance productivity and quality in drug development, ultimately to support the optimization and long-term sustainability of future social security expenditures.

Reference Material

Key Implementation Examples of SUSMED SDS

  • Industry-sponsored clinical trials9,10 not only for pharmaceuticals, but also for medical devices and regenerative medicine products.
  • A wide spectrum of clinical research11, including specified clinical research and investigator initiated trials12 for prescription digital therapeutics (PDTx).
  • A disease registry13 designed to support post-marketing database studies aimed at leveraging Real-World Data (RWD), expected to facilitate efficient data collection, enhance the value of post-marketing surveillance, and promote the strategic utilization of clinical data for label expansions and the development of proper-use guidelines.

9: SUSMED’s Blockchain-Based Clinical Trial System Implemented in Heartseed’s Clinical Trial

https://susmed.co.jp/wp-content/uploads/2025/10/cyfot8ygufgypuigouhphui.pdf

10: Announcement on the Operational Launch of Blockchain-Based Clinical Trial System in Pivotal Clinical Trial for Tinnitus Digital Therapeutic

https://susmed.co.jp/wp-content/uploads/2026/05/20260529_kTYLyEBa.pdf

11: Launch of Specified Clinical Research for Digital Therapeutic Targeting Premenstrual Syndrome (PMS) and Premenstrual Dysphoric Disorder (PMDD)

https://susmed.co.jp/wp-content/uploads/2025/01/PR20250128_PMSPMDD_ja.pdf

12: Announcement on the Operational Launch of SUSMED’s Clinical Trial System Utilizing Blockchain Technology in Investigator-Initiated Trial at the National Center of Neurology and Psychiatry (NCNP)

https://susmed.co.jp/wp-content/uploads/2024/12/susmed_press20241225_26.pdf

13: Construction of an Integrated Venous Disease Registry System to Promote Utilization of Post-Marketing Surveillance for Medical Devices, Streamlining Site Operations while Ensuring Data Integrity

https://susmed.co.jp/wp-content/uploads/2024/12/susmed_press20241211.pdf

New Technology Demonstration system (The Regulatory Sandbox system)

The “New Technology Demonstration system” (commonly known as the Regulatory Sandbox system) was established under the Act on Special Measures for Productivity Improvement to promote business activities utilizing novel technologies and innovative business models. By limiting participants and timeframes, this framework creates a controlled environment that allows companies to conduct real-world demonstrations without being bound by existing regulations. This system enables rapid verification, and allows to leverage the resulting data and evidence to drive regulatory reform.

About SUSMED, Inc.

SUSMED, Inc. is a R&D driven enterprise dedicated to advancing digital healthcare. In addition to developing digital therapeutics (DTx), including a therapeutic application for insomnia disorder. SUSMED provides a clinical development support system powered by blockchain technology. Grounded in robust technology, SUSMED is driving the evolution of digital healthcare through a proprietary intellectual property portfolio covering various patents in digital therapeutics and medical applications of blockchain technology.

Top page: https://www.susmed.co.jp/en/
SUSMED SDS: https://www.susmed.co.jp/en/system/

Notes

The information regarding pharmaceutical products contained in this release is intended solely for corporate disclosure purposes and does not constitute an advertisement or promotion of them.

Contacts

SUSMED, Inc.

E-mail: support@susmed.co.jp
E.SUN Debuts at COP17, Showcasing Taiwan’s Nature Finance in Action

E.SUN Debuts at COP17, Showcasing Taiwan’s Nature Finance in Action


E.SUN Debuts at COP17, Showcasing Taiwan’s Nature Finance in Action

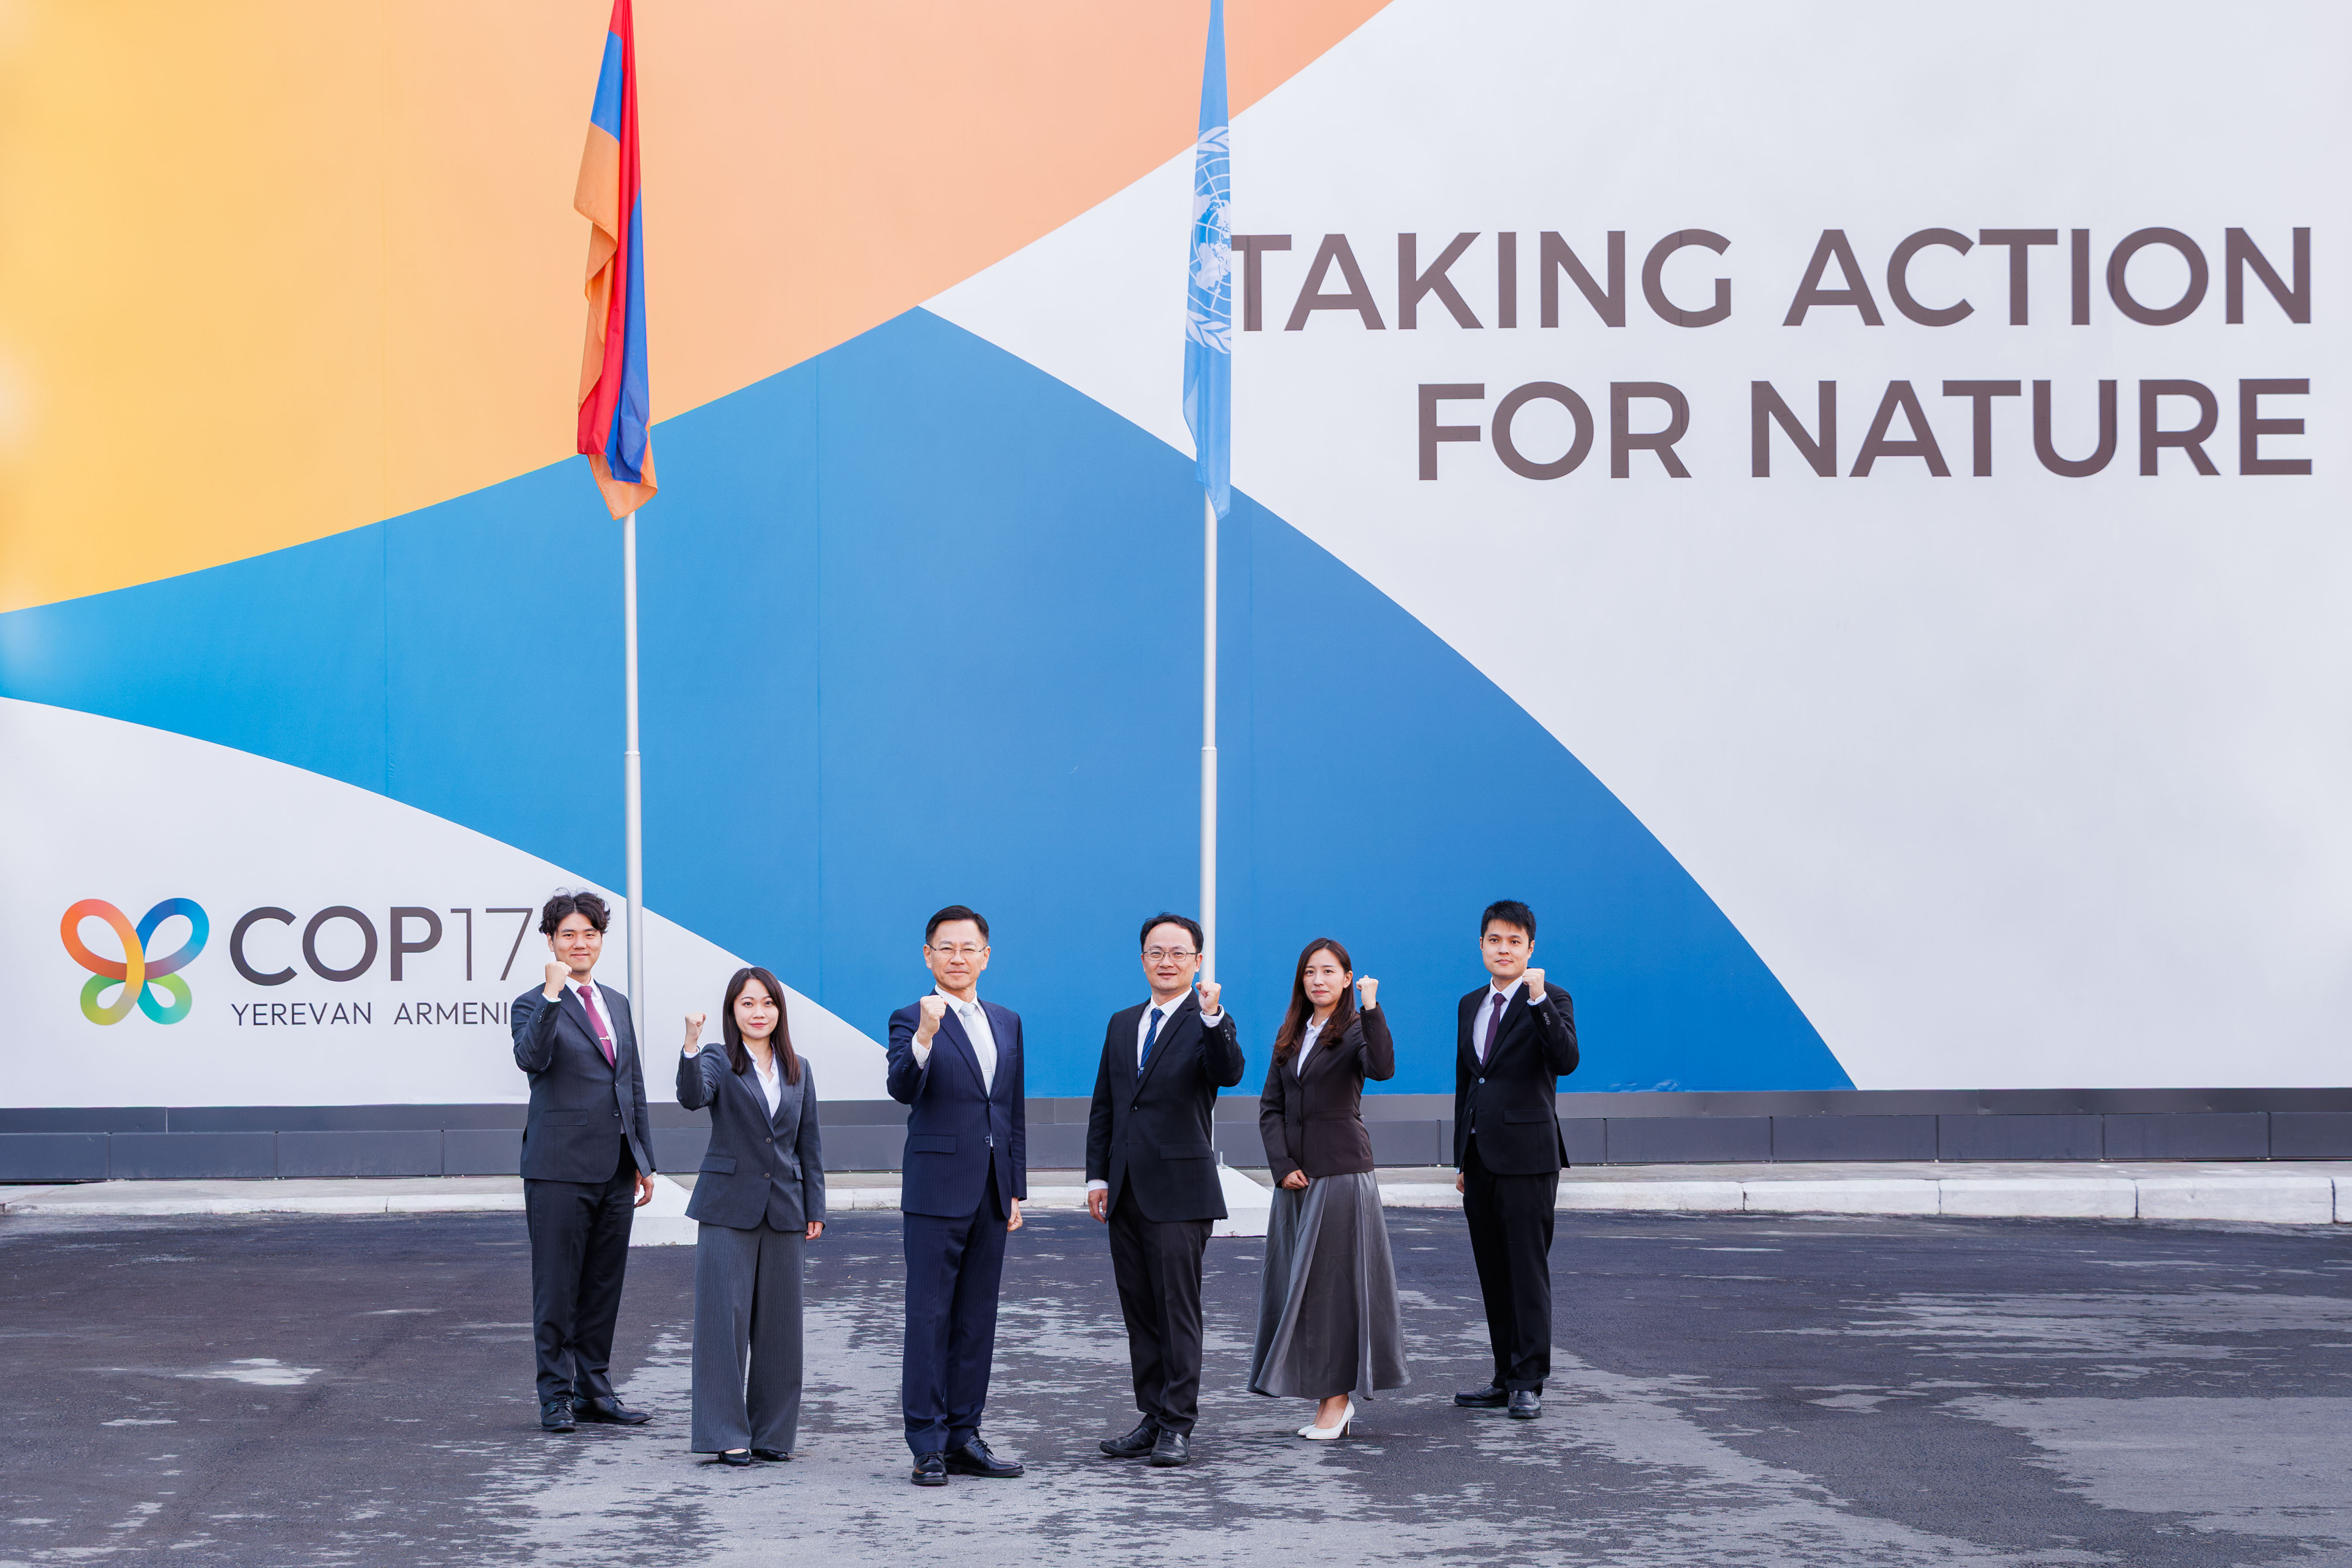
TAIPEI, Taiwan–(BUSINESS WIRE)–E.SUN Financial Holding made its debut at the 17th Conference of the Parties to the Convention on Biological Diversity (CBD COP17), bringing Taiwan’s nature finance experience to the global stage. E.SUN Chief Sustainability Officer Louis L.Y. Chang was invited by the World Climate Foundation (WCF) and the Finance for Biodiversity Foundation (FfB) to speak in the Blue Zone on October 26 and 27 about “Bridging the Biodiversity Financing Gap: E.SUN’s Practice and Vision for Nature Capital.”


Held in Yerevan, Armenia, from October 19 to 30 under the theme “Taking Action for Nature,” CBD COP17 reviewed progress on the Kunming-Montreal Global Biodiversity Framework and explored ways to accelerate action and mobilize finance for nature. As the only Taiwanese financial institution attending COP17, E.SUN shared how financial institutions can connect finance, businesses, and local communities to advance biodiversity. Based on E.SUN’s keynote, representatives from UNEP FI, the Inter-American Development Bank, Danske Bank Asset Management and Storebrand Asset Management further elaborated the discussion on strengthening stakeholder collaboration, supporting business transition, and scaling biodiversity finance.

E.SUN’s biodiversity efforts span conservation, education, and culture. Since 2014, E.SUN’s Malavi Project has supported Indigenous farmers in adopting organic farming practices, with more than 19 hectares of farmland now organically certified. In 2026, the project became one of Taiwan’s first certified Other Effective Area-Based Conservation Measures (OECMs), demonstrating how agricultural production and conservation can mutually reinforce one another.

In 2022, E.SUN partnered with National Taiwan University to launch the ESG Centenary Project, which aims to plant 100,000 native Taiwanese conifers over ten years. The partnership also revived 28 indigenous millet varieties in Xinyi Township, Nantou County, contributing to both native plant conservation and Indigenous cultural preservation.

E.SUN also uses finance to translate biodiversity ambitions into business action. Its “Farm to Table Sustainable Food and Agriculture Value Chain Financing” initiative connects agriculture, food processing, logistics, hospitality and other industries to support sustainable food systems. E.SUN has also established biodiversity and OECM-linked loans with Yulon Motor and CPC Corporation, Taiwan, incorporating species restoration and habitat conservation into financing arrangements.

Through a Public-Private-Philanthropic Partnership (PPPP), E.SUN has further brought together Okuma Fishing Tackle, the Kaohsiung Wild Bird Society, the Kaohsiung City Government Tourism Bureau, and Taiwan Water Corporation to remove invasive species and restore habitats at Niaosong Wetland through Nature-based Solutions.

“Without nature and biodiversity, climate action cannot succeed,” said Chang. “Finance can serve as a bridge between conservation and development, between local action and global impact, and between today’s investment and a nature-positive future.”

E.SUN will continue to harness the positive impact of finance and cross-sector collaboration to mobilize capital, expertise, and talent for nature-positive action, while bringing Taiwan’s experience in connecting finance, business, communities, and conservation to the global stage. ***

Contacts

Public Relations, E.SUN FHC

Virginia Lin

pr@esunfhc.com
(+8862)2175-1335

Public Relations, E.SUN FHC

Alice Chen

pr@esunfhc.com
(+8862)2175-1313 #8190

Alnylam Announces $1 Billion Share Repurchase Program and Change in Commercial Leadership

Alnylam Announces $1 Billion Share Repurchase Program and Change in Commercial Leadership


Alnylam Announces $1 Billion Share Repurchase Program and Change in Commercial Leadership

Board authorizes share repurchase program reflecting confidence in Alnylam’s long-term growth prospects and commitment to disciplined capital allocation

Commercial leadership transition supports the Company’s next phase of growth

CAMBRIDGE, Mass.–(BUSINESS WIRE)–Alnylam Pharmaceuticals, Inc. (Nasdaq: ALNY), the leading RNAi therapeutics company, today announced that its Board of Directors has authorized a $1 billion share repurchase program and that the Company will transition to new commercial leadership as it enters its next phase of growth.


“Today we announced two important decisions, both of which demonstrate our conviction in Alnylam’s future,” said Amy Schulman, Chair of Alnylam’s Board of Directors. “Alnylam has a distinctive and robust scientific foundation, a strong commercial opportunity and a fulsome pipeline. We will continue to focus on translating those strengths into sustained performance and long-term value for shareholders. Today’s repurchase authorization reflects our confidence in that future and our commitment to disciplined capital allocation. We are committed to continuing to evaluate opportunities to build on Alnylam’s strengths and create value for our shareholders.”

$1 Billion Share Repurchase Program

Alnylam’s Board of Directors has authorized the repurchase of up to $1 billion of the Company’s common stock, reflecting its confidence in the Company’s long-term growth prospects and its belief that the current share price does not fully reflect Alnylam’s intrinsic value. Pursuant to this authorization, the Company expects to repurchase approximately $500 million of its common stock by the end of 2026 and the balance during 2027, subject to market conditions.

Returning capital to shareholders represents an important evolution in Alnylam’s financial profile. Today’s share repurchase authorization allows us to enhance shareholder returns while continuing to invest in our ongoing ATTR-CM launch, robust pipeline and select external innovation.

Commercial Leadership Transition

Alnylam also announced that Tolga Tanguler will depart his role as Chief Commercialization Officer. The Company is working with an executive search firm to support the selection of its next Chief Commercialization Officer and anticipates providing an update in due course.

During Mr. Tanguler’s tenure, Alnylam significantly expanded its global commercial organization and marketed portfolio, establishing a strong foundation for future growth. As the Company looks ahead, it remains focused on expanding the reach of its medicines, building on its leadership in TTR and preparing for future launches.

“I’m grateful for Tolga’s leadership and the contributions he has made over the past several years,” said Yvonne Greenstreet, M.D., Chief Executive Officer of Alnylam. “As we look ahead, we have significant opportunities to reach more patients with our medicines, advance our pipeline and continue building a sustainable, high-performing business.”

About Alnylam Pharmaceuticals

Alnylam (Nasdaq: ALNY) is a leading global biopharmaceutical company and the pioneer of the RNA interference (RNAi) revolution. The Company is focused on developing transformative therapies with the potential to prevent, halt, or reverse disease. For more than two decades, Alnylam has advanced the Nobel-prize-winning science of RNAi, delivering critical breakthroughs and six approved medicines. Alnylam has medicines available in more than 70 countries and a rapidly expanding and robust pipeline, in addition to consistently being recognized as an exceptional workplace and socially responsible organization. The Company is executing on its Alnylam 2030 strategy to accelerate innovation and scale impact to transform human health. For more information, please visit www.alnylam.com or follow Alnylam on X, LinkedIn, Facebook, Instagram, or YouTube.

Forward-Looking Statements

This press release contains forward-looking statements. Forward-looking statements include statements regarding Alnylam’s expectations, beliefs, goals, plans or prospects, including, without limitation, statements regarding: Alnylam’s long-term growth prospects; Alnylam’s ability to build on its strengths, achieve sustained performance and create value for its shareholders; Alnylam’s ability to advance its pipeline and the potential it will pursue external innovation; the timing of Alnylam’s selection of its next Chief Commercialization Officer and any public announcement thereof; Alnylam’s ability to expand the reach of its medicines, build on its leadership in TTR, launch products in the future and continue to build a sustainable, high-performing business; the value of shares of Alnylam’s common stock that it will repurchase under its $1 billion share repurchase program, and the timing of any such repurchases; and Alnylam’s ability to achieve the goals in its Alnylam 2030 strategy. Actual results and future plans may differ materially from those indicated by these forward-looking statements as a result of various important risks, uncertainties and other factors, including, without limitation, risks and uncertainties relating to: Alnylam’s ability to successfully execute on its Alnylam 2030 strategy; Alnylam’s ability to successfully launch, market and sell Alnylam’s approved products globally, including AMVUTTRA; Alnylam’s ability to discover and develop novel drug candidates and delivery approaches and successfully demonstrate the efficacy and safety of its product candidates; the pre-clinical and clinical results for Alnylam’s product candidates; actions or advice of regulatory agencies and Alnylam’s ability to obtain and maintain regulatory approval for its product candidates, as well as favorable pricing and reimbursement; delays, interruptions or failures in the manufacture and supply of Alnylam’s marketed products or its product candidates; obtaining, maintaining and protecting intellectual property; Alnylam’s ability to manage its growth and operating expenses through disciplined investment in operations; Alnylam’s ability to maintain strategic business collaborations; Alnylam’s dependence on third parties for the development and commercialization of certain products; the outcome of litigation and government investigations; the risk of future litigation and government investigations; and unexpected expenditures; as well as those risks and uncertainties more fully discussed in the “Risk Factors” filed with Alnylam’s most recent periodic report (Quarterly Report on Form10-Q or Annual Report on Form 10-K) filed with the SEC and in its other SEC filings. Alnylam explicitly disclaims any obligation, except to the extent required by law, to update any forward-looking statements.

Contacts

Investor Relations
investors@alnylam.com

Media Relations
media@alnylam.com

Evommune Announces $30 Million Private Placement

Evommune Announces $30 Million Private Placement


Evommune Announces $30 Million Private Placement

PALO ALTO, Calif. & NEW YORK–(BUSINESS WIRE)–Evommune, Inc. (“Evommune” or the “Company”) (NYSE: EVMN), a clinical-stage biotechnology company developing innovative therapies that target key drivers of chronic inflammatory diseases, today announced that it has entered into a securities purchase agreement to sell 4,966,889 shares of its common stock to a select group of dedicated healthcare institutional investors in a private placement. The purchase price of each share of common stock is $6.04. Evommune anticipates the gross proceeds from the private placement to be approximately $30 million, before deducting any transaction-related expenses. The private placement is expected to close on or about November 6, 2026, subject to the satisfaction of customary closing conditions.


The financing includes participation from both new and existing institutional investors, including Adage Capital Partners, L.P., Balyasny Asset Management, Defilade Capital Management, OrbiMed, Squadron Capital Management and other fundamental healthcare dedicated funds.

Morgan Stanley & Co. LLC and Leerink Partners LLC acted as placement agents for the private placement. Oppenheimer & Co. Inc. acted as capital markets advisor for the private placement.

Evommune intends to use the net proceeds from the private placement to advance its clinical development programs, including EVO570, its newly in-licensed oral VAV1-directed molecular glue degrader, and for general corporate purposes.

The offer and sale of the securities to be sold in the private placement have not been registered under the Securities Act of 1933, as amended (the “Securities Act”), or any state or other applicable jurisdiction’s securities laws, and such securities may not be offered or sold in the United States absent registration or an applicable exemption from the registration requirements of the Securities Act and applicable state or other jurisdictions’ securities laws. Concurrently with entering into the securities purchase agreement, Evommune and the investors entered into a registration rights agreement pursuant to which Evommune has agreed to file a registration statement with the Securities and Exchange Commission registering the resale of the shares of common stock issued in the private placement.

This press release shall not constitute an offer to sell or the solicitation of an offer to buy these securities, nor shall there be any sale of these securities in any jurisdiction in which such offer, solicitation or sale would be unlawful prior to the registration or qualification under the securities laws of any such jurisdiction.

About Evommune, Inc.

Evommune, Inc., is a clinical-stage biotechnology company developing innovative therapies that target key drivers of chronic inflammatory diseases. The Company’s mission is to improve patients’ daily lives and prevent the long-term effects of uncontrolled inflammation that are a consequence of the limitations of existing therapies. To achieve this, Evommune is advancing a portfolio of differentiated product candidates that target key drivers of chronic inflammation.

Forward-Looking Statements

This press release contains “forward-looking statements” within the meaning of the “safe harbor” provisions of the Private Securities Litigation Reform Act of 1995. All statements in this press release other than statements of historical facts are “forward-looking statements.” These statements may be identified by words such as “aims,” “anticipates,” “believes,” “could,” “estimates,” “expects,” “forecasts,” “goal,” “intends,” “may,” “plans,” “possible,” “potential,” “seeks,” “will” and variations of these words or similar expressions that are intended to identify forward-looking statements, although not all forward-looking statements contain these words. Forward-looking statements in this press release include, but are not limited to, statements concerning expected proceeds from the private placement, expected use of proceeds, and expected closing of the private placement. These forward-looking statements are based on the Company’s expectations and assumptions as of the date of this press release. Each of these forward-looking statements involves risks and uncertainties that could cause the Company’s clinical development programs, future results or performance to differ materially from those expressed or implied by the forward-looking statements. Many factors may cause differences between current expectations and actual results, including: the Company’s limited operating history and historical losses; the potential that success in preclinical testing and earlier clinical trials does not ensure that later clinical trials will generate the same results or otherwise provide adequate data to demonstrate the efficacy and safety of a product candidate; the Company’s ability to obtain regulatory approval of and successfully commercialize its product candidates; the impacts of macroeconomic conditions, including heightened inflation and uncertain credit and financial markets, on the Company’s business, clinical trials and financial position; unexpected safety or efficacy data observed during preclinical studies or clinical trials; clinical trial site activation or enrollment rates that are lower than expected; the Company’s ability to realize the benefits of its collaborations and license agreements; changes in expected or existing competition; changes in the regulatory environment; the Company’s ability to obtain, maintain and protect its intellectual property; and unexpected litigation or other disputes. Other factors that may cause the Company’s actual results to differ from those expressed or implied in the forward-looking statements in this press release are identified under the heading “Risk Factors” in the Company’s Quarterly Report on Form 10-Q, filed with the SEC on August 6, 2026, and in other filings that the Company makes and will make with the SEC in the future. The Company expressly disclaims any obligation to update any forward-looking statements contained herein, whether as a result of any new information, future events, changed circumstances or otherwise, except as otherwise required by law.

Contacts

Investors and Media:
Megan Humphreys

megan.humphreys@evommune.com

Evommune Enters into Agreement to In-License EVO570, a Potential Best-in-Class Oral VAV1 Molecular Glue Degrader, from Hinova Pharmaceuticals

Evommune Enters into Agreement to In-License EVO570, a Potential Best-in-Class Oral VAV1 Molecular Glue Degrader, from Hinova Pharmaceuticals


Evommune Enters into Agreement to In-License EVO570, a Potential Best-in-Class Oral VAV1 Molecular Glue Degrader, from Hinova Pharmaceuticals

– Oral, highly selective and potent VAV1 molecular glue degrader designed to deliver a differentiated clinical profile across a broad range of indications in autoimmune disease

– Phase 1 expected to begin imminently; Evommune plans to initiate a Phase 2 trial in rheumatoid arthritis in 2027

– Evommune to pay $20MM upfront in cash and $10MM in the form of pre-funded warrants and up to approximately $1.1B in development, regulatory and commercial milestones, plus tiered royalties

PALO ALTO, Calif. & NEW YORK & CHENGDU, China–(BUSINESS WIRE)–Evommune, Inc. (NYSE: EVMN) (the “Company” or “Evommune”), a clinical-stage biotechnology company developing therapies for chronic inflammatory diseases, and Hinova Pharmaceuticals Inc. (SSE: 688302) today announced an exclusive license agreement granting Evommune worldwide rights outside of Greater China to develop, manufacture and commercialize EVO570 (formerly HC118 and known as HP570 in Greater China), an oral VAV1-directed molecular glue degrader (MGD).


VAV1 is a signaling protein that prominently drives T- and B-cell activation across a wide range of chronic inflammatory diseases. EVO570, discovered by Hinova, is an investigational oral MGD designed to selectively eliminate VAV1, quieting the immune activity underlying many autoimmune diseases. Unlike JAK enzymes, which are broadly expressed across many tissues and organs, VAV1 is largely restricted to immune cells, a core rationale for its potential to offer a much-needed, favorable safety and tolerability profile. In preclinical studies, EVO570 has demonstrated promising activity in animal models of multiple immune-mediated conditions and greater potency in human immune cells than competitor VAV1 degraders. Investigational New Drug (IND) applications have been submitted in China and accepted in the United States, and a Phase 1 trial evaluating safety, pharmacokinetics and target engagement/pharmacodynamics is on track to begin imminently.

“VAV1 is a compelling target in immunology today, offering a potential path to JAK-like efficacy without the safety and tolerability challenges of other available therapies,” said Dr. Jeegar Patel, Chief Scientific Officer of Evommune. “With a differentiated scaffold and structure-guided optimization resulting in sub-nanomolar potency, high selectivity and a profile supporting once-daily oral dosing, EVO570 has the potential to be a best-in-class approach for patients with rheumatoid arthritis and other T- and B-cell-driven autoimmune diseases. We look forward to reporting Phase 1 results and plan to advance it into its first Phase 2 trial in rheumatoid arthritis in 2027.”

“Adding this development-stage program is a strategic step for Evommune, expanding our pipeline into T- and B-cell-driven autoimmune diseases with a differentiated oral degrader that complements our existing pipeline,” said Greg Moss, Chief Business and Legal Officer of Evommune. “We have been impressed by Hinova’s innovative drug discovery and execution capabilities, and we are pleased to partner with a team that shares our commitment to developing differentiated therapies for patients. With this transaction, we are adding a novel therapeutic candidate with wide-ranging potential, while maintaining a disciplined portfolio management approach, and continuing to position Evommune to create meaningful long-term value for patients with chronic inflammatory diseases and for our stockholders.”

“We are proud to partner with Evommune, a leading immunology company with a proven track record of advancing differentiated therapies for chronic inflammatory diseases,” said Dr. Yuanwei Chen, Founder and Chairman of Hinova. “EVO570’s advancement reflects the strength of Hinova’s targeted protein degrader and translational medicine platforms, and we believe Evommune’s development expertise and global reach make it the ideal collaborator to bring this potential best-in-class VAV1 degrader to patients worldwide.”

Terms of the agreement

As consideration for the license, Evommune will pay Hinova $30 million upfront, consisting of $20 million in cash and $10 million in the form of pre-funded warrants to purchase Evommune common stock. Hinova is eligible to receive up to approximately $1.1 billion in development, regulatory and commercial milestones, as well as tiered royalties on net sales outside Greater China. BFC Group served as exclusive financial advisor and Fangda Partners served as exclusive legal advisor to Hinova.

About VAV1 and EVO570

VAV1 is a guanine nucleotide exchange factor largely restricted to immune cells that transmits signals from the T-cell and B-cell receptors to drive activation, cytokine production and migration. EVO570 is an orally administered MGD that directs VAV1 to the cell’s protein-disposal machinery for degradation. EVO570 is an investigational drug candidate and has not yet been approved by any regulatory authority.

About Evommune

Evommune, Inc. is a clinical-stage biotechnology company developing innovative therapies that target key drivers of chronic inflammatory diseases. The Company’s mission is to improve patients’ daily lives and prevent the long-term effects of uncontrolled inflammation that are a consequence of the limitations of existing therapies. To achieve this, Evommune is advancing a portfolio of differentiated product candidates that target key drivers of chronic inflammation. For more information, please visit www.evommune.com and follow us on LinkedIn.

About Hinova

Hinova Pharmaceuticals Inc. (SSE: 688302), headquartered in Chengdu, China, is an international, clinical-stage biopharmaceutical company focused on discovering, developing, manufacturing and commercializing innovative therapies for treating cancer, autoimmune diseases and metabolic diseases. Hinova has one product that has been approved in China for prostate cancer. The company’s pipeline includes multiple investigational drugs, including traditional small molecules, deuterated drug, PROTAC degraders, molecular glue degrader, targeting a variety of cancer types, autoimmune diseases and metabolic disorders, all aimed at addressing significant unmet medical needs and improving patient outcomes.

Forward-Looking Statements

This press release contains “forward-looking statements” within the meaning of the “safe harbor” provisions of the Private Securities Litigation Reform Act of 1995. All statements in this press release other than statements of historical facts are “forward-looking statements.” These statements may be identified by words such as “aims,” “anticipates,” “believes,” “could,” “estimates,” “expects,” “forecasts,” “goal,” “intends,” “may,” “plans,” “possible,” “potential,” “seeks,” “will” and variations of these words or similar expressions that are intended to identify forward-looking statements, although not all forward-looking statements contain these words. Forward-looking statements in this press release include, but are not limited to, statements regarding: the potential therapeutic benefit and best-in-class potential of EVO570, including the safety and tolerability profile of EVO570; the anticipated benefits of the license agreement with Hinova, including Evommune’s expectations regarding its worldwide rights (outside of Greater China) to develop, manufacture and commercialize EVO570; the design, objectives, initiation, timing and progress of the planned Phase 1 trial and the planned global Phase 2 trial of EVO570 in rheumatoid arthritis expected to begin in 2027; the timing and amount of upfront, milestone and royalty payments payable to Hinova under the license agreement, and Evommune’s ability to achieve the development, regulatory and commercial milestones that would trigger such payments; the Company’s ability to create meaningful long-term value for patients with chronic inflammatory diseases and for its stockholders; and the Company’s business strategies, plans and prospect and continued advancement of the Company’s portfolio. These forward-looking statements are based on the Company’s expectations and assumptions as of the date of this press release. Each of these forward-looking statements involves risks and uncertainties that could cause the Company’s clinical development programs, future results or performance to differ materially from those expressed or implied by the forward-looking statements. Many factors may cause differences between current expectations and actual results, including: the Company’s limited operating history and historical losses; the potential that success in preclinical testing and earlier clinical trials does not ensure that later clinical trials will generate the same results or otherwise provide adequate data to demonstrate the efficacy and safety of a product candidate; the Company’s ability to obtain regulatory approval of and successfully commercialize its product candidates, including EVO570; the impacts of macroeconomic conditions, including heightened inflation and uncertain credit and financial markets, on the Company’s business, clinical trials and financial position; unexpected safety or efficacy data observed during preclinical studies or clinical trials; clinical trial site activation or enrollment rates that are lower than expected; the Company’s ability to realize the anticipated benefits of the license agreement with Hinova and its other collaborations and license agreements, and to achieve the milestones and make the related payments contemplated by such agreements; changes in expected or existing competition; changes in the regulatory environment, including with respect to products licensed from and regulatory applications pending in China; the Company’s ability to obtain, maintain and protect its intellectual property; and unexpected litigation or other disputes. Other factors that may cause the Company’s actual results to differ from those expressed or implied in the forward-looking statements in this press release are identified under the heading “Risk Factors” in the Company’s Annual Report on Form 10-K, filed with the SEC on March 5, 2026, as supplemented by the Company’s Quarterly Report on Form 10-Q, filed with the SEC on August 6, 2026, and in other filings that the Company makes and will make with the SEC in the future. The Company expressly disclaims any obligation to update any forward-looking statements contained herein, whether as a result of any new information, future events, changed circumstances or otherwise, except as otherwise required by law.

Contacts

Evommune Media and Investors: Megan Humphreys, megan.humphreys@evommune.com

Hinova: Investor Relations: ir@hinovapharma.com

Heidelberg Pharma to Present New Clinical Data on its ATAC Candidates at the ASH Annual Meeting 2026

Heidelberg Pharma AG

/ Key word(s): Conference

Heidelberg Pharma to Present New Clinical Data on its ATAC Candidates at the ASH Annual Meeting 2026

04.11.2026 / 16:36 CET/CEST

The issuer is solely responsible for the content of this announcement.


PRESS RELEASE

Heidelberg Pharma to Present New Clinical Data on its ATAC Candidates at the ASH Annual Meeting 2026

Ladenburg, Germany, 4 November 2026 – Heidelberg Pharma AG (FSE: HPHA), a clinical stage company developing innovative Antibody Drug Conjugates (ADCs), will present new data from its Phase I/IIa clinical study with its lead Amanitin-based ADC candidate, pamlectabart tismanitin (HDP-101), in an oral presentation, as well as data from its second clinical candidate, HDP-102, during the poster abstract session of the 68th Annual Meeting of the American Society of Hematology (ASH), being held in New Orleans, USA, from 12 to 15 December 2026.

Details of the oral presentation (HDP-101):

Presentation title: First-in-human phase 1/2a study of pamlectabart tismanitin (HDP-101), an amanitin-based BCMA-targeting antibody-drug conjugate (ADC): Recommended phase 2 dose (RP2D) selection and preliminary antitumor activity in relapsed/refractory multiple myeloma (RRMM)

Presenter: Professor Marc-Steffen Raab, MD
Publication number: 270
Session: 654. Multiple Myeloma: Pharmacologic Therapies: Novel Approaches in Relapsed Myeloma: Immunotherapies and Small Molecules
Session time: Saturday, 12 December 2026, 02:00 – 03:30 pm (CDT)
Presentation: Saturday, 12 December 2026, 03:15 – 03:30 pm (CDT)
Room: Ernest N. Morial Convention Center (MCCNO) –
La Nouvelle A-B
Link to the abstract: ASH® ANNUAL PROGRAM

Pamlectabart tismanitin is an anti-BCMA antibody-Amanitin drug conjugate for the treatment of relapsed or refractory multiple myeloma (RRMM), a bone marrow cancer with a high unmet medical need. Phase I of the trial was a dose-escalation study to determine a dose level of the candidate for the Phase IIa expansion part of the study.

The Phase I part of the study included ten patient cohorts and was completed in April 2026, and the recommended dose for the Phase IIa expansion part (Recommended Phase 2 Dose; RP2D) was determined. The Phase IIa is progressing as planned.

Pamlectabart tismanitin demonstrated consistent clinical activity with a favorable safety profile in heavily pretreated RRMM patients supporting the potential of the candidate as a novel payload BCMA-targeted ADC therapy.

Details of the poster presentation (HDP-102):

Presentation title: HDP-102, a first-in-class CD37-targeted amanitin-based antibody-drug conjugate, demonstrates translational activity, favorable early safety and efficacy in relapsed/refractory (R/R) B-cell malignancies

Presenter: Dr. Irina Mocanu, MD
Publication number: 3717
Session: 629: Aggressive Lymphomas, Non-Cellular Immunotherapy including Bispecific Antibodies: Poster II
Session time: Sunday, 13 December 2026, 06:00 – 08:00 pm (CDT)
Presentation: Sunday, 13 December 2026, 06:00 – 08:00 pm (CDT)
Room: Ernest N. Morial Convention Center (MCCNO) –
Hall F
Link to the abstract: ASH® ANNUAL PROGRAM

HDP-102 is an ADC targeting CD37, which is overexpressed on B-cell lymphoma cells. Preclinical studies have shown excellent anti-tumor efficacy in in vivo studies as well as good tolerability. HDP-102 is intended to be developed for specific indications of non-Hodgkin lymphoma.

In 2025, Heidelberg Pharma announced the dosing of the first patient with HDP-102. Three patients were treated with 40 µg/kg in the first cohort. The treatment was well tolerated, and initial signs of biological activity have been observed already at the very low dose. Two patients showed stabilization of the disease with regression and reduction of lymph nodes. The safety committee recommended increasing the dose of HDP-102 to 65 µg/kg in cohort 2.

In line with Heidelberg Pharma’s strategic decision from September 2025 to discontinue the study, the Company is exploring partnership opportunities to support the continuation of the program.


About Heidelberg Pharma

Heidelberg Pharma is the first company to develop cancer therapies using Amanitin, a compound derived from the green death cap mushroom. The biological mechanism of action of the toxin represents a new therapeutic modality and is used as a compound in the Amanitin-based ADC technology, the so-called ATAC technology.

Lead candidate HDP-101 (INN: pamlectabart tismanitin) is a BCMA ATAC in clinical development for multiple myeloma. The candidate has been granted Orphan Drug Designation and Fast Track Designation from the FDA. A second ATAC candidate, HDP-102 is in clinical development stage in Non-Hodgkin Lymphoma. HDP-103 against metastatic castration-resistant prostate cancer and HDP-104 targeting gastrointestinal tumors such as colorectal cancer have completed preclinical development. These programs are available for partnering.

The company is based in Ladenburg, Germany, and is listed on the Frankfurt Stock Exchange: ISIN DE000A11QVV0 / WKN A11QVV / Symbol HPHA. More information is available at http://www.heidelberg-pharma.com/.

ATAC® is a registered trademark of Heidelberg Pharma Research GmbH.

Contact
Heidelberg Pharma AG
Sylvia Wimmer
Senior Director Corporate Communications
Tel.: +49 6203 1009 1004
E-mail: investors@hdpharma.com
Wallstadter Straße 59, 68526 Ladenburg
IR/PR-Support
MC Services AG
Katja Arnold (CIRO)
Managing Director & Partner
Tel.: +49 89 210 228-40
E-mail: katja.arnold@mc-services.eu  


This communication contains certain forward-looking statements relating to the Company’s business, which can be identified by the use of forward-looking terminology such as “estimates”, “believes”, “expects”, “may”, “will” “should” “future”, “potential” or similar expressions or by a general discussion of the Company’s strategy, plans or intentions. Such forward-looking statements involve known and unknown risks, uncertainties and other factors, which may cause our actual results of operations, financial condition, performance, or achievements, or industry results, to be materially different from any future results, performance or achievements expressed or implied by such forward-looking statements. Given these uncertainties, prospective investors and partners are cautioned not to place undue reliance on such forward-looking statements. We disclaim any obligation to update any such forward-looking statements to reflect future events or developments.


04.11.2026 CET/CEST Dissemination of a Corporate News, transmitted by EQS News – a service of EQS Group.
The issuer is solely responsible for the content of this announcement.

The EQS Distribution Services include Regulatory Announcements, Financial/Corporate News and Press Releases.


Language: English
Company: Heidelberg Pharma AG
Wallstadter Straße 59
68526 Ladenburg
Germany
Phone: +49 6203 10090
E-mail: investors@hdpharma.com
Internet: www.heidelberg-pharma.com
ISIN: DE000A11QVV0
WKN: A11QVV
Listed: Regulated Market in Frankfurt (Prime Standard); Regulated Unofficial Market in Dusseldorf, Munich, Stuttgart, Tradegate BSX
LEI Code: 391200E09XYBYITR1W32
EQS News ID: 2428588

 
End of News EQS News Service

2428588  04.11.2026 CET/CEST

Viromed Medical AG Receives Approval for ViroCAP® med in Turkey and Announces Further Operational Progress

Viromed Medical AG

/ Key word(s): Miscellaneous

Viromed Medical AG Receives Approval for ViroCAP® med in Turkey and Announces Further Operational Progress

04.11.2026 / 16:20 CET/CEST

The issuer is solely responsible for the content of this announcement.


Viromed Medical AG Receives Approval for ViroCAP® med in Turkey and Announces Further Operational Progress

Rellingen, November 4, 2026 – Viromed Medical AG (“Viromed”; ISIN: DE000A3MQR65), a medical technology company and pioneer of cold plasma technology, has received approval in Turkey for its medical device ViroCAP® med (MDR Class IIa). The product is now approved for medical applications on the Turkish market, in particular for the treatment of open wounds and intraoperative use.

Uwe Perbandt, CEO of Viromed Medical AG: “The approval of ViroCAP® med in Turkey is a significant success for our international market development. Following the registration of ViroCAP® derma with the ÜTS (National Medical Device Tracking System) in the summer, we have already received approval for ViroCAP® med, which we had expected no earlier than February or March 2027. This marks another important milestone for the commercial launch of our ViroCAP® technology in a healthcare market that is strategically important for us.”

Viromed already has binding minimum orders for 7,000 ViroCAP® derma/med devices through the end of 2028 for the Turkish market. Deliveries will be made on the basis of the approval now granted.

The required process for reimbursement by Turkish health insurers is already running in parallel. With this approval, Viromed continues its international expansion strategy and lays the foundation for the further development of the Turkish market.

ViroCAP® med is designed for professional use by qualified medical personnel and supports the treatment of open wounds as well as intraoperative use of cold atmospheric plasma. The application is contactless and mobile. Each application is planned to last one to two minutes, while skin temperature remains below 40 degrees Celsius.

Additional Local Approval Procedures Initiated

In parallel, Viromed together with its partner Umeco has started the approval process for ViroCAP® med in additional Asian countries. A prerequisite for this was likewise the prior approval of the product as an MDR Class IIa medical device in the EU. Outside the EU, a local approval in each target country is required, which is being pursued together with the distribution partners.

First Reimbursement by a Health Insurer – Selective Contract in Lower Saxony

Viromed has also achieved its first reimbursement of ViroCAP® by a statutory health insurer. A selective contract was concluded with a health insurer in Lower Saxony for an indication in the intensive care area: the treatment of patients with a left ventricular assist device (LVAD). The focus is on preventing infections at the driveline exit site.

Viromed is also in advanced discussions for further selective contracts with health insurers in three additional German federal states.

Further Studies

In November, a successfully completed retrospective study on the prevention of postoperative wound infections will be published. In addition, Viromed is starting a randomized study on the use of ViroCAP® in LVAD patients as well as a further study on the use of the technology in the field of heart valves.


About Viromed Medical AG

Viromed Medical AG specializes in the development, manufacture and distribution of medical products. The operating business of the company, which has been listed on the stock exchange since October 2022, focuses on the distribution of innovative cold plasma technology for medical applications via its wholly owned subsidiary Viromed Medical GmbH. Viromed can draw on a broad customer base in the DACH region and beyond. Viromed is pursuing the goal of further advancing the use of cold plasma technology in medicine in the coming years and realizing the corresponding growth potential.

www.viromed-medical-ag.de

Contact Viromed

E-Mail: kontakt@viromed-medical.de
 

Press contact

E-mail: viromed@kirchhoff.de


04.11.2026 CET/CEST Dissemination of a Corporate News, transmitted by EQS News – a service of EQS Group.
The issuer is solely responsible for the content of this announcement.

The EQS Distribution Services include Regulatory Announcements, Financial/Corporate News and Press Releases.


Language: English
Company: Viromed Medical AG
Hauptstraße 105
25462 Rellingen
Germany
E-mail: kontakt@viromed-medical.de
Internet: https://www.viromed-medical-ag.de/
ISIN: DE000A40ZVN7
WKN: A40ZVN
Listed: Regulated Unofficial Market in Dusseldorf, Frankfurt, Hamburg, Tradegate BSX
LEI Code: 894500XR05MYCVCCR171
EQS News ID: 2428580

 
End of News EQS News Service

2428580  04.11.2026 CET/CEST

Lonza CEO Wolfgang Wienand to present at Jefferies Global Healthcare Conference

Basel, Switzerland, 4 November 2026 – Lonza is pleased to announce that it will participate in the Jefferies Global Healthcare Conference in London on 17 and 18 November 2026.

As part of the conference, Lonza CEO Wolfgang Wienand will host a presentation and subsequent Q&A on 18 November from 15:30 – 15:55 GMT (16:30 – 16:55 CET). 

The session will be webcast live, and a replay will be available on the Lonza Investor Relations website, remaining accessible for 90 days.

The live audio webcast can be accessed via this link.

BRAIN Biotech and Syngenta form strategic cooperation applying synthetic biology to drive innovations in sustainable farming solutions  

BRAIN Biotech AG

/ Key word(s): Alliance

BRAIN Biotech and Syngenta form strategic cooperation applying synthetic biology to drive innovations in sustainable farming solutions  

04.11.2026 / 10:45 CET/CEST

The issuer is solely responsible for the content of this announcement.


BRAIN Biotech and Syngenta form strategic cooperation applying synthetic biology to drive innovations in sustainable farming solutions

  • Strategic Cooperation uses synthetic biology to drive innovations for biologicals in agriculture
  • BRAIN Biotech provides access to its proprietary BioArchive and its expertise in advanced microbial strain development

ZWINGENBERG, Germany, November 4, 2026 – BRAIN Biotech AG, a leading provider of specialized enzymes and innovative biosolutions for industry, and Syngenta, a global leader in agricultural innovation, join forces to accelerate the development of biologicals.

The companies strategically combine BRAIN Biotech´s scientific expertise in synthetic biology with Syngenta’s global leadership in biologicals to support the development of next-generation agricultural solutions and address key challenges in sustainable farming.

BRAIN Biotech will contribute its advanced microbial strain development capabilities to support Syngenta’s research initiatives aimed at expanding the potential of novel biologicals. These nature-inspired tools shall complement traditional crop protection solutions and aim to enhance plant health, resilience and productivity.

“Partnering with a global leader like Syngenta underscores the importance of biological innovation for sustainable agriculture,” says Martin Langer, EVP at BRAIN Biotech. “We see this collaboration as a powerful way to combine our extensive industrial biotechnology toolset with Syngenta’s scale and field-driven insights to create solutions which positively impact farmers as well as the environment.”

“At Syngenta, we believe agricultural innovation unlocks the next-generation of sustainable farming solutions and are committed to working with partners who share the same vision,” says Jérôme Cassayre, Syngenta’s Head Biologicals Research. “We’re excited to work with BRAIN Biotech to push back the boundaries of science in agricultural biologicals.”

BRAIN Biotech offers highly specialized microbiological services, including the identification and optimization of strains to produce enzymes and biomolecules. The company’s extensive portfolio of biotechnological methods and customer-oriented service are key to delivering innovative, next-generation solutions and products. These are now combined under the new industrial biotechnology brand BRAINBiocatalysts. Customers also get access to the BRAIN BioArchive, a collection of tens of thousands of unique microbial strains and can search for specific strains tailored to their applications.

+++

Contact Investor Relations and Media

Martina Schuster

Phone: +49 6251 9331-69

Email: martina.schuster@brainbio.com


BRAINBiocatalysts

BRAINBiocatalysts is the industrial biotechnology brand of the BRAIN Biotech Group. BRAINBiocatalysts combines expertise across discovery, development, scale-up and manufacturing to deliver scalable bio-based solutions for complex challenges and specialty applications. Its capabilities span enzyme discovery and development, microbial strain development, bioprocess development, application expertise and commercial manufacturing.

BRAINBiocatalysts brings together the capabilities of BRAIN Biotech AG, Biocatalysts Ltd., Breatec B.V., WeissBiotech GmbH and Biocatalysts Inc. The respective legal entities provide products and services and enter into contractual relationships with customers. For more information visit: www.brainbiocatalysts.com


BRAIN Biotech Group

The BRAIN Biotech Group develops and provides bio-based products and solutions for industry, with activities organized in two business segments: BRAINBiocatalysts (development, production, and distribution of specialty enzymes, microorganisms, and ingredients) and BRAINBioIncubator (research-intensive development projects and pharmaceuticals). BRAIN Biotech AG, the parent company of the BRAIN Biotech Group operates its own fermentation facilities in the UK and has additional production sites in continental Europe and the US. BRAIN Biotech AG has been listed on the Frankfurt Stock Exchange since February 9, 2016 (Ticker: BNN; ISIN: DE0005203947 / WKN: 520394). In fiscal year 2024/25, the group generated revenue of €49.6 million with around 280 employees.

For more information, visit: www.brain-biotech-group.com


Disclaimer

This press release contains forward-looking statements. These statements reflect the current views, expectations, and assumptions of the management of BRAIN Biotech AG, and are based on information currently available to the management.

Forward-looking statements are no guarantees of future performance, and entail both known and unknown risks as well as uncertainties that could cause actual results, performance or events to differ materially from those expressed or implied in such statements. Numerous factors exist that could influence the future performance of and future developments at BRAIN Biotech AG and the BRAIN Biotech Group. Such factors include, but are not limited to, changes in the general economic and competitive environment, risks associated with capital markets, currency exchange rate fluctuations, changes in international and national laws and regulations, in particular with respect to tax laws and regulations, as well as other factors.

BRAIN Biotech AG does not undertake any obligation to update or revise any forward-looking statements.


04.11.2026 CET/CEST Dissemination of a Corporate News, transmitted by EQS News – a service of EQS Group.
The issuer is solely responsible for the content of this announcement.

The EQS Distribution Services include Regulatory Announcements, Financial/Corporate News and Press Releases.


Language: English
Company: BRAIN Biotech AG
Darmstädter Straße 34-36
64673 Zwingenberg
Germany
Phone: +49 (0) 62 51 / 9331-0
Fax: +49 (0) 62 51 / 9331-11
E-mail: ir@brainbio.com
Internet: www.brain-biotech-group.com
ISIN: DE0005203947
WKN: 520394
Listed: Regulated Market in Frankfurt (Prime Standard); Regulated Unofficial Market in Dusseldorf, Hamburg, Munich, Stuttgart, Tradegate BSX
LEI Code: 391200JKPVHLD6JLZ107
EQS News ID: 2428236

 
End of News EQS News Service

2428236  04.11.2026 CET/CEST

Pentixapharm Announces Progress of Pivotal Phase 3 PANDA Study; Will Host Third Quarter Webcast on November 12, 2026

Pentixapharm Holding AG

/ Key word(s): Study/Quarterly / Interim Statement

Pentixapharm Announces Progress of Pivotal Phase 3 PANDA Study; Will Host Third Quarter Webcast on November 12, 2026

04.11.2026 / 08:00 CET/CEST

The issuer is solely responsible for the content of this announcement.


Pentixapharm Announces Progress of Pivotal Phase 3 PANDA Study;

Will Host Third Quarter Webcast on November 12, 2026

  • No FDA comments on the study protocol submitted in August 2026, including the primary endpoints and the planned number of approximately 325 patients; central U.S. ethics committee has approved the protocol and consent documents
  • First patient enrollment continues to be expected in the first quarter of 2027; no material impact on the further study timeline expected
  • Webcast on third quarter and first nine months 2026 results and PANDA study on November 12, 2026, at 3 p.m. CET

Berlin, Germany, November 4, 2026 – Pentixapharm Holding AG (Frankfurt Prime Standard: PTP), a clinical-stage biotechnology company developing innovative radiopharmaceutical diagnostics and therapeutics, has completed further key preparations towards the start of its pivotal Phase 3 PANDA study. The Company will publish its third quarter and first nine months of 2026 results on Thursday, November 12, 2026.

The PANDA study is designed to enroll approximately 325 patients. More than ten U.S. study sites have already been selected. The study network includes leading academic medical centers across several U.S. regions and a specialized adrenal center. Clinical investigators have also been identified to conduct the study. Agreements with participating study sites are currently being negotiated so that the sites can open for patient enrollment.

The FDA confirmed in June 2026 that the PANDA study could proceed and granted Fast Track designation in July 2026. The FDA raised no concerns regarding the study protocol submitted in August 2026, which specifies the primary endpoints and the planned number of patients; the study will therefore be conducted on the basis of this protocol.

The central U.S. ethics committee (Institutional Review Board) has since approved the study protocol and the documents used to inform prospective participants about the study and obtain their consent.

As announced on October 19, 2026, first patient enrollment is expected in the first quarter of 2027. The Company currently does not expect any material impact on the further timeline of the study.

Pentixapharm welcomes participants to the webcast at 3:00 p.m. CET (2:00 p.m. GMT / 9:00 a.m. EST) on November 12, 2026. The Management Board will discuss the results, progress on PANDA, the remaining steps towards first patient enrollment and the wider development portfolio. Please register here: https://www.nuways-ag.com/events/pentixapharm-earnings-call-2026-11-12

A recording will be available following the event on the Company’s website under Investors.

About PANDA

PANDA is designed to evaluate [68Ga]Ga-PentixaFor PET/CT for non-invasive subtyping in adults with confirmed primary aldosteronism in a registrational setting. Primary aldosteronism is a common, underdiagnosed hormonal cause of high blood pressure. Patients with one-sided disease can benefit from surgical removal of the affected adrenal gland, but identifying them currently requires adrenal vein sampling, an invasive catheter procedure. Independent studies from Europe, Asia and Australia, most recently presented at EANM 2026, support the approach of PANDA. PentixaFor is an investigational radiopharmaceutical candidate.

About Pentixapharm

Pentixapharm Holding AG (Frankfurt Prime Standard: PTP) is a clinical-stage biotechnology company headquartered in Berlin developing innovative nuclear medicine approaches to help physicians detect, characterize, and treat serious diseases with greater precision. The Company’s compounds are designed to deliver medical radioisotopes specifically to defined biological targets – either for high-resolution molecular imaging or for targeted radioligand therapy.

Pentixapharm’s pipeline is built on CXCR4 targeting, a receptor with strong scientific validation across multiple disease areas, and encompasses diagnostic and therapeutic programs in endocrine, cardiovascular, and oncological indications. The lead program, [68Ga]Ga-PentixaFor, is being advanced into the registrational Phase 3 PANDA study for non-invasive subtyping of primary aldosteronism. The PENTHERA program advances the CXCR4-directed theranostic platform into Phase I/II in oncology. Complementing the CXCR4 platform, Pentixapharm is also developing a next-generation antibody platform targeting CD24, an emerging immune checkpoint in difficult-to-treat cancers.

Forward-looking statements

This press release contains forward-looking statements based on the current beliefs, expectations and assumptions of the management of Pentixapharm Holding AG. These statements relate to future events, prospects or financial performance and are subject to known and unknown risks, uncertainties and other factors that may cause actual results, performance or achievements to differ materially, positively or negatively, from those expressed or implied by such forward-looking statements. Such factors include, among others, changes in general economic and business conditions, competitive pressures, the introduction of competing products, limited market acceptance of new products, services or technologies, regulatory developments and changes in business strategy. Pentixapharm Holding AG does not undertake any obligation to update or revise any forward-looking statements, whether as a result of new information, future events or otherwise, except as required by applicable law. Forward-looking statements are made only as of the date of this press release. All information provided in this press release is supplied without any warranty or guarantee as to its accuracy, completeness or timeliness. To the extent permitted by law, Pentixapharm Holding AG assumes no liability in connection with this press release.

Pentixapharm Investor and Media Contact

Robert-Rössle-Straße 10

13125 Berlin

Germany

E-mail: ir@pentixapharm.com

Website: www.pentixapharm.com


04.11.2026 CET/CEST Dissemination of a Corporate News, transmitted by EQS News – a service of EQS Group.
The issuer is solely responsible for the content of this announcement.

The EQS Distribution Services include Regulatory Announcements, Financial/Corporate News and Press Releases.


Language: English
Company: Pentixapharm Holding AG
Robert-Rössle-Straße 10
13125 Berlin
Germany
E-mail: info@pentixapharm.com
Internet: https://www.pentixapharm.com/
ISIN: DE000A40AEG0
WKN: A40AEG
Listed: Regulated Market in Frankfurt (Prime Standard); Regulated Unofficial Market in Dusseldorf, Hamburg, Hanover, Munich, Stuttgart, Tradegate BSX
LEI Code: 3912005VBOVXNDXEQZ36
EQS News ID: 2427900

 
End of News EQS News Service

2427900  04.11.2026 CET/CEST