Akemi Hair Glow Claims Evaluated: 2026 Hair Growth Transparency Report Examines DHT-Blocking Spray Research, “56% Thicker Hair” Data, and Ingredient Research for Women
Akemi Hair Glow Claims Evaluated: 2026 Hair Growth Transparency Report Examines DHT-Blocking Spray Research, “56% Thicker Hair” Data, and Ingredient Research for Women


Akemi Hair Glow Claims Evaluated: 2026 Hair Growth Transparency Report Examines DHT-Blocking Spray Research, “56% Thicker Hair” Data, and Ingredient Research for Women

A 2026 informational report presenting Akemi Hair Glow’s DHT-blocking claims alongside ingredient-level research context, topical delivery positioning, pricing structure, and key verification factors for consumers evaluating hair growth products

Elizabeth, NJ, March 21, 2026 (GLOBE NEWSWIRE) — This article contains affiliate links. If a purchase is made through these links, a commission may be earned at no additional cost to the buyer. This report is an informational overview and does not constitute medical, health, or dermatological advice. All product details described below are stated as presented by the company and should be verified directly on the official website before any purchasing decision.

In this report, the term “effectiveness” refers strictly to how the product’s marketing language describes potential outcomes. It does not indicate that the finished product has been clinically proven effective, and no published clinical trial appears to evaluate Akemi Hair Glow as a proprietary formula.

As consumer interest in topical hair growth solutions continues to grow, Akemi Hair Glow has become one of the products generating attention across social media platforms and wellness communities. The product is marketed primarily toward women experiencing thinning hair, increased shedding, or visible scalp areas that have become harder to conceal over time.

The product’s claims around DHT blocking, follicle reactivation, and “56% thicker hair in 18 weeks” have contributed to that visibility. These types of claims are common in the topical hair growth category and are presented in this report alongside published ingredient-level research for contextual reference.

Akemi Hair Glow Claims Evaluated 2026 Hair Growth Transparency Report Examines DHT-Blocking Spray Research, "56% Thicker Hair" Data, and Ingredient Research for Women

This report presents how the product is described alongside relevant research considerations. It outlines what the product page states, what published ingredient-level research supports under controlled study conditions, and where gaps remain between the marketing language and clinical evidence for the finished product.

Current product details, pricing, and terms can be confirmed by viewing the current Akemi Hair Glow offer (official Akemi Hair Glow page).

Individual results vary. Topical hair products are not substitutes for professional dermatological evaluation, balanced nutrition, or medical treatment for diagnosed hair loss conditions. Consult a qualified healthcare provider before starting any new topical hair regimen, especially if you are experiencing sudden or severe hair loss.

What Is Akemi Hair Glow

Akemi Hair Glow is a topical hair spray positioned as a DHT-blocking formula designed to support hair growth, reduce shedding, and improve hair thickness and texture. The product is sold exclusively through its official website, which operates on a Shopify-based platform under the domain buyskyline.co. The product page describes it as a leave-in daily treatment that delivers active ingredients directly to the scalp rather than through oral ingestion.

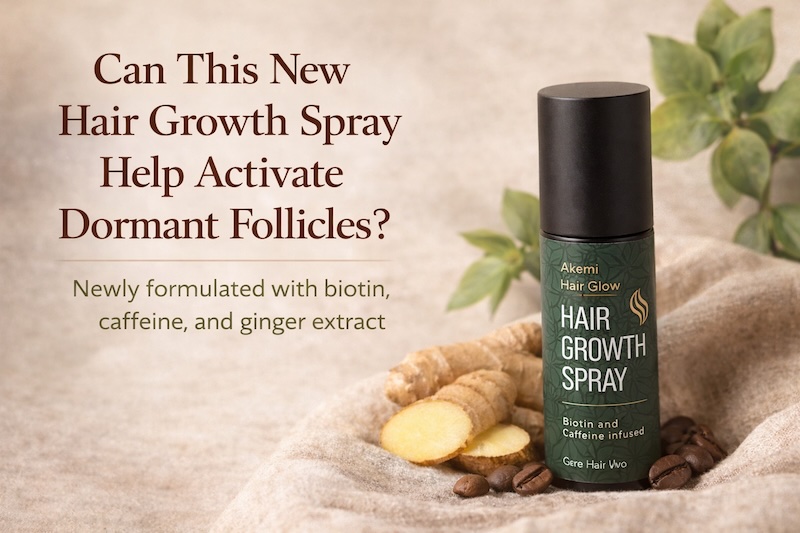
The product page lists six primary active ingredients: Biotin (Vitamin B7), Aminexil, Castor Oil, Caffeine, He Shou Wu (Fo-Ti), and Ginger Root Extract. The formula is described as free from parabens and toxins, with natural preservatives. The specific concentrations of each ingredient per application are not disclosed on the product page or in publicly available materials.

Per the FAQ section on the official website, orders ship from a New Jersey-based warehouse via USPS, FedEx, UPS, or DHL for international customers. The site states that products ship within 48 business hours, with standard delivery taking between five and seven days.

How Akemi Hair Glow Describes Its DHT-Blocking Mechanism

The product page centers its marketing narrative on DHT (dihydrotestosterone) as the primary driver of hair loss, particularly in women. The sales page describes DHT as a hormone that attacks hair follicles, shrinks them over time, and eventually causes them to stop producing hair entirely. The product is positioned as a formula that neutralizes DHT directly at the scalp.

The report presents what published dermatological research supports about that underlying mechanism, and where the product’s marketing language extends beyond the evidence currently available for this specific formula.

DHT’s role in androgenetic alopecia (pattern hair loss) is well established in published research. DHT is a metabolite of testosterone produced by the enzyme 5-alpha reductase. In genetically susceptible individuals, DHT binds to androgen receptors in hair follicles, triggering miniaturization — a process in which follicles progressively shrink and produce thinner, shorter hairs until they eventually stop producing visible hair. This mechanism is recognized across published dermatological literature as a primary contributor to pattern hair loss in both men and women.

The underlying science connecting DHT to hair loss is well documented. The question the report addresses is whether this specific spray, at its undisclosed ingredient concentrations, delivers DHT-blocking activity at levels sufficient to produce the results described on the product page.

Among the listed ingredients, caffeine has the most directly relevant published research regarding DHT interaction at the follicular level. A 2007 study published in the International Journal of Dermatology found that caffeine counteracted testosterone-suppressed hair growth in vitro — meaning in laboratory conditions using human hair follicles. A separate 2018 multicenter clinical study published in Skin Pharmacology and Physiology reported that a 0.2% caffeine-based topical liquid performed comparably to 5% minoxidil solution over six months in men with androgenetic alopecia. A February 2025 systematic review analyzing multiple clinical trials concluded that caffeine can stimulate cell growth and effectively penetrate hair follicles, positioning it as a promising topical agent against hair loss.

Those are meaningful findings. However, every one of those studies tested caffeine at disclosed concentrations under controlled conditions — not this specific multi-ingredient spray at undisclosed concentrations. The report references this research to provide context, not to confirm that Akemi Hair Glow replicates these study conditions.

He Shou Wu (Fo-Ti) is described on the product page as an ingredient that inhibits DHT production while stimulating dormant follicles. He Shou Wu has a long history in traditional Chinese medicine, and some in vitro studies have explored its effects on 5-alpha reductase activity. However, robust clinical trials specifically evaluating topical He Shou Wu for DHT blocking in human hair loss are limited in the published literature.

This is ingredient-level research context. Akemi Hair Glow as a finished product has not been evaluated in any published clinical trial appearing in peer-reviewed journals.

The “56% Thicker Hair in 18 Weeks” Claim: What the Data Trail Shows

The product page prominently states that customers see “56% thicker, fuller hair in just 18 weeks.” This figure appears alongside the name “Dr. Sarah Mitchell, Certified MD & Published Medical Author.”

The report outlines several questions about the evidence trail behind that number that consumers may want to consider before making a purchasing decision.

The source methodology behind the 56% figure is not disclosed on the product page. The site does not specify whether this number comes from a controlled clinical study, an internal consumer survey, instrument-measured trichoscopy data, or another methodology. Without knowing how the measurement was taken, what baseline was used, what sample size was involved, and whether a placebo control was included, the figure cannot be independently compared against published hair growth research standards.

The association with “Dr. Sarah Mitchell” is presented without a linked publication, institutional affiliation, or verifiable credential trail on the product page. Consumers researching this endorsement may want to verify the clinician’s credentials independently before weighting the claim.

For context, published clinical research on topical caffeine — one of the listed ingredients — has measured outcomes such as anagen-to-telogen hair ratios, hair density counts, and hair pull test improvements over periods of three to six months. These studies report improvements using standardized trichogram measurements. A claim of “56% thicker hair” would represent a notably strong result if measured by those clinical standards. That does not mean the claim is false — it means the methodology matters, and that methodology is not publicly available.

The product page also references a “93% success rate” and a “95% success rate” in different sections of the same page and describes “3,758 Reviews,” “3,000+ Verified Reviews,” and “4,000+ satisfied users” in different locations. These inconsistencies across the same product page are worth noting when forming an assessment of the company’s published statistics.

Akemi Hair Glow Ingredient Profile: What Published Research Supports at the Compound Level

The product page lists six primary active ingredients. The report outlines what published research supports at the individual ingredient level — and why the distinction between ingredient research and finished-product evidence matters when evaluating this type of product.

Biotin (Vitamin B7): Biotin is one of the most widely marketed hair growth ingredients. However, a 2024 systematic review published in the Journal of Clinical and Aesthetic Dermatology examined published clinical studies specifically on biotin for hair growth and found that the highest-quality study — a double-blind, placebo-controlled trial — showed no difference between the biotin and placebo groups for hair growth. The review concluded that a significant gap exists between public perception of biotin’s effectiveness and what the scientific literature actually demonstrates. Akemi Hair Glow uses topical rather than oral biotin, and the product page suggests this delivery method bypasses digestive limitations. Limited published data exists on topical biotin specifically for hair growth outcomes.

Caffeine: Among the listed ingredients, caffeine has the strongest published clinical evidence for topical hair applications. In vitro research demonstrated that caffeine counteracts DHT-induced hair follicle miniaturization and stimulates follicle proliferation. A clinical study found that a 0.2% caffeine topical performed comparably to 5% minoxidil over six months. A 2025 systematic review further supported caffeine’s potential as a topical hair loss agent while noting that most studies suffered from methodological limitations and significant gender imbalance — only 4.4% of study participants were women, despite hair loss products frequently targeting female consumers. The concentration of caffeine in Akemi Hair Glow is not disclosed, making direct comparison to published study dosages impossible.

Aminexil: The product page describes aminexil as an anti-fibrosis molecule that prevents DHT from stiffening tissue around follicles. Aminexil was originally developed and studied primarily by L’Oréal for their Dercos line. Published research has explored its potential to prevent collagen hardening around hair follicles, which can contribute to permanent hair loss. The published research base is more established than that of some cosmetic hair ingredients, though independent replication outside company-sponsored studies remains limited.

Castor Oil: Castor oil is a traditional hair care ingredient valued for its moisturizing properties. Ricinoleic acid in castor oil has demonstrated anti-inflammatory properties in published research. However, clinical evidence specifically connecting topical castor oil application to measurable hair regrowth is sparse. Most evidence supporting castor oil’s hair benefits comes from traditional use and mechanistic reasoning rather than controlled clinical trials.

He Shou Wu (Fo-Ti): This traditional Chinese medicinal herb has been used for centuries for hair-related concerns, including premature graying and hair loss. Some in vitro research has explored its effects on 5-alpha reductase activity and hair follicle cell proliferation. Clinical evidence for topical He Shou Wu in human hair growth remains limited. Additionally, oral He Shou Wu preparations have been associated with hepatotoxicity concerns in published case reports, though topical application presents a different risk profile.

Ginger Root Extract: Ginger has published evidence supporting anti-inflammatory and circulatory-boosting properties. Improved scalp microcirculation can theoretically support healthier follicle environments. However, one study published in the Journal of Dermatological Science suggested that 6-gingerol, an active compound in ginger, may actually suppress hair growth in certain contexts. The relationship between ginger and hair growth appears more nuanced than the product’s marketing suggests.

These are individual ingredient findings from published research. They do not represent clinical outcomes for Akemi Hair Glow as a finished formula, and individual experiences differ based on the type and cause of hair loss, genetics, hormonal factors, and consistency of use.

Topical Delivery: Does the Spray Format Offer a Research-Supported Advantage

The product page emphasizes that topical application delivers ingredients “directly to your follicles” rather than having them “dissolved in stomach acid where 97% of supplements get destroyed.” This positioning is central to how the product differentiates itself from oral hair supplements.

Published evidence supports the general principle that hair follicles serve as an effective penetration pathway for topically applied substances. Research published in the British Journal of Clinical Pharmacology demonstrated that caffeine penetrates hair follicles rapidly when applied topically and that follicular accumulation of caffeine can be significantly higher than in surrounding skin tissue. For topical caffeine specifically, the delivery advantage has a documented scientific basis.

The “97% destroyed in stomach acid” claim, however, oversimplifies digestive pharmacology. Oral bioavailability varies dramatically between compounds. Biotin, for example, is well absorbed orally — it has high oral bioavailability, which makes the blanket claim misleading when applied to that specific ingredient.

The more practical question is whether these specific ingredients, at their undisclosed concentrations, penetrate the scalp in sufficient quantities to produce the effects described on the product page. Without disclosed ingredient concentrations, this question cannot be answered from publicly available information.

Pricing, Guarantee, and Purchase Details

The product page includes promotional pricing ranges and inventory-related messaging that may vary over time and are determined by the company. At the time of this report, the site references “up to 50% off” and “up to 60% off” in different sections of the same page. Specific pricing tiers are presented during checkout.

The product page describes a 30-day money-back guarantee. The FAQ section states: “Simply return your product and we’ll give you a refund for 30 days.” However, the Terms of Service page includes a disclosure about a minimum 15% restocking fee for returns. This gap between the marketing presentation and the published terms is worth clarifying directly with the company before ordering.

The Terms of Service identify the operator as “Akemi Hair Glow” and reference New York for certain legal proceedings, while the Governing Law section references the laws of Florida.

Verify current pricing, refund terms, and restocking fee details before purchasing by viewing the current Akemi Hair Glow offer (official Akemi Hair Glow page).

What the Product Page Presents as Customer Feedback

The official website includes customer testimonials describing outcomes such as visible new hair growth at the crown and temples, reduced shedding, bald patches filling in, and hair texture improvements within specific timeframes. These testimonials are attributed to named individuals with locations.

The company does not disclose whether these testimonials reflect typical outcomes or controlled evaluation conditions. The site does not describe how testimonials are collected or verified, or whether any compensation or incentive is provided. People who write reviews are self-selected — satisfied customers are significantly more likely to post feedback than those with neutral or negative experiences.

Testimonial results are individual experiences and should not be interpreted as typical or guaranteed outcomes. Published research on hair growth interventions consistently shows that individual outcomes depend on the specific type and cause of hair loss, genetic factors, hormonal profiles, and treatment consistency.

Consumer Considerations Based on Product Characteristics

This section outlines general considerations based on the product’s described characteristics and publicly available ingredient research. It is not intended to recommend the product for any individual use case.

Consumers evaluating topical products may consider Akemi Hair Glow if they:

Prefer topical over oral approaches. Consumers who have tried oral hair supplements without satisfaction or who prefer applying active ingredients directly to the scalp may find a spray format aligns with their preference. Published research on topical caffeine shows promising follicular penetration and, in at least one clinical study, comparable performance to topical minoxidil.

Are exploring complementary options alongside professional care. Consumers already working with a dermatologist and looking for an additional daily topical may note that ingredients like caffeine and aminexil have published research supporting their potential roles in hair maintenance as part of a broader approach.

Want a non-prescription starting point. For consumers not yet ready for prescription options like minoxidil or finasteride, a botanical-based spray may serve as an entry point while evaluating whether professional intervention is appropriate for their specific situation.

Other options may be preferable for consumers who:

Require transparent ingredient dosing. Consumers who need exact ingredient concentrations for comparing against published research dosages should note that those amounts are not disclosed for this product.

Are experiencing sudden or severe hair loss. Rapid, sudden, or patchy hair loss may indicate medical conditions such as alopecia areata, thyroid disorders, or nutritional deficiencies that require professional diagnosis and treatment rather than a topical cosmetic product.

Expect results comparable to FDA-approved treatments. Published clinical data for FDA-approved treatments like minoxidil represents a different evidence standard than what is currently available for this proprietary formula.

Questions to consider before purchasing: What type of hair loss am I experiencing, and has a healthcare professional evaluated the cause? Have I researched the specific ingredients at the concentrations used in published studies? Am I comfortable purchasing a product without disclosed ingredient concentrations? Have I reviewed the full refund terms, including the potential restocking fee? These questions can help determine whether this product’s characteristics match a consumer’s specific situation.

Verification Checklist: What to Confirm Before Ordering

Verify ingredient concentrations. Individual ingredient amounts per application are not disclosed. If dosing relative to published research matters to the purchasing decision, contact the company directly to request this information.

Separate ingredient research from product research. Published studies on caffeine, biotin, aminexil, and other listed ingredients examined those compounds individually under controlled conditions. No published clinical trial appears to evaluate Akemi Hair Glow as a finished proprietary formula.

Trace the “56% thicker hair” claim. The methodology behind this figure is not publicly disclosed. Without understanding the measurement approach, sample size, and control conditions, the number cannot be independently verified against clinical standards.

Review the refund terms carefully. The product page describes a 30-day money-back guarantee, but the Terms of Service disclose a potential 15% restocking fee. Clarify exact refund terms before purchasing.

Consult a healthcare provider. This is especially important for women experiencing hair loss related to hormonal changes, thyroid conditions, nutritional deficiencies, or medical treatments. A dermatologist can help identify the underlying cause and recommend evidence-based interventions. This report is not a substitute for professional dermatological evaluation.

Consumer Questions About Akemi Hair Glow

What is DHT, and why does it matter for hair loss?

DHT (dihydrotestosterone) is a hormone derived from testosterone that plays a recognized role in androgenetic alopecia (pattern hair loss). In genetically susceptible individuals, DHT causes hair follicle miniaturization over time. This mechanism is well established in published dermatological research. However, not all hair loss is DHT-related — conditions such as telogen effluvium, alopecia areata, and nutritional deficiency-related hair loss involve entirely different mechanisms.

Is Akemi Hair Glow a medication or FDA-approved product?

Akemi Hair Glow is marketed as a topical cosmetic hair spray, not a medication. It is not FDA-approved for the treatment of any medical condition. The product has not undergone the clinical trial process required for FDA drug approval. Consumers seeking FDA-approved treatments for hair loss should discuss options such as topical minoxidil or oral finasteride with a qualified healthcare provider.

How does the caffeine in this product compare to what has been studied clinically?

Published clinical research has tested topical caffeine at disclosed concentrations — for example, a 0.2% caffeine-based topical that performed comparably to 5% minoxidil over six months. Akemi Hair Glow’s caffeine concentration is not disclosed, which makes direct comparison to published study dosages impossible. These are different products with different evidence bases, even though they share a common ingredient.

Will this product address my type of hair loss?

The product’s marketing focuses on DHT-related hair thinning. If hair loss has a different underlying cause — such as thyroid dysfunction, iron deficiency, autoimmune conditions, or medication side effects — a topical DHT-blocking spray may not address the underlying issue. A healthcare professional can help determine the cause and recommend appropriate interventions.

How long before results might be noticed?

The product page describes a week-by-week timeline suggesting changes beginning at week 2 and continuing through week 16. Published research on topical hair growth interventions generally suggests that meaningful results, when they occur, typically require three to six months of consistent use. Individual timelines depend on factors including the type of hair loss, its severity, and individual biological response. The timeline descriptions on the product page represent the company’s marketing positioning rather than clinically validated benchmarks for this specific product.

What should consumers know about the return policy before ordering?

The product page describes a 30-day money-back guarantee. The Terms of Service page discloses a potential minimum 15% restocking fee for returns. The exact refund process, applicable fees, and return shipping requirements should be confirmed directly before purchasing. Complete terms are available by viewing current Akemi Hair Glow terms (official Akemi Hair Glow page).

Summary of Key Considerations

Akemi Hair Glow is a topical hair spray positioned around DHT-blocking and follicle reactivation claims. The product contains ingredients — particularly caffeine — with published research at the individual compound level supporting potential hair growth benefits when applied topically. Caffeine’s evidence base is the strongest among the listed ingredients, with at least one clinical study demonstrating comparable performance to topical minoxidil at a disclosed concentration.

However, the finished formula has not been evaluated as a proprietary product in any published clinical trial. Ingredient concentrations are not disclosed. Several marketing statistics — including “56% thicker hair in 18 weeks,” a “93% success rate,” and varying review counts across the same page — lack publicly available methodology or present inconsistencies.

The biotin evidence is notably weaker than the product’s positioning suggests. A 2024 systematic review found no difference between biotin and placebo for hair growth in the highest-quality study examined. This is worth factoring into any evaluation of the formula’s overall research backing.

The product page describes a 30-day money-back guarantee, though the Terms of Service disclose a potential 15% restocking fee. Orders ship from a New Jersey warehouse, and customer support is available by email and phone.

Consumers who have completed their own research and want to review the full product details can do so. Complete product details, current pricing, and published terms are available by viewing the current Akemi Hair Glow offer (official Akemi Hair Glow page).

Contact Information

Per the company’s published contact information:

Company: Akemi Hair Glow

Email: support@buyskyline.co

Phone: +1 (866) 697-4823

Shipping Origin: New Jersey, United States (per the company’s FAQ)

Website: buyskyline.co

Disclaimers

Content and Consumer Information Disclaimer: This article is an informational overview and does not constitute medical, health, or dermatological advice. All product details, ingredient information, pricing, and policy terms described in this article are stated as presented on the publicly available product website and related materials. Readers are encouraged to verify all claims directly on the official website and to consult a qualified healthcare professional before beginning any topical hair treatment.

Health and Hair Loss Notice: Hair loss can result from numerous causes, including genetics, hormonal changes, nutritional deficiencies, medical conditions, medications, and stress. Topical cosmetic products are not substitutes for professional dermatological evaluation and treatment. If you are experiencing sudden, severe, or patchy hair loss, consult a healthcare provider. Individual results vary based on factors including the type and cause of hair loss, genetic factors, hormonal profiles, consistency of use, and other individual variables. While some consumers report improvements, results are not guaranteed.

Results May Vary: Individual results will vary based on numerous factors, including age, baseline hair condition, the type and underlying cause of hair loss, hormonal factors, consistency of use, genetic factors, current medications, and other individual variables. While some customers report improvements, results are not guaranteed. People who write reviews are self-selected — satisfied customers are more likely to post feedback than those with neutral or negative experiences.

FTC Affiliate Disclosure: This article contains affiliate links. If a product is purchased through these links, a commission may be earned at no additional cost to the buyer. This compensation does not influence the accuracy, neutrality, or integrity of the information presented. All descriptions are based on published research and publicly available information from the company’s official website.

Pricing Disclaimer: All prices, discounts, and promotional offers mentioned were based on information published on the official product website at the time of publication (March 2026) and are subject to change without notice. Always verify current pricing, promotions, and terms on the official website before making a purchase.

Publisher Responsibility: The publisher of this article has made every effort to ensure accuracy at the time of publication. The publisher does not accept responsibility for errors, omissions, or outcomes resulting from the use of the information provided. Readers are encouraged to verify all details directly with the company and their healthcare provider before making decisions.

CONTACT: Email: support@buyskyline.co
Phone: +1 (866) 697-4823

Prothena Partners Present Data Supporting Next Generation Treatments for Parkinson’s and Alzheimer’s Disease at AD/PD™ 2026

Prothena Partners Present Data Supporting Next Generation Treatments for Parkinson’s and Alzheimer’s Disease at AD/PD™ 2026


Prothena Partners Present Data Supporting Next Generation Treatments for Parkinson’s and Alzheimer’s Disease at AD/PD™ 2026

DUBLIN–(BUSINESS WIRE)–$PRTA #Prothena–Prothena Corporation plc (NASDAQ:PRTA), a late-stage clinical biotechnology company with a robust pipeline of investigational therapeutics built on protein dysregulation expertise, today announced partner presentations on clinical updates from prasinezumab for the treatment of Parkinson’s disease and BMS-986446 for the treatment of Alzheimer’s disease at the International Conference on Alzheimer’s and Parkinson’s Diseases and Related Neurological Disorders (AD/PD™ 2026) in Copenhagen, Denmark, and online.


Roche Presentations on Prasinezumab for the Potential Treatment of Parkinson’s Disease

Industry Symposium – Pathways to Progress: Exploring Innovations in AD and PD for Future Practice

  • Chair: Malú G. Tansey, Ph.D., Indiana University School of Medicine Stark Neuroscience Institute
  • Date: Tuesday March 17, 2026

This symposium reviewed the evolving understanding of the molecular pathophysiology and disease heterogeneity across Alzheimer’s disease (AD) and Parkinson’s disease (PD). Explored current and emerging treatment pathways, e.g. alpha-synuclein, amyloid-beta and neuroinflammation; including diagnostics, biomarkers and therapeutics. Considered how the advancing understanding of AD and PD informs innovative clinical development approaches and clinical practice.

Oral Presentation – Modeling Parkinson’s Disease Progression to Quantify Long-Term Treatment Effects via the Concept of ‘Time Saved’

  • Presenter: Benjamin Ribba, Roche
  • Date: Thursday March 19, 2026

The comparison of PASADENA open-label extension (OLE) data with PPMI-based model predictions supports potential disease-modifying efficacy with an estimated two years of ‘time saved’ providing an intuitive measure of long-term benefit. The observed PASADENA OLE outcomes consistently deviated from the model-predicted progression, suggesting a sustained treatment effect. On average, participants were approximately two years less advanced in disease severity five years after the start of the trial compared to the virtual comparator.

Oral Presentation – Prasinezumab in Early-Stage Parkinson’s Disease: Additional Data from the PADOVA Study

  • Presenter: Tania Nikolcheva, M.D., Ph.D., Roche
  • Date: Saturday March 21, 2026

Longer term data from the PADOVA OLE study in early-stage PD showed a sustained effect of prasinezumab in slowing Parkinson’s progression on top of effective symptomatic therapies. The totality of the evidence suggests a possible clinical benefit of prasinezumab and informed the initiation of the Phase III PARAISO study.

Poster Presentation – Prasinezumab’s Impact on Neuromelanin- and Iron-Sensitive MRI Biomarkers in Parkinson’s Disease: Findings from the PADOVA Phase IIb Study

Exploratory biomarker analysis of PADOVA suggests that prasinezumab is biologically active. This is supported by imaging biomarkers crucial to PD pathology, showing a slowing in the progressive loss of neuromelanin signal in substania nigra pars compacta and reduced iron accumulation in the putamen.

Poster Presentation – Sustained Effect on Prasinezumab on Parkinson’s Disease Motor Progression in the Open-Label Extension of the PASADENA Trial, 5-Year Update

At Year 5, the combined PASADENA arm (delayed- and early-start groups) showed less disease progression compared to the PPMI cohort. This lower progression was observed across multiple measures.

Poster Presentation – Digital Health Technology Detects Group Differences in Practically-Defined OFF L-DOPA State: Results of PADOVA Phase IIb Study of Prasinezumab

Post-hoc Digital Health Technology analyses showed consistent trends favoring prasinezumab in digital data collected in the practically-defined OFF L-DOPA state, in line with the PASADENA Phase 2a Simple Sum digital finding and clinical PADOVA readout.

Bristol Myers Squibb Presentation on BMS-986446 for the Potential Treatment of Alzheimer’s Disease

Oral Presentation – Randomized, Double-Blind, Placebo-Controlled Study Evaluating Safety, Tolerability, Pharmacokinetics, and Immunogenicity of BMS-986446 in Healthy Participants, Including Those of Japanese Ethnicity

  • Presenter: Ilena George, M.D., Bristol Myers Squibb
  • Date: Saturday March 21, 2026

Single-dose BMS-986446 was safe and well tolerated in all participants, including those of Japanese ethnicity. Plasma exposure of BMS-986446 increased dose proportionally. No anti-drug antibodies were detected. These results support BMS-986446 dosing in ongoing clinical studies without adjustments for Japanese ethnicity.

About Prasinezumab

Prasinezumab is an investigational monoclonal antibody designed to bind aggregated alpha-synuclein and thereby reduce neuronal toxicity. By reducing the build-up of alpha-synuclein protein in the brain, prasinezumab can potentially prevent further accumulation and spreading between cells, which may slow progression of the disease.

About Parkinson’s Disease

Parkinson’s disease is a chronic, progressive and debilitating neurodegenerative disease characterized by the gradual loss of neurons that make dopamine and other nerve cells. Today, Parkinson’s disease affects over 10 million people worldwide. The prevalence of Parkinson’s disease is increasing, and it has become one of the fastest-growing neurological disorders. Currently, symptomatic treatments that effectively alleviate motor symptoms are available. However, no therapies slow down or stop the clinical progression of Parkinson’s disease.

About BMS-986446

BMS-986446 is a humanized monoclonal antibody that targets multiple domains of the microtubule binding region of tau, a highly pathogenic tau fragment associated with neurofibrillary tangle formation and cognitive decline in Alzheimer’s disease. BMS-986446 binds to specific regions of the tau protein (R1–R3 within the microtubule-binding domain) to stop cell-to-cell spread of tau and tau uptake into cells. It also activates microglia—the brain’s immune cells—through its Fc receptor function, promoting the clearance of tau via phagocytosis.

About Alzheimer’s Disease

Alzheimer’s disease is a progressive, multifaceted and devastating neurodegenerative disease and the most common type of dementia in adults. Changes in the brain disrupt communication between neurons, impacting memory, cognition and behavior. As a result, Alzheimer’s disease has a significant impact on the day-to-day lives of those it directly affects, as well as on their families, caregivers and friends, resulting in considerable shifts in interpersonal relationships. There remains a critical need for disease-modifying therapies that can slow or delay the progression of Alzheimer’s disease as well as therapies that manage and ease neurobehavioral symptoms.

About Prothena

Prothena Corporation plc is a late-stage clinical biotechnology company with expertise in protein dysregulation with the potential to change the course of devastating neurodegenerative and rare peripheral amyloid diseases. Fueled by its deep scientific expertise built over decades of research, Prothena is advancing a pipeline of therapeutic candidates for a number of indications and novel targets for which its ability to integrate scientific insights around neurological dysfunction and the biology of misfolded proteins can be leveraged. Prothena’s pipeline includes both wholly-owned and partnered programs being developed for the potential treatment of diseases including Parkinson’s disease, ATTR amyloidosis with cardiomyopathy, Alzheimer’s disease, Amyotrophic lateral sclerosis (ALS) and a number of other neurodegenerative diseases. Prothena is developing and applying CYTOPE®, a novel technology that incorporates a cell-internalizing domain to drive efficient cytosolic delivery with highly specific macromolecular effectors. For more information, please visit the Company’s website at www.prothena.com and follow the Company on X (formerly Twitter) @ProthenaCorp.

Forward-Looking Statements

This press release contains forward-looking statements. These statements relate to, among other things, the treatment potential, designs, proposed mechanisms of action, and potential administration of prasinezumab and BMS-986446; and the continued advancement of our preclinical and clinical pipeline, including the potential and advancement of CYTOPE. These statements are based on estimates, projections and assumptions that may prove not to be accurate, and actual results could differ materially from those anticipated due to known and unknown risks, uncertainties and other factors, as well as those described in the “Risk Factors” sections of our Annual Report on Form 10-K filed with the Securities and Exchange Commission (SEC) on February 27, 2026, and discussions of potential risks, uncertainties, and other important factors in our subsequent filings with the SEC. We undertake no obligation to update publicly any forward-looking statements contained in this press release as a result of new information, future events, or changes in our expectations.

Contacts

Mark Johnson, CFA

Senior Vice President, Head of Investor Relations and Corporate Communications

650-837-8550

IR@prothena.com
Media@prothena.com

IBA Launches myQAMatriXXAiR to Advance Patient‑Specific QA in Particle Therapy

IBA Launches myQAMatriXXAiR to Advance Patient‑Specific QA in Particle Therapy


IBA Launches myQAMatriXXAiR to Advance Patient‑Specific QA in Particle Therapy

The first wireless ionization chamber array for particle therapy PSQA delivers fast, reliable verification for proton and carbon ion treatments, supporting emerging techniques such as FLASH and ARC radiotherapy.

Louvain-la-Neuve, Belgium, 21 March 2026 – IBA (Ion Beam Applications S.A., EURONEXT), the world leader in particle accelerator technology and a global provider of dosimetry and quality assurance (QA) solutions, announces the launch of myQA MatriXX AiR, a high resolution, wireless 2D ionization chamber array designed for patient specific quality assurance (PSQA) in particle therapy, including proton and carbon ion treatments. This new solution further demonstrates IBA’s role as a driving force in innovation and reinforces its leadership in quality assurance for particle‑beam therapies.

In radiation therapy, quality assurance is the critical safeguard that ensures increasingly complex treatments are delivered exactly as planned, translating technological innovation into clinical confidence and patient safety.

As particle therapy expands worldwide, proton and carbon‑ion centers are adopting increasingly advanced treatment modalities—from high‑precision conformal therapies to emerging beam‑delivery techniques such as FLASH and ARC radiotherapy. These innovations accelerate clinical potential, but they also elevate the need for robust, future‑proof quality assurance. With myQA MatriXX AiR, IBA sets a new industry standard as the first QA solution capable of providing full patient‑specific verification across both established particle‑therapy treatments and next‑generation delivery modes. This is not an adaptation to change—it is a decisive step forward, positioning IBA as the only partner offering uncompromised QA readiness for the future of particle‑beam therapy.

Engineered specifically for the demands of modern particle therapy, myQA MatriXX AiR features 1,521 high resolution air‑vented ionization chambers and delivers a complete 2D dose distribution readout within seconds, enabling fast and reliable verification of increasingly complex treatment plans. As treatment techniques become more complex, this level of detail directly translates into greater confidence, higher patient safety, and more reliable plan approval.

As the first commercially available wireless detector array for particle therapy PSQA, MatriXX AiR simplifies setup, reduces QA time, and integrates easily into existing workflows. The system is also FLASH‑ready, with hardware designed to support ultra‑high dose rate measurements as clinics prepare for next‑generation radiotherapy techniques.

Commenting on the solution Dr. Loïc Grevillot, Deputy Head of Medical Physics at the MedAustron Ion Therapy Center, stated: “We have been pleased to test the latest MatriXX detector from IBA Dosimetry, which offers a user-friendly interface and — most importantly for our needs —compatibility with carbon ion beams. The detector has demonstrated robust performance in dose evaluation and shows potential for clinical implementation in carbon ion therapy beam delivery QA and PSQA workflows.”

Jean-Marc Bothy, President of IBA Dosimetry, added: “We are proud to offer products that significantly enhance safety in patient plan deliveries in proton therapy. It’s exciting to see a QA solution designed specifically for particle therapy. MatriXX AiR builds on the strengths of our trusted MatriXX  PT and ONE, bringing a new level of precision, speed, and wireless flexibility. We’ve worked closely with clinical teams, and this evolution reflects their needs of effective QA that keeps pace with the future of treatment modalities. With this launch, we further strengthen our leadership in quality assurance for particle‑beam therapies.”

***ENDS***

About IBA

IBA (Ion Beam Applications S.A.) is the world leader in particle accelerator technology. The company is the leading supplier of equipment and services in the fields of proton therapy, considered as one of the most advanced forms of radiation therapy available today, as well as industrial sterilization, radiopharmaceuticals and dosimetry. The company, based in Louvain-la-Neuve, Belgium, employs approximately 2,100 people worldwide. IBA is a certified B Corporation (B Corp) meeting the highest standards of verified social and environmental performance.

IBA is listed on the pan-European stock exchange EURONEXT (IBA: Reuters IBAB.BR and Bloomberg IBAB.BB).

More information can be found at: www.iba-worldwide.com

CONTACTS

Olivier Lechien
Corporate Communication Director
+32 10 475 890
communication@iba-group.com

Attachment

Financière de Tubize – Annual report 2025

Financière de Tubize – Annual report 2025


Financière de Tubize – Annual report 2025

Annual Report 2025
Regulated information 21 March 2026

The board of directors of Financière de Tubize has established the 2025 annual report. This report is available on the website www.financiere-tubize.be

  • Profit for the financial year: € 93.2 million (€ 90.1 million in 2024, + 3.5%),
  • Full repayment of bank debt in May 2025 (€ 41.3 million at 31 December 2024),
  • Acquisition, in 2025, of 24,487 UCB shares, at an average price of €178.27, for a total amount of €4.4 million, increasing the holding of the Company in UCB from 36.27% on 31 December 2024 to 36.28% on 31 December 2025.

If the general shareholders meeting of 24 April 2026 approves the 2025 annual accounts, including the proposed result appropriation, a gross dividend of € 1.08 (compared to €1.04 for financial year 2024, an increase of 3.85%) will be payable as from 11 May 2026 onwards at the offices, seats and branches of BNP Paribas Fortis, in exchange of coupon n° 21.

Ex-dividend            7 May 2026
Record date           8 May 2026
Payment date        11 May 2026

Omeros Announces Upcoming Presentation at EBMT 2026 Highlighting Advances in TA-TMA Treatment

Omeros Announces Upcoming Presentation at EBMT 2026 Highlighting Advances in TA-TMA Treatment


Omeros Announces Upcoming Presentation at EBMT 2026 Highlighting Advances in TA-TMA Treatment

SEATTLE–(BUSINESS WIRE)–Omeros Corporation (NASDAQ: OMER) today announced that it will host an industry session at the 52nd Annual Meeting of the European Society for Blood and Marrow Transplantation (EBMT) on March 22, 2026, from 2:30pm – 4pm CET in Madrid, Spain. The session, titled “Advances in TA-TMA Treatment: Evaluating the Role of a Novel Targeted Therapy,” will be co-chaired by Rafael Duarte, MD, PhD, Hospital Universitario Puerta de Hierro Majadahonda, and Mohamad Mohty, MD, PhD, Hôpital Saint-Antoine, AP-HP, and will feature speakers Miguel-Angel Perales, MD, Memorial Sloan Kettering Cancer Center, and Michelle L. Schoettler, MD, MS, Children’s Healthcare of Atlanta.


The EBMT Annual Meeting is a key event for professionals in transplantation and cellular therapy, bringing together stakeholders from around the world.

About Omeros Corporation

Omeros is an innovative, commercial-stage biotechnology company that discovers and develops first-in-class protein and small-molecule therapeutics for large-market and orphan indications, with particular emphasis on complement-mediated diseases, cancers, and addictive or compulsive disorders. Omeros’ lead lectin pathway inhibitor YARTEMLEA® (narsoplimab-wuug), which inhibits the pathway’s effector enzyme MASP-2, is FDA-approved and commercially available in the U.S. for the treatment of hematopoietic stem cell transplant-associated thrombotic microangiopathy (TA-TMA) in adult and pediatric patients aged two years and older. A marketing authorization application for YARTEMLEA in TA-TMA is currently under review at the European Medicines Agency. OMS1029, Omeros’ long-acting MASP-2 inhibitor, has successfully completed Phase 1 clinical trials.

Under a recently announced asset purchase and licensing agreement, Novo Nordisk acquired global rights to zaltenibart (formerly OMS906), a MASP-3 inhibitor in clinical development for PNH and other alternative pathway indications, along with associated intellectual property and related assets. Omeros’ pipeline also includes OMS527, a phosphodiesterase 7 inhibitor in clinical development for cocaine use disorder that is fully funded by the National Institute on Drug Abuse, as well as a growing portfolio of novel molecular and cellular oncology programs. For more information about Omeros and its programs, visit www.omeros.com.

Contacts

Jennifer Cook Williams

Cook Williams Communications, Inc.

Investor and Media Relations

IR@omeros.com

Curi Bio and Battelle Announce Strategic Partnership to Accelerate the Adoption of New Approach Methods in Neuromuscular Pharmacology

Curi Bio and Battelle Announce Strategic Partnership to Accelerate the Adoption of New Approach Methods in Neuromuscular Pharmacology


Curi Bio and Battelle Announce Strategic Partnership to Accelerate the Adoption of New Approach Methods in Neuromuscular Pharmacology

SEATTLE & COLUMBUS, Ohio–(BUSINESS WIRE)–#BioTech–Curi Bio, a world leader in human-relevant 3D tissue technology, and Battelle, the world’s largest independent nonprofit research and development organization, today announced a strategic partnership to commercialize next-generation Neuromuscular Junction (NMJ) assay technology. This collaboration combines Curi Bio’s innovative microphysiological systems with Battelle’s extensive GLP-ready infrastructure to offer a transformative, human-relevant alternative to traditional pharmacology methodologies.


Advancing the 3Rs: At the core of this partnership is a commitment to the 3Rs (Replacement, Reduction, and Refinement). By providing functional, 3D human-tissue models that replicate complex neuromuscular physiology, the Curi Bio-Battelle partnership enables researchers to replace long-standing animal-based bioassays with highly reproducible new approach methods.

Innovation Meets Infrastructure: The partnership leverages Curi Bio’s proven Mantarray™ ecosystem which provides real-time functional readouts of human heart, muscle, and neuromuscular junction contractility. By offering a ready-to-use commercial solution for BoNT potency, Curi Bio enables customers to generate data with their specific drug products, accelerating the adoption of human-relevant, cell-based NMJ assays. Battelle serves as the scale and integration partner, offering the regulatory expertise and specialized laboratory facilities necessary to implement these technologies across global drug substance and medical countermeasure (MCM) development pipelines.

“We are excited to partner with Battelle to scale our technology for applications with significant commercial and governmental utility,” said Elliot Fisher, Chief Business Officer and Co-Founder of Curi Bio. “With the FDA’s new Non-Animal Models (NAMs) guidance and the NIH’s $150M commitment to human-relevant alternatives, the regulatory landscape is clearly shifting toward the scalable platforms we’ve built. This partnership is a high priority as we set the new gold standard for potency testing with our human-relevant NMJ assay and position Curi Bio at the forefront of animal-free drug development.”

“The partnership shows how technology innovators and technology integrators can combine strengths to accelerate real-world adoption of new approach methods,” said Bob Moyer, Research Leader at Battelle. “Battelle brings R&D scale, regulatory expertise, and world-class laboratory capabilities, while Curi Bio contributes cutting-edge technology and transformative human relevant models—together enabling innovation to translate into practical impact.”

About Curi Bio: Curi Bio provides a suite of human-relevant 3D tissue models and instrumentation to accelerate drug discovery and development. By integrating human iPSCs, tissue engineering, and automated data analysis, Curi Bio helps researchers build more predictive models of human disease. For more information, visit www.curibio.com or contact Marketing Director Jacqueline De Rose at jacqueline@curibio.com.

About Battelle: Every day, the people of Battelle apply science and technology to solving what matters most. At major technology centers and national laboratories around the world, Battelle conducts research and development, designs and manufactures products, and delivers critical services for government and commercial customers. For more information, visit www.battelle.org or contact Media Relations Director Katy Delaney at (614) 424-7208 or at delaneyk@battelle.org.

Contacts

Curi Bio

Jacqueline De Rose

Marketing Director

jacqueline@curibio.com
www.curibio.com

Battelle

Katy Delaney

Media Relations Director

(614) 424-7208

delaneyk@battelle.org
www.battelle.org

Bristol Myers Squibb Transforms the Classical Hodgkin Lymphoma Treatment Paradigm with Expanded U.S. and EMA Approvals for Opdivo® (nivolumab)

Bristol Myers Squibb Transforms the Classical Hodgkin Lymphoma Treatment Paradigm with Expanded U.S. and EMA Approvals for Opdivo® (nivolumab)


Bristol Myers Squibb Transforms the Classical Hodgkin Lymphoma Treatment Paradigm with Expanded U.S. and EMA Approvals for Opdivo® (nivolumab)

In the U.S., FDA approval establishes Opdivo in combination with doxorubicin, vinblastine and dacarbazine (AVD) as the first immunotherapy combination approved for adult and pediatric patients 12 years and older with previously untreated, Stage III or IV classical Hodgkin Lymphoma (cHL)

With approval in the EU, Opdivo in combination with brentuximab vedotin is now the first immunotherapy combination approved to treat certain pediatric and adult patients with relapsed or refractory cHL

PRINCETON, N.J.–(BUSINESS WIRE)–$BMY #CheckMate—Bristol Myers Squibb (NYSE: BMY) today announced that Opdivo® (nivolumab) has received approval for two new classical Hodgkin Lymphoma (cHL) indications in the U.S. and the European Union (EU). The U.S. Food and Drug Administration (FDA) granted approval of Opdivo in combination with doxorubicin, vinblastine and dacarbazine (AVD) for the treatment of adult and pediatric patients 12 years and older with previously untreated, Stage III or IV cHL.1 In the EU, the European Commission (EC) approved Opdivo in combination with brentuximab vedotin for the treatment of children 5 years of age and older, adolescents, and adults up to 30 years of age with relapsed or refractory cHL after one prior line of therapy.2


“These approvals represent a defining moment for people living with classical Hodgkin Lymphoma,” said Monica Shaw, MD, Senior Vice President of Oncology Commercialization. “In the U.S., we are particularly proud that Opdivo in combination with AVD now stands as an immunotherapy combination available for adults and pediatric patients, ages 12 and older, with previously untreated advanced disease.1 Concurrently, in the EU, Opdivo in combination with brentuximab vedotin has also achieved a milestone as the first immunotherapy combination for certain relapsed or refractory patients.2 These milestones reflect our continued commitment to advancing science that meaningfully improves the lives of patients and families worldwide.”

The U.S. approval is based on the Phase 3 SWOG 1826 (CA2098UT) study, evaluating Opdivo in combination with AVD for adult and pediatric (12 years and older) patients with previously untreated Stage III or IV cHL.3 A submission based on SWOG 1826 study is also currently under evaluation by the European Medicines Agency (EMA).

Opdivo and Yervoy are associated with the following Warnings and Precautions: severe and fatal immune-mediated adverse reactions including pneumonitis, colitis, hepatitis and hepatotoxicity, endocrinopathies, nephritis with renal dysfunction, dermatologic adverse reactions, other immune-mediated adverse reactions; infusion-related reactions; complications of allogeneic hematopoietic stem cell transplantation (HSCT); embryo-fetal toxicity; and increased mortality in patients with multiple myeloma when Opdivo is added to a thalidomide analogue and dexamethasone, which is not recommended outside of controlled clinical trials. Please see the Important Safety Information section below.

The EU approval is based on the Phase 2 CheckMate -744 (CA209744) study, evaluating Opdivo in combination with brentuximab vedotin for the treatment of children 5 years of age and older, adolescents and adults up to 30 years of age with relapsed or refractory cHL after one prior line of therapy.4

“For decades, treatment approaches in classical Hodgkin Lymphoma have presented significant challenges, both for newly diagnosed patients and those facing relapse,”5,6 said Alex Herrera, M.D., Chief of Division of Lymphoma, Department of Hematology & Hematopoietic Cell Transplantation, City of Hope National Medical Center. “In the U.S., the nivolumab-based combination for patients with previously untreated Stage III or IV cHL demonstrated improved progression-free survival compared with standard of care, BV-AVD. The SWOG 1826 study provides data for frontline use of this immunotherapy-based regimen.”5

“The availability of another treatment option for people living with certain types of Hodgkin lymphoma can make a real difference,” says Gwen Nichols, M.D., Chief Medical Officer of Blood Cancer United. “Each new FDA-approved therapy brings renewed hope for patients and their families, and advances like this one signal meaningful progress in improving outcomes for people facing this disease.”5

SWOG 1826 (Study CA209-8UT) demonstrated a 58% reduction in the risk of disease progression or death as determined per investigator (Hazard Ratio [HR] 0.42; 95% Confidence Interval [CI] 0.27–0.67; P=<0.0001). The trial demonstrated a statistically significant improvement in the primary endpoint of progression-free survival (PFS) for patients who received Opdivo in combination with AVD, which reflect a median follow-up of 13.7 months in the intention to treat population. After a median follow-up of 36.7 months, the median overall survival (OS) had not been reached in either treatment arm with a total of 26 deaths: 9 (1.8%) deaths in the Opdivo in combination with AVD arm and 17 (3.4%) deaths in the BV plus AVD arm.7

Select Safety Profile from SWOG 1826 (CA2098UT)

Serious adverse reactions occurred in 39% of patients receiving Opdivo in combination with doxorubicin, vinblastine, and dacarbazine (AVD) (n=490). The most frequent serious adverse reactions reported in ≥5% patients who received Opdivo in combination with AVD were peripheral neuropathy (41%), neutropenia (7%), pyrexia (7%), febrile neutropenia (6%), and nausea (6%). Fatal adverse reactions occurred in 3 patients (0.6%), all from sepsis. The most common adverse reactions were nausea (70%), neutropenia (61%), fatigue (59%), anemia (51%), constipation (49%), leukopenia (44%), musculoskeletal pain (42%), transaminases increase (41%), vomiting (33%), and stomatitis (30%).

About SWOG 1826 (CA2098UT)

SWOG 1826, also known as CA2098UT, is a randomized, multicenter, Phase 3 study evaluating Opdivo® (nivolumab) in combination with doxorubicin, vinblastine and dacarbazine (AVD) for adult and pediatric (12 years and older) patients with previously untreated Stage III or IV classical Hodgkin Lymphoma (cHL).3 The study is designed to assess progression-free survival as the primary endpoint, with key secondary endpoints that include overall survival and other measures of efficacy and safety.3 The SWOG 1826 study is sponsored by the National Cancer Institute (NCI), part of the National Institutes of Health (NIH) under a Cooperative Research and Development Agreement with Bristol Myers Squibb and conducted in the NCI National Clinical Trials Network (NCTN) led by the SWOG Cancer Research Network in collaboration with the Children’s Oncology Group (COG).3 It is the largest cHL study conducted in the NCTN.3 Bristol Myers Squibb co-sponsored the study and supplied Opdivo to the NCI through a Cooperative Research and Development agreement.3

About CheckMate -744 (CA209744)

CheckMate -744, also known as CA209744, is a risk-based, response-adapted, open-label, Phase 2 study investigating Opdivo® (nivolumab) in combination with brentuximab vedotin for children, adolescents, and young adults (between 5 and 30 years old) with CD30+ classical Hodgkin lymphoma (cHL) that has relapsed or is refractory after first-line treatment.4 The study aimed to determine the safety and efficacy of nivolumab plus brentuximab vedotin, with a subsequent treatment arm of brentuximab vedotin plus bendamustine for patients with a suboptimal response.4 The trial evaluated the overall effectiveness and tolerability of these regimens in this younger, relapsed/refractory patient population.4

Data from the Phase 2 CheckMate -744 study were presented at the American Society of Clinical Oncology (ASCO) Annual Meeting in 2023 and demonstrated that Opdivo in combination with brentuximab vedotin achieved high complete metabolic response rates in children, adolescents, and young adults with relapsed or refractory cHL after one prior line of therapy.6 The response-adapted regimen enabled the majority of patients to proceed to consolidation while maintaining a manageable safety profile.6 Responses were durable at follow-up, supporting the potential of Opdivo-based, chemotherapy-sparing approaches in this population.6

About Classical Hodgkin Lymphoma

Hodgkin lymphoma (HL), also known as Hodgkin disease, is a cancer that starts in white blood cells called lymphocytes, which are part of the body’s immune system.8 HL is the most common cancer diagnosed in adolescents (ages 15-19).9 It is most often diagnosed in early adulthood (ages 20-39) and late adulthood (older than 55 years of age).10 Classical Hodgkin lymphoma is the most common type of HL, accounting for 95% of cases.11 Despite progress in frontline therapy, advanced-stage HL still carries a substantial risk of relapse, highlighting the need for innovative approaches.10

About Opdivo

Opdivo is a programmed death-1 (PD-1) immune checkpoint inhibitor that is designed to uniquely harness the body’s own immune system to help restore anti-tumor immune response. By harnessing the body’s own immune system to fight cancer, Opdivo has become an important treatment option across multiple cancers.

Opdivo’s leading global development program is based on Bristol Myers Squibb’s scientific expertise in the field of Immuno-Oncology and includes a broad range of clinical trials across all phases, including Phase 3, in a variety of tumor types. To date, the Opdivo clinical development program has treated more than 35,000 patients.

The Opdivo trials have contributed to gaining a deeper understanding of the potential role of biomarkers in patient care, particularly regarding how patients may benefit from Opdivo across the continuum of PD-L1 expression.

In July 2014, Opdivo was the first PD-1 immune checkpoint inhibitor to receive regulatory approval anywhere in the world. Opdivo is currently approved in more than 65 countries, including the United States, the European Union, Japan and China. In October 2015, the Company’s Opdivo and Yervoy combination regimen was the first Immuno-Oncology combination to receive regulatory approval for the treatment of metastatic melanoma and is currently approved in more than 50 countries, including the United States and the European Union.

INDICATIONS

OPDIVO® (nivolumab), as a single agent, is indicated for the treatment of adult and pediatric patients 12 years and older with unresectable or metastatic melanoma.

OPDIVO® (nivolumab), in combination with YERVOY® (ipilimumab), is indicated for the treatment of adult and pediatric patients 12 years and older with unresectable or metastatic melanoma.

OPDIVO® is indicated for the adjuvant treatment of adult and pediatric patients 12 years and older with completely resected Stage IIB, Stage IIC, Stage III, or Stage IV melanoma.

OPDIVO® (nivolumab), in combination with platinum-doublet chemotherapy, is indicated as neoadjuvant treatment of adult patients with resectable (tumors ≥4 cm or node positive) non-small cell lung cancer (NSCLC).

OPDIVO® (nivolumab) in combination with platinum-doublet chemotherapy, is indicated for neoadjuvant treatment of adult patients with resectable (tumors ≥4 cm or node positive) non-small cell lung cancer (NSCLC) and no known epidermal growth factor receptor (EGFR) mutations or anaplastic lymphoma kinase (ALK) rearrangements, followed by single-agent OPDIVO® as adjuvant treatment after surgery.

OPDIVO® (nivolumab), in combination with YERVOY® (ipilimumab), is indicated for the first-line treatment of adult patients with metastatic non-small cell lung cancer (NSCLC) whose tumors express PD-L1 (≥1%) as determined by an FDA-authorized test, with no EGFR or ALK genomic tumor aberrations.

OPDIVO® (nivolumab), in combination with YERVOY® (ipilimumab) and 2 cycles of platinum-doublet chemotherapy, is indicated for the first-line treatment of adult patients with metastatic or recurrent non-small cell lung cancer (NSCLC), with no EGFR or ALK genomic tumor aberrations.

OPDIVO® (nivolumab) is indicated for the treatment of adult patients with metastatic non-small cell lung cancer (NSCLC) with progression on or after platinum-based chemotherapy. Patients with EGFR or ALK genomic tumor aberrations should have disease progression on FDA-approved therapy for these aberrations prior to receiving OPDIVO.

OPDIVO® (nivolumab), in combination with YERVOY® (ipilimumab), is indicated for the first-line treatment of adult patients with unresectable malignant pleural mesothelioma (MPM).

OPDIVO® (nivolumab), in combination with YERVOY® (ipilimumab), is indicated for the first-line treatment of adult patients with intermediate or poor risk advanced renal cell carcinoma (RCC).

OPDIVO® (nivolumab), in combination with cabozantinib, is indicated for the first-line treatment of adult patients with advanced renal cell carcinoma (RCC).

OPDIVO® (nivolumab) is indicated for the treatment of adult patients with advanced renal cell carcinoma (RCC) who have received prior anti-angiogenic therapy.

OPDIVO® (nivolumab), as a single agent, is indicated for the treatment of adult patients with classical Hodgkin lymphoma (cHL) that has relapsed or progressed after autologous hematopoietic stem cell transplantation (HSCT) and brentuximab vedotin or after 3 or more lines of systemic therapy that includes autologous HSCT.

OPDIVO® (nivolumab), in combination with doxorubicin, vinblastine, and dacarbazine (AVD), is indicated for the treatment of adult and pediatric patients 12 years and older with previously untreated, Stage III or IV classical Hodgkin lymphoma (cHL).

OPDIVO® (nivolumab) is indicated for the treatment of adult patients with recurrent or metastatic squamous cell carcinoma of the head and neck (SCCHN) with disease progression on or after platinum-based therapy.

OPDIVO® (nivolumab), as a single agent, is indicated for the adjuvant treatment of adult patients with urothelial carcinoma (UC) who are at high risk of recurrence after undergoing radical resection of UC.

OPDIVO® (nivolumab), in combination with cisplatin and gemcitabine, is indicated for the first-line treatment for adult patients with unresectable or metastatic urothelial carcinoma.

OPDIVO® (nivolumab) is indicated for the treatment of adult patients with locally advanced or metastatic urothelial carcinoma who have disease progression during or following platinum-containing chemotherapy or have disease progression within 12 months of neoadjuvant or adjuvant treatment with platinum-containing chemotherapy.

OPDIVO® (nivolumab), in combination with YERVOY® (ipilimumab), is indicated for the treatment of adult and pediatric patients 12 years and older with microsatellite instability-high (MSI-H) or mismatch repair deficient (dMMR) unresectable or metastatic colorectal cancer (CRC).

OPDIVO® (nivolumab), as a single agent, is indicated for the treatment of adult and pediatric (12 years and older) patients with microsatellite instability-high (MSI-H) or mismatch repair deficient (dMMR) metastatic colorectal cancer (CRC) that has progressed following treatment with a fluoropyrimidine, oxaliplatin, and irinotecan.

OPDIVO® (nivolumab), in combination with YERVOY® (ipilimumab), is indicated for the first-line treatment of adult patients with unresectable or metastatic hepatocellular carcinoma (HCC).

OPDIVO® (nivolumab), in combination with YERVOY® (ipilimumab), is indicated for the treatment of adult patients with unresectable or metastatic hepatocellular carcinoma (HCC) who have been previously treated with sorafenib.

OPDIVO® (nivolumab) is indicated for the adjuvant treatment of completely resected esophageal or gastroesophageal junction cancer with residual pathologic disease in adult patients who have received neoadjuvant chemoradiotherapy (CRT).

OPDIVO® (nivolumab), in combination with fluoropyrimidine- and platinum-containing chemotherapy, is indicated for the first-line treatment of adult patients with unresectable advanced or metastatic esophageal squamous cell carcinoma (ESCC) whose tumors express PD-L1 (≥1).

OPDIVO® (nivolumab), in combination with YERVOY® (ipilimumab), is indicated for the first-line treatment of adult patients with unresectable advanced or metastatic esophageal squamous cell carcinoma (ESCC) whose tumors express PD-L1 (≥1).

OPDIVO® (nivolumab) is indicated for the treatment of adult patients with unresectable advanced, recurrent or metastatic esophageal squamous cell carcinoma (ESCC) after prior fluoropyrimidine- and platinum-based chemotherapy.

OPDIVO® (nivolumab), in combination with fluoropyrimidine- and platinum-containing chemotherapy, is indicated for the treatment of adult patients with advanced or metastatic gastric cancer, gastroesophageal junction cancer, and esophageal adenocarcinoma whose tumors express PD-L1 (≥1).

IMPORTANT SAFETY INFORMATION

Severe and Fatal Immune-Mediated Adverse Reactions

Immune-mediated adverse reactions listed herein may not include all possible severe and fatal immune-mediated adverse reactions.

Immune-mediated adverse reactions, which may be severe or fatal, can occur in any organ system or tissue. While immune-mediated adverse reactions usually manifest during treatment, they can also occur after discontinuation of OPDIVO or YERVOY. Early identification and management are essential to ensure safe use of OPDIVO and YERVOY. Monitor for signs and symptoms that may be clinical manifestations of underlying immune-mediated adverse reactions. Evaluate clinical chemistries including liver enzymes, creatinine, adrenocorticotropic hormone (ACTH) level, and thyroid function at baseline and periodically during treatment with OPDIVO and before each dose of YERVOY. In cases of suspected immune-mediated adverse reactions, initiate appropriate workup to exclude alternative etiologies, including infection. Institute medical management promptly, including specialty consultation as appropriate.

Withhold or permanently discontinue OPDIVO and YERVOY depending on severity (please see section 2 Dosage and Administration in the accompanying Full Prescribing Information). In general, if OPDIVO or YERVOY interruption or discontinuation is required, administer systemic corticosteroid therapy (1 to 2 mg/kg/day prednisone or equivalent) until improvement to Grade 1 or less. Upon improvement to Grade 1 or less, initiate corticosteroid taper and continue to taper over at least 1 month. Consider administration of other systemic immunosuppressants in patients whose immune-mediated adverse reactions are not controlled with corticosteroid therapy. Toxicity management guidelines for adverse reactions that do not necessarily require systemic steroids (e.g., endocrinopathies and dermatologic reactions) are discussed below.

Immune-Mediated Pneumonitis

OPDIVO and YERVOY can cause immune-mediated pneumonitis. The incidence of pneumonitis is higher in patients who have received prior thoracic radiation. In patients receiving OPDIVO monotherapy, immune-mediated pneumonitis occurred in 3.1% (61/1994) of patients, including Grade 4 (<0.1%), Grade 3 (0.9%), and Grade 2 (2.1%). In patients receiving OPDIVO 1 mg/kg with YERVOY 3 mg/kg every 3 weeks, immune-mediated pneumonitis occurred in 7% (31/456) of patients, including Grade 4 (0.2%), Grade 3 (2.0%), and Grade 2 (4.4%). In patients receiving OPDIVO 3 mg/kg with YERVOY 1 mg/kg every 3 weeks, immune-mediated pneumonitis occurred in 3.9% (26/666) of patients, including Grade 3 (1.4%) and Grade 2 (2.6%). In NSCLC patients receiving OPDIVO 3 mg/kg every 2 weeks with YERVOY 1 mg/kg every 6 weeks, immune-mediated pneumonitis occurred in 9% (50/576) of patients, including Grade 4 (0.5%), Grade 3 (3.5%), and Grade 2 (4.0%). Four patients (0.7%) died due to pneumonitis.

Immune-Mediated Colitis

OPDIVO and YERVOY can cause immune-mediated colitis, which may be fatal. A common symptom included in the definition of colitis was diarrhea. Cytomegalovirus (CMV) infection/reactivation has been reported in patients with corticosteroid-refractory immune-mediated colitis. In cases of corticosteroid-refractory colitis, consider repeating infectious workup to exclude alternative etiologies. In patients receiving OPDIVO monotherapy, immune-mediated colitis occurred in 2.9% (58/1994) of patients, including Grade 3 (1.7%) and Grade 2 (1%). In patients receiving OPDIVO 1 mg/kg with YERVOY 3 mg/kg every 3 weeks, immune-mediated colitis occurred in 25% (115/456) of patients, including Grade 4 (0.4%), Grade 3 (14%) and Grade 2 (8%). In patients receiving OPDIVO 3 mg/kg with YERVOY 1 mg/kg every 3 weeks, immune-mediated colitis occurred in 9% (60/666) of patients, including Grade 3 (4.4%) and Grade 2 (3.7%).

Immune-Mediated Hepatitis and Hepatotoxicity

OPDIVO and YERVOY can cause immune-mediated hepatitis. In patients receiving OPDIVO monotherapy, immune-mediated hepatitis occurred in 1.8% (35/1994) of patients, including Grade 4 (0.2%), Grade 3 (1.3%), and Grade 2 (0.4%). In patients receiving OPDIVO 1 mg/kg with YERVOY 3 mg/kg every 3 weeks, immune-mediated hepatitis occurred in 15% (70/456) of patients, including Grade 4 (2.4%), Grade 3 (11%), and Grade 2 (1.8%). In patients receiving OPDIVO 3 mg/kg with YERVOY 1 mg/kg every 3 weeks, immune-mediated hepatitis occurred in 7% (48/666) of patients, including Grade 4 (1.2%), Grade 3 (4.9%), and Grade 2 (0.4%).

OPDIVO in combination with cabozantinib can cause hepatic toxicity with higher frequencies of Grade 3 and 4 ALT and AST elevations compared to OPDIVO alone. Consider more frequent monitoring of liver enzymes as compared to when the drugs are administered as single agents. In patients receiving OPDIVO and cabozantinib, Grades 3 and 4 increased ALT or AST were seen in 11% (35/320) of patients.

Immune-Mediated Endocrinopathies

OPDIVO and YERVOY can cause primary or secondary adrenal insufficiency, immune-mediated hypophysitis, immune-mediated thyroid disorders, and Type 1 diabetes mellitus, which can present with diabetic ketoacidosis. Withhold OPDIVO and YERVOY depending on severity (please see section 2 Dosage and Administration in the accompanying Full Prescribing Information). For Grade 2 or higher adrenal insufficiency, initiate symptomatic treatment, including hormone replacement as clinically indicated. Hypophysitis can present with acute symptoms associated with mass effect such as headache, photophobia, or visual field defects. Hypophysitis can cause hypopituitarism; initiate hormone replacement as clinically indicated. Thyroiditis can present with or without endocrinopathy. Hypothyroidism can follow hyperthyroidism; initiate hormone replacement or medical management as clinically indicated. Monitor patients for hyperglycemia or other signs and symptoms of diabetes; initiate treatment with insulin as clinically indicated.

In patients receiving OPDIVO monotherapy, adrenal insufficiency occurred in 1% (20/1994), including Grade 3 (0.4%) and Grade 2 (0.6%). In patients receiving OPDIVO 1 mg/kg with YERVOY 3 mg/kg every 3 weeks, adrenal insufficiency occurred in 8% (35/456) of patients, including Grade 4 (0.2%), Grade 3 (2.4%), and Grade 2 (4.2%).In patients receiving OPDIVO 3 mg/kg with YERVOY 1 mg/kg every 3 weeks, adrenal insufficiency occurred in 7% (48/666) of patients, including Grade 4 (0.

Contacts

Bristol Myers Squibb
Media Inquiries:
media@bms.com

Investors:
investor.relations@bms.com

Read full story here

Pfizer Recommends Shareholders Reject the Mini-Tender Offer by Tutanota LLC

Pfizer Recommends Shareholders Reject the Mini-Tender Offer by Tutanota LLC


Pfizer Recommends Shareholders Reject the Mini-Tender Offer by Tutanota LLC

NEW YORK–(BUSINESS WIRE)–Pfizer Inc. (NYSE: PFE) today announced that it has received notice of an unsolicited mini-tender offer by Tutanota LLC (“Tutanota”) to purchase up to 1 million shares of Pfizer common stock at a price of $32.00 per share in cash. The offer price of $32.00 per share is conditioned on, among other things, the closing price per share of Pfizer common stock exceeding $32.00 per share on the last trading day before the offer expires. This means that unless this condition is waived by Tutanota, Pfizer shareholders who tender their shares in the offer will receive a below-market price. The offer states that as of the date of the offer, Tutanota expects to extend the offer until the market price of Pfizer’s common stock exceeds the offer price. The offer is for approximately 0.02% of the shares of Pfizer common stock outstanding as of the March 9, 2026 offer date.


Pfizer recommends that shareholders do not tender their shares in response to Tutanota’s offer because the offer requires that the closing stock price for Pfizer’s common stock exceed the offer price, and it is subject to numerous additional conditions, including Tutanota obtaining financing for the offer, which Tutanota states it does not currently have. There is no guarantee the conditions of the offer will be satisfied. Under the terms of the offer, Tutanota can extend the offer and delay payment beyond the expiration date of the offer, currently scheduled for 5:00 p.m., New York City time, on Monday, April 13, 2026. The offer states that Tutanota intends to extend the offer until Pfizer’s stock price exceeds $32.00 per share. Per the terms of the offer, any shareholders who tender (or have already tendered) their shares can withdraw them prior to the expiration of the offer, currently scheduled for April 13, 2026, in accordance with the offering documents.

Pfizer does not endorse Tutanota’s unsolicited mini-tender offer and is not affiliated or associated in any way with Tutanota, its mini-tender offer or its offer documentation.

Tutanota has made many similar mini-tender offers for shares of other companies. Mini-tender offers seek to acquire less than 5 percent of a company’s shares outstanding, thereby avoiding many disclosure and procedural requirements of the U.S. Securities and Exchange Commission (“SEC”) that apply to offers for more than 5 percent of a company’s shares outstanding. As a result, mini-tender offers do not provide investors with the same level of protections as provided for larger tender offers under U.S. securities laws.

The SEC has cautioned investors that some bidders making mini-tender offers at below-market prices are “hoping that they will catch investors off guard if the investors do not compare the offer price to the current market price.” More on the SEC’s guidance to investors on mini-tender offers is available at SEC.gov | Mini-Tender Offers: Tips for Investors.

Pfizer urges investors to obtain current market quotations for their shares, to consult with their broker or financial advisor and to exercise caution with respect to Tutanota’s offer. Pfizer recommends that stockholders who have not responded to Tutanota’s offer take no action. Stockholders who have already tendered their shares may withdraw them at any time prior to the expiration of the offer, in accordance with Tutanota’s offer documentation. The offer is currently scheduled to expire at 5:00 p.m., New York City time, on April 13, 2026. Tutanota may extend the offering period at its discretion.

Pfizer encourages brokers and dealers, as well as other market participants, to review the SEC’s letter regarding broker-dealer mini-tender offer dissemination and disclosure at Letter to SIA re: Broker-Dealer Mini-Tender Offer Dissemination and Disclosures and the NASD Notice to Members 99-53 issued in July 1999 regarding guidance to members forwarding mini-tender offers to their customers, which can be found at https://www.finra.org/sites/default/files/NoticeDocument/p004221.pdf.

Pfizer requests that a copy of this news release be included with all distributions of materials relating to Tutanota’s mini-tender offer related to shares of Pfizer common stock.

About Pfizer: Breakthroughs That Change Patients’ Lives

At Pfizer, we apply science and our global resources to bring therapies to people that extend and significantly improve their lives. We strive to set the standard for quality, safety and value in the discovery, development and manufacture of health care products, including innovative medicines and vaccines. Every day, Pfizer colleagues work across developed and emerging markets to advance wellness, prevention, treatments and cures that challenge the most feared diseases of our time. Consistent with our responsibility as one of the world’s premier innovative biopharmaceutical companies, we collaborate with health care providers, governments and local communities to support and expand access to reliable, affordable health care around the world. For 175 years, we have worked to make a difference for all who rely on us. We routinely post information that may be important to investors on our website at www.Pfizer.com. In addition, to learn more, please visit us on www.Pfizer.com and follow us on X at @Pfizer and @Pfizer News, LinkedIn, YouTube and like us on Facebook at Facebook.com/Pfizer.

Category: Financial

Contacts

Media Contact:
PfizerMediaRelations@Pfizer.com

Investor Contact:
IR@Pfizer.com

Garaherb Claims Evaluated: Complete Scientific Investigation of Ingredients’ Effectiveness

Garaherb Claims Evaluated: Complete Scientific Investigation of Ingredients’ Effectiveness


Garaherb Claims Evaluated: Complete Scientific Investigation of Ingredients’ Effectiveness

An informational report compiling GaraHerb’s publicly available product materials, ingredient-level research references, formulation transparency considerations, and consumer details for 2026

Aurora, CO, March 20, 2026 (GLOBE NEWSWIRE) — Editor’s Note: The terms “Complete Scientific Investigation” and “Effectiveness” in the headline refer to a compilation of ingredient-level research and publicly cited studies related to compounds in the GaraHerb formulation. These terms describe how ingredients are positioned within available research literature and do not indicate independent clinical evaluation of the finished GaraHerb product.

This informational report is based on publicly available materials provided by GARAPLUS, including official product descriptions, ingredient references, and company disclosures. It compiles how GaraHerb is presented by the company alongside relevant ingredient-level research context. This report contains affiliate links. If a purchase is made through these links, a commission may be earned at no additional cost to the buyer. This content does not constitute medical, health, or dietary advice. All product details described below are stated as presented by the company and should be verified directly on the official website before any purchasing decision.

In this report, the term “effectiveness” refers strictly to how GaraHerb’s marketing language describes potential outcomes. It does not indicate that the finished product has been clinically proven effective, and no published clinical trial appears to evaluate GaraHerb as a proprietary formula.

Garaherb Claims Evaluated Complete Scientific Investigation of Ingredients' Effectiveness

GaraHerb has been generating attention across social media and wellness sites among men in their 30s through 70s looking for natural approaches to energy, stamina, and overall vitality. Consumers often look to understand how products in this category are described and supported by available information before making a purchasing decision — and with that visibility come questions worth exploring through the company’s own materials.

This report compiles what the company presents in its official materials alongside ingredient-level research that has been published in peer-reviewed journals. The information presented reflects the company’s publicly available product descriptions and cited research on individual compounds.

Current product details, pricing, and terms can be confirmed by View the current GaraHerb offer (official GaraHerb page).

Individual results vary. Dietary supplements are not substitutes for balanced nutrition, regular physical activity, or professional medical guidance. Consult a qualified healthcare provider before starting any new supplement regimen.

What Is GaraHerb

According to the company’s published materials, GaraHerb is a dietary supplement marketed as a natural male vitality formula. The company positions it as a once-daily capsule and describes it as a plant-based approach to supporting energy, stamina, and male performance. The product is distributed by GARAPLUS out of Aurora, Colorado.

The company states that GaraHerb is manufactured in the United States in a facility described as FDA-registered and GMP-certified. The label identifies the supplement as gluten-free and vegetarian. Each container provides 30 capsules at one capsule per serving.

The formula includes two individually dosed ingredients — Vitamin B3 (as Niacin) at 20 mg and Zinc (as Zinc Oxide) at 11 mg — along with a 570 mg proprietary blend containing L-Citrulline, L-Carnitine, Pine Bark Extract, Velvet Bean Seed Extract, Maca Root Extract, Grape Skin Extract, and Saffron Stigmas Extract. Other ingredients include the vegetable capsule shell, microcrystalline cellulose, magnesium stearate, and silicon dioxide.

How GaraHerb Is Described in Company Materials and Related Research

The company’s sales page describes GaraHerb as helping men “reclaim their vitality” and references renewed energy, confidence, and performance. According to the company’s marketing materials, results unfold in stages — with improvements in energy and mental clarity described in the first stage, firmer muscles and erections in the second, and lasting results by the third stage when taken continuously for at least three months.

These are the company’s marketing descriptions. No published clinical trial appears to have evaluated GaraHerb as a finished multi-ingredient supplement to verify these staged outcome claims. The staged timeline framework functions as a consumer-facing narrative presented by the company rather than a clinically validated progression.

Several individual ingredients within the proprietary blend have been studied in published research. The following section compiles what that research covers and the dosages used in those studies, which provides useful context for anyone looking into this product.

Ingredient-Level Research Context

In this context, “scientific investigation” refers to a compilation of ingredient-level research and publicly cited studies rather than clinical trials conducted on the finished GaraHerb formulation. The studies referenced below evaluate individual compounds under controlled conditions and are not presented as clinical validation of the GaraHerb product itself.

This is ingredient-level research; GaraHerb as a finished product has not been clinically studied.

L-Citrulline has been studied for its role in nitric oxide production and blood flow support. Published research, including a 2011 study in the journal Urology, examined L-Citrulline supplementation at 1,500 mg daily and reported improvements in certain measures of erectile function in a small group of men with mild concerns. The dosage used in that study — 1,500 mg of L-Citrulline alone — is more than double GaraHerb’s entire 570 mg proprietary blend shared across seven ingredients.

Pine Bark Extract (Pinus pinaster) has been reviewed in studies examining its antioxidant properties and potential effects on circulation. Some research has combined pine bark extract with L-Arginine at specific dosages and reported positive outcomes related to blood flow. These studies typically used pine bark extract at 40–120 mg daily alongside other compounds — not as one component among seven in a shared blend.

Maca Root Extract (Lepidium meyenii) has a body of published research examining its effects on libido, energy, and stamina. A 2009 systematic review noted limited but suggestive evidence for maca’s effects on sexual desire. Clinical studies generally used maca at dosages between 1,500 and 3,000 mg daily — significantly more than what a 570 mg shared blend could deliver.

Velvet Bean Seed Extract (Mucuna pruriens) contains L-DOPA, a precursor to dopamine, and has been studied for its potential effects on testosterone levels and reproductive health. Published research on Mucuna pruriens typically used dosages of 5,000 mg of seed powder daily in studies involving male fertility parameters. The extract form may concentrate active compounds, but the specific amount present in GaraHerb’s blend is not disclosed.

Saffron Stigmas Extract (Crocus sativus) has been examined in published studies for potential mood and libido-related effects. A 2012 study published in Psychopharmacology reported modest improvements in certain sexual function parameters using 30 mg of saffron daily. Whether GaraHerb’s blend delivers a comparable amount of saffron is unknown based on the label.

Zinc is the one ingredient dosed individually at 11 mg (100% Daily Value). Zinc plays a well-established role in testosterone production, immune function, and reproductive health. Published research, including a widely cited 1996 study, demonstrated that zinc restriction in young men was associated with decreased testosterone levels. The 11 mg dosage in GaraHerb aligns with the recommended daily intake and falls within ranges used in published studies.

Vitamin B3 (Niacin) at 20 mg supports energy metabolism and circulation. This dosage falls within standard supplementation ranges, though it is significantly lower than the therapeutic doses (500–3,000 mg) used in research examining cardiovascular effects.

These individual findings do not mean GaraHerb replaces prescribed treatment. Each study examined individual compounds at known dosages under controlled conditions — not GaraHerb’s specific proprietary formulation.

The Proprietary Blend Dosing Question

This is the most important transparency factor for anyone comparing the company’s ingredient references against published research.

According to the product label, the proprietary blend totals 570 mg across seven botanical ingredients. FDA labeling rules require ingredients to appear by weight from highest to lowest, meaning L-Citrulline is present in the largest amount and Saffron in the smallest. But the exact amount of each ingredient is not disclosed.

Here’s the practical math. If even two ingredients were dosed at their published research ranges, the remaining five would share a very small fraction of the blend. For context, published L-Citrulline research used 1,500 mg alone. Published maca studies used 1,500–3,000 mg. A single 570 mg blend containing seven ingredients cannot simultaneously deliver research-level dosages for multiple compounds.

This isn’t unique to GaraHerb — many supplements use proprietary blends. But understanding this structure helps set realistic expectations when the company’s marketing references potential outcomes without disclosing individual ingredient amounts.

A Note About the Scientific References on the GaraHerb Website

The official website lists 15 scientific references at the bottom of the sales page. When comparing these citations against the product label, several of the listed studies examine compounds that do not appear on GaraHerb’s Supplement Facts label.

For example, the reference list includes studies on magnesium and testosterone, saw palmetto, hawthorn berry, Cissus quadrangularis, chrysin, horny goat weed (icariin), and Tribulus terrestris. None of these ingredients are listed on the GaraHerb product label. The actual label ingredients — L-Citrulline, L-Carnitine, Pine Bark Extract, Velvet Bean Seed Extract, Maca Root Extract, Grape Skin Extract, and Saffron — are a different set of compounds.

The presence of citations for ingredients not found in the product is a detail consumers may want to be aware of. It does not necessarily indicate intent to mislead, but it means the reference list should be cross-checked against the actual Supplement Facts panel rather than taken as direct research support for the specific formula.

GaraHerb Pricing and Purchase Details

According to pricing published on the official website at the time of this report, GaraHerb is available in three package options: a single bottle (30-day supply) listed at $89 plus $9.99 shipping, a three-bottle package (90-day supply) listed at $59 per bottle with free shipping, and a six-bottle package (180-day supply) listed at $49 per bottle with free shipping. The company states that multi-bottle packages include bonus digital materials described on the sales page.

According to the company, all purchases are one-time payments with no subscriptions or automatic rebilling. The company states that orders ship within 1–2 business days via premium carriers, with delivery typically within 5–10 business days for domestic orders. Pricing and availability can change, so readers should verify current terms by View the current GaraHerb offer (official GaraHerb page).

GaraHerb Refund Policy Details

According to the company’s published policy, purchases are covered by a 60-day satisfaction period from the original purchase date, subject to specific return conditions, processing terms, and applicable fees.

The policy states that refund requests require a Return Merchandise Authorization (RMA) number obtained through customer support before returning product. All bottles — including empty, partially used, and any bonus or free bottles — must be returned within the 60-day window. According to the company, approved refunds are subject to a return-processing fee of up to 20% of the order value. Shipping charges are described as non-refundable, and return shipping is the buyer’s responsibility. The company limits refunds to one per product per household within a 12-month period. Full details are available on the official website.

Consumer Considerations Based on Company Materials

Cross-check the scientific references against the actual label. The references listed on the company’s sales page include studies on ingredients not present in the GaraHerb formula. Confirming which studies apply to ingredients actually in the product is a useful step.

Understand the proprietary blend structure. Without individual ingredient dosages disclosed, the formula cannot be directly compared to published research dosages. If exact amounts matter to your evaluation, the company’s contact information is listed below.

Read the full refund policy before ordering. According to the company’s published terms, the 60-day satisfaction period includes conditions such as a return-processing fee of up to 20%, an RMA requirement, and buyer-paid return shipping. Full terms are available on the official website.

Consider the staged results timeline in context. The three-stage progression described on the company’s website is a marketing framework, not a clinically validated timeline. According to published research, individual responses to any supplement depend on numerous personal health factors.

Talk to your healthcare provider. This is especially relevant for men taking medications for blood pressure, cardiovascular conditions, diabetes, hormonal imbalances, or erectile dysfunction. Several ingredients in GaraHerb’s blend interact with biological pathways that may overlap with prescription medications. Professional medical guidance should come before any supplement purchase.

Consumer Context Described in Company Materials

Based on the company’s published positioning and the product’s ingredient profile, GaraHerb may be of interest to men who prefer plant-based supplement options and who are exploring natural vitality support as one component of a broader wellness approach that includes balanced nutrition and regular physical activity.

The product may be less suited for consumers who need transparent individual ingredient dosing for clinical comparison. It may also not align with those seeking product-level clinical trial evidence rather than ingredient-level research, or individuals expecting rapid results from a single supplement without accompanying lifestyle changes.

As with any dietary supplement, individual outcomes depend on a wide range of personal health factors. Supplements are regulated differently than pharmaceutical drugs, and the FDA does not evaluate dietary supplement claims for efficacy before products reach the market. The FDA disclaimer printed on GaraHerb’s own label confirms that the product’s statements have not been evaluated by the Food and Drug Administration and that the product is not intended to diagnose, treat, cure, or prevent any disease.

Readers who want to confirm the full ingredient label, pricing tiers, and current availability can do so by View the current GaraHerb offer (official GaraHerb page).

Common Consumer Questions About GaraHerb

Is GaraHerb FDA approved?

GaraHerb is a dietary supplement. Under current federal regulations, dietary supplements do not require FDA approval before being sold. The company states the product is manufactured in an FDA-registered, GMP-certified facility, which relates to manufacturing standards — not product approval or efficacy verification.

Does GaraHerb have clinical trials?

No published clinical trial appears to have evaluated GaraHerb as a finished proprietary formula. The research cited on the product’s website pertains to individual ingredients or compounds not in the formula, often at dosages that may differ from what the proprietary blend delivers.

Why does GaraHerb use a proprietary blend?

Proprietary blends allow manufacturers to disclose which ingredients are included without revealing exact amounts for each one. This protects formulation trade secrets but limits a consumer’s ability to compare individual ingredient dosages against published research ranges.

How long does GaraHerb take to show results?

According to the company’s FAQ, most people notice differences within the first week, with optimal outcomes after three or more months of consistent use. These are the company’s marketing statements. Individual timelines depend on factors including baseline health, dietary habits, and physical activity levels.

What is GaraHerb’s refund process?

According to the company’s published policy, refund requests must be submitted within 60 days of the original purchase date. An RMA number is required, all bottles (including empty and bonus bottles) must be returned, and return shipping is the buyer’s responsibility. A processing fee of up to 20% may apply to approved refunds.

Do the scientific references on the website match the ingredients?

Not entirely. Several references cite studies on ingredients — including magnesium, saw palmetto, hawthorn berry, chrysin, horny goat weed, Tribulus terrestris, and Cissus quadrangularis — that are not listed on the GaraHerb Supplement Facts panel. Cross-referencing the citations with the actual label is recommended.

Additional Consumer Research

Consumers researching GaraHerb may benefit from additional publicly available information on this product. A 2026 informational overview of GaraHerb supplement facts, ingredient label details, and product disclosures provides additional context on what the Supplement Facts panel does and does not reveal. Readers may also find it helpful to explore the label-verified ingredient breakdown and manufacturing transparency details compiled earlier this year.

Evaluating multiple sources is recommended before making any purchasing decision. Consumers may also wish to review the FDA’s general guidance on dietary supplement labeling and claims.

Summary of Key Considerations

GaraHerb is a dietary supplement that the company positions as a natural male vitality formula built around botanical ingredients. The product contains individual compounds that have published research at the ingredient level — particularly zinc for testosterone support and L-Citrulline for nitric oxide production. However, the proprietary blend structure limits visibility into individual dosing, several of the scientific references on the sales page cite ingredients not present in the formula, and no published clinical trial has evaluated the finished product as formulated.

According to the company, GaraHerb is backed by a 60-day refund period with specific conditions including a processing fee of up to 20%, one-time purchase pricing with no subscriptions, and manufacturing under GMP standards in an FDA-registered facility.

Complete product details, current pricing, and published terms are available by View the current GaraHerb offer (official GaraHerb page).

Contact Information

According to the company’s published contact details:

Company: GARAPLUS

Return Address: 19655 E 35th Drive, Suite 100, Aurora, CO 80011, USA

Email: support@garaherb.com

U.S. Support: 1-302 455 7162

E.U. Support: +44 1704 320405

Website: https://garaherb.com/

Disclaimers

Content and Consumer Information Disclaimer: This is an informational report compiled from publicly available product materials, ingredient-level research references, and the company’s published disclosures. It does not constitute medical, health, dietary, financial, or legal advice. All product details, ingredient information, pricing, and policy terms described in this report are stated as presented by the company on its publicly available website and product labeling. This content has not been independently audited or verified unless specifically noted. Readers are encouraged to verify all claims directly with the manufacturer and to consult a qualified healthcare professional before beginning any dietary supplement.

FDA Health Disclaimer: These statements have not been evaluated by the Food and Drug Administration. This product is not intended to diagnose, treat, cure, or prevent any disease. Always consult your physician before starting any new supplement, especially if you have existing health conditions, take medications, or are pregnant or nursing.

Professional Medical Disclaimer: This content is educational and does not constitute medical advice. GaraHerb is a dietary supplement, not a medication. If you are currently taking medications, have existing health conditions, are pregnant or nursing, or are considering any major changes to your health regimen, consult your physician before starting GaraHerb or any new supplement. Do not change, adjust, or discontinue any medications or prescribed treatments without your physician’s guidance and approval.

Results May Vary: Individual results will vary based on factors including age, baseline health condition, lifestyle factors, consistency of use, genetic factors, current medications, and other individual variables. Results are not guaranteed. These are individual experiences and should not be interpreted as typical or guaranteed results.

FTC Affiliate Disclosure: This report contains affiliate links. If a purchase is made through these links, a commission may be earned at no additional cost to the buyer. This compensation does not influence the accuracy, neutrality, or integrity of the information presented. All descriptions are based on published research and publicly available information.

Pricing Disclaimer: All prices, discounts, and promotional offers mentioned were accurate at the time of publication (March 2026) but are subject to change without notice. Always verify current pricing and terms on the official GaraHerb website before making your purchase.

Publisher Responsibility: The publisher of this report has made every effort to ensure accuracy at the time of publication. No responsibility is assumed for errors, omissions, or outcomes resulting from the use of the information provided. Readers are encouraged to verify all details directly with GaraHerb and their healthcare provider before making decisions.

CONTACT: Email: support@garaherb.com
U.S Support 1-302 455 7162
E.U. Support +44 1704 320405 

Zevra Therapeutics Reports Inducement Grant to New Chief Financial Officer Under Nasdaq Listing Rule 5635(c)(4)

Zevra Therapeutics Reports Inducement Grant to New Chief Financial Officer Under Nasdaq Listing Rule 5635(c)(4)


Zevra Therapeutics Reports Inducement Grant to New Chief Financial Officer Under Nasdaq Listing Rule 5635(c)(4)

BOSTON, March 20, 2026 (GLOBE NEWSWIRE) — Zevra Therapeutics, Inc. (NasdaqGS: ZVRA) (Zevra, or the Company), a commercial-stage company focused on providing therapies for people living with rare disease, today announced that the Company has granted as of March 18, 2026 an equity inducement award to Justin Renz, the Company’s new Chief Financial Officer pursuant to the Company’s 2023 Employment Inducement Award Plan (as amended and/or restated, the “Inducement Award Plan”). The equity award was approved by the Compensation Committee of the Board of Directors in accordance with Nasdaq Rule 5635(c)(4) and made as a material inducement to Mr. Renz upon acceptance of employment with Zevra.

The Company granted Mr. Renz a new hire option to purchase 300,000 shares of Zevra’s common stock. The option has a 10-year term and an exercise price per share equal to $9.55, which was the closing price of Zevra’s common stock on March 18, 2026. The option vests over four years, subject to Mr. Renz’s continued service through the applicable vesting dates.

About Zevra Therapeutics, Inc.

Zevra Therapeutics, Inc. is a commercial-stage company with a late-stage pipeline committed to redefining what is possible in bringing life-changing therapies to people living with rare diseases. The Company is focused on broadening access through geographic expansion opportunities, progressing its pipeline toward key milestones, and delivering meaningful therapeutics. The commercialization of its lead product, marketed in the U.S. for Niemann-Pick disease type C (NPC), a rare, progressive neurodegenerative disease, provides a strong corporate foundation and validates its ability to advance therapies from development to market. Zevra’s vision is realized through disciplined execution of its strategic plan and core values — patient centricity, integrity, accountability, innovation, and courage — which guide its efforts to deliver long-term value.

For more information, please visit www.zevra.com or follow us on X and LinkedIn.

Investor Contact

Nichol Ochsner 
+1 (732) 754-2545 
nochsner@zevra.com  

Media Contact

Julie Downs
+1 (508) 246-3230
jdowns@zevra.com