BIO-Europe 2026 to Convene Global Biopharma Leaders in Cologne as Dealmaking Confidence Grows

Informa Connect

/ Key word(s): Conference

BIO-Europe 2026 to Convene Global Biopharma Leaders in Cologne as Dealmaking Confidence Grows

27.10.2026 / 15:00 CET/CEST

The issuer is solely responsible for the content of this announcement.


BIO-Europe 2026 to Convene Global Biopharma Leaders in Cologne as Dealmaking Confidence Grows

London, UK – October 27, 2026 – BIO-Europe 2026 will bring together more than 6,000 senior executives from the global biotech, pharmaceutical, and investment sectors in Cologne, Germany, November 9–11, 2026. Now in its 32nd edition, the event comes at a time of renewed confidence in biopharma dealmaking, despite continued financing challenges.

“While 75% of senior biopharma leaders expect appetite for partnering and M&A to increase over the next 12 months. The industry continues to face fundamental questions about how scientific innovation is financed, developed, and brought to patients,” said Claire Macht, Senior Portfolio Director, Europe, Informa Connect. “BIO-Europe provides a neutral platform for senior leaders to challenge established approaches, exchange perspectives, and address the strategic decisions shaping the future of biopharma.”

The findings, from the inaugural Biopharma C-Suite Agenda Survey 2026 (81 senior executives, including 50 CEOs), will anchor BIO-Europe’s Big Picture Plenary Series.

Rethinking Europe’s Biotech Business Models and Financing

As European biotech companies navigate changing capital markets and regulatory environments, industry leaders are increasingly questioning whether established business models can support the next generation of breakthrough therapies.

“CEOs need to speak up: when governments make drug development harder, patients pay the price. Titans must take responsibility,” said Jeremy Levin, Prior CEO Teva Pharmaceuticals. “Big venture funds must back young, science-based companies, or Europe will keep exporting the science and buying back the medicines. Now is the time to invest in innovation.”

“There is now a critical pre-clinical funding gap in Europe as bigger VC funds focus on later-stage clinical development,” said Edwin Moses, Chairman of NanoSyrinx, LabGenius, and Avantium. “These big funds should consider creating new agile vehicles with specialist managers and risk appetite to ensure there is a biotech product pipeline for the future. Innovation in speeding up regulatory pathways is a major opportunity for Europe to lead the way – let’s take the lead and simplify.”

Levin and Moses will debate these issues further at the Big Picture plenary, “Building a Biotech in a Changing World – Europe’s Choices,” examining whether traditional biotech business models can sustain breakthrough innovation amid shifting capital markets, platform-based company creation, and advances in AI.

These challenges will be explored further in a second Big Picture plenary, “Building a Competitive and Impactful European Biotech Community: Perspectives from the European Life Sciences Coalition and CEOs.” The session will introduce the newly established European Life Sciences Coalition (ELSC) and invite participants to support an open letter to European leaders. Moderated by Melanie Senior, the discussion will feature representatives from Andera Partners, Angelini Ventures, Tubulis, and argenx, examining how institutional capital, regulatory pathways, and deeper public capital markets can support biotech company creation and growth in Europe.

Building Globally Competitive Biotech Ecosystems

Beyond financing and regulatory reform, the conference will examine how innovation ecosystems can help biotech companies grow and compete internationally. The session “Building the Next Global Biotech Hubs: What Makes an Innovation Ecosystem Work?” will explore how talent, research infrastructure, investment, and industry collaboration combine to build successful life sciences clusters.

As host region, North Rhine-Westphalia provides an example of an established life sciences ecosystem, bringing together industry, research institutions, and healthcare organizations.

“North Rhine-Westphalia combines scientific excellence with a strong and diverse life sciences industry. Hosting BIO-Europe in Cologne puts our regional capabilities on an international stage,” said Dr. Stefan Rensch of Bio.NRW. “For our companies and research institutions, the event offers direct access to potential investors, pharmaceutical partners, and international markets, helping turn the region’s research and innovation strengths into commercial opportunities.”

Partnering and Registration for BIO-Europe 2026

One-to-one partnering is now open through partneringONE®.

A Digital Partnering Pass offers virtual partnering days on November 17–18, 2026, plus on-demand content, including company presentations and panel discussions.

Registration is open via the the BIO-Europe website, with discounted rates available through November 6, 2026.

About Informa Connect

Informa Connect, a division of Informa PLC, creates global platforms where industries connect, learn, and collaborate through events, digital services, and professional communities. Through its life sciences portfolio, Informa Connect brings together senior executives, investors, and strategic partners to exchange insights, build relationships, and drive innovation across global markets.

For more information, please visit: https://informaconnect.com/bioeurope/

Follow BIO-Europe 2026 on LinkedIn or X @EBDGroup (hashtag: #BIOEurope2026)

Media Contacts:
Informa Connect contacts

Ryan Hui
lifesciencepress@informa.com

MC Services AG contacts

Brittney Sojeva
Senior Consultant
brittney.sojeva@mc-services.eu
+49 211 529 252 14


27.10.2026 CET/CEST Dissemination of a Corporate News, transmitted by EQS News – a service of EQS Group.
The issuer is solely responsible for the content of this announcement.

The EQS Distribution Services include Regulatory Announcements, Financial/Corporate News and Press Releases.


2423008  27.10.2026 CET/CEST

Enhertu® Approved in the EU as Adjuvant Treatment for Patients with Residual Disease After Neoadjuvant Treatment for HER2 Positive Early Breast Cancer

Enhertu® Approved in the EU as Adjuvant Treatment for Patients with Residual Disease After Neoadjuvant Treatment for HER2 Positive Early Breast Cancer


Enhertu® Approved in the EU as Adjuvant Treatment for Patients with Residual Disease After Neoadjuvant Treatment for HER2 Positive Early Breast Cancer

  • Approval based on DESTINY-Breast05 phase 3 trial results that showed Daiichi Sankyo and AstraZeneca’s Enhertu reduced the risk of invasive disease recurrence or death by 53% versus T-DM1
  • First EU approval of Enhertu in the curative-intent early breast cancer setting
  • Third approval for Enhertu in the EU in four months

TOKYO–(BUSINESS WIRE)–Enhertu® (trastuzumab deruxtecan) has been approved in the European Union (EU) as a monotherapy for the adjuvant treatment of adult patients with resected HER2 positive (immunohistochemistry [IHC] 3+ or in-situ hybridization [ISH]+) breast cancer who have residual invasive disease after neoadjuvant taxane-based and HER2 targeted treatment.
Enhertu is a specifically engineered HER2 directed DXd antibody drug conjugate (ADC) discovered by Daiichi Sankyo (TSE: 4568) and being jointly developed and commercialized by Daiichi Sankyo and AstraZeneca (LSE/STO/NYSE: AZN).

The approval by the European Commission follows the positive opinion of the Committee for Medicinal Products for Human Use of the European Medicines Agency and is based on results from the DESTINY-Breast05 phase 3 trial presented at the 2025 European Society for Medical Oncology (#ESMO25) Congress and subsequently published in The New England Journal of Medicine.

In DESTINY-Breast05, Enhertu significantly reduced the risk of invasive disease recurrence or death (invasive disease-free survival [IDFS]) by 53% (hazard ratio [HR]=0.47; 95% confidence interval [CI]: 0.34-0.66; p<0.0001) compared to trastuzumab emtansine (T-DM1) in patients with HER2 positive breast cancer with residual invasive disease following neoadjuvant therapy. Enhertu demonstrated a three-year IDFS rate of 92.4% (95% CI: 89.7-94.4) and 83.7% with T-DM1 (95% CI: 80.2-86.7). Data also showed that Enhertu significantly reduced the risk of disease recurrence or death (disease-free survival [DFS]) by 53% (HR=0.47; 95% CI: 0.34-0.66; p<0.0001) compared to T-DM1. Results showed a three-year DFS rate of 92.3% (95% CI: 89.5-94.3) in the Enhertu arm and 83.5% with T-DM1 (95% CI: 79.9-86.4).

“For patients with HER2 positive early breast cancer and residual invasive disease after neoadjuvant therapy, reducing the risk of disease recurrence is critical for sustained long-term outcomes,” said Sibylle Loibl, MD, PhD, GBG (German Breast Group) and Goethe University Medicine Frankfurt, Frankfurt am Main and lead investigator of the DESTINY-Breast05 trial. “The EU approval of trastuzumab deruxtecan, which demonstrated a more than 50% reduction in risk of recurrence compared to standard of care in DESTINY-Breast05, has the potential to change how we treat eligible patients in this setting.”

In DESTINY-Breast05, the safety profile of Enhertu was consistent with the known profile with no new safety concerns identified. Grade 3 or grade 4 adverse reactions from a pooled safety analysis of 4,146 patients who received at least one dose of Enhertu (5.4 mg/kg) in clinical studies across multiple tumor types included neutropenia (18.7%), anemia (8.2%), fatigue (7.4%), leukopenia (6.6%), thrombocytopenia (5.4%), lymphopenia (5.0%), nausea (4.5%), increased transaminases (3.3%), hypokalemia (3.1%), pancytopenia (2.8%), diarrhea (2.1%), vomiting (2.0%), decreased appetite (1.4%) and pneumonia (1.2%). Grade 5 adverse reactions occurred in 0.9% of patients, including interstitial lung disease (ILD)/pneumonitis (0.8%).

“By bringing Enhertu into the adjuvant setting, we are advancing this treatment option earlier in the course of disease for patients with HER2 positive early breast cancer,” said Ken Keller, Global Head of Oncology Business, and President and CEO, Daiichi Sankyo, Inc. “This first approval of Enhertu in early breast cancer in the EU reflects our continued commitment to bringing innovative medicines into earlier treatment settings, where there is significant potential to improve long-term outcomes for patients.”

“The goal of treatment in early breast cancer is to help as many patients as possible achieve cure. Unfortunately, despite treatment advances, some patients still face recurrence in the long-term,” said Dave Fredrickson, Executive Vice President, Oncology Hematology Business Unit, AstraZeneca. “The approval of Enhertu in the adjuvant setting for patients with residual disease after neoadjuvant treatment has the potential to establish a new benchmark in this curative-intent setting, with more than 92% of patients alive and free of invasive disease at three years.”

Enhertu (5.4 mg/kg) is now approved in more than 35 countries/regions worldwide for the adjuvant treatment of adult patients with HER2 positive breast cancer who have residual invasive disease following neoadjuvant trastuzumab (with or without pertuzumab) and taxane-based treatment based on the DESTINY-Breast05 trial.

Financial Considerations

Following this approval in the EU, an amount of $75 million is due from AstraZeneca to Daiichi Sankyo as a milestone payment for the adjuvant HER2 positive early breast cancer indication. Sales of Enhertu in most EU territories are recognized by Daiichi Sankyo. For further details on the financial arrangements, please consult the collaboration agreement from March 2019.

About DESTINY-Breast05

DESTINY-Breast05 is a global, multicenter, randomized, open-label, phase 3 trial evaluating the efficacy and safety of Enhertu (5.4 mg/kg) versus T-DM1 in patients with HER2 positive early breast cancer with residual invasive disease in breast or axillary lymph nodes following neoadjuvant therapy and a high risk of recurrence. High risk of recurrence was defined as presentation with inoperable cancer (prior to neoadjuvant therapy) or pathologically positive axillary lymph nodes following neoadjuvant therapy.

The primary endpoint of DESTINY-Breast05 is investigator-assessed IDFS, which is defined as the time from randomization until first invasive local, axillary or distant recurrence or death from any cause. The key secondary endpoint is investigator-assessed DFS. Other secondary endpoints include overall survival, distant recurrence-free interval, brain metastases-free interval and safety.

DESTINY-Breast05 enrolled 1,635 patients in Asia, Europe, North America, Oceania and South America. For more information about the trial, visit ClinicalTrials.gov.

About HER2 Positive Early Breast Cancer

Breast cancer is the most common cancer in women worldwide and the leading cause of cancer-related deaths among women.1 Approximately 2.4 million breast cancer cases were diagnosed in 2024, with more than 690,000 deaths globally.1 In Europe, approximately 540,000 cases of breast cancer were diagnosed in 2024, with more than 140,000 deaths.2

HER2 is a tyrosine kinase receptor growth-promoting protein expressed on the surface of many types of tumors including breast cancer.3 HER2 protein overexpression may occur as a result of HER2 gene amplification and is often associated with aggressive disease and poor prognosis in breast cancer.3 Approximately one in five cases of breast cancer is considered HER2 positive.4

Approximately one in three patients with HER2 positive early-stage breast cancer is considered high-risk, meaning they are more likely to experience disease recurrence and have a poor prognosis.5 Despite receiving additional treatment with current standard of care for residual disease, some patients still experience invasive disease or death.6 Once patients are diagnosed with metastatic disease, the five-year survival rate drops from nearly 100% to approximately 34%.7

Adjuvant therapy represents a key opportunity to minimize the risk of recurrence and prevent progression to metastatic disease for patients with residual disease.8,9,10 Prior to the approval of Enhertu in the HER2 positive adjuvant (after surgery) setting for patients with residual invasive disease, the standard of care was T-DM1.11

About Enhertu

Enhertu (trastuzumab deruxtecan; fam-trastuzumab deruxtecan-nxki in the U.S. only) is a HER2 directed ADC. Designed using the proprietary DXd ADC Technology of Daiichi Sankyo, Enhertu is the lead ADC in the oncology portfolio of Daiichi Sankyo and the most advanced program in AstraZeneca’s ADC scientific platform. Enhertu consists of a HER2 monoclonal antibody attached to a number of topoisomerase I inhibitor payloads (an exatecan derivative, DXd) via tetrapeptide-based cleavable linkers.

Enhertu (5.4 mg/kg) followed by THP is approved in more than five countries/regions worldwide as a neoadjuvant treatment for adult patients with HER2 positive (IHC 3+ or ISH+) stage 2 or stage 3 breast cancer based on the results from the DESTINY-Breast11 trial.

Enhertu (5.4 mg/kg) is approved in more than 35 countries/regions worldwide for the adjuvant treatment of adult patients with HER2 positive breast cancer who have residual invasive disease following neoadjuvant trastuzumab (with or without pertuzumab) and taxane-based treatment based on the DESTINY-Breast05 trial.

Enhertu (5.4 mg/kg) in combination with pertuzumab is approved in more than 40 countries/regions worldwide as a first-line treatment for adult patients with unresectable or metastatic HER2 positive (IHC 3+ or ISH+) breast cancer, as determined by a locally or regionally approved test, based on the results from the DESTINY-Breast09 trial.

Enhertu (5.4 mg/kg) is approved in more than 100 countries/regions worldwide for the treatment of adult patients with unresectable or metastatic HER2 positive (IHC 3+ or ISH+) breast cancer who have received a prior anti-HER2-based regimen, either in the metastatic setting or in the neoadjuvant or adjuvant setting, and have developed disease recurrence during or within six months of completing therapy based on the results from the DESTINY-Breast03 trial.

Enhertu (5.4 mg/kg) is approved in more than 75 countries/regions worldwide for the treatment of adult patients with unresectable or metastatic hormone receptor (HR) positive, HER2 low (IHC 1+ or IHC 2+/ ISH-) or HER2 ultralow (IHC 0 with membrane staining) breast cancer, as determined by a locally or regionally approved test, that have progressed on one or more endocrine therapies in the metastatic setting based on the results from the DESTINY-Breast06 trial.

Enhertu (5.4 mg/kg) is approved in more than 100 countries/regions worldwide for the treatment of adult patients with unresectable or metastatic HER2 low (IHC 1+ or IHC 2+/ISH-) breast cancer who have received a prior systemic therapy in the metastatic setting or developed disease recurrence during or within six months of completing adjuvant chemotherapy based on the results from the DESTINY-Breast04 trial.

Enhertu (5.4 mg/kg) is approved in more than 80 countries/regions worldwide for the treatment of adult patients with unresectable or metastatic NSCLC whose tumors have activating HER2 (ERBB2) mutations, as detected by a locally or regionally approved test, and who have received a prior systemic therapy based on the results from the DESTINY-Lung02 and/or DESTINY-Lung05 trials. Continued approval in China and the U.S. for this indication may be contingent upon verification and description of clinical benefit in a confirmatory trial.

Enhertu (6.4 mg/kg) is approved in more than 90 countries/regions worldwide for the treatment of adult patients with locally advanced or metastatic HER2 positive (IHC 3+ or IHC 2+/ISH+) gastric or gastroesophageal junction (GEJ) adenocarcinoma who have received a prior trastuzumab-based regimen based on the results from the DESTINY-Gastric01, DESTINY-Gastric02 and/or DESTINY-Gastric04 trials.

Enhertu (5.4 mg/kg) is approved in more than 50 countries/regions worldwide for the treatment of adult patients with unresectable or metastatic HER2 positive (IHC 3+) solid tumors who have received prior treatment and have no satisfactory alternative treatment options based on efficacy results from the DESTINY-PanTumor02, DESTINY-Lung01, DESTINY-CRC02, DESTINY-PanTumor03 and/or HERALD trials. Continued approval in China and the U.S. for this indication may be contingent upon verification and description of clinical benefit in a confirmatory trial.

About the Enhertu Clinical Development Program

A comprehensive global clinical development program is underway evaluating the efficacy and safety of Enhertu as a monotherapy or in combination or sequentially with other cancer medicines across multiple HER2 targetable cancers.

About the Daiichi Sankyo and AstraZeneca Collaboration

Daiichi Sankyo and AstraZeneca entered into a global collaboration to jointly develop and commercialize Enhertu in March 2019 and Datroway® in July 2020, except in Japan where Daiichi Sankyo maintains exclusive rights for each ADC. Daiichi Sankyo is responsible for the manufacturing and supply of Enhertu and Datroway.

About the ADC Portfolio of Daiichi Sankyo

The Daiichi Sankyo ADC portfolio consists of nine ADCs in clinical development crafted from ADC technology discovered in-house by Daiichi Sankyo.

The DXd ADC Technology platform of Daiichi Sankyo consists of seven ADCs in clinical development where each ADC is comprised of a monoclonal antibody attached to a number of topoisomerase I inhibitor payloads (an exatecan derivative, DXd) via tetrapeptide-based cleavable linkers. The DXd ADCs include Enhertu and Datroway, which are being jointly developed and commercialized globally with AstraZeneca, and ifinatamab deruxtecan (I-DXd), raludotatug deruxtecan (R-DXd) and patritumab deruxtecan (HER3-DXd), which are being jointly developed and commercialized globally with Merck & Co., Inc., Rahway, NJ, USA. DS-3939 and DS3790 are being developed by Daiichi Sankyo.

Additional ADCs being developed by Daiichi Sankyo include DS3610, which consists of an antibody attached to a novel payload that acts as an agonist of STING, and DS1025, which consists of a CD25 directed antibody attached to an immuno-oncology optimized cytotoxic payload.

Ifinatamab deruxtecan, raludotatug deruxtecan, patritumab deruxtecan, DS-3939, DS3610, DS3790 and DS1025 are investigational medicines that have not been approved for any indication in any country. Safety and efficacy have not been established.

About Daiichi Sankyo

Daiichi Sankyo (TSE: 4568) is a global healthcare company committed to becoming a trusted healthcare innovator, transforming the lives of people through its strength in science and technology. The company discovers and develops new standards of care to address diverse medical needs to fulfill its purpose of contributing to the enrichment of quality of life around the world. With a strategic focus on oncology, Daiichi Sankyo is advancing an industry-leading antibody drug conjugate portfolio along with identifying new breakthrough generating technologies to deliver practice-changing medicines to patients, healthcare professionals and society. For more information, please visit www.daiichisankyo.com.

REFERENCES

1 World Health Organization. Global Status Report On Cancer 2026: The Future We Choose Together. Accessed October 2026.

2 World Health Organization. Breast Fact Sheet. Accessed October 2026.

3 Cheng X. Genes. 2024;15(7):903.

4 Tarantino P, et al. Ann Oncol. 2023;34(8):645-659.

5 Mahtani R, et al. Cancers (Basel). 2025;17(11):1848.

6 Geyer CE, et al. N Engl J Med. 2025;392(3):249-257.

7 National Cancer Institute. SEER Cancer Stat Facts: Female Breast Cancer Subtypes. Accessed October 2026.

8 von Minckwitz G, et al. N Engl J Med. 2019;380(7):617-628.

9 Zaborowski AM and Wong SM. BJS. 2023;110(7):765-772.

10 Loibl S, et al. N Engl J Med. 2026;394(9):845-857.

11 Loibl S, et al. ESMO Clinical Practice Guidelines. 2024.

Contacts

MEDIA CONTACTS:
Global:
Jennifer Brennan

jennifer.brennan@daiichisankyo.com
+1 908 900 3183 (mobile)

EU:
Simone Jendsch-Dowé

simone.jendsch-dowe@daiichisankyo.com
+49 (89) 78080 (office)

Japan:
DS-PR_jp@daiichisankyo.com

INVESTOR RELATIONS CONTACT:
DaiichiSankyoIR_jp@daiichisankyo.com
Avanzanite Bioscience Expands UK Presence and Appoints Sarah O’Connell as Country Manager

Avanzanite Bioscience Expands UK Presence and Appoints Sarah O’Connell as Country Manager


Avanzanite Bioscience Expands UK Presence and Appoints Sarah O’Connell as Country Manager

AMSTERDAM–(BUSINESS WIRE)–#Avanzanite–Avanzanite Bioscience B.V., a European pharmaceutical company focused on rare diseases, today announced the expansion of its United Kingdom (UK) presence and the appointment of Sarah O’Connell as UK Country Manager. The move strengthens Avanzanite’s long-term commitment to the UK and its ability to help eligible patients access innovative rare disease medicines.


“The UK is essential to our ambition of building Avanzanite into a European rare disease champion. We feel a deep and lasting connection to this market and there is no European reach without the UK,” said Adam Plich, co-founder and CEO, Avanzanite Bioscience. “At a time when parts of the life sciences sector are reassessing their investment and presence in the UK, we are choosing to commit and build for the long term. With Sarah’s extensive rare disease, market access and commercial leadership experience, we are strengthening the local capabilities and execution needed to ensure no patient in need is left behind.”

Sarah’s pharmaceutical career spans more than 25 years, including senior strategic roles at Amicus Therapeutics and Chiesi. At Amicus, she led a small, nimble team that achieved market leadership within the treatable population just eight months after launch.

“Scientific innovation is only the beginning; patient impact is the goal,” said Sarah O’Connell, UK Country Manager, Avanzanite Bioscience. “The UK combines exceptional scientific expertise with a health system capable of delivering innovation at scale, but turning that potential into patient impact requires partnership, innovative thinking, and disciplined Champions League-level execution. I am excited to build an agile UK organisation and work closely with the NHS, clinicians, and patient communities to help innovative rare disease medicines reach the people who need them.”

The UK expansion builds on Avanzanite’s existing team of 13 UK-based employees, including three executives. It is the latest step in Avanzanite’s rapid development of an integrated pan-European platform designed to turn Europe’s complexity into commercial opportunity. With Sarah’s appointment, Avanzanite’s “Champions League” team now includes more than 100 employees operating across 32 countries, including the European Economic Area, Switzerland and the UK. Together, these dedicated country teams and integrated capabilities support Avanzanite’s partners reach eligible patients, including in countries with challenging access landscapes as well as those in smaller and underserved markets.

About Avanzanite Bioscience

Avanzanite is a next-generation pharmaceutical company purpose-built to unlock the full potential of rare disease medicines across Europe. Founded in 2022 and headquartered in Amsterdam, the Netherlands, Avanzanite was created around a belief that scientific innovation creates patient impact only when medicines reach those who can benefit from them. Through tailored partnerships, biopharma innovators entrust Avanzanite with the European future of their medicines. Avanzanite combines pan-European expertise, bold commercial and access strategy, together with high-performance execution, to transform Europe’s complexity into opportunity, ensuring no patient is left behind and creating lasting long-term value for partners.

For more information, visit www.avanzanite.com.

Contacts

Avanzanite Bioscience B.V.

Phone: +31 20 301 21 13

Email: media@avanzanite.com
Website: www.avanzanite.com

Innate Pharma: PACIFIC-9 Phase 3 Trial Readout Timing Updated to H1 2027

Innate Pharma: PACIFIC-9 Phase 3 Trial Readout Timing Updated to H1 2027


Innate Pharma: PACIFIC-9 Phase 3 Trial Readout Timing Updated to H1 2027

  • PACIFIC-9 Phase 3 trial led by AstraZeneca evaluates durvalumab in combination with monalizumab or oleclumab in unresectable Stage III NSCLC

MARSEILLE, France–(BUSINESS WIRE)–$IPH #ADC–Regulatory News:


Innate Pharma SA (Euronext Paris: IPH; Nasdaq: IPHA) (“Innate” or the “Company”) today announced that AstraZeneca has updated the expected timing for the PACIFIC-9 Phase 3 trial high-level data readout to the first half of 2027, from the second half of 2026 previously. The revised timing reflects the event-driven nature of the study.

PACIFIC-9 is an AstraZeneca-led Phase 3 trial evaluating durvalumab in combination with monalizumab or oleclumab in patients with unresectable Stage III non-small cell lung cancer (NSCLC) who have not progressed following platinum-based chemoradiation therapy. Enrollment in the trial is complete.

Monalizumab is an anti-NKG2A antibody developed by Innate Pharma in collaboration with AstraZeneca. AstraZeneca obtained full oncology rights to monalizumab in October 2018 through a co-development and commercialisation agreement initiated in 2015.

About Innate Pharma

Innate Pharma S.A. is a global, clinical-stage biotechnology company developing immunotherapies for cancer patients. Leveraging its expertise in antibody engineering and innovative target identification, Innate Pharma is developing innovative and differentiated next-generation antibody therapeutics.

Innate Pharma is advancing a portfolio of differentiated potential first- and/or best-in-class assets, focused on areas of high unmet medical need. Its proprietary pipeline is centered on antibody-drug conjugates (ADCs), led by IPH4502, a differentiated Nectin-4 ADC in clinical development for solid tumors, and supported by a preclinical portfolio of next-generation ADC candidates. In parallel, Innate is advancing two partnered late-stage assets: lacutamab, developed with Sobi for T-cell lymphomas, and monalizumab, developed with AstraZeneca for non-small cell lung cancer (NSCLC).

Innate Pharma has established collaborations with leading biopharmaceutical companies, including Sobi, Sanofi and AstraZeneca, as well as renowned academic and research institutions, to advance innovation in immuno-oncology.

Headquartered in Marseille, France, Innate Pharma is listed on Euronext Paris and Nasdaq in the US.

Learn more about Innate Pharma at www.innate-pharma.com and follow us on LinkedIn and X.

Information about Innate Pharma shares

ISIN code: FR0010331421

Ticker code: Euronext Paris: IPH | Nasdaq: IPHA

LEI: 9695002Y8420ZB8HJE29

Disclaimer on forward-looking information and risk factors

For a discussion of risks and uncertainties, please refer to the Risk Factors (“Facteurs de Risque”) section of the Universal Registration Document filed with the French Financial Markets Authority (“AMF”), which is available on the AMF website http://www.amf-france.org or on Innate Pharma’s website, and public filings and reports filed with the U.S. Securities and Exchange Commission (“SEC”), including the Company’s Annual Report on Form 20-F for the year ended December 31, 2025, and subsequent filings and reports filed with the AMF or SEC, or otherwise made public by the Company. References to the Company’s website and the AMF website are included for information only and the content contained therein, or that can be accessed through them, are not incorporated by reference into, and do not constitute a part of, this press release.

This press release and the information contained herein do not constitute an offer to sell or a solicitation of an offer to buy or subscribe to shares in Innate Pharma in any country.

Contacts

Stéphanie Cornen

VP, Investor Relations & Corporate Communications

stephanie.cornen@innate-pharma.fr

Investor Relations

investors@innate-pharma.fr

Media

communication@innate-pharma.fr

MedTech Innovator Asia Pacific Names Ardens Lifescience 2026 Grand Prize Winner

MedTech Innovator Asia Pacific Names Ardens Lifescience 2026 Grand Prize Winner


MedTech Innovator Asia Pacific Names Ardens Lifescience 2026 Grand Prize Winner

South Korea medical technology company developing a transbronchial RF ablation system with a conforming electrode for peripheral lung cancer awarded USD 175,000 Grand Prize at the 2026 MedTech Innovator Asia Pacific Showcase & Grand Finals in Singapore

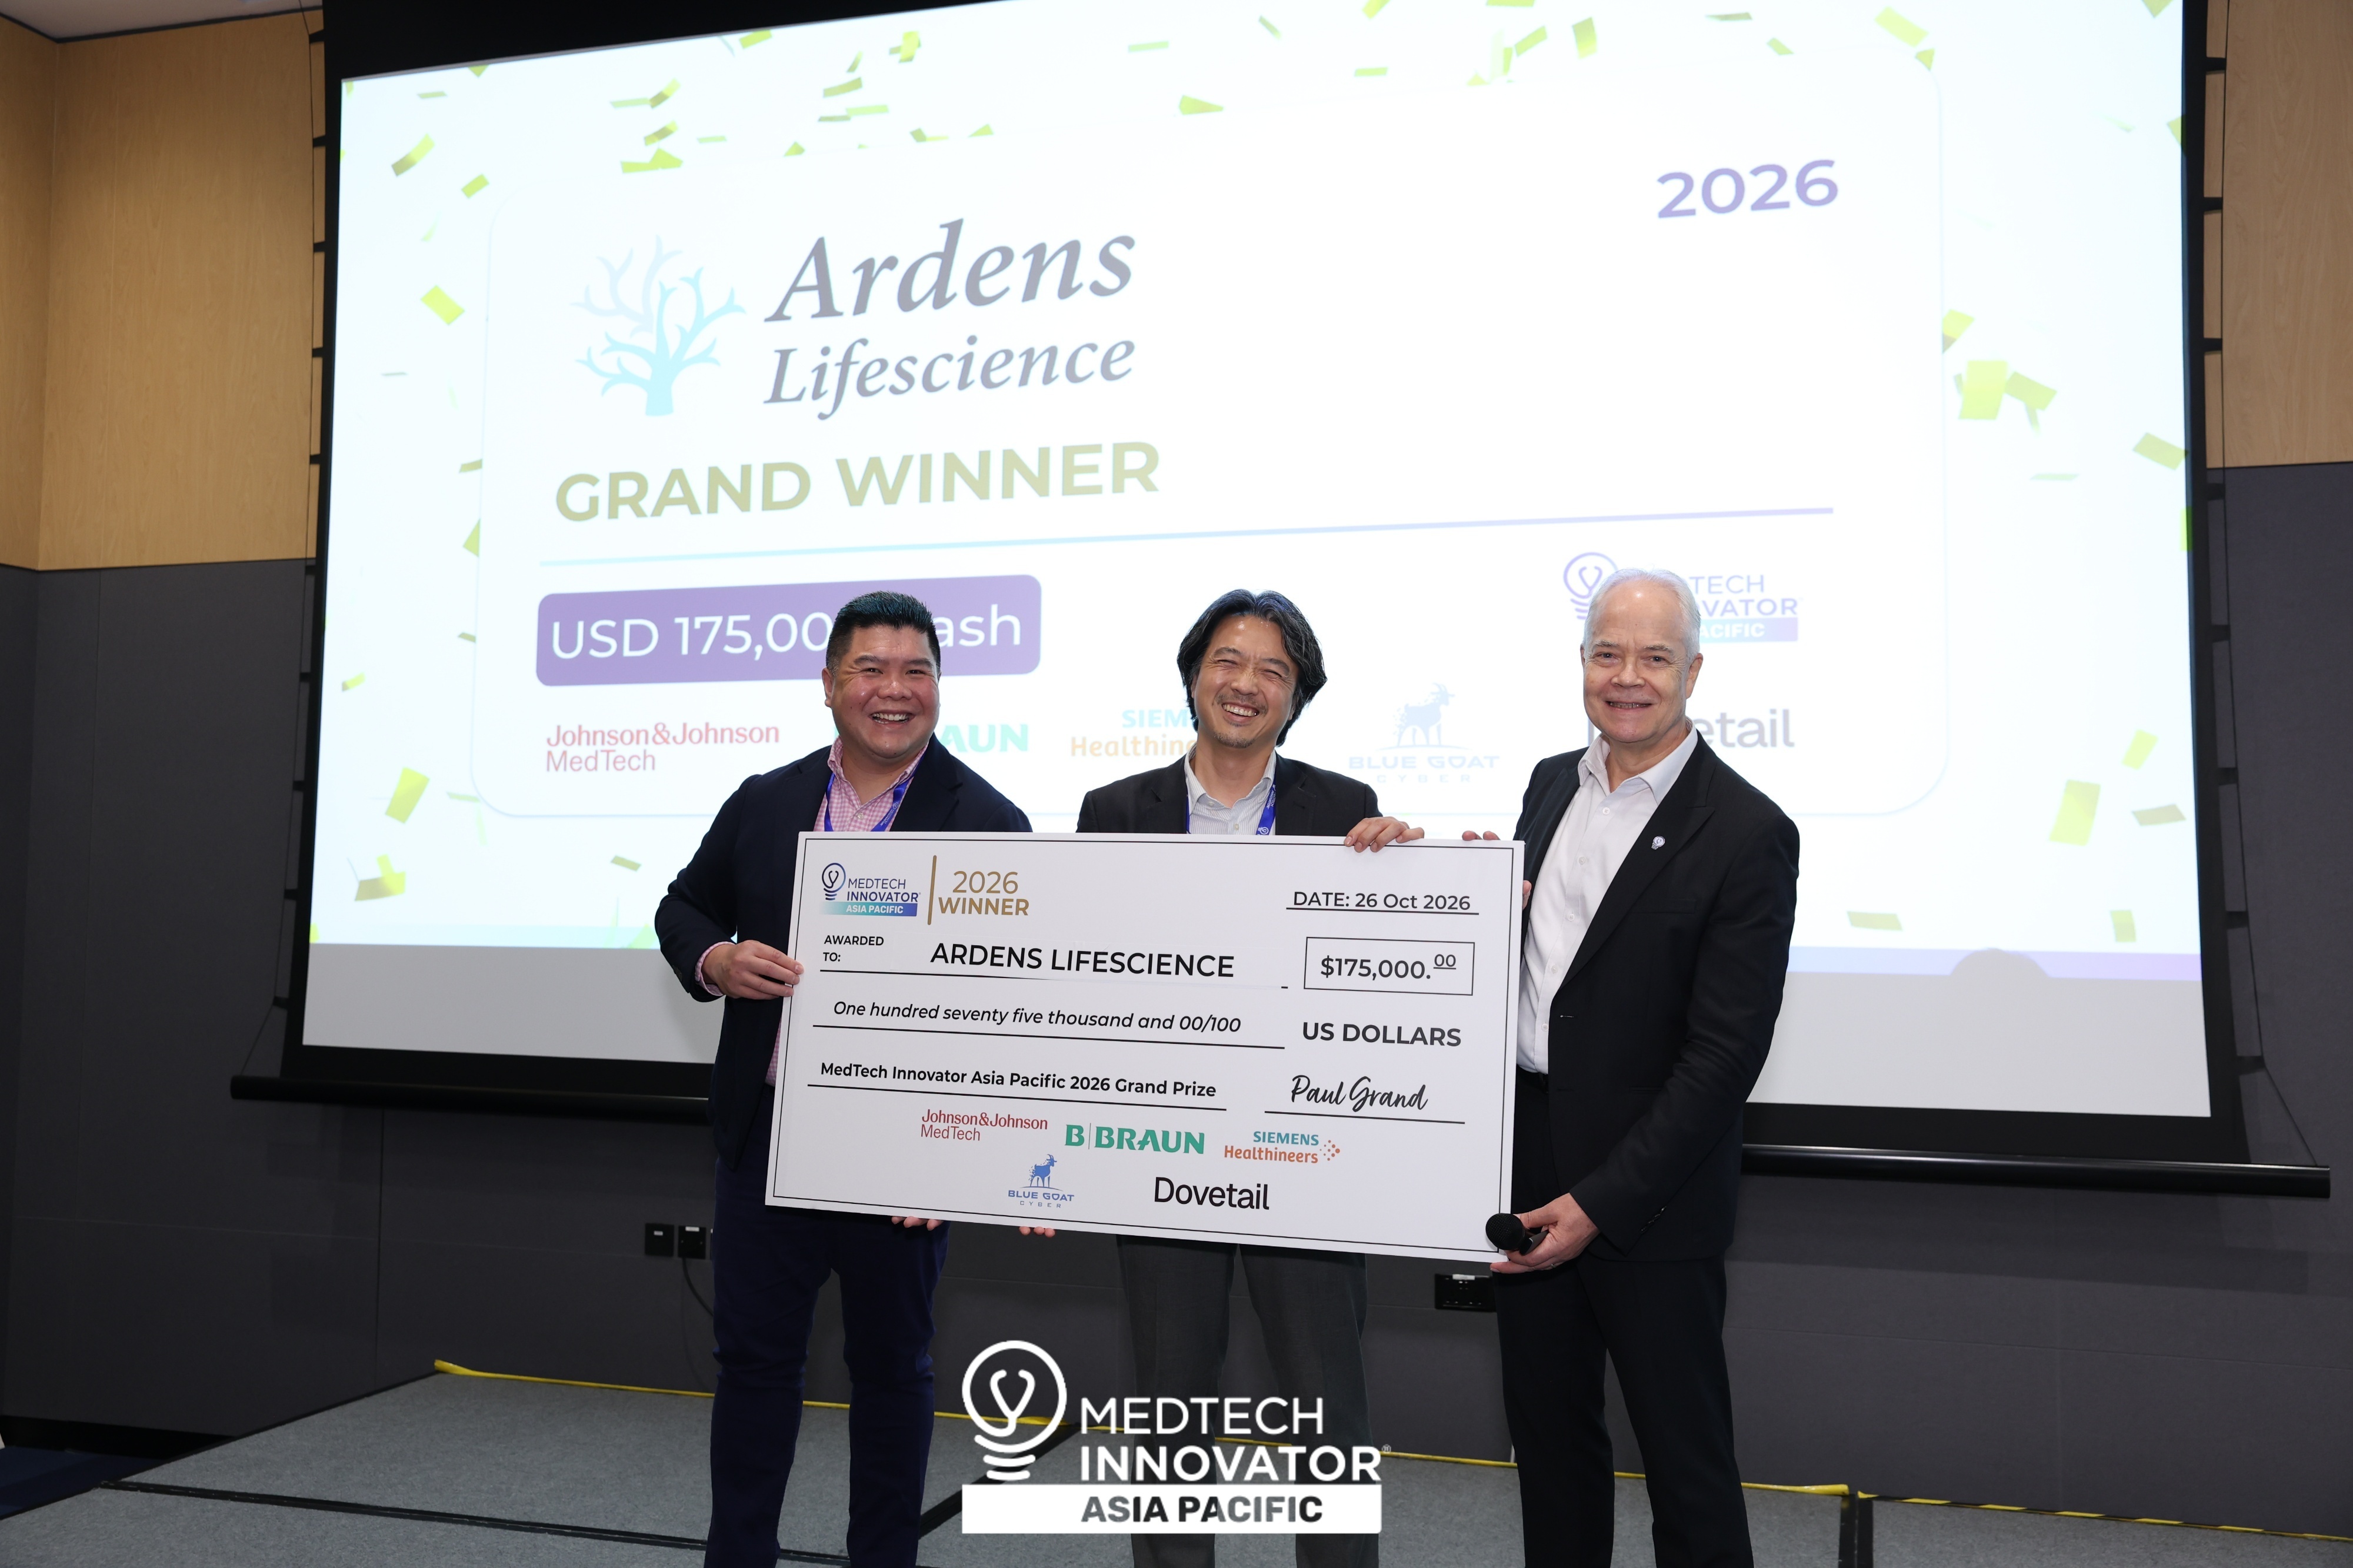
LOS ANGELES & SINGAPORE–(BUSINESS WIRE)–#Innovation—MedTech Innovator (MTI), the world’s largest accelerator of medical device, digital health, and diagnostic companies, today announced Ardens Lifescience as the 2026 MedTech Innovator Asia Pacific Grand Prize Winner, awarding the company USD 175,000 cash prize for its transbronchial radiofrequency (RF) ablation system with a conforming electrode for peripheral lung cancer.


The announcement was made at the MedTech Innovator Asia Pacific Showcase & Grand Finals in Singapore, where four finalists – Ardens Lifescience, BaiYi Healthcare Technology, Eyes of AI, and Physiologas Technology – competed before an audience of industry leaders, investors, healthcare stakeholders and innovators. Following the finalist presentations and a live Q&A with a panel of medtech executives, the audience selected Ardens Lifescience as the 2026 Grand Prize Winner. The three runners-up each received USD 10,000.

“We are incredibly honored to be named the 2026 MedTech Innovator Asia Pacific Grand Prize Winner,” said Alex Park, CEO of Ardens Lifescience. “This recognition is a significant milestone for our team. I believe we were able to become the winner because all of you resonated with and supported our mission to offer more effective, yet less invasive, treatments for patients suffering from lung cancer. With an even greater sense of responsibility, Ardens’ teams will do our best to bring CAROL to the world quickly to provide hope to many lung cancer patients.”

“Ardens Lifescience is advancing an important innovation in the treatment of peripheral lung cancer,” said Paul Grand, founder and CEO of MedTech Innovator. “Its conforming radiofrequency ablation technology has the potential to overcome key limitations of existing treatment approaches, enabling more precise tumor targeting while preserving healthy lung tissue. As earlier detection creates new opportunities to intervene, innovations like CAROL could play an important role in expanding less invasive treatment options for patients. We congratulate Ardens on this well-deserved recognition as our 2026 Asia Pacific Grand Prize Winner.”

The 2026 Grand Finals marked the culmination of MedTech Innovator Asia Pacific’s four-month accelerator program, which brought together 20 promising medical technology companies selected from more than 680 applications across the region and beyond. Representing a diverse range of healthcare technologies, the cohort featured innovations spanning medical devices, diagnostics, digital health, artificial intelligence, surgical technologies, and other emerging areas of healthcare.

The Grand Finals judging panel included Aditya Vaidya, General Manager, Asia Pacific – Heart Recovery, Johnson & Johnson, Santosh Nair, Head of Data & AI Portfolio, Siemens Healthineers, and Esther Kong, Manager, Innovation Research, B.Braun Medical Industries.

In addition to the Grand Prize and runner-up awards, several honors were presented to recognize outstanding achievements among the 2026 cohort:

EnterpriseSG STARTUP SG Grant (SGD 50,000): BaiYi Healthcare Technology, China

Blue Goat Cyber Cybersecurity Award (USD 25,000): mediVR Inc., Japan

Dovetail Regulatory Award (USD 25,000): Lumino Medison, South Korea

Best Video Competition Award (USD 5,000): Ardens Lifescience, South Korea

Throughout the four-month accelerator, participating companies received tailored mentorship, strategic guidance, and opportunities to engage with leading medtech corporations, investors, healthcare stakeholders, and industry experts. Through MedTech Innovator’s global network, the program connected companies with the expertise, partnerships, and resources needed to support clinical development, commercialization, and international growth.

Now in its eighth year, MedTech Innovator Asia Pacific has established a strong regional platform for identifying and advancing promising medical technology companies, fostering collaboration across the healthcare ecosystem, and connecting innovators with strategic partners and market opportunities across Asia Pacific and beyond.

MedTech Innovator Asia Pacific’s 2026 program partners include Johnson & Johnson, B.Braun Medical Industries, Siemens Healthineers, Blue Goat Cyber, Dovetail, Cambridge Consultants, The Mullings Group, and Enterprise Singapore joining as a 2026 partner to support regional expansion and innovation initiatives.

About MedTech Innovator

MedTech Innovator is the world’s largest accelerator of medical device, digital health, and diagnostic companies. Its mission is to improve human health by accelerating the growth of companies transforming patient care. MTI has been a catalyst for groundbreaking healthcare solutions, sourcing more than 20,000 applicants and fostering the growth of 933 graduates. Alumni have collectively raised $12 billion in follow-on funding, seen 66 acquisitions, and introduced 600+ products to the market, improving the health of millions worldwide. For more information about MedTech Innovator Asia Pacific, its annual programs, portfolio of industry-leading startups, and insights on trends, visit MTI’s website, follow MTI on LinkedIn, and subscribe to its monthly newsletter.

Contacts

Media Contact:
Bernice Tan

Program & Marketing Manager

bernice@medtechinnovator.org

AimedBio and Boehringer Ingelheim initiate Phase I study of antibody-drug conjugate BI 4060107

AimedBio and Boehringer Ingelheim initiate Phase I study of antibody-drug conjugate BI 4060107


AimedBio and Boehringer Ingelheim initiate Phase I study of antibody-drug conjugate BI 4060107

First patient dosed in first-in-human study evaluating BI 4060107 in people with advanced solid tumors

Clinical development milestone triggers a payment to AimedBio under the companies’ 2025 global licensing agreement

SEOUL, South Korea–(BUSINESS WIRE)–#ADC—AimedBio today announced that the first patient has been dosed in a Phase I study of BI 4060107, an investigational antibody-drug conjugate (ADC) discovered by AimedBio and now being developed by Boehringer Ingelheim under the companies’ global collaboration and licensing agreement. The first-in-human study (NCT07827144) is evaluating different doses of BI 4060107 in people with advanced solid tumors, with a focus on how well the treatment is tolerated.


Despite significant advances in cancer treatment, many people with advanced solid tumors still lack precision treatment options. ADCs are a promising approach to addressing this challenge, combining the specificity of an antibody with a cancer-cell-killing payload to potentially deliver treatment more precisely to tumor cells while limiting exposure to healthy tissue. BI 4060107 was developed using AimedBio’s proprietary P-ADC™ platform, which takes a patient-driven approach to identifying and validating targets for targeted cancer therapies.

“Dosing the first patient marks an important step for BI 4060107, as the program moves from discovery into clinical evaluation,” said Nam-Gu Her, CEO of AimedBio. “BI 4060107 represents another potential first-in-class ADC discovered by AimedBio to enter clinical development. We look forward to its continued development for patients facing difficult-to-treat cancers.”

AimedBio and Boehringer Ingelheim entered into a global collaboration and licensing agreement in October 2025 for BI 4060107. The investigational compound targets a tumor-selective marker that is highly expressed across a broad spectrum of cancers, minimally present in normal tissues and associated with tumor growth, metastasis and resistance to therapy.

“Starting clinical development is an important step toward our goal of bringing BI 4060107 to people with cancer,” said Vittoria Zinzalla, Global Head of Experimental Medicine at Boehringer Ingelheim. “It reflects our broader ambition in oncology to create unprecedented impact for people facing cancer by pioneering novel approaches and advancing first-in-class treatments.”

With the achievement of this development milestone, AimedBio is eligible to receive a milestone payment from Boehringer Ingelheim under the agreement. Under the collaboration and license agreement, AimedBio is eligible to receive up to $991 million in upfront, development and regulatory, and commercial milestone payments, plus royalties on net sales. The amount of the payment associated with this milestone is not being disclosed.

About AimedBio

AimedBio is a KOSDAQ-listed, clinical-stage biotech company focused on developing first-in-class ADCs, with core strengths in target discovery and validation, antibody discovery, and translational research. Founded in 2018 as a spin-off from Samsung Medical Center, the company integrates patient-derived data and samples throughout discovery and development to advance differentiated cancer therapies.

Contacts

AimedBio

Munyeong Huh

munyeong.huh@aimedbio.com

Mirum Pharmaceuticals Announces Positive Results from Cohort 2 of the Pivotal PROGRESS Study of ATEBRIOZ™ (zilurgisertib) in Pediatric Patients Aged 6 to <12 Years with Fibrodysplasia Ossificans Progressiva

Mirum Pharmaceuticals Announces Positive Results from Cohort 2 of the Pivotal PROGRESS Study of ATEBRIOZ™ (zilurgisertib) in Pediatric Patients Aged 6 to <12 Years with Fibrodysplasia Ossificans Progressiva


Mirum Pharmaceuticals Announces Positive Results from Cohort 2 of the Pivotal PROGRESS Study of ATEBRIOZ™ (zilurgisertib) in Pediatric Patients Aged 6 to <12 Years with Fibrodysplasia Ossificans Progressiva

– Zilurgisertib reduced total HO lesion volume and improved clinician-assessed joint mobility; no zilurgisertib-treated patients developed new HO lesions

– Mirum plans to submit an sNDA in H1 2027 to expand the ATEBRIOZ indication to pediatric patients aged 6 to <12 years

– Safety and efficacy findings consistent with PROGRESS Cohort 1; no new safety signals

FOSTER CITY, Calif.–(BUSINESS WIRE)–Mirum Pharmaceuticals, Inc. (NASDAQ: MIRM), a leading rare disease company, today announced positive results from Cohort 2 of the pivotal PROGRESS study of ATEBRIOZ™ (zilurgisertib), an oral activin receptor-like kinase 2 (ALK2) inhibitor, in pediatric patients aged 6 to <12 years with fibrodysplasia ossificans progressiva (FOP). At Week 24, zilurgisertib reduced total heterotopic ossification (HO) lesion volume and improved clinician-assessed joint mobility. No zilurgisertib-treated patients developed new HO lesions.


The results build on data from PROGRESS Cohort 1, which supported the U.S. Food and Drug Administration (FDA) approval of ATEBRIOZ in September 2026 for adult and pediatric patients aged 12 years and older with FOP. Mirum plans a supplemental New Drug Application (sNDA) submission in H1 2027 to expand the indication to pediatric patients aged 6 to <12 years.

“These results represent an important advance for younger children living with FOP,” said Rob Myers, M.D., Chief Medical Officer of Mirum. “The reduction in total HO volume and absence of new lesions seen with zilurgisertib build on the positive results previously observed in adolescents and adults in Cohort 1 of PROGRESS. We look forward to discussing these findings with the FDA to expand access to ATEBRIOZ for younger children living with this debilitating disease.”

PROGRESS Cohort 2 Results

Cohort 2 of the PROGRESS study enrolled 25 pediatric patients aged 6 to <12 years with FOP. Patients were randomized 1:1 to receive once-daily oral zilurgisertib 75 mg (n=12) or placebo (n=13) during a 24-week double-blind period, followed by an open-label extension. The mean age at baseline was 8.3 years, with a mean of 5.0 years since initial diagnosis. All patients completed the double-blind period and entered the open-label extension.

At Week 24, mean total HO lesion volume decreased by 8.09 cm³ in patients receiving zilurgisertib, compared with an increase of 9.14 cm³ in patients receiving placebo (nominal p=0.003). No zilurgisertib-treated patients developed new HO lesions.

Improvements in clinician-assessed joint mobility were also observed. Joint mobility was assessed using the Cumulative Analogue Joint Involvement Scale (CAJIS), a validated measure of physical function in FOP scored from 0 (no impairment) to 30 (maximum impairment). At Week 24, CAJIS scores improved by a mean of 0.80 points in patients receiving zilurgisertib, compared with 0.09 points in patients receiving placebo. In untreated patients with FOP, CAJIS scores are reported to worsen by approximately 0.5 points per year.1 Key efficacy data from Cohort 2 are shown below in the Efficacy Results table.

Efficacy Results

Endpoint

Zilurgisertib

75 mg QD

Placebo

Key Finding

Mean (SD) change from baseline in total HO lesion volume, cm³

-8.09 (15.58)

n=12

9.14 (11.66)

n=13

Treatment differencea of -14.0 cm³ favoring zilurgisertib (nominal p=0.003).b

Patients with new HO lesions, n (%)

0 (0)

n=12

1 (7.7)

n=13

No new HO lesions with zilurgisertib.

Mean (SD) change from baseline in CAJIS score

-0.80 (1.1)

n=10

-0.09 (1.0)

n=11

Improvement in clinician-assessed joint mobility with zilurgisertib.

a Hodges-Lehmann estimate of treatment difference

b p-value calculated using a Wilcoxon rank-sum test

Zilurgisertib was generally well tolerated, with a safety profile consistent with that observed in PROGRESS Cohort 1. No new safety signals were identified. No patients discontinued treatment during the double-blind period, and no zilurgisertib-treated patients experienced growth plate closure. Additional safety findings included:

  • Most adverse events were mild or moderate in severity.
  • Serious adverse events and Grade ≥3 adverse events were infrequent and occurred less frequently in patients receiving zilurgisertib than in those receiving placebo.
  • Adverse events occurring in more than 10% of zilurgisertib-treated patients and at a higher frequency than with placebo included arthralgia, cough, pyrexia, and vomiting (3 subjects each), and alopecia, epistaxis, fall, iron deficiency, and nasopharyngitis (2 subjects each).

“FOP is a devastating genetic disease that leads to progressive loss of mobility, independence, quality of life, and shortened lifespan,” said Frederick S. Kaplan, M.D., Isaac and Rose Nassau Professor of Orthopaedic Molecular Medicine at the University of Pennsylvania Perelman School of Medicine. “Childhood is a critical window in FOP, so a therapy targeting the underlying cause of the disease is especially important for this young age group. Results such as these offer hope that we may be able to slow disease progression in children before its most serious consequences arise.”

ATEBRIOZ is approved by the U.S. FDA to reduce the volume of total new HO in adult and pediatric patients aged 12 years and older with FOP. Total new HO lesion volume includes expansion of baseline HO lesion burden as well as any new discrete HO that developed during the 24-week double-blind period.

Full Cohort 2 results will be presented at an upcoming medical meeting. Cohort 3 of the PROGRESS study, which is evaluating zilurgisertib in pediatric patients aged 2 to <12 years, is currently enrolling.

ATEBRIOZ is available in the U.S. through Mirum Access Plus (MAP), a patient support program designed to help patients, families, and healthcare providers navigate treatment access. MAP provides insurance coverage and access support, financial assistance for eligible patients, personalized patient support, and educational resources for patients and caregivers. Eligible patients can pay as little as $0 per month. To learn more, call 855-MRM-4YOU (1-855-676-4968) or visit atebrioz.com.

About ATEBRIOZ™ (zilurgisertib) tablets

ATEBRIOZ™ (zilurgisertib) tablets are a once-daily oral activin receptor-like kinase 2 (ALK2) inhibitor approved by the U.S. Food and Drug Administration (FDA) to reduce the volume of total new heterotopic ossification in adult and pediatric patients aged 12 years and older with Fibrodysplasia Ossificans Progressiva (FOP). In people living with FOP, pathogenic variants in the ACVR1 gene result in abnormal activation of ALK2, leading to the formation of bone in muscles, tendons, ligaments and other soft tissues through a process known as heterotopic ossification (HO). For more information, please visit atebrioz.com.

Mirum Pharmaceuticals, Inc. licensed zilurgisertib from Incyte for worldwide development and commercialization.

IMPORTANT SAFETY INFORMATION

ATEBRIOZ is contraindicated in pregnancy. ATEBRIOZ can cause fetal harm based on data from animal studies. Patients of reproductive potential should use effective contraception and should immediately discontinue ATEBRIOZ and contact their healthcare provider if pregnancy occurs.

US Prescribing Information

About the PROGRESS Study

PROGRESS is a global, randomized, double-blind, placebo-controlled Phase 2 study evaluating the efficacy and safety of zilurgisertib in patients with fibrodysplasia ossificans progressiva (FOP). PROGRESS Cohort 1 enrolled 63 patients 12 years of age and older who were randomized 1:1 to receive zilurgisertib 100 mg once daily or placebo during a 24-week double-blind treatment period, followed by an open-label extension. Results from Cohort 1 have been reported through Week 48.

The PROGRESS study is also evaluating the safety and efficacy of zilurgisertib in younger patients with FOP. Cohort 2 enrolled 25 patients aged 6 to <12 years who were randomized 1:1 to receive zilurgisertib 75 mg once daily or placebo during a 24-week double-blind treatment period, followed by an open-label extension. Cohort 3, which is evaluating zilurgisertib in patients aged 2 to <12 years, is currently enrolling.

About Fibrodysplasia Ossificans Progressiva (FOP)

Fibrodysplasia ossificans progressiva (FOP) is an ultra-rare, progressive genetic disease affecting approximately 300 people in the United States and 900 worldwide. FOP is characterized by heterotopic ossification (HO), a process in which bone forms in muscles, tendons, ligaments and other soft tissues. Symptoms typically become apparent in early childhood, and the number and volume of HO lesions increase over time, progressively restricting movement and limiting mobility, daily function, and independence.

About Mirum Pharmaceuticals

Mirum Pharmaceuticals (NASDAQ: MIRM) is a leading rare disease company with a global footprint of approved products and a broad pipeline of investigational medicines. Purpose-built to bring forward breakthrough medicines for people with overlooked conditions, Mirum focuses on rare liver and rare genetic diseases, where it has built deep expertise and strong connections to patient communities. The company’s commercial portfolio includes LIVMARLI® (maralixibat) for Alagille syndrome (ALGS) and progressive familial intrahepatic cholestasis (PFIC), ATEBRIOZ™ (zilurgisertib) for fibrodysplasia ossificans progressiva (FOP), CHOLBAM® (cholic acid) for bile-acid synthesis disorders and CTEXLI® (chenodiol) for cerebrotendinous xanthomatosis (CTX).

Mirum’s clinical-stage pipeline includes volixibat, an IBAT inhibitor in late-stage development for primary sclerosing cholangitis (PSC) and primary biliary cholangitis (PBC), brelovitug, a fully human monoclonal antibody in late-stage development for chronic hepatitis delta virus (HDV) and MRM-3379, a PDE4D inhibitor being evaluated for Fragile X syndrome (FXS).

Mirum’s success is driven by a team dedicated to advancing high impact medicines through strategic development, disciplined execution and purposeful collaboration across the rare disease ecosystem. Learn more at https://mirumpharma.com/ and follow Mirum on Facebook, LinkedIn, Instagram and X.

Mirum Forward-Looking Statements

Statements contained in this press release regarding matters that are not historical facts are “forward-looking statements” within the meaning of the Private Securities Litigation Reform Act of 1995. Such forward-looking statements include statements regarding, among other things, expectations regarding the timing, enrollment or results of clinical trials for zilurgisertib, the potential benefit of zilurgisertib in real world settings versus clinical trial settings, including the potential to slow disease progression in children before FOP’s most serious consequences arise, the timing of regulatory engagement with the FDA and any resulting submission of an sNDA for ages 6 to <12, the importance of ATEBRIOZ™ as an additional therapy for the treatment of FOP, the potential presentation of zilurgisertib clinical results at an upcoming medical meeting, cost to patients and methods of availability. Because such statements are subject to risks and uncertainties, actual results may differ materially from those expressed or implied by such forward-looking statements. Words such as “plan,” “may,” “expected,” “will,” “could,” “would,” “guidance,” “potential,” “continue” and similar expressions are intended to identify forward-looking statements. These forward-looking statements are based upon Mirum’s current expectations and involve assumptions that may never materialize or may prove to be incorrect. Actual results could differ materially from those anticipated in such forward-looking statements as a result of various risks and uncertainties, which include, without limitation, risks and uncertainties associated with the development of investigational medicines generally, including the failure of future studies to generate the same or similar data as prior studies, risks inherent in the regulatory submission process in the U.S., risks and uncertainties associated with Mirum’s business in general, the impact of geopolitical and macroeconomic events, and the other risks described in Mirum’s Quarterly Report for the quarter ended June 30, 2026, filed with the Securities and Exchange Commission on August 5, 2026, and subsequent filings with the Securities and Exchange Commission, which are available at www.sec.gov. All forward-looking statements contained in this press release speak only as of the date on which they were made and are based on management’s assumptions and estimates as of such date. Mirum undertakes no obligation to update such statements to reflect events that occur or circumstances that exist after the date on which they were made, except as required by law.

Mirum and the Mirum logo are trademarks of Mirum Pharmaceuticals, Inc.

References

1Pignolo RJ, et al. Orphanet J Rare Dis. 2019;14:113.

Contacts

Mirum Investor Contact:
Andrew McKibben

ir@mirumpharma.com

Mirum Media Contact:
Meredith Kiernan

media@mirumpharma.com

Aptar Pharma Acquires Pulmotree, Expanding Growth Opportunities in Next-Generation Pulmonary Drug Delivery

Aptar Pharma Acquires Pulmotree, Expanding Growth Opportunities in Next-Generation Pulmonary Drug Delivery


Aptar Pharma Acquires Pulmotree, Expanding Growth Opportunities in Next-Generation Pulmonary Drug Delivery

Acquisition is expected to accelerate development and commercialization of Kolibri™ and expand Aptar Pharma’s respiratory drug delivery platform

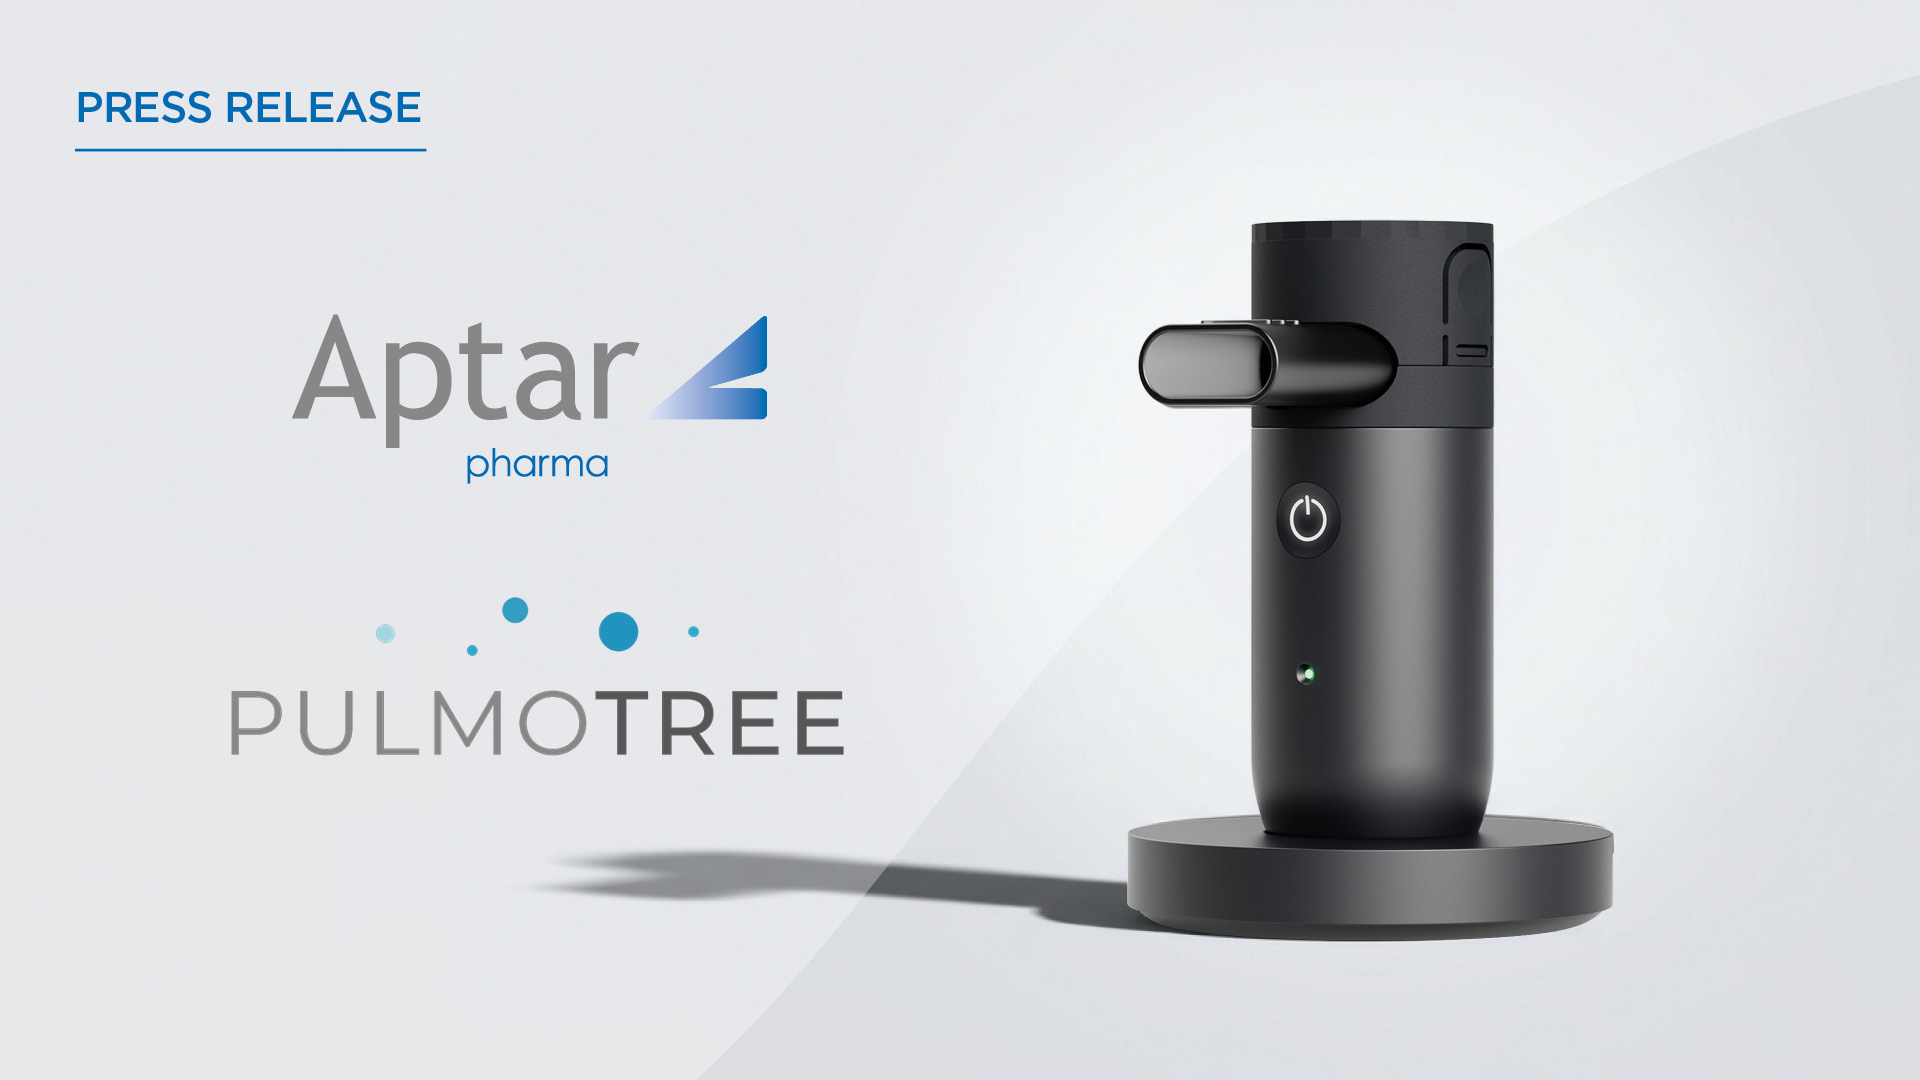
CRYSTAL LAKE, Ill.–(BUSINESS WIRE)–Aptar Pharma, a global leader in drug delivery, dosing, protection technologies and services, today announced the strategic acquisition of PULMOTREE, a Munich, Germany-based life science company specializing in pulmonary drug delivery systems for targeted delivery to the lungs. The acquisition is expected to accelerate the development, industrialization and commercialization of PULMOTREE’s Kolibri™ Non-Propellant Liquid Inhaler (NPLI) platform.


The acquisition is intended to expand Aptar Pharma’s opportunities for growth in respiratory drug delivery by adding a differentiated non-propellant platform capable of supporting the delivery of complex molecules, including biologics.

“The acquisition of PULMOTREE is an important step in expanding Aptar Pharma’s leadership in respiratory drug delivery and broadening the opportunities we can address with our pharmaceutical partners,” said Gael Touya, President and CEO of Aptar. “Kolibri™ brings a differentiated technology to our portfolio with the potential to support increasingly complex inhaled therapies, including biologics. By combining this platform with Aptar Pharma’s formulation, device development, regulatory, manufacturing and patient-support capabilities, we can help customers move innovative therapies from development toward commercialization while creating new avenues for long-term growth.”

Jonathan Mulpas, Director of Business Development, Pulmonary Category, Aptar Pharma, stated, “Our collaboration with PULMOTREE over the past two years has enabled pharmaceutical partners to evaluate and gain hands-on experience with the Kolibri™ platform. The interest we have seen reinforces our belief that there is a need for this differentiated patient-centric technology designed to support the next generation of inhaled therapies.”

Ulf Krüger, CEO of PULMOTREE, stated, “The partnership between PULMOTREE and Aptar Pharma has been built on close, trust-based collaboration, and joining Aptar is the natural next step on that shared path. For our customers and partners, this acquisition is intended to provide continuity in the technology – now backed by the reach and resources of a global organization.”

Beyond expanding Aptar Pharma’s respiratory device portfolio, the acquisition is expected to create opportunities to deepen customer relationships and capture greater value across the drug development lifecycle. By leveraging offerings within Aptar Pharma companies such as Nanopharm’s formulation development and analytical expertise, Noble’s Human Factors support and patient onboarding, and Aptar Digital Health’s services, Aptar Pharma can customize drug formulation development, optimize device design, and support efforts to improve patient outcomes to provide their customers with a value-added finished product.

Non-Propellant Liquid Inhalers encompass nebulizers and soft-mist inhalers and are being developed as an alternative to propellant-based systems. PULMOTREE’s Kolibri™ utilizes a robust breath-triggered Vibrating Mesh aerosolization engine with unique breath-guidance capabilities designed to enhance targeted lung deposition. The award-winning Kolibri™ technology incorporates advanced connectivity features that can support treatment monitoring, patient engagement and the generation of real-world therapy insights.

Building on the exclusive collaboration agreement established with PULMOTREE in 2024, Aptar Pharma intends to leverage its comprehensive capabilities and services, including regulatory, manufacturing and commercial expertise, with a goal of providing pharmaceutical partners with a clear pathway from development through clinical evaluation and market launch for Kolibri™.

About Aptar Pharma

Aptar Pharma is part of AptarGroup, Inc. (NYSE:ATR), a global leader in drug delivery, dosing and protection technologies, and consumer product dispensing. Aptar partners with the world’s top healthcare and consumer brands to deliver medicines and create exceptional user experiences. Serving diverse markets, from pharmaceutical to beauty to food and beverage, Aptar combines market expertise with proprietary design, engineering and science to develop innovative solutions that help improve lives worldwide. Headquartered in Crystal Lake, Illinois, Aptar employs 14,000 dedicated people across 20 countries. Learn more at http://www.aptar.com.

This press release contains forward-looking statements, including statements regarding the anticipated benefits of the acquisition of PULMOTREE; the development, industrialization and commercialization of the Kolibri™ Non-Propellant Liquid Inhaler platform; expected growth opportunities in respiratory drug delivery; the integration of PULMOTREE’s technology with Aptar Pharma’s capabilities and services; and customer interest in and demand for the platform. Forward-looking statements generally can be identified by the fact that they do not relate strictly to historical or current facts and by use of words such as “expects,” “anticipates,” “believes,” “intends,” “aims,” “designed,” “potential,” “opportunities,” “accelerate,” “expand,” “create,” “future” and other similar expressions, or future or conditional verbs such as “will,” “can,” “should,” “would” and “could,” which are intended to identify such forward-looking statements. Forward-looking statements are made pursuant to the safe harbor provisions of Section 27A of the Securities Act of 1933 and Section 21E of the Securities Exchange Act of 1934 and are based on our beliefs as well as assumptions made by and information currently available to us. Accordingly, our actual results or other events may differ materially from those expressed or implied in such forward-looking statements due to known or unknown risks and uncertainties that exist in our operations and business environment including, but not limited to: our ability to successfully integrate PULMOTREE and to realize the anticipated benefits of the acquisition; the timing, cost and success of the development, industrialization, regulatory approval and commercialization of the Kolibri™ platform and of our customers’ products incorporating it; customer adoption of and demand for non-propellant inhalation technologies; our ability to retain PULMOTREE’s key personnel and customer relationships; our ability to protect and enforce intellectual property relating to the Kolibri™ platform; the regulatory environment, including regulation of drug-device combination products and connected-health features; and competition, including technological advances. For additional information on these and other risks and uncertainties, please see our filings with the Securities and Exchange Commission, including the discussion under “Risk Factors” and “Management’s Discussion and Analysis of Financial Condition and Results of Operations” in our Form 10-Ks and Form 10-Qs. We undertake no obligation to update publicly any forward-looking statements, whether as a result of new information, future events or otherwise, except as required by law.

Contacts

Media Contact
Ciara Jackson

Aptar Pharma

ciara.jackson@aptar.com

PolyPeptide announces results of the extraordinary general meeting of shareholders

PolyPeptide Group

/ Key word(s): AGMEGM

PolyPeptide announces results of the extraordinary general meeting of shareholders

26.10.2026 / 19:10 CET/CEST


Media Release

PolyPeptide announces results of the extraordinary general meeting of shareholders

Baar, 26 October 2026 – PolyPeptide Group AG (“PolyPeptide”) (SIX: PPGN), a specialized global CDMO for peptide-based active pharmaceutical ingredients, announced that the shareholders approved all proposals by its board of directors (the “Board”) at today’s extraordinary general meeting of shareholders.

In particular, the shareholders elected John Chongbo Rim, Seungho Ryu, Cheol Woo Ryu, Mihyun Yang and Joonkyung Seong as members of the Board for a term of office ending at the conclusion of the next annual general meeting. John Chongbo Rim was elected as chair of the Board, and Cheol Woo Ryu and Seungho Ryu were elected as members of the Remuneration and Nomination Committee, in each case for a term of office ending at the conclusion of the next annual general meeting. All elections are subject to and effective as of the settlement of the public tender offer by Samsung Peptide AG, a wholly owned subsidiary of Samsung Biologics Co., Ltd., for all publicly held registered shares of PolyPeptide.

The shareholders also resolved, subject to the satisfaction or waiver of all of the offer conditions set forth in the offer prospectus of Samsung Peptide AG dated 31 August 2026 (the “Offer Prospectus”), other than the offer condition set forth in section B.7.1(e) of the Offer Prospectus, to remove from the articles of association of PolyPeptide the transfer restrictions (Vinkulierung) and the voting limitations (Stimmrechtsbeschränkungen).


Contact

PolyPeptide Group AG   
Strategic Marketing and Corporate Communications
Laura Rosso    
mediateam@polypeptide.com
T: +41 43 502 0580

PolyPeptide Group AG 
Finance / Investor Relations
Tim Brandl
investorrelations@polypeptide.com 
T: +41 43 502 0580 
 

About PolyPeptide

PolyPeptide Group AG and its consolidated subsidiaries (“PolyPeptide”) is a specialized Contract Development & Manufacturing Organization (CDMO) for peptide-based active pharmaceutical ingredients. By supporting its customers mainly in pharma and biotech, it contributes to the health of millions of patients across the world. PolyPeptide serves a fast-growing market, offering products and services from pre-pre-clinical to commercial stages. Its broad portfolio reflects the opportunities in drug therapies across areas and with significant exposure to metabolic diseases, including GLP-1. Dating back to 1952, PolyPeptide today runs a global network of six GMP-certified facilities in Europe, the U.S. and India. PolyPeptide’s shares (SIX: PPGN) are listed on SIX Swiss Exchange. For more information, please visit polypeptide.com.

@PolyPeptide — follow us on LinkedIn

Forward-looking Information and Statements

This media release has been prepared by PolyPeptide Group AG and contains certain forward-looking statements that reflect the current views of management. Such statements are subject to known and unknown risks, uncertainties and other factors that may cause actual developments to differ materially from those expressed or implied in this release. PolyPeptide Group AG is providing the information in this release as of this date and, except as required by applicable laws or regulations, does not undertake any obligation to update any statements contained in it as a result of new information, future events or otherwise.


Additional features:

File: PolyPeptide_EGM Results 2026_ENG


End of Media Release


Language: English
Company: PolyPeptide Group
Neuhofstrasse 24
6340 Baar
Switzerland
Phone: +41435020580
E-mail: mediateam@polypeptide.com
Internet: www.polypeptide.com
ISIN: CH1110760852
Valor: 111076085
Listed: SIX Swiss Exchange
EQS News ID: 2422544

 
End of News EQS News Service

2422544  26.10.2026 CET/CEST

Viromed Medical AG: Successful Completion of Capital Increase

Viromed Medical AG

/ Key word(s): Capital Increase

Viromed Medical AG: Successful Completion of Capital Increase

26.10.2026 / 17:52 CET/CEST

The issuer is solely responsible for the content of this announcement.


Viromed Medical AG: Successful Completion of Capital Increase

Rellingen, October 26, 2026 – Viromed Medical AG (“Viromed”; ISIN: DE000A40ZVN7) has successfully completed the capital increase announced today. A total of 450,000 new registered no-par value shares were placed at a placement price of EUR 4.50 per share. The gross proceeds amount to approximately EUR 2 million and will be used to finance the company’s working capital.

Subject to the registration of the implementation of the capital increase in the commercial register, the share capital will increase from EUR 21,250,000 by EUR 450,000 to EUR 21,700,000. The capital increase was carried out by partially utilizing the Authorized Capital 2025 and with the exclusion of the shareholders’ subscription rights.


About Viromed Medical AG

Viromed Medical AG specializes in the development, manufacture and distribution of medical products. The operating business of the company, which has been listed on the stock exchange since October 2022, focuses on the distribution of innovative cold plasma technology for medical applications via its wholly owned subsidiary Viromed Medical GmbH. Viromed can draw on a broad customer base in the DACH region and beyond. Viromed is pursuing the goal of further advancing the use of cold plasma technology in medicine in the coming years and realizing the corresponding growth potential.

www.viromed-medical-ag.de

Contact Viromed

E-Mail: kontakt@viromed-medical.de
 

Press contact

E-mail: viromed@kirchhoff.de


26.10.2026 CET/CEST Dissemination of a Corporate News, transmitted by EQS News – a service of EQS Group.
The issuer is solely responsible for the content of this announcement.

The EQS Distribution Services include Regulatory Announcements, Financial/Corporate News and Press Releases.


Language: English
Company: Viromed Medical AG
Hauptstraße 105
25462 Rellingen
Germany
E-mail: kontakt@viromed-medical.de
Internet: https://www.viromed-medical-ag.de/
ISIN: DE000A40ZVN7
WKN: A40ZVN
Listed: Regulated Unofficial Market in Dusseldorf, Frankfurt, Hamburg, Tradegate BSX
LEI Code: 894500XR05MYCVCCR171
EQS News ID: 2422458

 
End of News EQS News Service

2422458  26.10.2026 CET/CEST