Fresenius Medical Care AG : Change in CEO – Shervin J. Korangy Succeeds Helen Giza

Fresenius Medical Care AG / Key word(s): Personnel decisions / Executive Board

Fresenius Medical Care AG : Change in CEO – Shervin J. Korangy Succeeds Helen Giza

07-Oct-2026 / 16:54 CET/CEST

Disclosure of an inside information acc. to Article 17 MAR of the Regulation (EU) No 596/2014, transmitted by EQS News – a service of EQS Group.

The issuer is solely responsible for the content of this announcement.


Bad Homburg, (October 7, 2026) – The Supervisory Board of Fresenius Medical Care AG unanimously resolved today to appoint Shervin J. Korangy as Chairman of the Management Board effective October 12, 2026. Shervin J. Korangy has resigned from his position as a member of the Company’s Supervisory Board. Helen Giza, the long-serving Chairwoman of the Management Board, will leave the Company with effect from the end of October 11, 2026. The Supervisory Board thanks Helen Giza for her service to the Company.

Contact:
Dr. Dominik K. Heger
Executive Vice President
Global Head of Investor Relations, Market & Competition, Sustainability
& Head of Investor Relations

Fresenius Medical Care AG
Else-Kroener-Strasse 1, 61352 Bad Homburg v.d. Hoehe, Germany
P +49 6172 2685822
Dominik.Heger@FreseniusMedicalCare.com
www.FreseniusMedicalCare.com

End of Inside Information


07-Oct-2026 CET/CEST The EQS Distribution Services include Regulatory Announcements, Financial/Corporate News and Press Releases.


Language: English
Company: Fresenius Medical Care AG
Else-Kröner-Straße 1
61352 Bad Homburg
Germany
Phone: +49 (0) 6172- 609 2525
Fax: +49 (0) 6172- 609 2301
E-mail: ir@freseniusmedicalcare.com
Internet: www.freseniusmedicalcare.com
ISIN: DE0005785802
WKN: 578580
Indices: DAX
Listed: Regulated Market in Frankfurt (Prime Standard); Regulated Unofficial Market in Dusseldorf, Hamburg, Hanover, Munich, Stuttgart, Tradegate BSX; NYSE, Luxembourg Stock Exchange
LEI Code: 549300CP8NY40UP89Q40
EQS News ID: 2412002

 
End of Announcement EQS News Service

2412002  07-Oct-2026 CET/CEST

Asahi Kasei Therapeutics Announces FDA Approval of HOVILPRI® (pritelivir) Tablets for Mucocutaneous HSV Lesions Refractory (With or Without Resistance) to Standard Antiviral Treatment in Immunocompromised Adults

Asahi Kasei Therapeutics Announces FDA Approval of HOVILPRI® (pritelivir) Tablets for Mucocutaneous HSV Lesions Refractory (With or Without Resistance) to Standard Antiviral Treatment in Immunocompromised Adults


Asahi Kasei Therapeutics Announces FDA Approval of HOVILPRI® (pritelivir) Tablets for Mucocutaneous HSV Lesions Refractory (With or Without Resistance) to Standard Antiviral Treatment in Immunocompromised Adults

  • HOVILPRI® (pritelivir), a novel oral helicase-primase inhibitor represents the first FDA approved HSV therapy with a novel mechanism of action in nearly 30 years.1-3
  • Approval is supported by results from the pivotal Phase 3 PRIOH-1 trial, in which HOVILPRI demonstrated statistically superior complete lesion healing up to day 28. By Day 28, 63% of HOVILPRI-treated participants achieved complete lesion healing compared with 34% of participants receiving Investigator’s Choice treatment, corresponding to an adjusted treatment difference of 28.4% (95% CI: 9.6, 47.3; p=0.0047).1,4
  • As an oral therapy, HOVILPRI provides a non-intravenous treatment option for eligible immunocompromised adult patients with refractory (with or without resistance) HSV infection.1
  • Pritelivir was granted Fast Track designation by the FDA along with Breakthrough Therapy designation and the New Drug Application received Priority Review.5,6

TOKYO–(BUSINESS WIRE)–Asahi Kasei Therapeutics, Inc., the global specialty pharmaceutical business of Tokyo-based Asahi Kasei, with U.S. and European business headquarters in Cary, North Carolina, today announced that the U.S. Food and Drug Administration (FDA) has approved HOVILPRI® (pritelivir) 100 mg tablets for the treatment of mucocutaneous lesions caused by herpes simplex virus (HSV) infection that is refractory (with or without documented resistance) to treatment with acyclovir, valacyclovir, or famciclovir in immunocompromised adults. HOVILPRI, a novel helicase-primase inhibitor, was developed by AiCuris Anti-infective Cures AG, which is now a part of Asahi Kasei Therapeutics.1,2


“Today’s approval delivers the first FDA-approved treatment for refractory mucocutaneous HSV lesions with a novel mechanism of action in nearly 30 years and marks an important advancement for immunocompromised adults,” said Stacy Wheeler, Chief Executive Officer of Asahi Kasei Therapeutics in the U.S. and Europe. “HOVILPRI offers a new oral treatment option for adult patients with a persistent, difficult-to-treat disease.”1,3,7-10

In immunocompetent adults, HSV infection is often asymptomatic or associated with mild disease. In immunocompromised adults, including hematopoietic stem cell and solid organ transplant recipients, people living with HIV, and patients with malignancies or autoimmune and inflammatory conditions, HSV infection can occur more frequently and may be more severe. Refractory HSV infection can result in prolonged and difficult-to-treat mucocutaneous disease.7-10 Because nucleoside analogue therapies such as acyclovir require activation by the viral thymidine kinase (TK) enzyme, mutations in TK can reduce susceptibility to these treatments in some patients.2,9 When HSV infection does not improve with nucleoside analogue therapy, available treatment options have often required intravenous administration and may present additional treatment-management challenges for patients and healthcare teams.8-10 HOVILPRI is a helicase-primase inhibitor with a mechanism of action distinct from nucleoside analogue therapies.1,2

The approval is supported by results from PRIOH-1 Part C, the pivotal Phase 3 randomized, open-label, comparator-controlled superiority trial in immunocompromised adults with mucocutaneous HSV infection that was refractory, with or without documented resistance, to acyclovir, valacyclovir, or famciclovir. The trial met its primary efficacy endpoint, demonstrating superiority of HOVILPRI over Investigator’s Choice treatment, after up to 28 days of treatment. By Day 28, HOVILPRI achieved a significantly higher rate of complete lesion healing vs Investigator’s Choice (63% vs 34% respectively, corresponding to an adjusted treatment difference of 28.4% (95% CI: 9.6, 47.3; p=0.0047). Up to day 42, the observed response rates were 82% for HOVILPRI and 42% for Investigator’s Choice treatment, corresponding to an adjusted treatment difference of 40.0% (95% CI: 22.7, 57.7).1,4

In Part C of the pivotal Phase 3 PRIOH-1 trial, adverse reactions were reported less frequently in participants treated with HOVILPRI compared with Investigator’s Choice treatment (22% vs. 54%). Discontinuations due to adverse drug reactions were also less frequent with HOVILPRI compared with Investigator’s Choice treatment (2% vs. 20%). The most common adverse reaction was headache, occurring in 6% of patients taking HOVILPRI and 4% of patients taking Investigators Choice.1

“In patients with blood cancers or those who have undergone stem cell transplantation, refractory HSV infections can be prolonged, painful, and difficult to treat, while available intravenous treatments carry substantial risks,” said Genovefa Papanicolaou, MD, Clinical Director of Infectious Disease Service at Memorial Sloan Kettering Cancer Center and Professor at Weill Cornell College of Medicine, and a PRIOH-1 investigator. “In PRIOH-1, pritelivir demonstrated superior lesion healing compared with Investigator’s Choice treatment up to day 28.”1,4,7-10

“Refractory HSV can be a serious clinical challenge for immunocompromised adults, a population with limited treatment options,” said Carl Kraus, Chief Medical Officer at Asahi Kasei Therapeutics. “The approval of HOVILPRI reflects years of clinical research in refractory HSV and reinforces our commitment to advancing scientifically differentiated medicines for patients whose conditions remain difficult to treat. We are grateful to the patients, investigators, clinical teams, and scientific community whose contributions made this milestone possible.”1,4,7-10

HOVILPRI is expected to be available in the United States before the end of 2026. U.S. Healthcare providers can learn more about HOVILPRI at www.hovilprihcp.com.

About the PRIOH-1 Trial

PRIOH-1 (NCT03073967 / Eudra-CT 2023-510088-37-00) is a global, controlled, open-label, comparative trial evaluating pritelivir in immunocompromised adults with refractory or resistant HSV infection. The registrational Phase 3 portion (Part C) randomized and treated 101 immunocompromised adults with mucocutaneous HSV infection that was refractory with or without documented resistance to acyclovir, valacyclovir, or famciclovir. This included hematopoietic cell and solid organ transplant recipients, patients with malignancies, autoimmune or inflammatory disorders, and people living with HIV. Participants received either oral pritelivir (400 mg loading dose on Day 1 followed by 100 mg once daily) or Investigator’s Choice treatment (IV foscarnet, IV/topical cidofovir, or 5% topical imiquimod) for up to 28 days, with treatment extended to 42 days if lesions were improving. Pritelivir met its primary endpoint demonstrating statistically significant superiority in lesion healing up to Day 28 compared with Investigator’s Choice treatment.1,4

About Herpes Simplex Virus

Herpes Simplex Virus (HSV) includes two types, HSV-1 and HSV-2, both of which cause lifelong infections; there is no cure. HSV-1 most commonly causes labial herpes, while HSV-2 is the primary cause of genital herpes. Although many people with HSV infection never develop symptoms, the virus can reactivate to cause recurrent, painful lesions and may cause complications in certain cases.11-12 HSV represents a substantial global public health burden: an estimated 3.8 billion people under the age of 50 (64% of the global population) were infected with HSV-1 in 2020, and approximately 520 million people aged 15 to 49 were living with HSV-2.11 The disease burden is particularly high in immunocompromised patients, whose weakened immune systems leave them at increased risk of more frequent, more severe, and treatment-refractory HSV infections.7,9-10

About HOVILPRI (pritelivir)

HOVILPRI is an orally administered HSV helicase-primase inhibitor indicated for the treatment of mucocutaneous lesions caused by HSV infection that is refractory, with or without documented resistance, to acyclovir, valacyclovir, or famciclovir in immunocompromised adults. HOVILPRI is active against both HSV-1 and HSV-2, including strains resistant to nucleoside analogs and foscarnet. In the pivotal Phase 3 PRIOH-1 trial, HOVILPRI demonstrated statistically significant superiority in lesion healing up to day 28 compared to Investigator’s Choice treatment. The most common adverse reaction reported with HOVILPRI was headache.1-2,4

INDICATION AND IMPORTANT SAFETY INFORMATION FOR US AUDIENCE

Indication

HOVILPRI (pritelivir) tablets is indicated in immunocompromised adults for the treatment of mucocutaneous lesions caused by herpes simplex virus (HSV) infection that is refractory (with or without documented resistance) to treatment with acyclovir, valacyclovir, or famciclovir.

Important Safety Information

Warnings and Precautions

Drug Interactions With Acid-reducing Agents: Concomitant use of HOVILPRI with certain acid-reducing agents may reduce pritelivir exposure, potentially resulting in reduced therapeutic effect and development of resistance. Avoid concomitant use with certain proton pump inhibitors, and stagger concomitant use with H2-receptor antagonists and antacids.

Adverse Reactions: In clinical studies, the adverse reaction (all grades) occurring in more than 5% of participants receiving HOVILPRI was headache, which occurred in 6% and 4% of the HOVILPRI and Investigator’s Choice participants, respectively.

Drug Interactions:

Pritelivir is a breast cancer resistance protein (BCRP) inhibitor. Co-administration of HOVILPRI with a BCRP substrate may increase the plasma concentrations of the BCRP substrate, which may increase the risk of adverse reactions.

Concomitant use of acid-reducing agents may elevate gastric pH and decrease the systemic bioavailability of HOVILPRI, which may reduce its effectiveness or cause drug resistance to HOVILPRI. Avoid concomitant use with esomeprazole or rabeprazole. Avoid concomitant use with omeprazole or lansoprazole at daily doses above 20 mg, or with pantoprazole at daily doses above 40 mg. Take HOVILPRI at least 2 hours before or 12 hours after H2-receptor antagonists. Take HOVILPRI at least 2 hours before or 2 hours after antacids.

Please see full Prescribing Information at https://www.hovilprihcp.com/pdf/prescribing-information.pdf.

About Asahi Kasei Therapeutics

Asahi Kasei Therapeutics is the global specialty pharmaceutical business of Tokyo-based Asahi Kasei. We are identifying, developing and commercializing innovative medicines for diseases with significant unmet medical needs. Drawing upon our strong heritage of scientific innovation, we are expanding therapeutic options and helping to positively impact health in immunology, nephrology, transplantation, and infectious diseases. For more information, visit: https://www.aktx.com.

About Asahi Kasei

Asahi Kasei is a diversified global company that contributes to life and living for people around the world. Since its foundation in 1922, with businesses in ammonia and cellulose fiber, Asahi Kasei has consistently grown through proactive portfolio transformation to meet the evolving needs of every age. With 50,000 employees worldwide, the company contributes to sustainability by providing solutions to the world’s challenges across its three business sectors: Healthcare, Homes, and Material. For more information, visit https://www.asahi-kasei.com/.

Asahi Kasei positions Pharmaceuticals as a First Priority business in its medium-term management plan, emphasizing value creation by expanding its research and development activities globally to deliver innovative therapeutic options worldwide.

References:

1. HOVILPRI (pritelivir) tablets. Prescribing information. Asahi Kasei Therapeutics, Inc.; 2026.

2. Birkmann A et al. J Med Chem. 2022;65(20):13614-13628. doi:10.1021/acs.jmedchem.2c00668

3. Birkmann A, Saunders R. Antiviral Res. 2025;237:106152. doi:10.1016/j.antiviral.2025.106152

4. Papanicolaou GA et al. Oral late-breaker presented at: Tandem Meetings; February 4-7, 2026; Salt Lake City, Utah.

5. AiCuris Anti-infective Cures AG. FDA Breakthrough Therapy designation and Priority Review correspondence. Data on file.

6. AiCuris Anti-infective Cures AG. Aicuris Receives FDA Priority Review for Pritelivir NDA and Presents New Phase 3 Data at ESCMID 2026. Published April 16, 2026.

7. Shafat T et al. Clin Microbiol Infect. 2025;31(5):761-772. doi:10.1016/j.cmi.2025.01.033

8. Hammond SP et al. Open Forum Infect Dis. 2024;11(3):ofae046. doi:10.1093/ofid/ofae046

9. Sallée L, Boutolleau D. Rev Med Virol. 2024;34(5):e2574. doi:10.1002/rmv.2574

10. Chemaly RF et al. Clin Infect Dis. 2025;81(3):593-601. doi:10.1093/cid/ciae638

11. Harfouche M et al. Sex Transm Infect. 2025;101(4):214-222. doi:10.1136/sextrans-2024-056307

12. Workowski KA et al. MMWR Recomm Rep. 2021;70(4):1-187. doi:10.15585/mmwr.rr7004a1

US-PRI-2600128

Contacts

Asahi Kasei Corporation
Christian Okeefe

christian.okeefe@ak-america.com

Asahi Kasei Therapeutics
Anna Michaels

Director, Corporate Communications

media@aktx.com

ME Therapeutics Extends Option Agreement for CD22 Nanobody Asset

ME Therapeutics Extends Option Agreement for CD22 Nanobody Asset


ME Therapeutics Extends Option Agreement for CD22 Nanobody Asset

VANCOUVER, British Columbia–(BUSINESS WIRE)–ME Therapeutics Holdings Inc. (“ME Therapeutics” or the “Company”) (CSE: METX) (FSE: Q9T), a publicly listed biotechnology company working on novel cancer-fighting drugs in the field of immuno-oncology, today announced an extension to its option agreement to license the National Research Council of Canada’s (NRC) proprietary nanobody-based CD22 binder for use in Chimeric Antigen Receptor (CAR) therapy. ME Therapeutics will continue developing the CD22 nanobody as part of its in vivo CAR program. The option to license the CD22 binder is now exercisable until February 10, 2027. All other terms remain the same.


About ME Therapeutics

ME Therapeutics is a publicly listed biotechnology company based in Vancouver focused on developing novel therapeutics designed to reprogramme immune cells in vivo to reshape the tumour microenvironment and to directly recognize and kill cancer cells. For more information, visit www.metherapeutics.com

Neither the Canadian Securities Exchange nor any Market Regulator (as that term is defined in the policies of the Canadian Securities Exchange) accepts responsibility for the adequacy or accuracy of this release.

Cautionary Statement Regarding Forward-Looking Statements

This news release includes certain “forward-looking statements” under applicable Canadian securities legislation. Forward-looking statements consist of statements that are not purely historical, including any statements regarding beliefs, plans, expectations or intentions regarding the future. Such forward-looking statements in this news release include, but are not limited to, statements regarding the Company’s patent protection, research plans, the intended outcomes of the research, the intended benefits and applications of the Company’s technology, the Company’s plans for development of its business, plans for potential first-in-human clinical trials, opportunities to explore earlier stage drug discovery and to enhance the Company’s drug pipeline, its reliance on third-party collaborators, regarding potential in-licensing and partnership opportunities, the Company’s plan for pursuing a U.S. listing and the projected benefits to shareholders, and as to the timing for the various plans set out in this news release. Such statements are subject to risks and uncertainties that may cause actual results, performance or developments to differ materially from those contained in the statements, including risks related to factors beyond the control of the Company, that the results of the testing are not favorable or consistent with results to date, that the Company’s mRNA or in vivo CAR candidates prove ineffective during testing, that the Company’s testing strategy does not yield the anticipated benefits, that the Company’s business may not develop as set out in this news release, that the Company does not proceed with human clinical trials or that the results of such trials, if any, are not favorable, that the results of testing are not received in a timely manner, that the Company does not acquire the necessary regulatory approvals, that the Company does not complete any licensing or partnership agreements or that if such agreements are completed that the terms may not be favorable to the Company, that the Company does not have sufficient funds to advance its business plan, that the research is not completed within the projected timelines, that a U.S. listing is not obtained, and such other risks described in the Company’s public disclosure and risks which are inherent to businesses of this nature. No assurance can be given that any of the events anticipated by the forward-looking statements will occur or, if they do occur, what benefits the Company will obtain from them. There can be no assurance that such statements will prove to be accurate, as actual results and future events could differ from forward-looking statements. The Company disclaims any intention or obligation to update or revise any forward-looking statements, whether as a result of new information, future events or otherwise, except as required by law.

Contacts

Company: Salim Dhanji, PhD, salim@metherapeutics.com, +1-236-516-7714

Media: Claire Piech, claire@magneticcomms.com, +1-604-698-6637

INB.bio Launches 2026 Growth Strategy With New Market Entries Across Africa, MENA and Southeast Asia

INB.bio Launches 2026 Growth Strategy With New Market Entries Across Africa, MENA and Southeast Asia


INB.bio Launches 2026 Growth Strategy With New Market Entries Across Africa, MENA and Southeast Asia

The direct advertiser and wellness infrastructure company targets five new countries as global wellness demand accelerates in emerging markets

INB.bio Launches 2026 Growth Strategy With New Market Entries Across Africa, MENA and Southeast Asia

BERLIN, March 22, 2026 (GLOBE NEWSWIRE) — Global demand for wellness products is climbing steadily, with the industry reaching a record $6.8 trillion and projected to hit $9.8 trillion by 2029, and a growing share of that demand is concentrated in emerging markets that global brands have historically underserved. INB.bio, a direct advertiser in dietary and wellness supplements operating across 15 countries in Africa, MENA, and Latin America, has launched its 2026 growth strategy. Plans include a market entry into Tanzania, expansion into three to five new countries across Africa and Southeast Asia, and the rollout of a rebuilt AI-powered affiliate CRM platform designed to support rapid multi-market operations.

The company, which manufactures and distributes an exclusive portfolio of wellness products covering men’s health, joint support, weight management, immunity, and cardiovascular health, enters 2026 having built one of the most operationally complete direct advertiser models in its category. Its infrastructure in each active market includes native call centers, owned logistics networks, and a proprietary technology platform built for cash-on-delivery economies where digital payment access is limited.

“Tanzania is the opening move of 2026,” said Rozhden Totskoinov, Founder of INB.bio. “Africa and Southeast Asia represent the largest untapped opportunity in wellness right now, and we are entering these markets the same way we have always entered them. With people on the ground, owned logistics and technology built for the reality of how these markets actually operate.“

Africa’s middle class is projected to reach 1.1 billion people by 2060, according to the African Development Bank, a demographic shift already driving increased consumer spending on health and lifestyle products across the continent. The global wellness economy recorded $6.8 trillion in 2024 and is on track to reach $9.8 trillion by 2029, per the Global Wellness Institute, with Sub-Saharan Africa and Southeast Asia among the regions where growth is accelerating most notably. INB.bio has positioned its entire operating model around exactly those market characteristics, prioritising regions with COD payment dominance, mobile-first populations, and limited existing competition from established global players.

The company is simultaneously launching a turnkey partner program through which high-performing affiliate partners can co-develop new country operations alongside INB.bio. Participants receive exclusive geographic access, native call center resources, owned logistics support, and live performance data through an upgraded affiliate dashboard. Payouts are processed twice weekly, on Tuesdays and Fridays, with a minimum threshold of fifty dollars in USDT TRC-20, and additional payment options, including wire transfer and PayPal, are in development.

“We are proving that the model we built over six years is repeatable at scale,” said Rozhden. “Tier-2 and tier-3 markets are not too difficult for serious operators. They are simply too difficult for operators unwilling to do the actual work, and 2026 is where we demonstrate what that means at full speed.”

Visit the company’s website for more information.

About INB.bio
INB.bio is a direct advertiser in dietary and wellness supplements operating across 15 countries in Africa, MENA and Latin America. The company manufactures and distributes an exclusive portfolio of wellness products supported by owned infrastructure in every active market, including native call centers, proprietary logistics networks and a custom-built technology platform designed for cash-on-delivery economies. Founded six years ago, INB.bio serves millions of end customers and a global network of affiliate partners through a twice-weekly payout model. The company’s 2026 expansion targets new geographic markets across Africa and Southeast Asia as part of a long-term strategy to build wellness infrastructure across 30 emerging markets worldwide. For more information, visit INB.bio.

Contact:
Paulina Dvor, Senior Content & PR Manager
INB.bio
p.dvor@inb.bio
marketing@inb.bio

A photo accompanying this announcement is available at https://www.globenewswire.com/NewsRoom/AttachmentNg/964e46f5-e16e-4577-9b87-f6d9a64eb290

HUTCHMED Initiates Phase III Trial of HMPL-760 in Patients with Relapsed/Refractory Diffuse Large B-cell Lymphoma in China

HUTCHMED Initiates Phase III Trial of HMPL-760 in Patients with Relapsed/Refractory Diffuse Large B-cell Lymphoma in China


HUTCHMED Initiates Phase III Trial of HMPL-760 in Patients with Relapsed/Refractory Diffuse Large B-cell Lymphoma in China

HONG KONG and SHANGHAI and FLORHAM PARK, N.J., March 23, 2026 (GLOBE NEWSWIRE) — HUTCHMED (China) Limited (“HUTCHMED”) (Nasdaq/AIM:​HCM; HKEX:​13) today announces that it has initiated a registrational Phase III clinical trial of HMPL-760 in combination with R-GemOx (rituximab, gemcitabine and oxaliplatin) in patients with relapsed/refractory diffuse large B-cell lymphoma (“DLBCL”) in China. The first patient received the first dose on March 20, 2026.

DLBCL is the most common form of aggressive non-Hodgkin lymphoma (“NHL”) worldwide, accounting for approximately 40% of all NHL cases in China.1  In 2022, approximately 81,000 new cases of NHL are estimated to have been diagnosed in China.2 Bruton’s tyrosine kinase (“BTK”) is considered a validated target for drugs that aim to treat certain hematological cancers. HMPL-760 is a highly potent, selective, and reversible inhibitor with long target engagement against BTK, including wild-type and C481S-mutated BTK.

The trial is a randomized, double-blind, positive controlled Phase III study to evaluate the efficacy, safety, and pharmacokinetics (“PK”) of HMPL-760 in combination with R-GemOx versus placebo in combination with R-GemOx in DLBCL patients who are relapsed or refractory after prior treatment with first-line systemic chemotherapy, immunotherapy, or immunochemotherapy regimens and ineligible for transplantation. Primary outcome measures include investigator-assessed progression-free survival (“PFS”) and overall survival (“OS”). Secondary outcome measures include independent review committee (“IRC”)-assessed PFS, IRC- and investigator-assessed objective response rate (“ORR”), complete response rate (“CRR”), duration of response (DoR), clinical benefit rate (CBR), time to response (TTR), safety and PK characteristics. Additional details may be found at clinicaltrials.gov, using identifier NCT07409428.

This registrational trial plans to enroll approximately 240 patients and is being led by principal investigator Professor Weili Zhao, Vice President of Ruijin Hospital Affiliated to Shanghai Jiao Tong University School of Medicine and Director of the Shanghai Institute of Hematology.

About HMPL-760

HMPL-760 is an investigational, non-covalent, third generation BTK inhibitor. It is a highly potent, selective, and reversible inhibitor with long target engagement against BTK, including wild-type and C481S-mutated BTK. BTK C481S mutation plays an important role in resistance to certain BTK inhibitors.3,4

A randomized, double-blind Phase II study (NCT06601504) evaluating HMPL-760 in combination with R-GemOx in patients with relapsed/refractory DLBCL has demonstrated encouraging improvements in ORR, CRR, PFS and OS compared to R-GemOx alone, with a manageable safety profile and no unexpected safety signal. These encouraging results supported the initiation of this registrational Phase III trial.

HUTCHMED currently retains all rights to HMPL-760 worldwide.

About HUTCHMED

HUTCHMED (Nasdaq/AIM:​HCM; HKEX:​13) is an innovative, commercial-stage, biopharmaceutical company. It is committed to the discovery and global development and commercialization of targeted therapies and immunotherapies for the treatment of cancer and immunological diseases. Since inception it has focused on bringing drug candidates from in-house discovery to patients around the world, with its first three medicines marketed in China, the first of which is also approved around the world including in the US, Europe and Japan. For more information, please visit: www.hutch-med.com or follow us on LinkedIn.

Forward-Looking Statements

This press release contains forward-looking statements within the meaning of the “safe harbor” provisions of the US Private Securities Litigation Reform Act of 1995. These forward-looking statements reflect HUTCHMED’s current expectations regarding future events, including its expectations regarding the therapeutic potential of HMPL-760 for the treatment of DLBCL and the further development of HMPL-760 in this and other indications. Forward-looking statements involve risks and uncertainties. Such risks and uncertainties include, among other things, assumptions regarding the timing and outcome of clinical studies and the sufficiency of clinical data to support a new drug application submission of HMPL-760 for the treatment of DLBCL or other indications in China or other jurisdictions, its potential to gain approvals from regulatory authorities on an expedited basis or at all, the efficacy and safety profile of HMPL-760, HUTCHMED’s ability to fund, implement and complete its further clinical development and commercialization plans for HMPL-760 and the timing of these events. In addition, as certain studies rely on the use of other drug products such as R-GemOx as combination therapeutics with HMPL-760, such risks and uncertainties include assumptions regarding the safety, efficacy, supply and continued regulatory approval of these therapeutics.  Existing and prospective investors are cautioned not to place undue reliance on these forward-looking statements, which speak only as of the date hereof. For further discussion of these and other risks, see HUTCHMED’s filings with the US Securities and Exchange Commission, The Stock Exchange of Hong Kong Limited and on AIM. HUTCHMED undertakes no obligation to update or revise the information contained in this press release, whether as a result of new information, future events or circumstances or otherwise.

CONTACTS

Investor Enquiries +852 2121 8200 / ir@hutch-med.com
   
Media Enquiries  
FTI Consulting – +44 20 3727 1030 / HUTCHMED@fticonsulting.com
  Ben Atwell / Tim Stamper +44 7771 913 902 (Mobile) / +44 7779 436 689 (Mobile)
Brunswick – Zhou Yi +852 9783 6894 (Mobile) / HUTCHMED@brunswickgroup.com
   
Panmure Liberum Nominated Advisor and Joint Broker
Atholl Tweedie / Emma Earl / Rupert Dearden +44 20 7886 2500
   
Cavendish Joint Broker
Geoff Nash / Nigel Birks +44 20 7220 0500
   
Deutsche Numis Joint Broker
Freddie Barnfield / Jeffrey Wong / Duncan Monteith +44 20 7260 1000

_____________________
1 Li XQ, Li GD, Gao ZF, et al. Distribution pattern of lymphoma subtypes in China: A nationwide multicenter study of 10002 cases. J Diagn Concepts Pract. 2012; 11(02):111-115.
2 The Global Cancer Observatory, China fact sheet. https://gco.iarc.who.int/media/globocan/factsheets/populations/160-china-fact-sheet.pdf. Accessed December 3, 2025.
3 Woyach JA, Ruppert AS, Guinn D, et al. BTKC481S-Mediated Resistance to Ibrutinib in Chronic Lymphocytic Leukemia. J Clin Oncol. 2017;35(13):1437-1443. doi:10.1200/JCO.2016.70.2282.
4 Woyach JA, Huang Y, Rogers K, et al.  Resistance to Acalabrutinib in CLL is Mediated Primarily by BTK Mutations. Blood.  2019;134 (Supplement_1): 504.  doi:10.1182/blood-2019-127674.

Apogee Therapeutics to Host Conference Call to Report Part A 52-Week Data from the Phase 2 APEX Trial of Zumilokibart (APG777) in Patients with Moderate-to-Severe Atopic Dermatitis on March 23, 2026

Apogee Therapeutics to Host Conference Call to Report Part A 52-Week Data from the Phase 2 APEX Trial of Zumilokibart (APG777) in Patients with Moderate-to-Severe Atopic Dermatitis on March 23, 2026


Apogee Therapeutics to Host Conference Call to Report Part A 52-Week Data from the Phase 2 APEX Trial of Zumilokibart (APG777) in Patients with Moderate-to-Severe Atopic Dermatitis on March 23, 2026

Webcast to be held Monday, March 23rd at 8:00 a.m. ET

SAN FRANCISCO and BOSTON, March 22, 2026 (GLOBE NEWSWIRE) — Apogee Therapeutics, Inc. (Nasdaq: APGE), a clinical-stage biotechnology company advancing optimized, novel biologics with potential for best-in-class profiles in the largest inflammatory and immunology (I&I) markets, today announced it will report Part A 52-week data from the Phase 2 APEX trial of zumilokibart on Monday, March 23, 2026. Following the announcement, the company will host a conference call and webcast at 8:00 a.m. ET to discuss the results.

Webcast Details
Apogee Therapeutics’ live webcast of the Phase 2 APEX Part A results will begin on Monday, March 23rd at 8:00 a.m. ET. The live webcast can be accessed via this link or the Investors section on the Company’s website at https://investors.apogeetherapeutics.com/news-events/events. A replay of the webcast will be available following the call.

About Apogee
Apogee Therapeutics is a clinical-stage biotechnology company advancing novel biologics with potential for differentiated efficacy and dosing in the largest I&I markets, including for the treatment of AD, asthma, EoE, Chronic Obstructive Pulmonary Disease (COPD) and other I&I indications. Apogee’s antibody programs are designed to overcome limitations of existing therapies by targeting well-established mechanisms of action and incorporating advanced antibody engineering to optimize half-life and other properties. Zumilokibart, the company’s most advanced program, is being initially developed for the treatment of AD, which is the largest and one of the least penetrated I&I markets, as well as asthma and EoE. With four validated targets in its portfolio, Apogee is seeking to achieve best-in-class efficacy and dosing through monotherapies and combinations of its novel antibodies. Based on a broad pipeline and depth of expertise, the company believes it can deliver value and meaningful benefit to patients underserved by today’s standard of care. For more information, please visit https://apogeetherapeutics.com.

Investor Contact:
Noel Kurdi
VP, Investor Relations
Apogee Therapeutics, Inc.
Noel.Kurdi@apogeetherapeutics.com

Media Contact:
Dan Budwick
1AB Media
dan@1abmedia.com

Akemi Hair Glow Claims Evaluated: 2026 Hair Growth Transparency Report Examines DHT-Blocking Spray Research, “56% Thicker Hair” Data, and Ingredient Research for Women

Akemi Hair Glow Claims Evaluated: 2026 Hair Growth Transparency Report Examines DHT-Blocking Spray Research, “56% Thicker Hair” Data, and Ingredient Research for Women


Akemi Hair Glow Claims Evaluated: 2026 Hair Growth Transparency Report Examines DHT-Blocking Spray Research, “56% Thicker Hair” Data, and Ingredient Research for Women

A 2026 informational report presenting Akemi Hair Glow’s DHT-blocking claims alongside ingredient-level research context, topical delivery positioning, pricing structure, and key verification factors for consumers evaluating hair growth products

Elizabeth, NJ, March 21, 2026 (GLOBE NEWSWIRE) — This article contains affiliate links. If a purchase is made through these links, a commission may be earned at no additional cost to the buyer. This report is an informational overview and does not constitute medical, health, or dermatological advice. All product details described below are stated as presented by the company and should be verified directly on the official website before any purchasing decision.

In this report, the term “effectiveness” refers strictly to how the product’s marketing language describes potential outcomes. It does not indicate that the finished product has been clinically proven effective, and no published clinical trial appears to evaluate Akemi Hair Glow as a proprietary formula.

As consumer interest in topical hair growth solutions continues to grow, Akemi Hair Glow has become one of the products generating attention across social media platforms and wellness communities. The product is marketed primarily toward women experiencing thinning hair, increased shedding, or visible scalp areas that have become harder to conceal over time.

The product’s claims around DHT blocking, follicle reactivation, and “56% thicker hair in 18 weeks” have contributed to that visibility. These types of claims are common in the topical hair growth category and are presented in this report alongside published ingredient-level research for contextual reference.
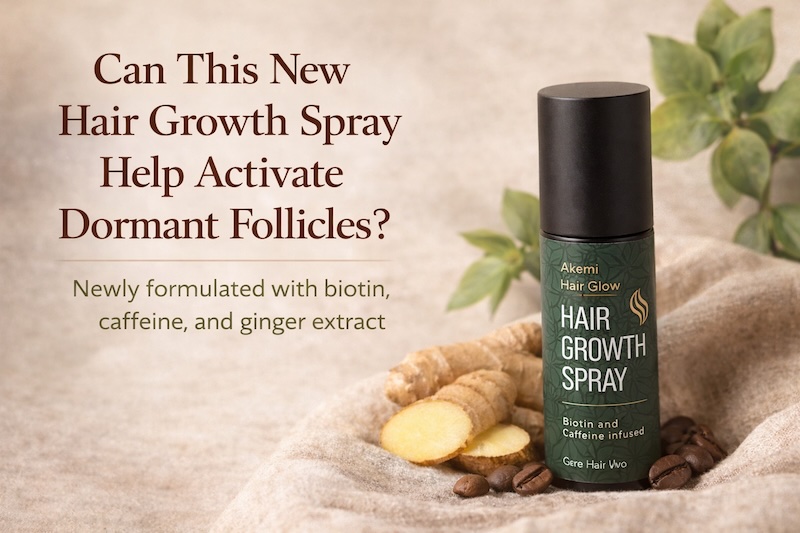
Akemi Hair Glow Claims Evaluated 2026 Hair Growth Transparency Report Examines DHT-Blocking Spray Research, "56% Thicker Hair" Data, and Ingredient Research for Women

This report presents how the product is described alongside relevant research considerations. It outlines what the product page states, what published ingredient-level research supports under controlled study conditions, and where gaps remain between the marketing language and clinical evidence for the finished product.

Current product details, pricing, and terms can be confirmed by viewing the current Akemi Hair Glow offer (official Akemi Hair Glow page).

Individual results vary. Topical hair products are not substitutes for professional dermatological evaluation, balanced nutrition, or medical treatment for diagnosed hair loss conditions. Consult a qualified healthcare provider before starting any new topical hair regimen, especially if you are experiencing sudden or severe hair loss.

What Is Akemi Hair Glow

Akemi Hair Glow is a topical hair spray positioned as a DHT-blocking formula designed to support hair growth, reduce shedding, and improve hair thickness and texture. The product is sold exclusively through its official website, which operates on a Shopify-based platform under the domain buyskyline.co. The product page describes it as a leave-in daily treatment that delivers active ingredients directly to the scalp rather than through oral ingestion.

The product page lists six primary active ingredients: Biotin (Vitamin B7), Aminexil, Castor Oil, Caffeine, He Shou Wu (Fo-Ti), and Ginger Root Extract. The formula is described as free from parabens and toxins, with natural preservatives. The specific concentrations of each ingredient per application are not disclosed on the product page or in publicly available materials.

Per the FAQ section on the official website, orders ship from a New Jersey-based warehouse via USPS, FedEx, UPS, or DHL for international customers. The site states that products ship within 48 business hours, with standard delivery taking between five and seven days.

How Akemi Hair Glow Describes Its DHT-Blocking Mechanism

The product page centers its marketing narrative on DHT (dihydrotestosterone) as the primary driver of hair loss, particularly in women. The sales page describes DHT as a hormone that attacks hair follicles, shrinks them over time, and eventually causes them to stop producing hair entirely. The product is positioned as a formula that neutralizes DHT directly at the scalp.

The report presents what published dermatological research supports about that underlying mechanism, and where the product’s marketing language extends beyond the evidence currently available for this specific formula.

DHT’s role in androgenetic alopecia (pattern hair loss) is well established in published research. DHT is a metabolite of testosterone produced by the enzyme 5-alpha reductase. In genetically susceptible individuals, DHT binds to androgen receptors in hair follicles, triggering miniaturization — a process in which follicles progressively shrink and produce thinner, shorter hairs until they eventually stop producing visible hair. This mechanism is recognized across published dermatological literature as a primary contributor to pattern hair loss in both men and women.

The underlying science connecting DHT to hair loss is well documented. The question the report addresses is whether this specific spray, at its undisclosed ingredient concentrations, delivers DHT-blocking activity at levels sufficient to produce the results described on the product page.

Among the listed ingredients, caffeine has the most directly relevant published research regarding DHT interaction at the follicular level. A 2007 study published in the International Journal of Dermatology found that caffeine counteracted testosterone-suppressed hair growth in vitro — meaning in laboratory conditions using human hair follicles. A separate 2018 multicenter clinical study published in Skin Pharmacology and Physiology reported that a 0.2% caffeine-based topical liquid performed comparably to 5% minoxidil solution over six months in men with androgenetic alopecia. A February 2025 systematic review analyzing multiple clinical trials concluded that caffeine can stimulate cell growth and effectively penetrate hair follicles, positioning it as a promising topical agent against hair loss.

Those are meaningful findings. However, every one of those studies tested caffeine at disclosed concentrations under controlled conditions — not this specific multi-ingredient spray at undisclosed concentrations. The report references this research to provide context, not to confirm that Akemi Hair Glow replicates these study conditions.

He Shou Wu (Fo-Ti) is described on the product page as an ingredient that inhibits DHT production while stimulating dormant follicles. He Shou Wu has a long history in traditional Chinese medicine, and some in vitro studies have explored its effects on 5-alpha reductase activity. However, robust clinical trials specifically evaluating topical He Shou Wu for DHT blocking in human hair loss are limited in the published literature.

This is ingredient-level research context. Akemi Hair Glow as a finished product has not been evaluated in any published clinical trial appearing in peer-reviewed journals.

The “56% Thicker Hair in 18 Weeks” Claim: What the Data Trail Shows

The product page prominently states that customers see “56% thicker, fuller hair in just 18 weeks.” This figure appears alongside the name “Dr. Sarah Mitchell, Certified MD & Published Medical Author.”

The report outlines several questions about the evidence trail behind that number that consumers may want to consider before making a purchasing decision.

The source methodology behind the 56% figure is not disclosed on the product page. The site does not specify whether this number comes from a controlled clinical study, an internal consumer survey, instrument-measured trichoscopy data, or another methodology. Without knowing how the measurement was taken, what baseline was used, what sample size was involved, and whether a placebo control was included, the figure cannot be independently compared against published hair growth research standards.

The association with “Dr. Sarah Mitchell” is presented without a linked publication, institutional affiliation, or verifiable credential trail on the product page. Consumers researching this endorsement may want to verify the clinician’s credentials independently before weighting the claim.

For context, published clinical research on topical caffeine — one of the listed ingredients — has measured outcomes such as anagen-to-telogen hair ratios, hair density counts, and hair pull test improvements over periods of three to six months. These studies report improvements using standardized trichogram measurements. A claim of “56% thicker hair” would represent a notably strong result if measured by those clinical standards. That does not mean the claim is false — it means the methodology matters, and that methodology is not publicly available.

The product page also references a “93% success rate” and a “95% success rate” in different sections of the same page and describes “3,758 Reviews,” “3,000+ Verified Reviews,” and “4,000+ satisfied users” in different locations. These inconsistencies across the same product page are worth noting when forming an assessment of the company’s published statistics.

Akemi Hair Glow Ingredient Profile: What Published Research Supports at the Compound Level

The product page lists six primary active ingredients. The report outlines what published research supports at the individual ingredient level — and why the distinction between ingredient research and finished-product evidence matters when evaluating this type of product.

Biotin (Vitamin B7): Biotin is one of the most widely marketed hair growth ingredients. However, a 2024 systematic review published in the Journal of Clinical and Aesthetic Dermatology examined published clinical studies specifically on biotin for hair growth and found that the highest-quality study — a double-blind, placebo-controlled trial — showed no difference between the biotin and placebo groups for hair growth. The review concluded that a significant gap exists between public perception of biotin’s effectiveness and what the scientific literature actually demonstrates. Akemi Hair Glow uses topical rather than oral biotin, and the product page suggests this delivery method bypasses digestive limitations. Limited published data exists on topical biotin specifically for hair growth outcomes.

Caffeine: Among the listed ingredients, caffeine has the strongest published clinical evidence for topical hair applications. In vitro research demonstrated that caffeine counteracts DHT-induced hair follicle miniaturization and stimulates follicle proliferation. A clinical study found that a 0.2% caffeine topical performed comparably to 5% minoxidil over six months. A 2025 systematic review further supported caffeine’s potential as a topical hair loss agent while noting that most studies suffered from methodological limitations and significant gender imbalance — only 4.4% of study participants were women, despite hair loss products frequently targeting female consumers. The concentration of caffeine in Akemi Hair Glow is not disclosed, making direct comparison to published study dosages impossible.

Aminexil: The product page describes aminexil as an anti-fibrosis molecule that prevents DHT from stiffening tissue around follicles. Aminexil was originally developed and studied primarily by L’Oréal for their Dercos line. Published research has explored its potential to prevent collagen hardening around hair follicles, which can contribute to permanent hair loss. The published research base is more established than that of some cosmetic hair ingredients, though independent replication outside company-sponsored studies remains limited.

Castor Oil: Castor oil is a traditional hair care ingredient valued for its moisturizing properties. Ricinoleic acid in castor oil has demonstrated anti-inflammatory properties in published research. However, clinical evidence specifically connecting topical castor oil application to measurable hair regrowth is sparse. Most evidence supporting castor oil’s hair benefits comes from traditional use and mechanistic reasoning rather than controlled clinical trials.

He Shou Wu (Fo-Ti): This traditional Chinese medicinal herb has been used for centuries for hair-related concerns, including premature graying and hair loss. Some in vitro research has explored its effects on 5-alpha reductase activity and hair follicle cell proliferation. Clinical evidence for topical He Shou Wu in human hair growth remains limited. Additionally, oral He Shou Wu preparations have been associated with hepatotoxicity concerns in published case reports, though topical application presents a different risk profile.

Ginger Root Extract: Ginger has published evidence supporting anti-inflammatory and circulatory-boosting properties. Improved scalp microcirculation can theoretically support healthier follicle environments. However, one study published in the Journal of Dermatological Science suggested that 6-gingerol, an active compound in ginger, may actually suppress hair growth in certain contexts. The relationship between ginger and hair growth appears more nuanced than the product’s marketing suggests.

These are individual ingredient findings from published research. They do not represent clinical outcomes for Akemi Hair Glow as a finished formula, and individual experiences differ based on the type and cause of hair loss, genetics, hormonal factors, and consistency of use.

Topical Delivery: Does the Spray Format Offer a Research-Supported Advantage

The product page emphasizes that topical application delivers ingredients “directly to your follicles” rather than having them “dissolved in stomach acid where 97% of supplements get destroyed.” This positioning is central to how the product differentiates itself from oral hair supplements.

Published evidence supports the general principle that hair follicles serve as an effective penetration pathway for topically applied substances. Research published in the British Journal of Clinical Pharmacology demonstrated that caffeine penetrates hair follicles rapidly when applied topically and that follicular accumulation of caffeine can be significantly higher than in surrounding skin tissue. For topical caffeine specifically, the delivery advantage has a documented scientific basis.

The “97% destroyed in stomach acid” claim, however, oversimplifies digestive pharmacology. Oral bioavailability varies dramatically between compounds. Biotin, for example, is well absorbed orally — it has high oral bioavailability, which makes the blanket claim misleading when applied to that specific ingredient.

The more practical question is whether these specific ingredients, at their undisclosed concentrations, penetrate the scalp in sufficient quantities to produce the effects described on the product page. Without disclosed ingredient concentrations, this question cannot be answered from publicly available information.

Pricing, Guarantee, and Purchase Details

The product page includes promotional pricing ranges and inventory-related messaging that may vary over time and are determined by the company. At the time of this report, the site references “up to 50% off” and “up to 60% off” in different sections of the same page. Specific pricing tiers are presented during checkout.

The product page describes a 30-day money-back guarantee. The FAQ section states: “Simply return your product and we’ll give you a refund for 30 days.” However, the Terms of Service page includes a disclosure about a minimum 15% restocking fee for returns. This gap between the marketing presentation and the published terms is worth clarifying directly with the company before ordering.

The Terms of Service identify the operator as “Akemi Hair Glow” and reference New York for certain legal proceedings, while the Governing Law section references the laws of Florida.

Verify current pricing, refund terms, and restocking fee details before purchasing by viewing the current Akemi Hair Glow offer (official Akemi Hair Glow page).

What the Product Page Presents as Customer Feedback

The official website includes customer testimonials describing outcomes such as visible new hair growth at the crown and temples, reduced shedding, bald patches filling in, and hair texture improvements within specific timeframes. These testimonials are attributed to named individuals with locations.

The company does not disclose whether these testimonials reflect typical outcomes or controlled evaluation conditions. The site does not describe how testimonials are collected or verified, or whether any compensation or incentive is provided. People who write reviews are self-selected — satisfied customers are significantly more likely to post feedback than those with neutral or negative experiences.

Testimonial results are individual experiences and should not be interpreted as typical or guaranteed outcomes. Published research on hair growth interventions consistently shows that individual outcomes depend on the specific type and cause of hair loss, genetic factors, hormonal profiles, and treatment consistency.

Consumer Considerations Based on Product Characteristics

This section outlines general considerations based on the product’s described characteristics and publicly available ingredient research. It is not intended to recommend the product for any individual use case.

Consumers evaluating topical products may consider Akemi Hair Glow if they:

Prefer topical over oral approaches. Consumers who have tried oral hair supplements without satisfaction or who prefer applying active ingredients directly to the scalp may find a spray format aligns with their preference. Published research on topical caffeine shows promising follicular penetration and, in at least one clinical study, comparable performance to topical minoxidil.

Are exploring complementary options alongside professional care. Consumers already working with a dermatologist and looking for an additional daily topical may note that ingredients like caffeine and aminexil have published research supporting their potential roles in hair maintenance as part of a broader approach.

Want a non-prescription starting point. For consumers not yet ready for prescription options like minoxidil or finasteride, a botanical-based spray may serve as an entry point while evaluating whether professional intervention is appropriate for their specific situation.

Other options may be preferable for consumers who:

Require transparent ingredient dosing. Consumers who need exact ingredient concentrations for comparing against published research dosages should note that those amounts are not disclosed for this product.

Are experiencing sudden or severe hair loss. Rapid, sudden, or patchy hair loss may indicate medical conditions such as alopecia areata, thyroid disorders, or nutritional deficiencies that require professional diagnosis and treatment rather than a topical cosmetic product.

Expect results comparable to FDA-approved treatments. Published clinical data for FDA-approved treatments like minoxidil represents a different evidence standard than what is currently available for this proprietary formula.

Questions to consider before purchasing: What type of hair loss am I experiencing, and has a healthcare professional evaluated the cause? Have I researched the specific ingredients at the concentrations used in published studies? Am I comfortable purchasing a product without disclosed ingredient concentrations? Have I reviewed the full refund terms, including the potential restocking fee? These questions can help determine whether this product’s characteristics match a consumer’s specific situation.

Verification Checklist: What to Confirm Before Ordering

Verify ingredient concentrations. Individual ingredient amounts per application are not disclosed. If dosing relative to published research matters to the purchasing decision, contact the company directly to request this information.

Separate ingredient research from product research. Published studies on caffeine, biotin, aminexil, and other listed ingredients examined those compounds individually under controlled conditions. No published clinical trial appears to evaluate Akemi Hair Glow as a finished proprietary formula.

Trace the “56% thicker hair” claim. The methodology behind this figure is not publicly disclosed. Without understanding the measurement approach, sample size, and control conditions, the number cannot be independently verified against clinical standards.

Review the refund terms carefully. The product page describes a 30-day money-back guarantee, but the Terms of Service disclose a potential 15% restocking fee. Clarify exact refund terms before purchasing.

Consult a healthcare provider. This is especially important for women experiencing hair loss related to hormonal changes, thyroid conditions, nutritional deficiencies, or medical treatments. A dermatologist can help identify the underlying cause and recommend evidence-based interventions. This report is not a substitute for professional dermatological evaluation.

Consumer Questions About Akemi Hair Glow

What is DHT, and why does it matter for hair loss?

DHT (dihydrotestosterone) is a hormone derived from testosterone that plays a recognized role in androgenetic alopecia (pattern hair loss). In genetically susceptible individuals, DHT causes hair follicle miniaturization over time. This mechanism is well established in published dermatological research. However, not all hair loss is DHT-related — conditions such as telogen effluvium, alopecia areata, and nutritional deficiency-related hair loss involve entirely different mechanisms.

Is Akemi Hair Glow a medication or FDA-approved product?

Akemi Hair Glow is marketed as a topical cosmetic hair spray, not a medication. It is not FDA-approved for the treatment of any medical condition. The product has not undergone the clinical trial process required for FDA drug approval. Consumers seeking FDA-approved treatments for hair loss should discuss options such as topical minoxidil or oral finasteride with a qualified healthcare provider.

How does the caffeine in this product compare to what has been studied clinically?

Published clinical research has tested topical caffeine at disclosed concentrations — for example, a 0.2% caffeine-based topical that performed comparably to 5% minoxidil over six months. Akemi Hair Glow’s caffeine concentration is not disclosed, which makes direct comparison to published study dosages impossible. These are different products with different evidence bases, even though they share a common ingredient.

Will this product address my type of hair loss?

The product’s marketing focuses on DHT-related hair thinning. If hair loss has a different underlying cause — such as thyroid dysfunction, iron deficiency, autoimmune conditions, or medication side effects — a topical DHT-blocking spray may not address the underlying issue. A healthcare professional can help determine the cause and recommend appropriate interventions.

How long before results might be noticed?

The product page describes a week-by-week timeline suggesting changes beginning at week 2 and continuing through week 16. Published research on topical hair growth interventions generally suggests that meaningful results, when they occur, typically require three to six months of consistent use. Individual timelines depend on factors including the type of hair loss, its severity, and individual biological response. The timeline descriptions on the product page represent the company’s marketing positioning rather than clinically validated benchmarks for this specific product.

What should consumers know about the return policy before ordering?

The product page describes a 30-day money-back guarantee. The Terms of Service page discloses a potential minimum 15% restocking fee for returns. The exact refund process, applicable fees, and return shipping requirements should be confirmed directly before purchasing. Complete terms are available by viewing current Akemi Hair Glow terms (official Akemi Hair Glow page).

Summary of Key Considerations

Akemi Hair Glow is a topical hair spray positioned around DHT-blocking and follicle reactivation claims. The product contains ingredients — particularly caffeine — with published research at the individual compound level supporting potential hair growth benefits when applied topically. Caffeine’s evidence base is the strongest among the listed ingredients, with at least one clinical study demonstrating comparable performance to topical minoxidil at a disclosed concentration.

However, the finished formula has not been evaluated as a proprietary product in any published clinical trial. Ingredient concentrations are not disclosed. Several marketing statistics — including “56% thicker hair in 18 weeks,” a “93% success rate,” and varying review counts across the same page — lack publicly available methodology or present inconsistencies.

The biotin evidence is notably weaker than the product’s positioning suggests. A 2024 systematic review found no difference between biotin and placebo for hair growth in the highest-quality study examined. This is worth factoring into any evaluation of the formula’s overall research backing.

The product page describes a 30-day money-back guarantee, though the Terms of Service disclose a potential 15% restocking fee. Orders ship from a New Jersey warehouse, and customer support is available by email and phone.

Consumers who have completed their own research and want to review the full product details can do so. Complete product details, current pricing, and published terms are available by viewing the current Akemi Hair Glow offer (official Akemi Hair Glow page).

Contact Information

Per the company’s published contact information:

Company: Akemi Hair Glow

Email: support@buyskyline.co

Phone: +1 (866) 697-4823

Shipping Origin: New Jersey, United States (per the company’s FAQ)

Website: buyskyline.co

Disclaimers

Content and Consumer Information Disclaimer: This article is an informational overview and does not constitute medical, health, or dermatological advice. All product details, ingredient information, pricing, and policy terms described in this article are stated as presented on the publicly available product website and related materials. Readers are encouraged to verify all claims directly on the official website and to consult a qualified healthcare professional before beginning any topical hair treatment.

Health and Hair Loss Notice: Hair loss can result from numerous causes, including genetics, hormonal changes, nutritional deficiencies, medical conditions, medications, and stress. Topical cosmetic products are not substitutes for professional dermatological evaluation and treatment. If you are experiencing sudden, severe, or patchy hair loss, consult a healthcare provider. Individual results vary based on factors including the type and cause of hair loss, genetic factors, hormonal profiles, consistency of use, and other individual variables. While some consumers report improvements, results are not guaranteed.

Results May Vary: Individual results will vary based on numerous factors, including age, baseline hair condition, the type and underlying cause of hair loss, hormonal factors, consistency of use, genetic factors, current medications, and other individual variables. While some customers report improvements, results are not guaranteed. People who write reviews are self-selected — satisfied customers are more likely to post feedback than those with neutral or negative experiences.

FTC Affiliate Disclosure: This article contains affiliate links. If a product is purchased through these links, a commission may be earned at no additional cost to the buyer. This compensation does not influence the accuracy, neutrality, or integrity of the information presented. All descriptions are based on published research and publicly available information from the company’s official website.

Pricing Disclaimer: All prices, discounts, and promotional offers mentioned were based on information published on the official product website at the time of publication (March 2026) and are subject to change without notice. Always verify current pricing, promotions, and terms on the official website before making a purchase.

Publisher Responsibility: The publisher of this article has made every effort to ensure accuracy at the time of publication. The publisher does not accept responsibility for errors, omissions, or outcomes resulting from the use of the information provided. Readers are encouraged to verify all details directly with the company and their healthcare provider before making decisions.

CONTACT: Email: support@buyskyline.co
Phone: +1 (866) 697-4823

Prothena Partners Present Data Supporting Next Generation Treatments for Parkinson’s and Alzheimer’s Disease at AD/PD™ 2026

Prothena Partners Present Data Supporting Next Generation Treatments for Parkinson’s and Alzheimer’s Disease at AD/PD™ 2026


Prothena Partners Present Data Supporting Next Generation Treatments for Parkinson’s and Alzheimer’s Disease at AD/PD™ 2026

DUBLIN–(BUSINESS WIRE)–$PRTA #Prothena–Prothena Corporation plc (NASDAQ:PRTA), a late-stage clinical biotechnology company with a robust pipeline of investigational therapeutics built on protein dysregulation expertise, today announced partner presentations on clinical updates from prasinezumab for the treatment of Parkinson’s disease and BMS-986446 for the treatment of Alzheimer’s disease at the International Conference on Alzheimer’s and Parkinson’s Diseases and Related Neurological Disorders (AD/PD™ 2026) in Copenhagen, Denmark, and online.


Roche Presentations on Prasinezumab for the Potential Treatment of Parkinson’s Disease

Industry Symposium – Pathways to Progress: Exploring Innovations in AD and PD for Future Practice

  • Chair: Malú G. Tansey, Ph.D., Indiana University School of Medicine Stark Neuroscience Institute
  • Date: Tuesday March 17, 2026

This symposium reviewed the evolving understanding of the molecular pathophysiology and disease heterogeneity across Alzheimer’s disease (AD) and Parkinson’s disease (PD). Explored current and emerging treatment pathways, e.g. alpha-synuclein, amyloid-beta and neuroinflammation; including diagnostics, biomarkers and therapeutics. Considered how the advancing understanding of AD and PD informs innovative clinical development approaches and clinical practice.

Oral Presentation – Modeling Parkinson’s Disease Progression to Quantify Long-Term Treatment Effects via the Concept of ‘Time Saved’

  • Presenter: Benjamin Ribba, Roche
  • Date: Thursday March 19, 2026

The comparison of PASADENA open-label extension (OLE) data with PPMI-based model predictions supports potential disease-modifying efficacy with an estimated two years of ‘time saved’ providing an intuitive measure of long-term benefit. The observed PASADENA OLE outcomes consistently deviated from the model-predicted progression, suggesting a sustained treatment effect. On average, participants were approximately two years less advanced in disease severity five years after the start of the trial compared to the virtual comparator.

Oral Presentation – Prasinezumab in Early-Stage Parkinson’s Disease: Additional Data from the PADOVA Study

  • Presenter: Tania Nikolcheva, M.D., Ph.D., Roche
  • Date: Saturday March 21, 2026

Longer term data from the PADOVA OLE study in early-stage PD showed a sustained effect of prasinezumab in slowing Parkinson’s progression on top of effective symptomatic therapies. The totality of the evidence suggests a possible clinical benefit of prasinezumab and informed the initiation of the Phase III PARAISO study.

Poster Presentation – Prasinezumab’s Impact on Neuromelanin- and Iron-Sensitive MRI Biomarkers in Parkinson’s Disease: Findings from the PADOVA Phase IIb Study

Exploratory biomarker analysis of PADOVA suggests that prasinezumab is biologically active. This is supported by imaging biomarkers crucial to PD pathology, showing a slowing in the progressive loss of neuromelanin signal in substania nigra pars compacta and reduced iron accumulation in the putamen.

Poster Presentation – Sustained Effect on Prasinezumab on Parkinson’s Disease Motor Progression in the Open-Label Extension of the PASADENA Trial, 5-Year Update

At Year 5, the combined PASADENA arm (delayed- and early-start groups) showed less disease progression compared to the PPMI cohort. This lower progression was observed across multiple measures.

Poster Presentation – Digital Health Technology Detects Group Differences in Practically-Defined OFF L-DOPA State: Results of PADOVA Phase IIb Study of Prasinezumab

Post-hoc Digital Health Technology analyses showed consistent trends favoring prasinezumab in digital data collected in the practically-defined OFF L-DOPA state, in line with the PASADENA Phase 2a Simple Sum digital finding and clinical PADOVA readout.

Bristol Myers Squibb Presentation on BMS-986446 for the Potential Treatment of Alzheimer’s Disease

Oral Presentation – Randomized, Double-Blind, Placebo-Controlled Study Evaluating Safety, Tolerability, Pharmacokinetics, and Immunogenicity of BMS-986446 in Healthy Participants, Including Those of Japanese Ethnicity

  • Presenter: Ilena George, M.D., Bristol Myers Squibb
  • Date: Saturday March 21, 2026

Single-dose BMS-986446 was safe and well tolerated in all participants, including those of Japanese ethnicity. Plasma exposure of BMS-986446 increased dose proportionally. No anti-drug antibodies were detected. These results support BMS-986446 dosing in ongoing clinical studies without adjustments for Japanese ethnicity.

About Prasinezumab

Prasinezumab is an investigational monoclonal antibody designed to bind aggregated alpha-synuclein and thereby reduce neuronal toxicity. By reducing the build-up of alpha-synuclein protein in the brain, prasinezumab can potentially prevent further accumulation and spreading between cells, which may slow progression of the disease.

About Parkinson’s Disease

Parkinson’s disease is a chronic, progressive and debilitating neurodegenerative disease characterized by the gradual loss of neurons that make dopamine and other nerve cells. Today, Parkinson’s disease affects over 10 million people worldwide. The prevalence of Parkinson’s disease is increasing, and it has become one of the fastest-growing neurological disorders. Currently, symptomatic treatments that effectively alleviate motor symptoms are available. However, no therapies slow down or stop the clinical progression of Parkinson’s disease.

About BMS-986446

BMS-986446 is a humanized monoclonal antibody that targets multiple domains of the microtubule binding region of tau, a highly pathogenic tau fragment associated with neurofibrillary tangle formation and cognitive decline in Alzheimer’s disease. BMS-986446 binds to specific regions of the tau protein (R1–R3 within the microtubule-binding domain) to stop cell-to-cell spread of tau and tau uptake into cells. It also activates microglia—the brain’s immune cells—through its Fc receptor function, promoting the clearance of tau via phagocytosis.

About Alzheimer’s Disease

Alzheimer’s disease is a progressive, multifaceted and devastating neurodegenerative disease and the most common type of dementia in adults. Changes in the brain disrupt communication between neurons, impacting memory, cognition and behavior. As a result, Alzheimer’s disease has a significant impact on the day-to-day lives of those it directly affects, as well as on their families, caregivers and friends, resulting in considerable shifts in interpersonal relationships. There remains a critical need for disease-modifying therapies that can slow or delay the progression of Alzheimer’s disease as well as therapies that manage and ease neurobehavioral symptoms.

About Prothena

Prothena Corporation plc is a late-stage clinical biotechnology company with expertise in protein dysregulation with the potential to change the course of devastating neurodegenerative and rare peripheral amyloid diseases. Fueled by its deep scientific expertise built over decades of research, Prothena is advancing a pipeline of therapeutic candidates for a number of indications and novel targets for which its ability to integrate scientific insights around neurological dysfunction and the biology of misfolded proteins can be leveraged. Prothena’s pipeline includes both wholly-owned and partnered programs being developed for the potential treatment of diseases including Parkinson’s disease, ATTR amyloidosis with cardiomyopathy, Alzheimer’s disease, Amyotrophic lateral sclerosis (ALS) and a number of other neurodegenerative diseases. Prothena is developing and applying CYTOPE®, a novel technology that incorporates a cell-internalizing domain to drive efficient cytosolic delivery with highly specific macromolecular effectors. For more information, please visit the Company’s website at www.prothena.com and follow the Company on X (formerly Twitter) @ProthenaCorp.

Forward-Looking Statements

This press release contains forward-looking statements. These statements relate to, among other things, the treatment potential, designs, proposed mechanisms of action, and potential administration of prasinezumab and BMS-986446; and the continued advancement of our preclinical and clinical pipeline, including the potential and advancement of CYTOPE. These statements are based on estimates, projections and assumptions that may prove not to be accurate, and actual results could differ materially from those anticipated due to known and unknown risks, uncertainties and other factors, as well as those described in the “Risk Factors” sections of our Annual Report on Form 10-K filed with the Securities and Exchange Commission (SEC) on February 27, 2026, and discussions of potential risks, uncertainties, and other important factors in our subsequent filings with the SEC. We undertake no obligation to update publicly any forward-looking statements contained in this press release as a result of new information, future events, or changes in our expectations.

Contacts

Mark Johnson, CFA

Senior Vice President, Head of Investor Relations and Corporate Communications

650-837-8550

IR@prothena.com
Media@prothena.com

IBA Launches myQAMatriXXAiR to Advance Patient‑Specific QA in Particle Therapy

IBA Launches myQAMatriXXAiR to Advance Patient‑Specific QA in Particle Therapy


IBA Launches myQAMatriXXAiR to Advance Patient‑Specific QA in Particle Therapy

The first wireless ionization chamber array for particle therapy PSQA delivers fast, reliable verification for proton and carbon ion treatments, supporting emerging techniques such as FLASH and ARC radiotherapy.

Louvain-la-Neuve, Belgium, 21 March 2026 – IBA (Ion Beam Applications S.A., EURONEXT), the world leader in particle accelerator technology and a global provider of dosimetry and quality assurance (QA) solutions, announces the launch of myQA MatriXX AiR, a high resolution, wireless 2D ionization chamber array designed for patient specific quality assurance (PSQA) in particle therapy, including proton and carbon ion treatments. This new solution further demonstrates IBA’s role as a driving force in innovation and reinforces its leadership in quality assurance for particle‑beam therapies.

In radiation therapy, quality assurance is the critical safeguard that ensures increasingly complex treatments are delivered exactly as planned, translating technological innovation into clinical confidence and patient safety.

As particle therapy expands worldwide, proton and carbon‑ion centers are adopting increasingly advanced treatment modalities—from high‑precision conformal therapies to emerging beam‑delivery techniques such as FLASH and ARC radiotherapy. These innovations accelerate clinical potential, but they also elevate the need for robust, future‑proof quality assurance. With myQA MatriXX AiR, IBA sets a new industry standard as the first QA solution capable of providing full patient‑specific verification across both established particle‑therapy treatments and next‑generation delivery modes. This is not an adaptation to change—it is a decisive step forward, positioning IBA as the only partner offering uncompromised QA readiness for the future of particle‑beam therapy.

Engineered specifically for the demands of modern particle therapy, myQA MatriXX AiR features 1,521 high resolution air‑vented ionization chambers and delivers a complete 2D dose distribution readout within seconds, enabling fast and reliable verification of increasingly complex treatment plans. As treatment techniques become more complex, this level of detail directly translates into greater confidence, higher patient safety, and more reliable plan approval.

As the first commercially available wireless detector array for particle therapy PSQA, MatriXX AiR simplifies setup, reduces QA time, and integrates easily into existing workflows. The system is also FLASH‑ready, with hardware designed to support ultra‑high dose rate measurements as clinics prepare for next‑generation radiotherapy techniques.

Commenting on the solution Dr. Loïc Grevillot, Deputy Head of Medical Physics at the MedAustron Ion Therapy Center, stated: “We have been pleased to test the latest MatriXX detector from IBA Dosimetry, which offers a user-friendly interface and — most importantly for our needs —compatibility with carbon ion beams. The detector has demonstrated robust performance in dose evaluation and shows potential for clinical implementation in carbon ion therapy beam delivery QA and PSQA workflows.”

Jean-Marc Bothy, President of IBA Dosimetry, added: “We are proud to offer products that significantly enhance safety in patient plan deliveries in proton therapy. It’s exciting to see a QA solution designed specifically for particle therapy. MatriXX AiR builds on the strengths of our trusted MatriXX  PT and ONE, bringing a new level of precision, speed, and wireless flexibility. We’ve worked closely with clinical teams, and this evolution reflects their needs of effective QA that keeps pace with the future of treatment modalities. With this launch, we further strengthen our leadership in quality assurance for particle‑beam therapies.”

***ENDS***

About IBA

IBA (Ion Beam Applications S.A.) is the world leader in particle accelerator technology. The company is the leading supplier of equipment and services in the fields of proton therapy, considered as one of the most advanced forms of radiation therapy available today, as well as industrial sterilization, radiopharmaceuticals and dosimetry. The company, based in Louvain-la-Neuve, Belgium, employs approximately 2,100 people worldwide. IBA is a certified B Corporation (B Corp) meeting the highest standards of verified social and environmental performance.

IBA is listed on the pan-European stock exchange EURONEXT (IBA: Reuters IBAB.BR and Bloomberg IBAB.BB).

More information can be found at: www.iba-worldwide.com

CONTACTS

Olivier Lechien
Corporate Communication Director
+32 10 475 890
communication@iba-group.com

Attachment

Financière de Tubize – Annual report 2025

Financière de Tubize – Annual report 2025


Financière de Tubize – Annual report 2025

Annual Report 2025
Regulated information 21 March 2026

The board of directors of Financière de Tubize has established the 2025 annual report. This report is available on the website www.financiere-tubize.be

  • Profit for the financial year: € 93.2 million (€ 90.1 million in 2024, + 3.5%),
  • Full repayment of bank debt in May 2025 (€ 41.3 million at 31 December 2024),
  • Acquisition, in 2025, of 24,487 UCB shares, at an average price of €178.27, for a total amount of €4.4 million, increasing the holding of the Company in UCB from 36.27% on 31 December 2024 to 36.28% on 31 December 2025.

If the general shareholders meeting of 24 April 2026 approves the 2025 annual accounts, including the proposed result appropriation, a gross dividend of € 1.08 (compared to €1.04 for financial year 2024, an increase of 3.85%) will be payable as from 11 May 2026 onwards at the offices, seats and branches of BNP Paribas Fortis, in exchange of coupon n° 21.

Ex-dividend            7 May 2026
Record date           8 May 2026
Payment date        11 May 2026