|
EQS Newswire / 16/03/2026 / 18:08 UTC+8 Focused on developing high-value inhalation therapies for pulmonary hypertension (PAH and PH-ILD), with clinical potential to expand into pulmonary fibrosis indications (PF-ILD, including IPF and PPF). Suzhou, China, March 13, 2026 — CF PHARMTECH, INC. (HKEX: 2652.HK, hereinafter referred to as “CF PHARMTECH, INC.” or “the Company”) today announced that the National Medical Products Administration (NMPA) has officially accepted the Investigational New Drug (IND) application for ICF001. Independently developed by the Company, ICF001 is an innovative inhalation powder for the treatment of pulmonary arterial hypertension (PAH) and pulmonary hypertension associated with interstitial lung disease (PH-ILD). It is classified as a Class 2.1 improved new chemical drug in China. Following the recent acceptance of ICF004, ICF001 is another candidate from the Company’s high-barrier respiratory pipeline to reach this milestone, signaling an accelerated harvest phase for CF PHARMTECH, INC.’s innovative R&D. ICF001 utilizes a prodrug-based mechanism designed to achieve long-acting efficacy. As drugs in this class have already demonstrated blockbuster potential in treating rare and serious pulmonary diseases, ICF001 is positioned to capture significant growth as it expands into new indications. Addressing Unmet Clinical Needs and Filling a Domestic Treatment GapICF001 targets two critical categories of pulmonary hypertension: WHO Group 1 pulmonary arterial hypertension (PAH) and WHO Group 3 pulmonary hypertension associated with interstitial lung disease (PH-ILD). Both conditions are associated with poor prognosis and urgently require better treatment options.
Notably, there are currently no approved targeted therapies for PH-ILD in China. The rapid development of ICF001 positions it to potentially become the first inhaled therapy approved for this indication in China, addressing a critical therapeutic void and offering a transformative treatment option for patients worldwide. Tackling Key Industry Challenges with a Globally Differentiated Improved MechanismWhile the industry is shifting toward long-acting therapies to reduce dosing frequency, existing approaches often face challenges, including single-dose burden, local tolerability, and titration complexity, all of which can affect dose escalation and long-term patient adherence. ICF001 is designed to address these clinical pain points through precise formulation and pharmacokinetic optimization, with the goal of delivering two key breakthroughs while demonstrating multi-indication expansion potential: Enhanced patient adherence through reduced dosing frequencyBy optimizing molecular structure and formulation, ICF001 increases drug loading efficiency and improves local tolerability, reducing the overall administration burden for long-term therapy. Optimized pharmacokinetics through a “peak-shaving and trough-filling” profile, balancing safety and efficacyDelivered directly to the lungs, ICF001 is designed to blunt peak plasma concentration (Cmax) while extending drug exposure (AUC). This “peak-shaving and trough-filling” profile improves systemic tolerability and may enhance clinical efficacy while maintaining safety. “One drug, Multiple indications” StrategyExpanding beyond PAH and PH-ILD, ICF001 utilizes a mechanism of action that targets pulmonary fibroblast activation, offering dual potential in pulmonary hypertension and pulmonary fibrosis. Backed by cutting-edge global research and clinical exploration of this drug class, ICF001 is expected to emerge as a next-generation blockbuster, addressing significant unmet needs in the broader respiratory market. These differentiated advantages represent the Company’s R&D goals and strategic direction based on translational medicine models. If improvements in tolerability and titration efficiency are confirmed in subsequent clinical studies, ICF001 is expected to improve long-term treatment adherence, strengthen efficacy potential, and further expand clinical accessibility. The rapid acceptance of this IND application marks another critical milestone in the Company’s clinical development of its high-barrier respiratory pipeline. It demonstrates the Company’s solid fundamentals, forward-looking strategic positioning, and efficient execution in innovative drug R&D. Furthermore, it established a strong foundation for the Company to further penetrate the global high-value inhalation therapy market to address unmet clinical needs. The market holds high expectations for the clinical application of such improved new drugs. Validating Platform Value: Extending from Complex Formulation Capabilities to Innovative Drug TranslationThe IND acceptance of ICF001 marks a key transition for CF PHARMTECH, INC.’s inhaled innovative drug program enters the regulatory phase. This progress represents a strategic breakthrough with long-term sustainable development potential:
By leveraging its integrated global capabilities, CF PHARMTECH is building a multi-layered product portfolio centered on the synergy of advanced complex formulations and innovative therapeutics. This strategic focus solidifies the Company’s position in the high-value global inhalation market while enabling the expansion of its proprietary delivery technology into broader therapeutic areas and innovative drug development.
16/03/2026 Dissemination of a Financial Press Release, transmitted by EQS News. Media archive at www.todayir.com |
Author Archives:
BIO-Europe Spring® 2026: Partnering Momentum Builds as Global Biopharma Leaders Set Course for Lisbon
|
EBD Group / Key word(s): Conference BIO-Europe Spring® 2026: Partnering Momentum Builds as Global Biopharma Leaders Set Course for Lisbon 16.03.2026 / 11:00 CET/CEST The issuer is solely responsible for the content of this announcement.
PRESS RELEASE BIO-Europe Spring® 2026: Partnering Momentum Builds as Global Biopharma Leaders Set Course for Lisbon MUNICH, GERMANY – March 16, 2026. The global biopharmaceutical community is preparing to gather in Lisbon, Portugal, for the 20th edition of BIO-Europe Spring®, the premier springtime partnering conference for the industry, taking place March 23–25, 2026. More than 3,700 attendees from over 2,000 companies, including leading pharmaceutical firms, innovative biotech companies, and global investors, are expected along Lisbon’s shores for one of Europe’s most important meeting points for life science innovation and dealmaking. Held alongside LSX Europe as part of Life Sciences Spring Innovation Week, this year’s event will create an expanded platform for strategic discussions across biotech, pharma, and investment communities. Partnering has opened and is now in full swing, with over 20,000 one-to-one meetings expected, reinforcing BIO-Europe Spring’s role as a vital hub for investment, collaboration, and cross-border engagement across the global life sciences ecosystem. “Portugal’s life sciences sector has developed into a dynamic and internationally connected ecosystem, driven by strong research, innovative biotech companies, and growing investment activity,” said Joana Branco, Director of Innovation and Ecosystem at Biocant Park, co-host sponsor of BIO-Europe Spring 2026. Filipa Sacadura, Director General of P-Bio, also co-host sponsor, added: “Hosting BIO-Europe Spring 2026 in Lisbon is a fantastic opportunity to showcase Portugal’s innovation landscape and connect our biotech community with global industry leaders, investors, and partners.” “We are excited to bring the 20th annual edition of BIO-Europe Spring to Lisbon, a city whose life sciences ecosystem continues to gain international recognition,” said Claire Macht, Director of BIO-Europe Spring. “With strong participation from biotech innovators, pharmaceutical leaders, and global investors, BIO-Europe Spring 2026 will provide an essential platform to explore new partnering opportunities and help shape the next wave of life science innovation.” BIO-Europe Spring 2026 will focus on partnering and dealmaking from early-stage innovators to mature biotech companies. The event recently strengthened its collaboration with YVC Collective, a network of more than 600 emerging life sciences venture capital professionals across Europe. Through this partnership, YVC Collective members help review applications for the conference’s Startup Spotlight and Presenting Company programs, ensuring promising early-stage companies gain exposure to investors and industry leaders at a pivotal stage of development. Key Sessions and Events at BIO-Europe Spring 2026: The 20th annual BIO-Europe Spring conference will feature an engaging program curated by EBD Group (an Informa company) offering company presentations, panel discussions, and networking opportunities designed to foster collaboration across the life sciences value chain. A featured panel, “Piecing Together the Therapeutic Landscape with Analyst Insights”, will leverage leading analyst intelligence and data-driven insights to provide a deep dive into the evolving biopharma ecosystem. Moderated by Daniel Chancellor (Evaluate), the panelists Samuel Bennett (Novo Nordisk), Hakan Goker (M Ventures), Laura Lane (Eli Lilly and Company), Toby Richardson (Johnson & Johnson), and Jan Van den Bossche (Andera Partners) will explore emerging trends, market dynamics, and investment opportunities across key therapeutic areas, offering strategic insights to navigate 2026 and beyond. Another ever-popular, recurring session, “Day in the Life of an Experienced Dealmaker”, will offer a firsthand look at the current state of the life sciences landscape. Anton Gueth (EVOLUTION Life Science Partners), Evonne Sepsis (ESC Advisors), Claudio Costa-Neto (Jeito Capital), Evan Lippman (Teva Pharmaceuticals), and Chris Sheldon (GSK) will share strategies for navigating the complex biotech and pharma ecosystem. The program will also spotlight emerging innovators through the Startup Spotlight, a live pitch competition showcasing promising biotech startups. Selected companies will present their innovations to the BIO-Europe Spring audience and a panel of expert judges, offering insight into the next generation of life science innovation. Partnering and Registration for BIO-Europe Spring 2026 One-to-one meetings will be powered by partneringONE®, an industry-standard platform that enables delegates to search, request, schedule, and conduct meetings efficiently. To enhance access and extend engagement beyond the in-person event, the conference will continue with two days of virtual partnering on March 31–April 1, allowing participants to connect regardless of time zone or travel constraints. Registration information for BIO-Europe Spring is available online. Discounted rates are still available until March 20, 2026. Additional Links and Information BIO-Europe Spring is organized by EBD Group, an Informa company. For more information and live updates, please visit the conference website at: https://informaconnect.com/bioeurope-spring and follow BIO-Europe Spring 2026 on LinkedIn. About Informa and EBD Group EBD Group’s mission is to help collaborations get started across the life science value chain. Our range of partnering conferences has grown to become the largest and most productive conference portfolio in the industry. Each one of our landmark events, held in key life science markets around the world, is powered by our state-of-the-art partnering software, partneringONE, that enables delegates to efficiently identify and engage with new opportunities via one-to-one meetings. Today, our events and those of our sister organization, LSX (BIO-Europe, BIO-Europe Spring®, LSX Europe, Biotech Showcase™, LSX USA, Investival Showcase USA, ChinaBio® Partnering Forum, Asia Bio Partnering Forum, LSX Nordic, BioEquity Europe, Investival Showcase EU, and the European Lifestars Awards), annually attract more than 15,000 senior life science executives who engage in over 50,000 one-to-one partnering meetings. These vital one-to-one engagements are the wellspring of deals that drive innovation in our industry. EBD is an Informa company. Informa is a leading international Live B2B Events, B2B Digital Services, and Academic Markets Group. Informa is a member of the FTSE 100 and works in over 30 countries. For more information, please visit: https://informaconnect.com/partnering-investment-strategy/ Media Contacts: MC Services AG Informa
16.03.2026 CET/CEST Dissemination of a Corporate News, transmitted by EQS News – a service of EQS Group. The EQS Distribution Services include Regulatory Announcements, Financial/Corporate News and Press Releases. |
2291418 16.03.2026 CET/CEST
NVision and Aarhus University Awarded Innovation Fund Denmark Grant to Advance Clinical Translation of Quantum-Enhanced MRI
|
NVision Imaging Technologies GmbH / Key word(s): Miscellaneous NVision and Aarhus University Awarded Innovation Fund Denmark Grant to Advance Clinical Translation of Quantum-Enhanced MRI 16.03.2026 / 10:15 CET/CEST The issuer is solely responsible for the content of this announcement. NVision and Aarhus University Awarded Innovation Fund Denmark Grant to Advance Clinical Translation of Quantum-Enhanced MRI The €5.4 million (40m DKK) investment for the MIRAQLE project supports the development of scalable, clinically deployable hyperpolarized MRI for early liver cancer diagnostics. ULM, Germany & AARHUS, Denmark — March 16, 2026 — NVision, a quantum technology company, and Aarhus University have secured a €5.4 million (40m DKK) grant for the MIRAQLE project under Denmark’s Grand Solutions in Quantum Technologies program. The award is a significant endorsement of NVision’s quantum hyperpolarization technology and establishes a direct route for quantum-enabled diagnostics to move from the lab to the clinic. Cancer is often detected too late. Although standard MRI scans are good at showing a tumor’s size and location, they miss the metabolic activity that reveals how aggressive the disease is or whether a treatment is working. MIRAQLE aims to change this. Researchers from the MR Research Centre at Aarhus University and NVision Imaging Technologies will develop a new type of MRI platform capable of visualizing cancer metabolism. By boosting the imaging agent’s MRI signal, metabolic activity can become visible during a routine MRI scan. This could make it possible to detect serious disease earlier and to monitor treatment effects far more precisely than is currently possible. Expanding Clinical Applications to Liver Cancer NVision’s research partnerships are rapidly expanding to cover a diverse range of medical applications. While research at Memorial Sloan Kettering Cancer Center is focused on monitoring treatment efficacy in various cancers, NVision’s recently established research hub at the University of Cambridge Department of Radiology allows for the exploration of both oncology and non-oncology applications, including general liver disease and multiple sclerosis. The new research project at Aarhus University builds on this momentum with a dedicated focus on liver cancer. In hepatocellular carcinoma, it can be difficult for doctors to distinguish between benign changes and early-stage liver cancer, often delaying diagnosis until the disease is advanced. With NVision’s technology, doctors will be able to not only detect the disease earlier but also assess how aggressive it is. “Our expanding network of research partnerships enables us to advance the use of hyperpolarized MRI across a broad spectrum of critical medical applications—from early disease detection and stratification to monitoring oncology treatments and tailoring therapies based on metabolic response,” said Prof. Myriam Chaumeil, Head of Research and MIRAQLE project lead at NVision. “Liver cancer is a compelling application because hyperpolarized MRI can reveal the metabolic activity that drives tumor growth in real time, providing insights that conventional imaging cannot offer.” “We will be able to see how the cancer behaves, not just where it is. This could change the feedback cycle from months to days, making a decisive difference to when and how patients are treated,” said Professor Christoffer Laustsen, project leader and Head of the MR Research Centre at Aarhus University. Up to 100,000 Times More Sensitive NVision’s proprietary technology, POLARIS, is based on quantum physics and can make MRI scans up to 100,000 times more sensitive than those available today. This means that even very small changes on the cellular level can be detected. At the same time, the scan is carried out entirely without radioactive radiation – unlike many current methods used in cancer diagnostics. POLARIS is being developed so that it can be used in regular hospitals and integrated into existing MRI infrastructure. Driving this clinical translation is the goal of the MIRAQLE project, which spans everything from laboratory research to preclinical and clinical studies in hepatocellular carcinoma to demonstrate the technology’s diagnostic value. Strong European Backing MIRAQLE strengthens NVision’s robust portfolio of publicly funded European research consortiums. The project joins a growing list of successful initiatives aimed at advancing and commercializing quantum medical technology, including CHARM, HYPERCELL, MAGSENSE, Q-AID, Que-MRT, and most recently, the AURORA project. Together, these consortiums represent more than €35 million in public grant funding secured since 2022 to support the transition of NVision’s technology from the lab to the clinic. About NVision Imaging NVision Imaging was founded in 2015 and is headquartered in Ulm, Germany. By leveraging advances in quantum physics, NVision enables real-time visualization of metabolism on standard MRI, aiming to set a new standard for faster drug development, earlier diagnosis, and better therapy decisions. Learn more at nvision-imaging.com. Contact: Leah Wiedenmann
16.03.2026 CET/CEST Dissemination of a Corporate News, transmitted by EQS News – a service of EQS Group. The EQS Distribution Services include Regulatory Announcements, Financial/Corporate News and Press Releases. |
2291810 16.03.2026 CET/CEST
Oculis to Present at Upcoming North American Neuro-Ophthalmology Society Annual Meeting
Oculis to Present at Upcoming North American Neuro-Ophthalmology Society Annual Meeting
Oculis to Present at Upcoming North American Neuro-Ophthalmology Society Annual Meeting

ZUG, Switzerland, March 16, 2026 (GLOBE NEWSWIRE) — Oculis Holding AG (Nasdaq: OCS / XICE: OCS) (Oculis), a global biopharmaceutical company focused on breakthrough innovations to address significant unmet medical needs in ophthalmology and neuro-ophthalmology, today announced the presentation of the ACUITY Phase 2 data with Privosegtor at the North American Neuro-Ophthalmology Society (NANOS) 52nd Annual Meeting.
This premier annual gathering of neuro‑ophthalmology experts worldwide provides a scientific forefront platform for presenting transformative clinical data amongst leading clinicians, researchers, and thought leaders.
As a Platinum Sponsor of the 2026 Congress, Oculis is proud to support the most influential global scientific meeting in neuro‑ophthalmology, reinforcing its commitment to advancing scientific progress in this field.
NANOS 2026
March 20-24, Boston, MA, USA
“Improved Low-Contrast Visual Acuity and Reduction in Retinal Ganglion Cell Loss with Privosegtor in Acute Optic Neuritis: Results from a Multicenter, Randomized, Placebo-Controlled, Double-Masked Trial”.
- Presenter: Professor Martin S. Zinkernagel, M.D., Ph.D., Chair of the Department of Ophthalmology at the University Hospital of Bern, Switzerland
- Date/Time: Monday March 23, 8:48 AM EDT
The presentation of the Phase 2 ACUITY trial results investigating Privosegtor in patients with optic neuritis (ON) will focus on the significant improvements in low contrast visual acuity and promising neuroprotective structural and biological effects observed in the study. Privosegtor, a peptoid small molecule with the ability to cross both the blood brain and retinal barriers, has the potential to become the first neuroprotective therapy for optic neuritis, with broad potential applicability in other neuro-ophthalmic and neurological diseases. Following Breakthrough Therapy designation from the U.S. Food and Drug Administration for ON, Oculis is advancing the PIONEER registrational program for Privosegtor in optic neuropathies, with the PIONEER-1 trial in optic neuritis initiated in Q4 2025.
Riad Sherif, M.D., Chief Executive Officer of Oculis, said “Our commitment to advancing the field of neuro‑ophthalmology has never been stronger. The ACUITY Phase 2 results with Privosegtor highlight meaningful progress toward delivering a much‑needed neuroprotective therapy to patients with optic neuritis and other optic neuropathies. The opportunity to present the groundbreaking data from this trial at NANOS, recognized as the most important global forum for neuro‑ophthalmic science, represents a significant milestone for Oculis and for patients. As a Platinum Sponsor for this year’s congress, we are honored to support and partner with the scientific community in shaping the future of neuroprotective therapies.”
Martin Zinkernagel, M.D., Ph.D., is Professor and Chair of the Department of Ophthalmology at the University Hospital of Bern, Switzerland, as well as director of the internationally renowned Bern Photographic Reading Center at the University of Bern. Professor Zinkernagel is highly recognized in the field with over 270 publications spanning his research interests in retinal imaging, ophthalmic data analysis using artificial intelligence, and the influence of the gut microbiome in retinal disease. He earned his M.D. at the University of Zurich and his PhD in ocular immunology and gene therapy from the University of Western Australia.
–END–
About Privosegtor
Privosegtor, a novel peptoid small-molecule candidate that crosses the blood-brain and retinal barriers, has the potential to become the first neuroprotective therapy for optic neuritis (ON) and other neuro-ophthalmic diseases. Positive results from the ACUITY Phase 2 trial demonstrated Privosegtor’s neuroprotective potential through anatomical preservation of the retina and improvements in visual function after an acute episode of optic neuritis. Consistent results were observed in animal models of neuroinflammation and neurodegeneration, where Privosegtor preserved retinal ganglion cell damage and was associated with improvements in mobility (clinical function disability). Privosegtor has received Breakthrough Therapy designation from the FDA and Orphan Drug designation from both the FDA and the EMA for ON and is now entering registrational trials for this indication, as well as a registrational trial in non-arteritic anterior ischemic optic neuropathy (NAION), as part of Oculis’ PIONEER (Privosegtor Investigation in Optic Neuropathies Efficacy Evaluation Research) program. In addition to its potential neuroprotective effect on the optic nerve, Privosegtor could also have wide applicability in treating other neuro-ophthalmic and neurological indications.
Privosegtor is an investigational drug and has not received regulatory approval for commercial use in any country.
About Optic Neuritis
Optic Neuritis (ON) is a rare condition characterized by an acute inflammation of the optic nerve that can lead to permanent visual impairment. It affects up to 8 in 100,000 people worldwide with a U.S. annual incidence estimated to be >30,000 and often represents the first sign of multiple sclerosis1,2. It mainly occurs in adults between the age of 20 and 40 years and is more frequent in women (2:1)3. ON is a type of neuropathy (nerve disease) that happens when acute inflammation of the optic nerve affects the signals traveling from the eyes through the brain, causing pain, vision loss and other symptoms. The cells that make up the optic nerve have a lipid protective coating called a myelin sheath, which is preferentially damaged in ON. Without myelin, the optic nerve cells can’t send signals properly and axons can be irreversibly lost. To date there is no specific therapy approved for acute optic neuritis and the unmet needs remain for therapies that can prevent vision loss after an acute episode by reducing nerve cell permanent damage or death.
About Oculis
Oculis is a global biopharmaceutical company (Nasdaq: OCS; XICE: OCS) focused on breakthrough innovations to address significant unmet medical needs in neuro-ophthalmology and ophthalmology. Oculis’ highly differentiated late-stage clinical pipeline includes three core product candidates: Privosegtor, a breakthrough neuroprotective candidate in the PIONEER program which consists of studies intended to support registration plans for treatment in optic neuropathies like optic neuritis (ON) and non-arteritic anterior ischemic optic neuropathy (NAION), with potentially broad clinical applications in various other neuro-ophthalmic and neurological diseases; OCS-01, an eye drop in pivotal registration studies, aiming to become the first non-invasive topical treatment for diabetic macular edema (DME); and Licaminlimab, a novel, topical anti-TNFα in registrational trial, which is being developed with a genotype-based approach to drive precision medicine in dry eye disease (DED). Headquartered in Switzerland with operations in the U.S. and Iceland, Oculis is led by an experienced management team with a successful track record and supported by leading international healthcare investors.
For more information, please visit: www.oculis.com
Oculis Contact
Ms. Sylvia Cheung, CFO
sylvia.cheung@oculis.com
Investor Relations
LifeSci Advisors
Corey Davis, Ph.D.
cdavis@lifesciadvisors.com
Media Relations
ICR Healthcare
Amber Fennell / David Daley / Sean Leous
oculis@icrhealthcare.com
Cautionary Statement Regarding Forward Looking Statements
This press release contains forward-looking statements and information. For example, statements regarding the potential benefits of the Company’s product candidates, the initiation, timing, progress and results of current and future clinical trials, Oculis’ research and development programs, regulatory and business strategy; and Oculis’ future development plans, are forward-looking. All forward-looking statements are based on estimates and assumptions that, while considered reasonable by Oculis and its management, are inherently uncertain and are inherently subject to risks, variability, and contingencies, many of which are beyond Oculis’ control. These forward-looking statements are provided for illustrative purposes only and are not intended to serve as, and must not be relied on by an investor as, a guarantee, assurance, prediction or definitive statement of a fact or probability. Actual events and circumstances are difficult or impossible to predict and will differ from assumptions. All forward-looking statements are subject to risks, uncertainties and other factors that may cause actual results to differ materially from those that we expected and/or those expressed or implied by such forward-looking statements. Forward-looking statements are subject to numerous conditions, many of which are beyond the control of Oculis, including those set forth in the Risk Factors section of Oculis’ annual report on Form 20-F and any other documents filed with the SEC. Copies of these documents are available on the SEC’s website, www.sec.gov. Oculis undertakes no obligation to update these statements for revisions or changes after the date of this release, except as required by law.
References:
- Martínez-Lapiscina EH, et al. (2014): Is the incidence of optic neuritis rising? Evidence from an epidemiological study in Barcelona (Spain) 2008-2012. J Neurol. 2014 Apr; 261(4): 759-767.
- Weidong Gu et al. (2023) Incidence of Optic Neuritis and the Associated Risk of Multiple Sclerosis for Service Members of U.S. Armed Forces, Military Medicine, vol. 188, March/April 2023
- Guier CP, Kaur K, Stokkermans TJ. Optic Neuritis. January 2025. StatPearls. https://www.ncbi.nlm.nih.gov/books/NBK557853
Oculis to Present at Upcoming North American Neuro-Ophthalmology Society Annual Meeting
Oculis to Present at Upcoming North American Neuro-Ophthalmology Society Annual Meeting
Oculis to Present at Upcoming North American Neuro-Ophthalmology Society Annual Meeting

ZUG, Switzerland, 16 March, 2026 — Oculis Holding AG (Nasdaq: OCS / XICE: OCS) (Oculis), a global biopharmaceutical company focused on breakthrough innovations to address significant unmet medical needs in ophthalmology and neuro-ophthalmology, today announced the presentation of the ACUITY Phase 2 data with Privosegtor at the North American Neuro-Ophthalmology Society (NANOS) 52nd Annual Meeting.
This premier annual gathering of neuro-ophthalmology experts worldwide provides a scientific forefront platform for presenting transformative clinical data amongst leading clinicians, researchers, and thought leaders.
As a Platinum Sponsor of the 2026 Congress, Oculis is proud to support the most influential global scientific meeting in neuro-ophthalmology, reinforcing its commitment to advancing scientific progress in this field.
NANOS 2026
March 20-24, Boston, MA, USA
“Improved Low-Contrast Visual Acuity and Reduction in Retinal Ganglion Cell Loss with Privosegtor in Acute Optic Neuritis: Results from a Multicenter, Randomized, Placebo-Controlled, Double-Masked Trial”.
- Presenter: Professor Martin S. Zinkernagel, M.D., Ph.D., Chair of the Department of Ophthalmology at the University Hospital of Bern, Switzerland
- Date/Time: Monday March 23, 8:48 AM EDT
The presentation of the Phase 2 ACUITY trial results investigating Privosegtor in patients with optic neuritis (ON) will focus on the significant improvements in low contrast visual acuity and promising neuroprotective structural and biological effects observed in the study. Privosegtor, a peptoid small molecule with the ability to cross both the blood brain and retinal barriers, has the potential to become the first neuroprotective therapy for optic neuritis, with broad potential applicability in other neuro-ophthalmic and neurological diseases. Following Breakthrough Therapy designation from the U.S. Food and Drug Administration for ON, Oculis is advancing the PIONEER registrational program for Privosegtor in optic neuropathies, with the PIONEER-1 trial in optic neuritis initiated in Q4 2025.
Riad Sherif, M.D., Chief Executive Officer of Oculis, said “Our commitment to advancing the field of neuro-ophthalmology has never been stronger. The ACUITY Phase 2 results with Privosegtor highlight meaningful progress toward delivering a much-needed neuroprotective therapy to patients with optic neuritis and other optic neuropathies. The opportunity to present the groundbreaking data from this trial at NANOS, recognized as the most important global forum for neuro-ophthalmic science, represents a significant milestone for Oculis and for patients. As a Platinum Sponsor for this year’s congress, we are honored to support and partner with the scientific community in shaping the future of neuroprotective therapies.”
Martin Zinkernagel, M.D., Ph.D., is Professor and Chair of the Department of Ophthalmology at the University Hospital of Bern, Switzerland, as well as director of the internationally renowned Bern Photographic Reading Center at the University of Bern. Professor Zinkernagel is highly recognized in the field with over 270 publications spanning his research interests in retinal imaging, ophthalmic data analysis using artificial intelligence, and the influence of the gut microbiome in retinal disease. He earned his M.D. at the University of Zurich and his PhD in ocular immunology and gene therapy from the University of Western Australia.
–END–
About Privosegtor
Privosegtor, a novel peptoid small-molecule candidate that crosses the blood-brain and retinal barriers, has the potential to become the first neuroprotective therapy for optic neuritis (ON) and other neuro-ophthalmic diseases. Positive results from the ACUITY Phase 2 trial demonstrated Privosegtor’s neuroprotective potential through anatomical preservation of the retina and improvements in visual function after an acute episode of optic neuritis. Consistent results were observed in animal models of neuroinflammation and neurodegeneration, where Privosegtor preserved retinal ganglion cell damage and was associated with improvements in mobility (clinical function disability). Privosegtor has received Breakthrough Therapy designation from the FDA and Orphan Drug designation from both the FDA and the EMA for ON and is now entering registrational trials for this indication, as well as a registrational trial in non-arteritic anterior ischemic optic neuropathy (NAION), as part of Oculis’ PIONEER (Privosegtor Investigation in Optic Neuropathies Efficacy Evaluation Research) program. In addition to its potential neuroprotective effect on the optic nerve, Privosegtor could also have wide applicability in treating other neuro-ophthalmic and neurological indications.
Privosegtor is an investigational drug and has not received regulatory approval for commercial use in any country.
About Optic Neuritis
Optic Neuritis (ON) is a rare condition characterized by an acute inflammation of the optic nerve that can lead to permanent visual impairment. It affects up to 8 in 100,000 people worldwide with a U.S. annual incidence estimated to be >30,000 and often represents the first sign of multiple sclerosis1,2. It mainly occurs in adults between the age of 20 and 40 years and is more frequent in women (2:1)3. ON is a type of neuropathy (nerve disease) that happens when acute inflammation of the optic nerve affects the signals traveling from the eyes through the brain, causing pain, vision loss and other symptoms. The cells that make up the optic nerve have a lipid protective coating called a myelin sheath, which is preferentially damaged in ON. Without myelin, the optic nerve cells can’t send signals properly and axons can be irreversibly lost. To date there is no specific therapy approved for acute optic neuritis and the unmet needs remain for therapies that can prevent vision loss after an acute episode by reducing nerve cell permanent damage or death.
About Oculis
Oculis is a global biopharmaceutical company (Nasdaq: OCS; XICE: OCS) focused on breakthrough innovations to address significant unmet medical needs in neuro-ophthalmology and ophthalmology. Oculis’ highly differentiated late-stage clinical pipeline includes three core product candidates: Privosegtor, a breakthrough neuroprotective candidate in the PIONEER program which consists of studies intended to support registration plans for treatment in optic neuropathies like optic neuritis (ON) and non-arteritic anterior ischemic optic neuropathy (NAION), with potentially broad clinical applications in various other neuro-ophthalmic and neurological diseases; OCS-01, an eye drop in pivotal registration studies, aiming to become the first non-invasive topical treatment for diabetic macular edema (DME); and Licaminlimab, a novel, topical anti-TNFα in registrational trial, which is being developed with a genotype-based approach to drive precision medicine in dry eye disease (DED). Headquartered in Switzerland with operations in the U.S. and Iceland, Oculis is led by an experienced management team with a successful track record and supported by leading international healthcare investors.
For more information, please visit: www.oculis.com
Oculis Contact
Ms. Sylvia Cheung, CFO
sylvia.cheung@oculis.com
Investor Relations
LifeSci Advisors
Corey Davis, Ph.D.
cdavis@lifesciadvisors.com
Media Relations
ICR Healthcare
Amber Fennell / David Daley / Sean Leous
oculis@icrhealthcare.com
Cautionary Statement Regarding Forward Looking Statements
This press release contains forward-looking statements and information. For example, statements regarding the potential benefits of the Company’s product candidates, the initiation, timing, progress and results of current and future clinical trials, Oculis’ research and development programs, regulatory and business strategy; and Oculis’ future development plans, are forward-looking. All forward-looking statements are based on estimates and assumptions that, while considered reasonable by Oculis and its management, are inherently uncertain and are inherently subject to risks, variability, and contingencies, many of which are beyond Oculis’ control. These forward-looking statements are provided for illustrative purposes only and are not intended to serve as, and must not be relied on by an investor as, a guarantee, assurance, prediction or definitive statement of a fact or probability. Actual events and circumstances are difficult or impossible to predict and will differ from assumptions. All forward-looking statements are subject to risks, uncertainties and other factors that may cause actual results to differ materially from those that we expected and/or those expressed or implied by such forward-looking statements. Forward-looking statements are subject to numerous conditions, many of which are beyond the control of Oculis, including those set forth in the Risk Factors section of Oculis’ annual report on Form 20-F and any other documents filed with the SEC. Copies of these documents are available on the SEC’s website, www.sec.gov. Oculis undertakes no obligation to update these statements for revisions or changes after the date of this release, except as required by law.
References:
- Martínez-Lapiscina EH, et al. (2014): Is the incidence of optic neuritis rising? Evidence from an epidemiological study in Barcelona (Spain) 2008-2012. J Neurol. 2014 Apr; 261(4): 759-767.
- Weidong Gu et al. (2023) Incidence of Optic Neuritis and the Associated Risk of Multiple Sclerosis for Service Members of U.S. Armed Forces, Military Medicine, vol. 188, March/April 2023
- Guier CP, Kaur K, Stokkermans TJ. Optic Neuritis. January 2025. StatPearls. https://www.ncbi.nlm.nih.gov/books/NBK557853
Dvorak Botanicals Launches Low-Dose Hemp-Derived THC Gummies Focused on Mindful Consumption
Dvorak Botanicals Launches Low-Dose Hemp-Derived THC Gummies Focused on Mindful Consumption
Dvorak Botanicals Launches Low-Dose Hemp-Derived THC Gummies Focused on Mindful Consumption
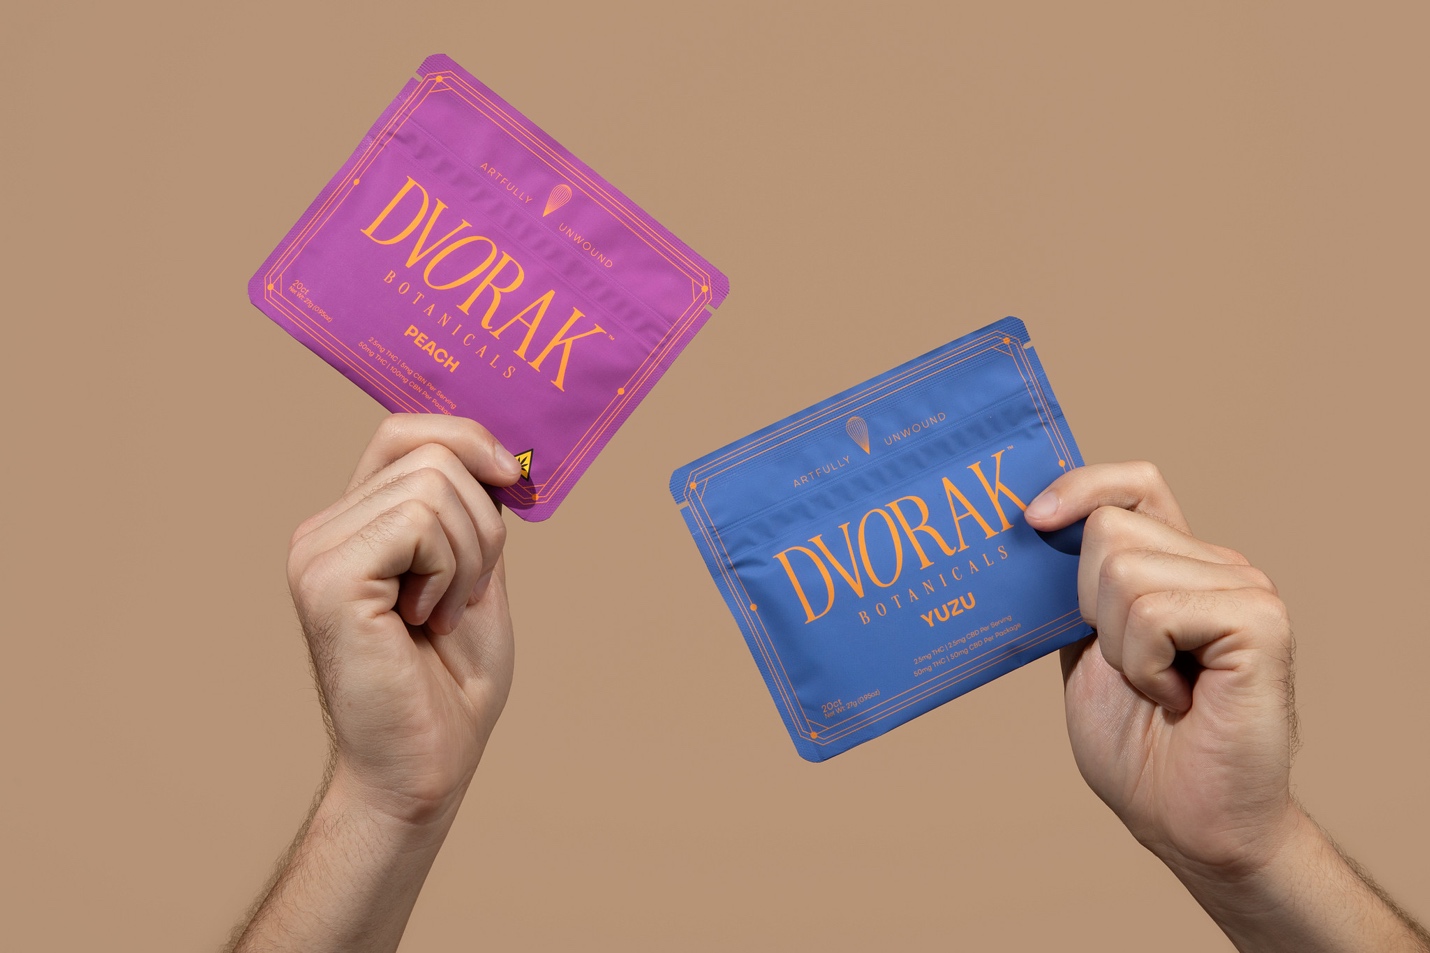
Atlanta, GA, March 16, 2026 (GLOBE NEWSWIRE) — Dvorak Botanicals, a new premium hemp-derived THC brand founded by Atlanta-based entrepreneur Dane Dvorak, has launched with a focus on controlled-dose hemp products that support more intentional and balanced ways for adults to relax.
Dvorak Botanicals’ 2.5mg hemp-derived THC gummies in Yuzu and Peach, designed for a controlled, low-dose experience.
The company’s initial product line features low-dose hemp-derived Delta-9 THC gummies. Each gummy contains 2.5 mg of THC, allowing users to control their experience through gradual dosing. The gummies are packed in 20-count containers and formulated with premium flavor profiles such as yuzu and peach, reflecting the company’s emphasis on elevated ingredients and approachable formulations.
The launch comes as consumer interest in mindful consumption continues to grow. Many adults are exploring alternatives to traditional alcohol-centered social experiences or high-potency cannabis products. By focusing on controlled, low-dose formats, Dvorak Botanicals aims to offer a more approachable option for adults looking to unwind, connect socially, and incorporate cannabis into their routines.
“Many cannabis products are designed for intensity, but not necessarily for balance,” said Dane Dvorak, founder of Dvorak Botanicals. “We wanted to create something that allows adults to ease into the experience, control their dosage, and enjoy THC thoughtfully.”
All Dvorak Botanicals products undergo third-party laboratory testing for potency and purity to help ensure product consistency and transparency for consumers. The gummies are formulated using hemp-derived Delta-9 THC in accordance with federal hemp regulations.
In addition to direct-to-consumer sales through the company’s website, Dvorak Botanicals products are currently carried by select independent boutiques across Georgia, including Soberish, Ghost Apothecary, and The Zero Co. The company partners with retailers that align with its design-focused and wellness-oriented brand philosophy and is actively expanding its retail presence through the region. Dvorak Botanicals plans to grow into a broader wellness platform built around intentional relaxation and thoughtful dosing practices.
“Our focus starts with cannabinoids,” Dvorak added, “but the larger goal is to build a brand people trust for premium products and more mindful ways to relax.”
To learn more, please visit https://dvorakbotanicals.com.
About Dvorak Botanicals
Dvorak Botanicals is a modern hemp-derived THC brand created for adults exploring more balanced ways to relax and socialize. The company combines thoughtful dosing with elevated product design to create approachable cannabinoid experiences. Dvorak Botanicals is focused on building a lifestyle brand that supports mindful consumption and contemporary wellness habits.
Media Contact
Company Name: Dvorak Botanicals
Contact Person: Dane Dvorak
Contact Number: 404-436-2280
Email: support@dvorakbotanicals.com
Country: United States
Website: https://dvorakbotanicals.com
Social Media Handles: @Dvorak.botanicals
CONTACT: Media Contact
Company Name: Dvorak Botanicals
Contact Person: Dane Dvorak
Contact Number: 404-436-2280
Email: support@dvorakbotanicals.com
Country: United States
Website: https://dvorakbotanicals.com
Social Media Handles: @Dvorak.botanicals

GN Store Nord A/S enters into agreement to sell its Hearing business to Amplifon S.p.A. for DKK 17.0 billion
GN Store Nord A/S enters into agreement to sell its Hearing business to Amplifon S.p.A. for DKK 17.0 billion
GN Store Nord A/S enters into agreement to sell its Hearing business to Amplifon S.p.A. for DKK 17.0 billion

Today, GN Store Nord A/S (“GN”) has entered into a definitive agreement for the sale of GN’s Hearing business to Amplifon S.p.A. (“Amplifon”), a leading global hearing care retailer. The transaction values the Hearing business at DKK 17.0 billion1 on a cash-free and debt-free basis and will be settled with DKK 12.6 billion2 in cash and 56 million shares in Amplifon. The transaction is subject to customary regulatory approvals and completion of the Hearing business separation from GN and is expected to complete by the end of 2026.
Amplifon and GN will together create a global integrated leader in audiology combining hearing technology leadership with high-quality, innovative hearing care to offer comprehensive solutions to both audiology professionals and patients. The combination will create the industry leader in hearing care solutions and services worldwide with benefits to audiologists, consumers, and investors.
Following the transaction, GN will fully focus to drive success in the large and attractive audio and video peripherals markets. GN is well positioned with leading brands, premium product differentiation, and technology leadership in sound processing and low-power edge AI. Financially, GN will drive margin expansion supported by the product differentiation and its scalable and agile operations setup, while the asset-light business model will support healthy cash flows.
As the proceeds from the transaction is larger than GN’s net interest-bearing debt, GN expects to use the proceeds to 1) significantly reduce debt, 2) invest in the ongoing business to support sustainable and profitable growth as well as 3) returning capital to shareholders, subject to applicable law and in line with the dividend policy.
GN Store Nord Chair Jukka Pekka Pertola said:
We are excited to transform GN into a pure technology company. We have unique positions and capabilities making us well positioned to further grow in the large audio and video peripherals market in a focused way. While we will miss all of our great employees in Hearing, we are proud that we, together with Amplifon, will create an industry leading player in global audiology. We are convinced that the transaction and steps ahead will be beneficial to our customers, employees and investors.
GN Store Nord CEO Peter Karlströmer said:
We are today acting on an opportunity to create a global leader in audiology together with Amplifon. The new company has every chance to develop into an industry leader driving further innovation and success in hearing care. For GN, this also creates an opportunity to further strengthen our position in the large and attractive audio and video peripherals markets. We have unique brands, talents, and capabilities and a rich set of opportunities. We have in the last few years strengthened GN significantly financially and operationally. As we open up this new chapter, we are focused on driving success as an innovative technology company with global reach and scale. This is very exciting for GN’s customers, employees, and investors.
Amplifon CEO Enrico Vita said: “Today marks a turning point for Amplifon. We are announcing the most transformative acquisition in our 75-year-long history, that will fundamentally change the future of the hearing care industry worldwide and create significant and long-term value for all our stakeholders. With combined revenues of approximately €3.3 billion, a presence in more than 100 countries, a cutting-edge R&D platform, leading manufacturing capabilities, and an unparalleled commercial network, we will be forming a truly global vertically integrated leader in audiology. Importantly, Amplifon will continue to preserve and enhance Hearing’s unique identity, heritage, and strengths, recognizing its brand value, culture, and distinct capabilities as key contributors to the future success of the combined organization. We look forward to working closely with the exceptional team at Hearing to improve hearing health for millions of people globally”
Background
Over the past years, GN has significantly strengthened its strategic focus and execution capabilities across the group, supported by substantial investments in innovation, technology, and its supply chain. These efforts have created a solid foundation for sustainable and profitable growth. During this period, the Board and Executive Management received several unsolicited expressions of interest, including from Amplifon, and subsequently engaged in exploratory discussions. One of these led to Amplifon making a financially and strategically compelling proposal with a high degree of deal certainty. After extensive deliberation and a thorough evaluation process, the Board and Executive Management unanimously decided to pursue and accept the proposal from Amplifon.
Transaction Highlights
- Consideration: DKK 17.0 billion3 on a cash-free and debt-free basis, comprised of DKK 12.64 billion in cash (subject to customary true up closing mechanism adjustments) and 56 million shares in Amplifon, allowing GN to retain upside to the strong strategic value proposition of the transaction. Once the transaction closes, GN will hold approximately 16%5 of the current outstanding share capital in Amplifon, subject to customary lock-up
- Hearing Perimeter: Includes the entire Hearing business comprising of c. 5,500 employees globally, the ReSound, Beltone and other hearing brands, all hearing product IP, R&D, manufacturing and operations (including relevant sites used by the Hearing business), and Beltone network partnerships. The transaction does not include the current financial investment in Nations Benefits LLC. In 2025, the Hearing business generated revenue of DKK 7,214 million and reported pro-forma EBITDA of DKK 1,185 million6.
- Shareholders’ agreement: At closing, GN and Ampliter S.r.I, Amplifon’s controlling shareholder, are expected to enter into a shareholders’ agreement providing for GN’s right to propose a representative for appointment to Amplifon’s Board of Directors, together with customary transfer restrictions and lock-up undertakings in respect of the Amplifon shares received by GN as consideration. Additionally, Ampliter S.r.l., and core shareholder, Tamburi Investment Partners S.P.A., confirmed their financial support to the Transaction, including investment in a future equity raise conducted by Amplifon.
- Carve-out: Prior to closing of the transaction, the Hearing business will be separated from GN through a carve-out, including a statutory demerger of the Hearing business in accordance with the Danish Companies Act. The de-merger is expected to take place as a taxable transaction.
- Transitional Arrangements: Certain shared services and support functions by GN will continue to be provided on a temporary basis following the closing of the transaction via customary transitional services agreements between the parties.
- Fairness Opinions: J.P. Morgan Securities plc has provided to the Board of Directors of GN an opinion as of 16 March 2026 based on the factors, assumption and procedures specified therein, which confirms that the transaction value is fair from a financial point of view (the “Fairness Opinion”). J.P. Morgan provided its Fairness Opinion solely for the benefit and information of the Board of Directors of GN in its evaluation of the transaction, and the Fairness Opinion may not be relied upon by any third party or used for any other purpose. Additionally, Nordea Corporate Finance, part of Nordea Danmark, Filial af Nordea Bank Abp, Finland (“Nordea”), has as Independent Financial Advisor, provided to the Board of Directors of GN an opinion as of 16 March 2026 based on the factors, assumptions and procedures specified therein, which confirms that the transaction value is fair for GN from a financial point of view (the “Independent Fairness Opinion”). Nordea provided its Independent Fairness Opinion solely for the benefit and information of the Board of Directors of GN Store Nord A/S in its evaluation of the transaction, and the Independent Fairness Opinion may not be relied upon by any third party or used for any other purpose.
- Closing conditions: The transaction is subject to customary conditions, including merger control and other regulatory approvals from relevant authorities and completion of the Hearing business separation from GN.
- Anticipated Closing Date: The transaction is expected to close by the end of 2026. GN remains committed to serving the customers of the Hearing business and will continue delivering high-quality, innovative products.
Financial implications and guidance
As a direct consequence of the transaction, Hearing will be classified as discontinued operations, and associated assets and liabilities as held-for-sale.
GN’s financial guidance for 2026 now excludes discontinued operations and therefore only reflects Enterprise and Gaming. Consequently, the organic revenue guidance is now expected to be 2-8%, based on unchanged divisional assumptions of 0-6% organic revenue growth in Enterprise, and 7-13% organic revenue growth in Gaming.
To drive the necessary changes of this significant transaction, GN will incur one-off costs, including but not limited to transaction and carve-out costs, which will impact group EBITA in 2026. Following the establishment of a standalone operating system structure, the company is expected to re-introduce a profitability guidance.
| Financial guidance 2026 | Organic revenue growth |
| GN | 2% to 8% |
The current long-term financial targets are suspended; however, the company’s ambition to drive strong profitable growth remains unchanged. GN is in a strong position to deliver healthy growth, strong profitability and an attractive cash conversion.
Capital Markets Day
GN intends to hold a Capital Markets Day post-closing of the transaction. The Capital Markets Day will cover among other subjects the future strategy, long-term financial targets, capital allocation policy, including further clarifications on distributing excess capital to shareholders.
Press Conference
GN will host a press conference at 09:30 CET on 16 March 2026.
Contact Group Media Manager Helge Coroli Frandsen at hcfrandsen@gn.com to sign up.
Investor and analyst conference call
GN will hold will host a teleconference at 12:00 CET on 16 March 2026. Please visit www.gn.com/investor to access the teleconference. Presentation material will be available on the website prior to the start of the teleconference.
Advisors
J.P. Morgan Securities plc is acting as exclusive financial advisor, and Gorrissen Federspiel Advokatpartnerselskab and Gianni & Origoni are acting as legal advisors, and PricewaterhouseCoopers has provided due diligence services to GN.
This announcement contains inside information within the meaning of Article 7 of Regulation (EU) No 596/2014 (the Market Abuse Regulation).
For further information, please contact:
Investor Relations
Rune Sandager +45 45 75 92 57
Media Relations
Steen Frentz Laursen +45 20 65 34 20
About GN
GN brings people closer through our leading intelligent hearing, audio, video, and gaming solutions. Inspired by people and driven by innovation, we deliver technologies that enhance the senses of hearing and sight. We help people with hearing loss overcome real-life challenges, improve communication and collaboration for businesses, and provide great experiences for audio and gaming enthusiasts. GN was founded more than 150 years ago with a vision to connect the world. Today, inspired by our strong heritage, GN touches more lives than ever with our unique expertise and the broadest portfolio of products and services in our history – bringing people closer to what is important to them.
We market our solutions with the brands Jabra, ReSound, SteelSeries, Beltone, Interton, BlueParrott, Danavox, and FalCom in 100 countries. Founded in 1869, GN Group employs more than 7,500 people and is listed on Nasdaq Copenhagen (GN.CO).
Visit our homepage GN.com – and connect with us on LinkedIn, Facebook and Twitter
About Amplifon
Amplifon, global leader in the hearing care retail market, empowers people to rediscover all the emotions of sound. Amplifon’s around 20,600 people worldwide strive every day to understand the unique needs of every customer, delivering exclusive, innovative and highly personalized products and services, to ensure everyone the very best solution and outstanding experience. The Group, with annual revenues of around 2.4 billion euros, operates through a network of around 10,000 locations in 25 Countries and 5 continents. More information about the Group is available at: https://corporate.amplifon.com.
The forward-looking statements in this announcement reflect the management’s current expectations of certain future events and financial results. Statements regarding the future are, naturally, subject to risks and uncertainties, which may result in considerable deviations from the outlook set forth. Furthermore, some of these expectations are based on assumptions regarding future events, which may prove incorrect. Changes to such expectations and assumptions will not be disclosed on an ongoing basis, unless required pursuant to general disclosure obligations to which GN is subject.
Factors that may cause actual results to deviate materially from expectations include – but are not limited to – general economic developments and developments in the financial markets as well as foreign exchange rates, risks related to the above mentioned transaction, technological developments, changes and amendments to legislation and regulations governing GN markets, changes in the demand for GN products, competition, fluctuations in sub-contractor supplies, and developments in ongoing litigation (including but not limited to class action and patent infringement litigation in the United States).
For more information, please see the “Management’s report” and “Risk management” sections in the Annual Report. This announcement should not be considered an offer to sell or buy securities in.
1 Share component value based on €10.52 per Amplifon share as of 13 March 2026
2 Cash component of EUR 1,690m converted to DKK at DKK/EUR 0.134
3 Share component value based on €10.52 per Amplifon share as of 13 March 2026
4 Cash component of EUR 1,690m converted to DKK at DKK/EUR 0.134
5 Assumes Amplifon completes up to EUR 0.75bn equity raise as per Amplifon’s communication to the market. Should Amplifon not complete any equity raise, GN’s shareholding at closing would amount to 19.8%
6 EBITDA includes group cost allocations and certain carve-out costs that the buyer will not take over, as well as other transaction adjustments of around DKK 230 million in total
RevnaBio Secures Triple International Laboratory Accreditation to Expand Precision Medicine and Clinical Research Infrastructure in Africa
RevnaBio Secures Triple International Laboratory Accreditation to Expand Precision Medicine and Clinical Research Infrastructure in Africa
RevnaBio Secures Triple International Laboratory Accreditation to Expand Precision Medicine and Clinical Research Infrastructure in Africa
A2LA accreditation across ISO 15189, ISO 20387, and ISO/IEC 17043 strengthens diagnostic quality for patients while enabling global pharmaceutical and research partnerships.
ACCRA, Ghana–(BUSINESS WIRE)–#A2LA–RevnaBio today announced that it has received triple accreditation from the American Association for Laboratory Accreditation (A2LA), validating its laboratory and biobanking operations under three internationally recognized standards: ISO 15189 for medical laboratories, ISO 20387 for biobanking, and ISO/IEC 17043 for proficiency testing providers.
The combined accreditation confirms the strength of RevnaBio’s integrated quality systems across clinical diagnostics, biospecimen management, and laboratory performance monitoring. These capabilities support the delivery of reliable molecular testing for patients while providing a trusted platform for pharmaceutical companies, diagnostics developers, and research organizations conducting biomedical research in Africa.
The milestone represents a significant step toward expanding precision medicine infrastructure in West Africa and strengthening access to internationally accredited diagnostic services.
Improving Diagnostic Quality for Patients
Accurate and reliable diagnostics are essential for effective treatment selection, disease monitoring, and improved clinical outcomes.
Through its ISO 15189–accredited medical laboratory, RevnaBio provides molecular and genomic testing systems that meet internationally recognized standards for quality, accuracy, and traceability. These capabilities help clinicians make better-informed treatment decisions while expanding access to advanced molecular testing that historically required samples to be sent overseas.
By strengthening local diagnostic capacity, RevnaBio contributes to improving healthcare delivery and patient outcomes across Ghana.
Building Research Infrastructure for the Region
RevnaBio’s ISO 20387 biobanking accreditation validates the company’s governance and operational standards for collecting, processing, storing, and managing research-grade biospecimens.
These systems ensure that biological samples are linked to high-quality clinical and molecular data, supporting scientific research aimed at improving understanding of diseases affecting African populations and people of African descent globally.
Strengthening biospecimen infrastructure within Africa allows research involving African populations to be conducted with greater scientific rigor, representation, and ethical oversight.
Advancing Laboratory Quality Across West Africa
RevnaBio’s ISO/IEC 17043 accreditation authorizes the company to design and deliver accredited proficiency testing programs that help laboratories measure and improve their performance.
In regions where locally delivered external quality assessment programs remain limited, this capability enables RevnaBio to support laboratory networks through standardized benchmarking and quality monitoring.
Expanding access to accredited proficiency testing strengthens diagnostic reliability across healthcare systems, ultimately benefiting patients, clinicians, and public health programs throughout the region.
Supporting Global Biomedical Research and Drug Development
The accreditation also strengthens RevnaBio’s ability to collaborate with biopharmaceutical companies, diagnostics developers, and international research institutions seeking high-quality laboratory and biospecimen infrastructure in Africa.
Accredited diagnostics, validated biobanking systems, and reliable laboratory quality monitoring are essential for enabling biomarker discovery, pharmacogenomic research, and clinical studies involving diverse populations.
RevnaBio’s integrated platform helps facilitate research that advances precision medicine while ensuring that African populations are appropriately represented in global biomedical research.
Leadership Perspective
Commenting on the milestone, Dr. Derrick Akpalu, CEO and Co-Founder of RevnaBio, said: “This accreditation reflects our commitment to building laboratory systems that meet the highest international standards while serving the needs of patients and clinicians in Ghana. By strengthening diagnostic quality, biospecimen infrastructure, and laboratory performance monitoring, we aim to support improved healthcare locally while enabling meaningful collaboration with global research partners working to advance precision medicine.”
Ms. Jennifer Dent, a board member of the company, remarked, “I am incredibly proud to see the company achieve this important milestone. Receiving triple accreditation from the American Association for Laboratory Accreditation reflects RevnaBio’s commitment to quality, scientific excellence, and global laboratory best practices. The triple accreditation reflects the team’s commitment to quality, scientific excellence, and global standards. It also strengthens partnerships and supports African scientists and clinicians in leading impactful research.”
About Revna Biosciences (RevnaBio)
Revna Biosciences (RevnaBio) is a precision medicine company with an operational base in Accra, Ghana that provides genomic testing, clinical research services, and biobanking infrastructure across Africa. Founded in 2017 and domiciled in Delaware, RevnaBio partners with hospitals, research institutions, and global pharmaceutical companies to generate high-quality genomic and clinical datasets that reflect Africa’s genetic diversity, an essential foundation for precision-medicine discovery and drug development.
The company operates advanced laboratory and biospecimen infrastructure that supports molecular diagnostics, biomarker research, and clinical research programs across the region. Through these capabilities, RevnaBio enables pharmaceutical partners and academic researchers to conduct high-quality studies and develop therapies informed by African genomic data.
RevnaBio holds triple accreditation from the American Association for Laboratory Accreditation (A2LA) to ISO 15189:2022 (Medical Laboratories), ISO 20387:2018 (Biobanking), and ISO/IEC 17043 (Proficiency Testing Providers). This rare combination of accreditations confirms adherence to internationally recognized standards for diagnostic accuracy, biospecimen governance, and laboratory quality systems.
By building trusted biomedical infrastructure and generating globally relevant genomic data, RevnaBio is helping expand Africa’s role in precision medicine, clinical research, and pharmaceutical innovation.
Contacts
Media Contact
Revna Biosciences
Communications Office
Email: info@revnabio.com
Phone: +233-59-161-2549 / +233-59-900-9977
Website: www.revnabio.com
KERENDIA® (finerenone) Meets Primary Endpoint in Investigational Phase III FIND-CKD Study in Patients with Non-Diabetic Chronic Kidney Disease
KERENDIA® (finerenone) Meets Primary Endpoint in Investigational Phase III FIND-CKD Study in Patients with Non-Diabetic Chronic Kidney Disease
KERENDIA® (finerenone) Meets Primary Endpoint in Investigational Phase III FIND-CKD Study in Patients with Non-Diabetic Chronic Kidney Disease
- KERENDIA® (finerenone) met its primary endpoint demonstrating a statistically significant improvement vs. placebo in the estimated glomerular filtration rate (eGFR) slope from baseline to Month 32 – a surrogate endpoint for slowing kidney disease progression1
- FIND-CKD is the fifth consecutive Phase III clinical trial where KERENDIA met its primary endpoint, adding to a clinical trial program of more than 20,000 patients across multiple patient populations with heart and kidney diseases
- FIND-CKD is the largest Phase III study to date focused on non-diabetic chronic kidney disease (CKD) and now expands KERENDIA’s clinical data in CKD to both diabetic and non-diabetic patients
- The clinical data from FIND-CKD will be presented at an upcoming scientific conference, and Bayer anticipates submitting the data to the U.S. Food and Drug Administration (FDA) to extend the indication of KERENDIA to this patient population
- KERENDIA is currently approved by the FDA for use in adults with CKD associated with type 2 diabetes (T2D) and heart failure with left ventricular ejection fraction (HF LVEF) ≥40%2
WHIPPANY, N.J.–(BUSINESS WIRE)–The Phase III study FIND-CKD (NCT05047263) — investigating the efficacy and safety of KERENDIA® (finerenone) versus placebo when added to standard of care in adult patients with non-diabetic chronic kidney disease (CKD) — has met its primary endpoint.1 The results demonstrated a statistically significant improvement versus placebo in the primary efficacy outcome of estimated glomerular filtration rate (eGFR) slope, defined as the mean annual rate of change from baseline to Month 32,1 a validated surrogate endpoint for kidney disease progression.3 The safety profile of KERENDIA in the FIND-CKD study was consistent with its established safety profile.1
The FIND-CKD clinical trial data will be presented at an upcoming scientific conference. Bayer anticipates submitting the data to the U.S. Food and Drug Administration (FDA) to extend the indication of KERENDIA to non-diabetic CKD patients.
“Patients with chronic kidney disease have substantial risk for cardiovascular events and kidney failure, so new treatments are needed to help slow kidney disease progression and improve outcomes,” said Hiddo L. Heerspink, Professor of Clinical Trials and Personalized Medicine, clinical trialist at the Department of Clinical Pharmacy and Pharmacology at the University Medical Center Groningen, Netherlands, and Co-Chair of the study’s Executive Committee. “The FIND-CKD topline results are encouraging because they now provide evidence for finerenone in a non-diabetic chronic kidney disease population, on top of its established evidence in diabetic chronic kidney disease.”
Since 2021, KERENDIA has been approved to reduce the risk of cardiovascular death, hospitalization for heart failure (HF), non-fatal myocardial infarction, sustained eGFR decline, and end-stage kidney disease in adult patients with CKD associated with type 2 diabetes (T2D). In July 2025, KERENDIA also received FDA approval for the treatment of heart failure with left ventricular ejection fraction (HF LVEF) ≥40%.2
Approximately 850 million people worldwide are living with CKD, and those with non-diabetic CKD represent more than half of these cases.4,5,6 In the U.S., more than 35 million people are estimated to have CKD.7 In 2023, CKD accounted for over 1.4 million deaths, ranking as the ninth leading cause of death.8
“The FIND‑CKD findings mark the fifth consecutive Phase III trial in the KERENDIA clinical development program to meet its primary endpoint and represent a major milestone for people living with non-diabetic chronic kidney disease,” said Carolina Aldworth, M.D., MSc, Executive Medical Director at Bayer. “When considered alongside the growing evidence base, this important trial adds to our understanding of KERENDIA across multiple patient populations with heart and kidney diseases.”
KERENDIA is a non-steroidal mineralocorticoid receptor antagonist (nsMRA) that selectively and potently blocks mineralocorticoid receptor overactivation in the heart and kidneys.2 FIND-CKD is the largest Phase III study to date focused on non-diabetic CKD and investigated KERENDIA in a population spanning different etiologies of non-diabetic CKD.
About FIND-CKD
The Phase III FIND-CKD9 study investigated finerenone compared to placebo in addition to standard of care in more than 1,500 patients with non-diabetic CKD etiologies, of which etiologies included hypertension and chronic glomerulonephritis (inflammation of the kidneys’ blood filters). Patients were randomized to receive either finerenone 10mg or 20mg, based on serum potassium levels and eGFR, or placebo on top of individually tolerated maximum labeled doses of a renin-angiotensin system (RAS)-blocking therapy such as an angiotensin-converting enzyme (ACE) inhibitor or an angiotensin II receptor blocker (ARB). The primary endpoint was the mean annual rate of change in eGFR from baseline to 32 months. The safety endpoints were the occurrence of treatment-emergent adverse events (AEs), treatment-emergent serious AEs, and hyperkalemia AEs.
About KERENDIA’s Clinical Trial Program
KERENDIA’s clinical trial program—called FINEOVATE—currently comprises 10 Phase III studies with dedicated programs in HF (MOONRAKER) and CKD (THUNDERBALL). The MOONRAKER program includes FINEARTS-HF10 as well as the ongoing, collaborative, investigator-sponsored studies REDEFINE-HF,11 CONFIRMATION-HF12 and FINALITY-HF.13 The THUNDERBALL CKD program consists of the completed studies FIDELIO-DKD,14 FIGARO-DKD,15 FINE-ONE16 and FIND-CKD17 as well as the ongoing investigational studies, FIONA18 and FIONA-OLE.19
About Chronic Kidney Disease
CKD is a common and potentially deadly condition that is widely underrecognized. CKD progresses silently and unpredictably, with many symptoms not appearing until the disease is well-advanced. CKD affects 850 million people worldwide. In the U.S., 1 in 3 adults is at risk for the disease. At advanced stages of CKD, patients may need dialysis or a kidney transplant to stay alive. Healthy kidneys act as the body’s filter, removing waste products from the blood. They also control how much water and electrolytes are in the body, regulating blood pressure. As kidney function goes down, patients may experience a range of symptoms including leg swelling, tiredness in the day, nausea, muscle cramps, joint pain, confusion, trouble focusing and memory problems. Major underlying causes of CKD include diabetes, hypertension and glomerulonephritis such as immunoglobulin A nephropathy, focal segmental glomerulonephritis and membranous nephropathy.
About KERENDIA® (finerenone)2
INDICATIONS:
KERENDIA (finerenone) is indicated to reduce the risk of:
- sustained estimated glomerular filtration rate (eGFR) decline, end-stage kidney disease, cardiovascular death, non-fatal myocardial infarction, and hospitalization for heart failure in adult patients with chronic kidney disease (CKD) associated with type 2 diabetes (T2D) (10mg, 20mg tablets)
- cardiovascular death, hospitalization for heart failure, and urgent heart failure visits in adult patients with heart failure with left ventricular ejection fraction (HF LVEF) ≥40% (10mg, 20mg, 40mg tablets)
IMPORTANT SAFETY INFORMATION
CONTRAINDICATIONS:
- Hypersensitivity to any component of this product
- Concomitant use with strong CYP3A4 inhibitors
- Patients with adrenal insufficiency
WARNINGS AND PRECAUTIONS:
-
Hyperkalemia: KERENDIA can cause hyperkalemia. The risk for developing hyperkalemia increases with decreasing kidney function and is greater in patients with higher baseline potassium levels or other risk factors for hyperkalemia.
Measure serum potassium and eGFR in all patients before initiation of treatment with KERENDIA and dose accordingly. Do not initiate KERENDIA if serum potassium is >5 mEq/L. Measure serum potassium periodically during treatment with KERENDIA and adjust dose accordingly. More frequent monitoring may be necessary for patients at risk for hyperkalemia, including those on concomitant medications that impair potassium excretion or increase serum potassium.
-
Worsening of Renal Function in Patients with Heart Failure: KERENDIA can cause worsening of renal function in patients with heart failure. Rarely, severe events associated with worsening renal function, including events requiring hospitalization, have been observed.
Measure eGFR in all patients before initiation of treatment or with dose titration of KERENDIA and dose accordingly. Initiation of KERENDIA in patients with heart failure and an eGFR <25 mL/min/1.73 m2 is not recommended. Measure eGFR periodically during maintenance treatment with KERENDIA in patients with heart failure. Consider delaying up-titration or interrupting treatment with KERENDIA in patients who develop clinically significant worsening of renal function.
MOST COMMON ADVERSE REACTIONS:
- CKD associated with T2D: From the pooled data of FIDELIO-DKD and FIGARO-DKD, the adverse reactions reported in ≥1% of patients on KERENDIA and more frequently than placebo were hyperkalemia (14% vs 6.9%), hypotension (4.6% vs 3%), and hyponatremia (1.3% vs 0.7%).
- HF LVEF ≥40%: From FINEARTS-HF, the adverse reactions reported in ≥1% of patients on KERENDIA and more frequently than placebo were hyperkalemia (9.7% vs 4.2%), hypotension (7.6% vs 4.7%), and hyponatremia (1.9% vs 0.9%).20 Events related to worsening renal function were reported more frequently in the KERENDIA group (18%) compared with placebo (12%).
DRUG INTERACTIONS:
- Strong CYP3A4 Inhibitors: Concomitant use of KERENDIA with strong CYP3A4 inhibitors is contraindicated. Avoid concomitant intake of grapefruit or grapefruit juice.
- Moderate and Weak CYP3A4 Inhibitors: Monitor serum potassium during drug initiation or dosage adjustment of either KERENDIA or the moderate or weak CYP3A4 inhibitor, and adjust KERENDIA dosage as appropriate.
- Strong and Moderate CYP3A4 Inducers: Avoid concomitant use of KERENDIA with strong or moderate CYP3A4 inducers.
- Sensitive CYP2C8 Substrates at KERENDIA 40mg: Monitor patients more frequently for adverse reactions caused by sensitive CYP2C8 substrates if KERENDIA 40mg is co-administered with such substrates, since minimal concentration changes may lead to serious adverse reactions.
USE IN SPECIFIC POPULATIONS:
- Lactation: Avoid breastfeeding during treatment with KERENDIA and for 1 day after treatment.
- Hepatic Impairment: Avoid use of KERENDIA in patients with severe hepatic impairment (Child Pugh C) and consider additional serum potassium monitoring with moderate hepatic impairment (Child Pugh B).
Please see the Prescribing Information for KERENDIA.
About Bayer’s Commitment in Cardiovascular and Kidney Diseases
Bayer’s legacy in cardiovascular care spans decades of scientific innovation and patient-focused research. As a long-standing leader in cardiology, Bayer has consistently advanced therapies that address the complex interplay between the heart and kidneys—two organs deeply connected in both health and disease. Today, that heritage continues to guide our commitment to developing innovative treatments for patients facing high unmet medical needs. With a growing portfolio of approved therapies and promising compounds in development, Bayer is shaping the future of cardiovascular care through precision medicine, scientific rigor, and a deep sense of purpose.
About Bayer
Bayer is a global enterprise with core competencies in the life science fields of health care and nutrition. In line with its mission, “Health for all, Hunger for none,” the company’s products and services are designed to help people and the planet thrive by supporting efforts to master the major challenges presented by a growing and aging global population. Bayer is committed to driving sustainable development and generating a positive impact with its businesses. At the same time, the Group aims to increase its earning power and create value through innovation and growth. The Bayer brand stands for trust, reliability and quality throughout the world. In fiscal 2025, the Group employed around 88,000 people and had sales of 45.6 billion euros. R&D expenses amounted to 5.8 billion euros. For more information, go to www.bayer.com.
Find more information at https://pharma.bayer.com/
Follow us on Facebook: http://www.facebook.com/bayer
Follow us on X: @BayerPharma
Forward-Looking Statements
This release may contain forward-looking statements based on current assumptions and forecasts made by Bayer management. Various known and unknown risks, uncertainties and other factors could lead to material differences between the actual future results, financial situation, development or performance of the company and the estimates given here. These factors include those discussed in Bayer’s public reports, which are available on the Bayer website at www.bayer.com. The company assumes no liability whatsoever to update these forward-looking statements or to conform them to future events or developments.
|
___________________________ 1 Data on file. 2 Bayer Pharmaceuticals. Kerendia (finerenone) [package insert]. U.S. Food and Drug Administration. Available at: https://labeling.bayerhealthcare.com/html/products/pi/Kerendia_PI.pdf. Accessed March 4, 2026 3 Research C for DE and. Table of Surrogate Endpoints That Were the Basis of Drug Approval or Licensure. FDA. Published online August 20, 2020. https://www.fda.gov/drugs/development-resources/table-surrogate-endpoints-were-basis-drug-approval-or-licensure 4 Jager KJ, et al. Kidney Int. 2019;96(5):1048-1050. 5 GBD 2017 Disease and Injury Incidence and Prevalence Collaborators. The Lancet. 2018;392(10159):1789-1858 6 Wanner C, et al. BMC Nephrol. 2025;26:1–11. / Webster AC, Nagler EV, Morton RL, Masson P. Chronic kidney disease. Lancet. 2017;389:1238–52. 7 NIDDK. Kidney disease statistics for the United States. National Institute of Diabetes and Digestive and Kidney Diseases. Published 2023. Accessed March 4, 2026. https://www.niddk.nih.gov/health-information/health-statistics/kidney-disease 8 GBD 2023 Chronic Kidney Disease Collaborators. The Lancet. 2025;406(10518):2461-2482. 9 Heerspink HJL, Agarwal R, Bakris GL, et al. Design and baseline characteristics of the Finerenone, in addition to standard of care, on the progression of kidney disease in patients with Non-Diabetic Chronic Kidney Disease (FIND-CKD) randomized trial. Nephrol Dial Transplant. 2025;40(2):308-319. doi:10.1093/ndt/gfae132 10 Study to Evaluate the Efficacy (Effect on Disease) and Safety of Finerenone in Participants With Heart Failure and Left Ventricular Ejection Fraction (Proportion of Blood Expelled Per Heart Stroke) Greater or Equal to 40% (FINEARTS-HF) Clinical trial registration No. NCT04435626. https://clinicaltrials.gov/study/NCT04435626 Accessed March 4, 2026 11 A Study to Determine the Efficacy and Safety of Finerenone on Morbidity and Mortality Among Hospitalized Heart Failure Patients (REDEFINE-HF). Clinical trial registration No. NCT 06008197. https://www.clinicaltrials.gov/study/NCT06008197. Accessed March 4, 2026. 12 A Study to Determine the Efficacy and Safety of Finerenone and SGLT2i in Combination in Hospitalized Patients with Heart Failure (CONFIRMATION-HF) (CONFIRMATION). Clinical trial registration No. NCT06024746. https://www.clinicaltrials.gov/study/NCT06024746. Accessed March 4, 2026. 13 A Study to Evaluate Finerenone on Clinical Efficacy and Safety in Patients with Heart Failure Who are Intolerant or Not Eligible for Treatment with Steroidal Mineralocorticoid Receptor Antagonists (FINALITY-HF). Clinical trial registration No. NCT06033950. https://www.clinicaltrials.gov/study/NCT06033950. Accessed March 4, 2026. 14 Efficacy and Safety of Finerenone in Subjects With Type 2 Diabetes Mellitus and Diabetic Kidney Disease (FIDELIO-DKD) Clinical trial registration No. NCT02540993. https://clinicaltrials.gov/study/NCT02540993. Accessed March 4, 2026. 15 Efficacy and Safety of Finerenone in Subjects With Type 2 Diabetes Mellitus and the Clinical Diagnosis of Diabetic Kidney Disease (FIGARO-DKD) Clinical trial registration No. NCT02545049 https://clinicaltrials.gov/study/NCT02545049. Accessed March 4, 2026. 16 A Study to Learn How Well the Study Treatment Finerenone Works and How Safe it is in People With Long-term Decrease in the Kidneys’ Ability to Work Properly (Chronic Kidney Disease) Together With Type 1 Diabetes (FINE-ONE). Clinical trial registration No. NCT05901831. https://www.clinicaltrials.gov/study/NCT05901831. Accessed March 4, 2026. 17 A Trial to Learn How Well Finerenone Works and How Safe it is in Adult Participants With Non-diabetic Chronic Kidney Disease (FIND-CKD). Clinical trial registration No. NCT05047263. https://www.clinicaltrials.gov/study/NCT05047263. Accessed March 4, 2026. 18 A Study to Learn More About How Well the Study Treatment Finerenone Works, How Safe it is, How it Moves Into, Through and Out of the Body, and the Effects it Has on the Body When Taken With an ACE Inhibitor or Angiotensin Receptor Blocker in Children with Chronic Kidney Disease and Proteinuria (FIONA). Clinical trial registration No. NCT05196035. https://www.clinicaltrials.gov/study/NCT05196035. Accessed March 4, 2026. 19 A Study to Learn More About How Safe the Study Treatment Finerenone is in Long-term Use When Taken With an ACE Inhibitor or Angiotensin Receptor Blocker Over 18 Months of Use in Children and Young Adults From 1 to 18 Years of Age With Chronic Kidney Disease and Proteinuria (FIONA OLE). Clinical trial registration No. NCT05457283. https://www.clinicaltrials.gov/study/NCT05457283. Accessed March 4, 2026. 20 Data on file. |
COR-KER-US-0185-1 3/26
Contacts
Media Contact:
Sarra Herzog
Bayer Media Relations
Sarra.Herzog@bayer.com
+1 862.460.8764
BioVersys Receives Green Light from US FDA for BV100 HABP/VABP Phase 3 Pivotal Trial Start
|
Basel, Switzerland. March 16, 2026, 7am CET
|
- BV100 is a potential best-in-class anti-infective agent in treating hospital-acquired bacterial pneumonia (HABP) or ventilator-associated bacterial pneumonia (VABP), caused by carbapenem-resistant Acinetobacter baumannii (CRAB).
- US FDA confirms that the BV100 global pivotal Phase 3 HABP/VABP trial may proceed, based on the initial IND submission.
- BV100 global pivotal Phase 3 is on track to read-out by the end of 2027.
BioVersys AG (SIX: BIOV), a multi-asset, clinical stage biopharmaceutical company focusing on research and development of novel antibacterial products for serious life-threatening infections caused by multi-drug resistant (MDR) bacteria, announced today that the US Food and Drug Administration (US FDA) has confirmed that the global Phase 3 pivotal trial to recruit US patients into the RIV-TARGET clinical trial (NCT07326540) can proceed. The Phase 3 trial will recruit patients with hospital-acquired or ventilator-associated bacterial pneumonia (HABP or VABP), due to carbapenem-resistant Acinetobacter baumannii-calcoaceticus Complex (CRABC).
CRABC is a highly-drug resistant Gram-negative pathogen that is recognized as a critical priority by global health authorities. BV100 is a novel intravenous formulation of rifabutin based on the newly identified mode of action for the active uptake of rifabutin into the Acinetobacter baumannii-calcoaceticus complex. BV100 is being developed for MDR hospital infections caused by Acinetobacter baumannii, including carbapenem-resistant Acinetobacter baumannii (CRAB) strains. BV100 has Qualified Infectious Disease Product (QIDP) Designation from the U.S. FDA, making BV100 eligible for priority FDA review, Fast Track designation, and a five-year extension of market exclusivity.
Dr. Glenn E Dale, Chief Development Officer: “This is a significant step towards our goal of completing the BV100 global Phase 3 trial by the end of 2027. HABP and VABP infections remain a leading cause of infectious disease hospital mortality and represent a substantial burden on healthcare systems. Our teams have already gathered interest from US-based clinical sites to participate in the trial, and this successful IND application will allow us to activate the clinical sites prior to enrolling US patients in the coming months.”
Dr. Marc Gitzinger, Chief Executive Officer of BioVersys: “We are excited about this regulatory development. Enrolling US patients into the trial is an important contribution towards the global nature of the study. This follows closely behind our announcement in December 2025 that we had initiated the BV100 global Phase 3 trial. I am incredibly proud of our team’s multi-pronged efforts in ensuring that this trial can proceed simultaneously in all the key regions of the world. We will bring this much-needed treatment option to patients in dire need.”
Phase 3 trial design
The global Phase 3 trial, is a randomized, active-controlled two-part parallel-group trial to evaluate the efficacy and safety of BV100 plus low-dose polymyxin B in patients with HABP or VABP suspected or confirmed to be due to CRABC infection (RIV-TARGET). Part A is the pivotal, randomized, comparative portion of the trial, employing a partially blinded design aiming to enroll approximately 300 HABP/VABP patients with suspected or confirmed CRABC infections. Patients will be randomized 1:1 to receive either [1] BV100 combined with low dose polymyxin B or [2] colistin combined with high-dose ampicillin-sulbactam, with both arms allowing meropenem as background in case of polymicrobial infections. The primary efficacy endpoint is defined as 28-day all-cause mortality (ACM) in the CRABC microbiological modified intention-to-treat (CRABC m-MITT) population. Secondary efficacy endpoints will include clinical cure status at the test of cure (ToC) in CRABC m-MITT, ventilator free days, time spent in intensive care unit (ICU) and time in hospital. As part of the study protocol, data safety monitoring boards (DSMB)[1] will be convened at regular intervals to review trial progress.
The Phase 3 trial also includes an open-label, non-randomized, additional single group (Part B) to evaluate the efficacy and safety of BV100 plus low-dose polymyxin B in patients with HABP or VABP due to CRABC known to be resistant to colistin or polymyxin B prior to study entry and patients where colistin or polymyxin B regimen has failed prior to study entry. Approximately 25 patients are expected to be enrolled in Part B.
This pivotal Phase 3 trial follows the successful completion of a Phase 2 trial in documented Acinetobacter VABP patients. BV100 combined with polymyxin B demonstrated a clear survival benefit, resulting in a 50% relative reduction in 28-day ACM compared with best available therapy (BAT), in VABP patients suffering from confirmed CRAB infections (28-day ACM: 60% for BAT vs 25% for BV100), and was generally safe and well tolerated. The current Phase 3 trial mimics the successful study design of the positive Phase 2 trial and is expected to read-out towards the end of 2027. Subsequent regulatory submissions aimed at commercial approval are planned in 2028, initially for the US, Europe and China.
In parallel to the Phase 3 pivotal trial an open-label Phase 2b differentiation trial (RIV-CARE) will be initiated in H1 2026 comparing BV100 with BAT in multiple geographies. The Phase 2b trial aims to provide real world evidence of clinical practices in settings with very high drug resistance levels. Interim analysis is planned for end of 2026.
About BV100
BV100 is a novel formulation of rifabutin suitable for intravenous administration, with a recently discovered novel mode of action showing an active uptake of rifabutin into the Gram-negative bacterial species, Acinetobacter baumannii. For the first time, BV100 allows for the targeting of the RNA-polymerase enzyme in Gram-negative bacteria with a human-suitable dose. BV100 is being developed for the treatment of infections caused by Acinetobacter baumannii calcoaceticus complex (ABC), including carbapenem-resistant ABC (CRABC) in critically important indications of ventilator associated bacterial pneumonia (VABP), hospital-acquired bacterial pneumonia (HABP) and bloodstream infections (BSI). BV100 was granted QIDP Designation by the U.S. FDA in May 2019 for use in the treatment of VABP, HABP and BSI, making BV100 eligible for priority FDA review, Fast Track designation, and a five-year extension of market exclusivity upon approval of the first QIDP indication.
About Acinetobacter baumannii
Acinetobacter baumannii calcoaceticus complex (ABC) are Gram-negative bacteria found in the environment (e.g., in soil and water) and an opportunistic pathogen in humans, typically infecting critically ill and immunocompromised patients, that can result in severe pneumonia and bloodstream infections in addition to affecting other parts of the body. ABC is considered a significant worldwide threat in the healthcare setting given its ability to survive for prolonged periods on surfaces, combined with its ability to develop or acquire resistance to standard of care antibiotics, e.g. carbapenems. Carbapenem-resistance as well as multi-drug resistance (MDR) rates for ABC are among the highest recorded for any bacteria in current times (The Lancet 2022; 399: 629–55). Incidence and resistance rates for ABC are trending upwards and COVID-19 has exacerbated this significantly. BioVersys forecasts the annual number of carbapenem-resistant A. baumannii infections in hospitals to have surpassed one million globally and due to the limited treatment options, such infections come with high (up to 50%) mortality rates.
About BioVersys
BioVersys AG is a multi-asset, clinical stage biopharmaceutical company focused on identifying, developing and commercializing novel antibacterial products for serious life-threatening infections caused by multi-drug resistant (“MDR”) bacteria. Derived from the company’s two internal technology platforms (TRIC and Ansamycin Chemistry), candidates are designed and developed to overcome resistance mechanisms, block virulence production and directly affect the pathogenesis of harmful bacteria towards the identification of new treatment options in the antimicrobial and microbiome fields. This enables BioVersys to address the high unmet medical need for new treatments against life-threatening resistant bacterial infections and bacteria-exacerbated chronic inflammatory microbiome disorders. The company’s most advanced research and development programs address nosocomial infections of Acinetobacter baumannii (BV100, Phase 3), and tuberculosis (alpibectir, Phase 2, in collaboration with GlaxoSmithKline (GSK) and a consortium of the University of Lille, France). BioVersys is located in the biotech hub of Basel, Switzerland.
BioVersys contact
Hernan Levett, CFO, Tel. +41 61 633 22 50; Mail: hernan.levett@bioversys.com
For media: media@bioversys.com Website: www.bioversys.com
Disclaimer
This communication expressly or implicitly contains certain forward-looking statements, such as “believe”, “assume”, “expect”, “forecast”, “project”, “may”, “could”, “might”, “will” or similar expressions concerning BioVersys and its business, including with respect to the progress, timing and completion of research, development and clinical studies for product candidates. Such statements involve certain known and unknown risks, uncertainties and other factors, which could cause the actual results, financial condition, performance or achievements of BioVersys to be materially different from any future results, performance or achievements expressed or implied by such forward-looking statements. BioVersys is providing this communication as of this date and does not undertake to update any forward-looking statements contained herein as a result of new information, future events or otherwise.
[1] The DSMB review is the periodic evaluation of unblinded or partially unblinded clinical trial data by an independent committee of experts to determine whether the study should continue, be modified, or be stopped based on safety, efficacy, or futility considerations.