Egle Therapeutics and Consortium Partners Awarded €8 Million Grant from Horizon Europe to Advance Clinical Development of EGL-001 in Neoadjuvant Head and Neck Cancer

Egle Therapeutics and Consortium Partners Awarded €8 Million Grant from Horizon Europe to Advance Clinical Development of EGL-001 in Neoadjuvant Head and Neck Cancer


Egle Therapeutics and Consortium Partners Awarded €8 Million Grant from Horizon Europe to Advance Clinical Development of EGL-001 in Neoadjuvant Head and Neck Cancer

PARIS, March 02, 2026 (GLOBE NEWSWIRE) — Egle Therapeutics SAS (Egle), a clinical-stage biotechnology company pioneering precision medicines that modulate regulatory T cells (Tregs) to rebalance immune function in patients with autoimmune diseases and cancer, today announced it was awarded approximately €8 million in grant funding by Horizon Europe, the European Commission’s key funding program for research and innovation. This funding from the European Union will support an Egle-led initiative in partnership with a consortium of four leading European scientific institutions to advance the clinical development of EGL-001, a novel anti-CTLA-4 x IL-2m fusion protein, and to facilitate a comprehensive translational research program to de-risk later-stage clinical development.

“This Horizon Europe funding strengthens our ability to advance EGL-001’s clinical development and significantly expand the translational research needed to guide patient selection alongside several leading European partners. The grant enables the evaluation of EGL-001 by studying therapy-naïve patients in the neoadjuvant setting and focusing on head and neck squamous cell carcinoma as a single, well-defined indication,” said John Celebi, CEO of Egle Therapeutics. “In parallel, we are advancing our ongoing Phase 1/2 study of EGL-001, where it has been well tolerated with early signs of single-agent activity in PD-(L)1-resistant disease. We remain on track to report top-line data from this study in the second half of this year and look forward to bringing our Treg-focused product candidates closer to patients in need.”

As part of this initiative, Egle will collaborate with University College London (United Kingdom), Vall d’Hebron Institute of Oncology (VHIO) (Spain), Gustave Roussy (France), and Technical University of Dresden (Germany) to conduct a multicenter, randomized, open-label Phase 1/2 neoadjuvant clinical trial in patients with head and neck squamous cell carcinoma (HNSCC), a high-incidence, high-mortality cancer with limited benefit from current immunotherapies. This trial will evaluate the safety, tolerability, and early efficacy of EGL-001 in combination with pembrolizumab in these patients.

This initiative also integrates a comprehensive translational research program, including multi-omics profiling, immune monitoring, and AI-driven pathomics modeling, to deliver predictive biomarkers for patient stratification, ultimately supporting regulatory alignment and de-risking later-stage development. The consortium also provides cutting-edge expertise in immunology, genomics, pathology, and computational biology, alongside established clinical trial capacity across EU member states.

“EGL-001 represents a highly differentiated, Treg-based approach to cancer treatment that is designed to selectively disarm and remove regulatory T cells inside tumors, making tumor types with prominent Treg-driven immune suppression, including HNSCC, compelling settings for clinical evaluation,” said Prof. Aurélien Marabelle, M.D., Ph.D., Gustave Roussy Cancer Center. “In addition, this project incorporates a rigorous, clinically actionable framework to evaluate a novel approach to tumor-driven immune suppression and to rapidly generate translational insights that can inform future development and patient selection. On behalf of the consortium partners, we are excited to partner with Egle to advance this work and generate the clinical and translational evidence needed to bring new options like EGL-001 to patients.”

About EGL-001
EGL-001 is an investigational anti-CTLA-4-IL-2m fusion protein designed to selectively disarm and remove regulatory T cells (Tregs) in the tumor microenvironment and to block CTLA-4 to help restore effector T-cell priming and function. This dual-target approach is intended to induce tumor-selective Treg apoptosis through an Fc receptor-independent mechanism, with the goal of strengthening anti-tumor immunity while supporting tolerability and use in combination regimens, including with PD-(L)1 inhibitors. EGL-001 is currently being evaluated in an ongoing multicenter, open-label, first-in-human Phase 1/2 clinical trial (NCT06622486) in patients with selected advanced and/or metastatic solid tumors, with dose escalation underway both as a single agent and in combination with an anti-PD-(L)1 therapy to assess safety, tolerability, and preliminary anti-tumor activity and to identify recommended doses for expansion.

About Egle Therapeutics
Egle Therapeutics is a clinical-stage biotechnology company pioneering precision medicines that modulate regulatory T cells (Tregs) to rebalance immune function in patients with autoimmune diseases and cancer. The company’s proprietary technology platform enables the engineering of highly selective IL-2 variant immuno-conjugates which are designed to overcome the efficacy and safety limitations of previous Treg-focused therapies. Its autoimmunity portfolio is led by EGL-003, an IL-2 agonist Treg engager currently being explored as a potential treatment for atopic dermatitis and other autoimmune diseases. Egle’s immuno-oncology portfolio includes EGL-001, a clinical-stage anti-CTLA-4-IL-2m fusion, and EGL-002, an anti-CCR8 x anti-CD25 bispecific. For more information, visit www.egle-tx.com and follow us on LinkedIn.

Contacts
contact@egle-tx.com / 0033 (0)1 86 64 08 57
investor.relations@egle-tx.com / 0033 (0)1 86 64 08

Paul Chaplin Steps Down as CEO of Bavarian Nordic

Paul Chaplin Steps Down as CEO of Bavarian Nordic


Paul Chaplin Steps Down as CEO of Bavarian Nordic

COPENHAGEN, Denmark, March 2, 2026 – Bavarian Nordic A/S (OMX: BAVA) announced today, that the Board of Directors has entered into an agreement with CEO, Paul Chaplin, who wishes to step down for personal reasons. Paul Chaplin will continue in his role for the remainder of 2026, or until a successor has been identified. The Board of Directors has initiated the process to identify a new CEO.

Paul Chaplin has been with Bavarian Nordic since 1999 and was appointed CEO in 2014. He has led the development of the Company’s smallpox and mpox vaccine, representing a significant contribution to public health globally. Under his leadership, Bavarian Nordic has grown from a small research-based company with a single government customer to a large vaccine manufacturer with strong commercial operations in key markets. This growth has been made possible through strategic acquisition of travel vaccines, excellent tech transfer capability, and a successful commercial transformation of Bavarian Nordic.

Reflecting on his tenure, Paul Chaplin said: “I am leaving Bavarian Nordic with great pride. I have truly enjoyed the experiences and professional challenges over nearly three decades and have had the privilege of leading many talented colleagues towards a shared mission of improving and saving lives. We have achieved a lot and created a strong foundation for the future of the company. My family wants to relocate back to Australia, but I remain fully committed to the company for the remainder of the year to allow for a stable transition to its next CEO.”

Chair of the Board of Directors, Anne Louise Eberhard, said: “On behalf of the Board, I would like to thank Paul for his strong leadership and dedication throughout nearly 30 years. Paul has made a valuable contribution, not only to Bavarian Nordic’s scientific advances, but also to recent years’ commercial success and to global public health. Paul has steadily led the company through a transformation to becoming a leading and profitable vaccine company. I wish Paul all the best in this next chapter of life and appreciate his commitment to ensure a smooth transition to a new CEO for Bavarian Nordic, a search process we have now initiated.”

About Bavarian Nordic
Bavarian Nordic is a global vaccine company with a mission to improve health and save lives through innovative vaccines. We are a preferred supplier of mpox and smallpox vaccines to governments to enhance public health preparedness and have a leading portfolio of travel vaccines. For more information, visit www.bavarian-nordic.com

Forward-looking statements
This announcement includes forward-looking statements that involve risks, uncertainties and other factors, many of which are outside of our control, that could cause actual results to differ materially from the results discussed in the forward-looking statements. Forward-looking statements include statements concerning our plans, objectives, goals, future events, performance and/or other information that is not historical information. All such forward-looking statements are expressly qualified by these cautionary statements and any other cautionary statements which may accompany the forward-looking statements. We undertake no obligation to publicly update or revise forward-looking statements to reflect subsequent events or circumstances after the date made, except as required by law.

Contact investors:
Europe: Disa Tuominen, IR Manager, detu@bavarian-nordic.com
US: Graham Morrell, Gilmartin Group, graham@gilmartinir.com, Tel: +1 781 686 9600

Contact media:
Nicole Seroff, Vice President Corporate Communications, nise@bavarian-nordic.com, Tel: +45 53 88 06 03

Company Announcement no. 08 / 2026

Attachment

Press Release: Sanofi’s rilzabrutinib earns orphan drug designation in Japan for IgG4-related disease

Press Release: Sanofi’s rilzabrutinib earns orphan drug designation in Japan for IgG4-related disease


Press Release: Sanofi’s rilzabrutinib earns orphan drug designation in Japan for IgG4-related disease

Sanofi’s rilzabrutinib earns orphan drug designation in Japan for IgG4-related disease

  • Designation based on positive data from a phase 2 study of rilzabrutinib in IgG4-RD
  • Third global orphan drug designation for rilzabrutinib in IgG4-RD, underpinning Sanofi’s commitment to rare immune-mediated diseases

Paris, March 2, 2026 – The Ministry of Health, Labour and Welfare (MHLW) in Japan has granted orphan drug designation to rilzabrutinib, a novel, oral, reversible covalent Bruton’s tyrosine kinase (BTK) inhibitor, for IgG4-related disease (IgG4-RD). There is still unmet medical need and limited treatment options in Japan for IgG4-RD, a rare, progressive, immune-mediated chronic condition in which the immune system attacks various tissues and organs leading to serious damage. The MHLW grants orphan designation to medicines that address rare diseases or conditions with unmet medical needs.

Rilzabrutinib for the treatment of IgG4-RD was evaluated in a phase 2 study (clinical study identifier: NCT04520451) and results were presented at the European Alliance of Associations for Rheumatology 2025 congress. In IgG4-RD patients, treatment with rilzabrutinib for 52 weeks led to reduction in disease flares and other disease markers and minimized the need for treatment with glucocorticoids. The safety profile of rilzabrutinib in the study was consistent with previous studies in other indications, with no new safety signals observed. Treatment-emergent adverse events reported by >10% of patients include diarrhea, COVID-19, dizziness, dry mouth and nausea. Currently, rilzabrutinib in IgG4-RD is being evaluated in the RILIEF phase 3 study (clinical study identifier: NCT07190196).

Rilzabrutinib is being studied across multiple rare immune-mediated diseases. In 2025, it received approval for immune thrombocytopenia (ITP) in the US, the EU, and the UAE. Additionally, rilzabrutinib is currently under regulatory review for ITP in Japan. Rilzabrutinib has received several expedited designations from global regulatory authorities for ITP, IgG4-RD, warm autoimmune hemolytic anemia (wAIHA), and sickle cell disease (SCD). Other than the approved ITP indications in the US, EU, and UAE, these uses of rilzabrutinib are investigational and have not been evaluated by any regulatory authority.

About rilzabrutinib
Rilzabrutinib, Wayrilz where approved, is a novel, oral, reversible covalent BTK inhibitor that has the potential to be an effective new medicine for several rare immune-mediated or inflammatory diseases by working to restore immune balance via multi-immune modulation. BTK, expressed in B cells, macrophages, and other innate immune cells, plays a critical role in multiple immune-mediated disease processes and inflammatory pathways. With the application of the TAILORED COVALENCY® technology, rilzabrutinib can selectively inhibit the BTK target. Rilzabrutinib is now approved for the treatment of immune thrombocytopenia (ITP) in the US, the EU, and the UAE. Regulatory review for use in ITP is currently ongoing in Japan.

About IgG4-RD
IgG4-RD is a progressive, relapsing, chronic immune-mediated rare disease, which can manifest in almost every organ and can lead to organ damage and irreversible dysfunction with a sometimes fatal outcome. People with IgG4-RD experience flare-ups of the condition characterized by periods of exacerbated symptoms. Due to its rarity and challenges with diagnosis, the global prevalence of IgG4-RD is unknown.

About Sanofi
Sanofi is an R&D driven, AI-powered biopharma company committed to improving people’s lives and delivering compelling growth. We apply our deep understanding of the immune system to invent medicines and vaccines that treat and protect millions of people around the world, with an innovative pipeline that could benefit millions more. Our team is guided by one purpose: we chase the miracles of science to improve people’s lives; this inspires us to drive progress and deliver positive impact for our people and the communities we serve, by addressing the most urgent healthcare, environmental, and societal challenges of our time.
Sanofi is listed on EURONEXT: SAN and NASDAQ: SNY

Media Relations
Sandrine Guendoul | +33 6 25 09 14 25 | sandrine.guendoul@sanofi.com
Evan Berland | +1 215 432 0234 | evan.berland@sanofi.com
Léo Le Bourhis | +33 6 75 06 43 81 | leo.lebourhis@sanofi.com
Victor Rouault | +1 617 356 4751 | victor.rouault@sanofi.com
Timothy Gilbert | +1 516 521 2929 | timothy.gilbert@sanofi.com
Léa Ubaldi | +33 6 30 19 66 46 | lea.ubaldi@sanofi.com
Ekaterina Pesheva | +1 410 926 6780 | ekaterina.pesheva@sanofi.com

Investor Relations
Thomas Kudsk Larsen | +44 7545 513 693 | thomas.larsen@sanofi.com
Alizé Kaisserian | +33 6 47 04 12 11 | alize.kaisserian@sanofi.com
Keita Browne | +1 781 249 1766 | keita.browne@sanofi.com
Nathalie Pham | +33 7 85 93 30 17 | nathalie.pham@sanofi.com
Nina Goworek | +1 908 569 7086 | nina.goworek@sanofi.com
Thibaud Châtelet | +33 6 80 80 89 90 | thibaud.chatelet@sanofi.com
Yun Li | +33 6 84 00 90 72 | yun.li3@sanofi.com

Sanofi forward-looking statements
This press release contains forward-looking statements within the meaning of applicable securities laws, including the Private Securities Litigation Reform Act of 1995, as amended. Forward-looking statements are statements that are not historical facts. These statements include projections and estimates and their underlying assumptions regarding the marketing and other potential of the product; regarding potential future events and revenues from the product. Words such as “expect,” “anticipate,” “believe,” “intend,” “estimate,” “plan,” “can,” “contemplate,” “could,” “is designed to,” “may,” “might,” “potential,” “objective,” “attempt,” “target,” “project,” “strategy,” “strive,” “desire,” “predict,” “forecast,” “ambition,” “guideline,” “seek,” “should,” “will,” “goal,” or the negative of these and similar expressions are intended to identify forward-looking statements. Although Sanofi’s management believes that the expectations reflected in such forward-looking statements are reasonable, investors are cautioned that forward-looking information and statements are subject to various risks and uncertainties, many of which are difficult to predict and generally beyond the control of Sanofi, that could cause actual results and developments to differ materially from those expressed in, or implied or projected by, the forward-looking information and statements. These risks, uncertainties and assumptions include among other things, unexpected regulatory actions or delays, or government regulation generally, that could affect the availability or commercial potential of the product, the fact that product may not be commercially successful; authorities’ decisions regarding whether and when to approve a product candidate; political pressure in the United States to mandate lower drug prices including “most favored nation” pricing for State Medicaid programs; the uncertainties inherent in research and development, including future clinical data and analysis of existing clinical data relating to the product, including post marketing, unexpected safety, quality or manufacturing issues; competition in general; risks associated with intellectual property and any related pending or future litigation and the ultimate outcome of such litigation, and volatile economic and market conditions, and the impact that global crises may have on us, our customers, suppliers, vendors, and other business partners, and the financial condition of any one of them, as well as on our employees and on the global economy as a whole. The risks and uncertainties also include the uncertainties discussed or identified in the public filings with the SEC and the French Markets Authority (AMF) made by Sanofi, including those listed under “Risk Factors” and “Cautionary Statement Regarding Forward-Looking Statements” in Sanofi’s annual report on Form 20-F for the year ended December 31, 2025 or contained in our periodic reports on Form 6-K. Other than as required by applicable law, Sanofi does not undertake any obligation to update or revise any forward-looking information or statements. In light of these risks, uncertainties and assumptions, you should not place undue reliance on any forward-looking statements contained herein.

All trademarks mentioned in this press release are the property of the Sanofi group.

Attachment

VERAXA Biotech Shareholders Approve Merger and Issuance of New Shares to Voyager Acquisition Corp.

VERAXA Biotech Shareholders Approve Merger and Issuance of New Shares to Voyager Acquisition Corp.


VERAXA Biotech Shareholders Approve Merger and Issuance of New Shares to Voyager Acquisition Corp.

ZURICH, SWITZERLAND – March 2, 2026 — VERAXA Biotech AG (“VERAXA”), an emerging leader in designing novel cancer therapies, today announced that its shareholders approved the merger between VERAXA and Veraxa Biotech Holding AG and the issuance of new shares of the combined company at the Extraordinary General Meeting (“EGM”) on February 27, 2026. Both resolutions are prerequisites for the closing of the proposed business combination (the “Business Combination”) among VERAXA, Veraxa Biotech Holding AG and Voyager Acquisition Corp. (NASDAQ: VACH, “Voyager”), a special purpose acquisition company sponsored by Cantor Fitzgerald & Co., Voyager Acquisition Sponsor Holdco LLC, and Odeon Capital Group LLC.

The merger will be carried out by means of an absorption merger. Veraxa Biotech Holding AG, as the acquiring company, will take over VERAXA as the transferring company and simultaneously change its name to Veraxa Biotech AG. VERAXA will continue to operate under its existing management team led by Chief Executive Officer, Christoph Antz. Additionally, the EGM approved an ordinary capital increase of a maximum of CHF 223,400.00 and offering the corresponding number of shares to the shareholders of Voyager in connection with the business combination.

“We appreciate our shareholders’ support and their approval to take the next steps in our business combination process with Voyager”, said Christoph Antz, Ph.D., Chief Executive Officer of VERAXA. “VERAXA is well-positioned to generate significant long-term value by addressing the growing need for safer and more effective cancer therapies with a focus on antibody-drug conjugates and bispecific
T cell engagers. We look forward to continuing our path to becoming a leading innovator in cancer medicine.”

Subject to the approval of Voyager’s shareholders, VERAXA and Voyager will commence final procedures towards closing the Business Combination and the expected trading of shares of the combined company on NASDAQ under the symbol “VRXA”. The resolutions of the EGM are subject to the condition of the approval of the Business Combination by Voyager’s shareholders.

About the Business Combination

On April 22, 2025, VERAXA entered into a definitive business combination agreement (the “Business Combination Agreement“) with Voyager Acquisition Corp., a Cayman Islands exempted company and special purpose acquisition company targeting the healthcare sector (NASDAQ: VACH, “Voyager”). Upon closing of the Business Combination, the combined company is expected to become a publicly traded company listed on NASDAQ trading under the symbol “VRXA”.

The description of the Business Combination contained herein is only a high-level summary and is qualified in its entirety by reference to the underlying documents filed with the Securities and Exchange Commission (the “SEC”). A more detailed description of the terms of the transaction has been provided in a proxy statement/prospectus filed with the SEC by Voyager on February 19, 2026.

Note to Shareholders of VERAXA Biotech AG

In preparation for the registration of shares and subsequent share trading, we ask all shareholders of VERAXA Biotech AG to do the following:

  • To prepare your registration within the US share register, please send us your current email address to info@veraxa.com by March 6, 2026.
  • Please ensure that your shares are or will be registered by your bank with the Computershare share register.
  • To avoid short selling, please ensure that your shares are not used for lending by your bank.

About VERAXA Biotech AG

At VERAXA, we are building a premier engine for the discovery and development of next-generation antibody-based therapeutics, including bispecific ADCs, bispecific T cell engagers and other innovative formats. Powered by a suite of transformative technologies and guided by rigorous quality-by-design principles, we are rapidly advancing our pipeline of ADCs and proprietary BiTAC formats into clinical development and beyond. VERAXA was founded on scientific breakthroughs made at the European Molecular Biology Laboratory, a world-renowned institution known for pioneering life science research and cutting-edge technology.

For regular updates about VERAXA Biotech, visit www.veraxa.com. You can also follow us on LinkedIn.

About Voyager Acquisition Corp.

Voyager is a special purpose acquisition company with a bold mission: to revolutionize the healthcare sector through a merger, stock purchase, or business combination. Our team of experienced executives includes unparalleled expertise in investing, operations, and medical innovation, supported by a vast network of connections. With these strengths, we not only seek to drive success but commit to scaling companies to unprecedented heights in the healthcare industry. For more information, please visit https://www.voyageracq.com.

Participants In the Solicitation

Voyager, VERAXA, and their respective directors, executive officers, other members of management, and employees may be deemed participants in the solicitation of proxies from Voyager’s stockholders with respect to the Business Combination. Investors and security holders may obtain more detailed information regarding the names and interests in the Business Combination of Voyager’s directors and officers in Voyager’s filings with the SEC, including the preliminary proxy statement/prospectus, the definitive proxy statement/prospectus, amendments and supplements thereto, and other documents filed with the SEC. Such information with respect to VERAXA’s directors and executive officers is also included in the proxy statement/prospectus. You may obtain free copies of these documents as described below under the heading “Additional Information and Where to Find It”.

Non-Solicitation

This press release is not a proxy statement or solicitation of a proxy, consent or authorization with respect to any securities or in respect of the potential transaction and shall not constitute an offer to sell or a solicitation of an offer to buy the securities of Voyager or VERAXA, nor shall there be any sale of any such securities in any state or jurisdiction in which such offer, solicitation, or sale would be unlawful prior to registration or qualification under the securities laws of such state or jurisdiction. No offer of securities shall be made except by means of a prospectus meeting the requirements of the Securities Act of 1933, as amended.

Forward-Looking Statements

This press release includes certain statements that may be considered forward-looking statements within the meaning of the federal securities laws. Forward-looking statements include, without limitation, statements about future events or Voyager’s or VERAXA’s future financial or operating performance. For example, statements regarding VERAXA’s anticipated growth and the anticipated growth and other metrics, statements regarding the benefits of the Business Combination, and the anticipated timing of the completion of the Business Combination are forward-looking statements. In some cases, you can identify forward-looking statements by terminology such as “may,” “should,” “could,” “might,” “plan,” “possible,” “project,” “strive,” “budget,” “forecast,” “expect,” “intend,” “will,” “estimate,” “anticipate,” “believe,” “predict,” “potential” or “continue,” or the negatives of these terms or variations of them or similar terminology.

These forward-looking statements regarding future events and the future results of Voyager and VERAXA are based on current expectations, estimates, forecasts, and projections about the industry in which VERAXA operates, as well as the beliefs and assumptions of Voyager’s management and VERAXA’s management. These forward-looking statements are only predictions and are subject to, without limitation, (i) known and unknown risks, including the risks and uncertainties indicated from time to time in the final prospectus of Voyager relating to its initial public offering filed with the SEC, and in the proxy statement/prospectus filed by Voyager and VERAXA on February 19, 2026, including those under “Risk Factors” therein, and other documents filed or to be filed with the SEC by Voyager; (ii) uncertainties; (iii) assumptions; and (iv) other factors beyond Voyager’s or VERAXA’s control that are difficult to predict because they relate to events and depend on circumstances that will occur in the future. They are neither statements of historical fact nor promises or guarantees of future performance. Therefore, VERAXA’s actual results may differ materially and adversely from those expressed or implied in any forward-looking statements and Voyager and VERAXA therefore caution against relying on any of these forward-looking statements.

These forward-looking statements are based upon estimates and assumptions that, while considered reasonable by Voyager and its management and VERAXA and its management, as the case may be, are inherently uncertain and are inherently subject to risks, variability and contingencies, many of which are beyond Voyager’s or VERAXA’s control. Factors that may cause actual results to differ materially from current expectations include, but are not limited to: (i) the occurrence of any event, change or other circumstances that could give rise to the termination of the Business Combination Agreement and any subsequent definitive agreements with respect to the Business Combination; (ii) the outcome of any legal proceedings that may be instituted against Voyager, VERAXA, or others following the announcement of the Business Combination and any definitive agreements with respect thereto; (iii) the inability to complete the Business Combination due to the failure to obtain consents and approvals of the shareholders of Voyager, to obtain financing to complete the Business Combination or to satisfy other conditions to closing, or delays in obtaining, adverse conditions contained in, or the inability to obtain necessary regulatory approvals required to complete the transactions contemplated by the Business Combination Agreement; (iv) the failure to realize estimated shareholder redemptions, purchase price and other adjustments; and (v) other risks and uncertainties set forth in the filings by Voyager with the SEC. There may be additional risks that neither Voyager nor VERAXA presently know or that Voyager and VERAXA currently believe are immaterial that could also cause actual results to differ from those contained in the forward-looking statements. Any forward-looking statements made by or on behalf of Voyager or VERAXA speak only as of the date they are made. None of Voyager or VERAXA undertakes any obligation to update any forward-looking statements to reflect any changes in their respective expectations with regard thereto or any changes in events, conditions or circumstances on which any such statement is based.

Additional Information and Where to Find It

In connection with the Business Combination Agreement, Voyager and VERAXA have filed a proxy statement/prospectus of Voyager, and will file other documents regarding the proposed transaction with the SEC. This communication is not intended to be, and is not, a substitute for the proxy statement/prospectus or any other document that Voyager has filed or may file with the SEC in connection with the proposed transaction. The definitive proxy statement and other relevant materials for the proposed transaction have been mailed or made available to stockholders of Voyager as of a record date to be established for voting on the proposed transaction.

Before making any voting or investment decision, investors and stockholders of Voyager are urged to carefully read the entire registration statement, the proxy statement/prospectus, and any other relevant documents filed with the SEC, as well as any amendments or supplements to these documents, and the documents incorporated by reference therein, because they will contain important information about Voyager, VERAXA, and the proposed transaction. Voyager’s investors and stockholders and other interested persons can also obtain copies of the registration statement, the preliminary proxy statement/prospectus, the definitive proxy statement/prospectus, other documents filed with the SEC that will be incorporated by reference therein, and all other relevant documents filed with the SEC by Voyager and/or VERAXA in connection with the transaction, without charge, once available, at the SEC’s website at www.sec.gov, or by directing a request to Voyager at the address set forth below.

Contact

VERAXA Biotech AG Voyager Acquisition Corp.
Dr. Christoph Antz
CEO
Email: antz@veraxa.com
Mr. Adeel Rouf
CEO, and Director
Email: adeel@voyageracq.com
   
For Media and Investors
Mario Brkulj
Valency Communications
Email: mbrkulj@valencycomms.eu
 

Attachment

Takeda and Protagonist Announce U.S. Food and Drug Administration Accepts New Drug Application and Grants Priority Review for Rusfertide as a Potential First-in-Class Therapy for Polycythemia Vera

Takeda and Protagonist Announce U.S. Food and Drug Administration Accepts New Drug Application and Grants Priority Review for Rusfertide as a Potential First-in-Class Therapy for Polycythemia Vera


Takeda and Protagonist Announce U.S. Food and Drug Administration Accepts New Drug Application and Grants Priority Review for Rusfertide as a Potential First-in-Class Therapy for Polycythemia Vera

  • Rusfertide Demonstrated Significant Improvements in Hematocrit Control, Phlebotomy Reduction and Patient Reported Outcomes for Patients with Polycythemia Vera in a Pivotal Study
  • Submission Primarily Based on Phase 3 VERIFY Study, in Which Rusfertide Plus Standard of Care More Than Doubled Clinical Response Rates, as Well as Four-Year Efficacy and Safety Data from Phase 2 REVIVE/THRIVE Studies
  • Prescription Drug User Fee Act (PDUFA) Target Action Date is in the Third Quarter of this Calendar Year

OSAKA, Japan & CAMBRIDGE, Mass. & NEWARK, Calif.–(BUSINESS WIRE)–Takeda (TSE:4502/NYSE:TAK) and Protagonist Therapeutics, Inc. (“Protagonist”) (NASDAQ:PTGX) today announced that the U.S. Food and Drug Administration (FDA) accepted the New Drug Application (NDA) and granted Priority Review for rusfertide. Rusfertide is an investigational, first-in-class hepcidin mimetic peptide therapeutic for the treatment of adults with polycythemia vera (PV). The FDA has set a Prescription Drug User Fee Act (PDUFA) goal date in the third quarter of this calendar year. In addition to Priority Review, rusfertide has received Breakthrough Therapy designation, Orphan Drug designation and Fast Track designation from the U.S. FDA.

PV is characterized by the overproduction of red blood cells (erythrocytosis), which increases blood viscosity, or thickness, and can result in life threatening thrombotic events. Hematocrit is the ratio of red blood cells to the total amount of blood in the body. Achieving and maintaining controlled hematocrit levels of <45% is the primary treatment goal in PV to prevent thrombotic events and alleviate burdensome symptoms.

“There is an urgent need for innovative treatment options in polycythemia vera, where patients currently face limited therapeutic choices to control their hematocrit and significant symptom burden,” said Andy Plump, M.D., Ph.D., president of R&D at Takeda. “The FDA’s acceptance of our NDA brings us closer to potentially offering a first-in-class therapy that could meaningfully improve clinical outcomes and quality of life. This milestone is a reflection of our successful partnership with Protagonist and Takeda’s unwavering commitment to advancing innovative treatments in hematologic cancers where significant unmet needs persist.”

The NDA for rusfertide was primarily based on the positive 32-week primary analysis and 52-week results from the Phase 3 global randomized VERIFY study (NCT05210790), as well as four-year efficacy and safety data from the Phase 2 REVIVE study (NCT04057040) and long-term extension THRIVE study (NCT06033586). In the VERIFY study, rusfertide met the primary endpoint and all four key secondary endpoints. Patients receiving rusfertide plus current standard of care demonstrated a higher response rate compared to current standard of care. This included hematocrit control, a reduction in phlebotomy requirements and improvement in pre-specified patient reported outcomes of fatigue and symptom burden. Rusfertide was generally well-tolerated through 52 weeks of treatment. The most common treatment-emergent adverse events (AEs) in rusfertide-treated patients were injection site reactions (47.4%), anemia (25.6%) and fatigue (19.6%), which were mainly grade 1 or 2. Serious AEs occurred in 8.1% of overall rusfertide-treated patients.

“Rusfertide exemplifies Protagonist’s end-to-end expertise, from exploring a novel hepcidin mimetic mechanism to address unmet needs in polycythemia vera to discovering the peptide and driving its clinical development through NDA filing. We are very pleased with the FDA granting rusfertide Priority Review and look forward to its potential approval in 2026,” said Dinesh V. Patel, Ph.D., Protagonist President and CEO. “We have identified a great partner in Takeda as rusfertide progresses toward this milestone, thereby bringing a successful closure to our more than decade-long journey from concept-to-commercialization.”

In January 2024, Protagonist and Takeda entered into a worldwide license and collaboration agreement for rusfertide. Protagonist discovered rusfertide and led its development through Phase 3 studies, with Takeda responsible for implementing the regulatory strategy for the U.S. NDA filing and for leading any future global regulatory filings. Protagonist holds an option to co-commercialize in the U.S. through a 50/50 profit and loss share structure or to opt-out of this structure, providing Takeda with a worldwide license pursuant to the license and collaboration agreement.

About Rusfertide

Rusfertide is a first-in-class investigational subcutaneous treatment that mimics the action of hepcidin, a natural hormone that regulates iron homeostasis and red blood cell production. By targeting the underlying mechanism of iron dysregulation in polycythemia vera, rusfertide aims to reduce excess red blood cell production and help patients achieve sustained hematocrit control. Rusfertide is administered once weekly via subcutaneous self-injection and has been generally well-tolerated in clinical trials to date.

About VERIFY

The Phase 3 VERIFY study (NCT05210790) is an ongoing, three-part, global, randomized, placebo-controlled study evaluating rusfertide in 293 patients with polycythemia vera over a 156-week period, with treatment extension for participants who are continuing to derive benefit from rusfertide beyond the 156-week treatment period. The study is evaluating the efficacy and safety of once-weekly, subcutaneously self-administered rusfertide in patients with uncontrolled hematocrit who are phlebotomy-dependent despite current standard of care treatment, which could include phlebotomy, hydroxyurea, interferon and/or ruxolitinib. The primary endpoint of the study was the proportion of patients achieving a response during Weeks 20-32, which was defined as the absence of “phlebotomy eligibility.” To meet phlebotomy eligibility, patients in the study were required to have: confirmed hematocrit ≥45% that was ≥3% higher than their baseline hematocrit value, or hematocrit ≥48%.

All patients have completed their participation in the randomized, placebo-controlled portion of the study evaluating the efficacy and safety of rusfertide plus current standard of care versus placebo plus current standard of care and are now in the open-label portions of the study.

About REVIVE and THRIVE

The Phase 2 REVIVE study (NCT04057040) evaluated rusfertide in adult patients with polycythemia vera and consisted of three parts, including 70 patients in the dose-finding Part 1 (28 weeks), 59 patients in the blinded, placebo-controlled, randomized withdrawal Part 2 (13 weeks) and 58 patients in the Part 3 open-label expansion (52 weeks). The THRIVE study (NCT06033586) is an ongoing, open-label extension study evaluating the long-term durability of response and safety profile of rusfertide in patients with polycythemia vera. The study includes 46 patients who previously participated in REVIVE. Patients eligible to transition to the THRIVE study completed the open-label extension portion of REVIVE, ≥12 months of rusfertide therapy and had an end-of-treatment visit. THRIVE is designed to further assess the maintenance of hematocrit control, reduction in the need for therapeutic phlebotomy and overall safety of once-weekly, subcutaneous rusfertide over an additional two-year treatment period.

About Polycythemia Vera (PV)

Polycythemia vera (PV) is characterized by the overproduction of red blood cells (erythrocytosis), which increases blood viscosity, or thickness, and can result in life threatening thrombotic events such as stroke, deep vein thrombosis and pulmonary embolism. Hematocrit is the ratio of red blood cells to the total amount of blood in the body. Achieving and maintaining controlled hematocrit levels of <45% is the primary treatment goal in PV to prevent thrombotic events and alleviate burdensome symptoms, including severe fatigue, difficulty in concentrating, night sweats and pruritus.

About Takeda

Takeda is focused on creating better health for people and a brighter future for the world. We aim to discover and deliver life-transforming treatments in our core therapeutic and business areas, including gastrointestinal and inflammation, rare diseases, plasma-derived therapies, oncology, neuroscience and vaccines. Together with our partners, we aim to improve the patient experience and advance a new frontier of treatment options through our dynamic and diverse pipeline. As a leading values-based, R&D-driven biopharmaceutical company headquartered in Japan, we are guided by our commitment to patients, our people and the planet. Our employees in approximately 80 countries and regions are driven by our purpose and are grounded in the values that have defined us for more than two centuries. For more information, visit www.takeda.com.

About Protagonist

Protagonist Therapeutics is a discovery through late-stage development biopharmaceutical company. Two novel peptides derived from Protagonist’s proprietary discovery platform are currently in advanced Phase 3 clinical development, with NDAs for both ICOTYDE™ (icotrokinra) and rusfertide under review at the FDA. ICOTYDE is a first-in-class investigational targeted oral peptide that selectively blocks the Interleukin-23 receptor (“IL-23R”), which is licensed to Janssen Biotech, Inc., a Johnson & Johnson company. Following ICOTYDE’s joint discovery by Protagonist and Johnson & Johnson scientists pursuant to the companies’ IL-23R collaboration, Protagonist was primarily responsible for the development of ICOTYDE through Phase 1, with Johnson & Johnson assuming responsibility for development in Phase 2 and beyond.

Rusfertide is a first-in-class hepcidin mimetic peptide that is being co-developed with Takeda Pharmaceuticals pursuant to a worldwide license and collaboration agreement entered in 2024. Protagonist holds an option to co-commercialize rusfertide in the U.S. through a 50/50 profit and loss share structure or can opt-out of this structure. The Company also has a number of preclinical stage drug discovery programs addressing clinically and commercially validated targets including an oral IL-17 peptide antagonist, obesity dual and triple agonists, an oral hepcidin functional mimetic, and the recently announced IL-4 and amylin programs.

More information on Protagonist, its pipeline drug candidates, and clinical studies can be found on the Company’s website at https://www.protagonist-inc.com.

Takeda Important Notice

For the purposes of this notice, “press release” means this document, any oral presentation, any question and answer session and any written or oral material discussed or distributed by Takeda Pharmaceutical Company Limited (“Takeda”) regarding this release. This press release (including any oral briefing and any question-and-answer in connection with it) is not intended to, and does not constitute, represent or form part of any offer, invitation or solicitation of any offer to purchase, otherwise acquire, subscribe for, exchange, sell or otherwise dispose of, any securities or the solicitation of any vote or approval in any jurisdiction. No shares or other securities are being offered to the public by means of this press release. No offering of securities shall be made in the United States except pursuant to registration under the U.S. Securities Act of 1933, as amended, or an exemption therefrom. This press release is being given (together with any further information which may be provided to the recipient) on the condition that it is for use by the recipient for information purposes only (and not for the evaluation of any investment, acquisition, disposal or any other transaction). Any failure to comply with these restrictions may constitute a violation of applicable securities laws.

The companies in which Takeda directly and indirectly owns investments are separate entities. In this press release, “Takeda” is sometimes used for convenience where references are made to Takeda and its subsidiaries in general. Likewise, the words “we”, “us” and “our” are also used to refer to subsidiaries in general or to those who work for them. These expressions are also used where no useful purpose is served by identifying the particular company or companies.

Takeda Forward-Looking Statements

This press release and any materials distributed in connection with this press release may contain forward-looking statements, beliefs or opinions regarding Takeda’s future business, future position and results of operations, including estimates, forecasts, targets and plans for Takeda. Without limitation, forward-looking statements often include words such as “targets”, “plans”, “believes”, “hopes”, “continues”, “expects”, “aims”, “intends”, “ensures”, “will”, “may”, “should”, “would”, “could”, “anticipates”, “estimates”, “projects”, “forecasts”, “outlook” or similar expressions or the negative thereof. These forward-looking statements are based on assumptions about many important factors, including the following, which could cause actual results to differ materially from those expressed or implied by the forward-looking statements: the economic circumstances surrounding Takeda’s global business, including general economic conditions in Japan and the United States and with respect to international trade relations; competitive pressures and developments; changes to applicable laws and regulations, including drug pricing, tax, tariff and other trade-related rules; challenges inherent in new product development, including uncertainty of clinical success and decisions of regulatory authorities and the timing thereof; uncertainty of commercial success for new and existing products; manufacturing difficulties or delays; fluctuations in interest and currency exchange rates; claims or concerns regarding the safety or efficacy of marketed products or product candidates; the impact of health crises, like the novel coronavirus pandemic; the success of our environmental sustainability efforts, in enabling us to reduce our greenhouse gas emissions or meet our other environmental goals; the extent to which our efforts to increase efficiency, productivity or cost-savings, such as the integration of digital technologies, including artificial intelligence, in our business or other initiatives to restructure our operations will lead to the expected benefits; and other factors identified in Takeda’s most recent Annual Report on Form 20-F and Takeda’s other reports filed with the U.S. Securities and Exchange Commission, available on Takeda’s website at: https://www.takeda.com/investors/sec-filings-and-security-reports/ or at https://www.sec.gov/. Takeda does not undertake to update any of the forward-looking statements contained in this press release or any other forward-looking statements it may make, except as required by law or stock exchange rule. Past performance is not an indicator of future results and the results or statements of Takeda in this press release may not be indicative of, and are not an estimate, forecast, guarantee or projection of Takeda’s future results.

Takeda Medical Information

This press release contains information about products that may not be available in all countries, or may be available under different trademarks, for different indications, in different dosages, or in different strengths. Nothing contained herein should be considered a solicitation, promotion or advertisement for any prescription drugs including the ones under development.

Protagonist Cautionary Note on Forward-Looking Statements

This press release contains forward-looking statements for purposes of the safe harbor provisions of the Private Securities Litigation Reform Act of 1995. Forward-looking statements include statements regarding the potential benefits of rusfertide. In some cases, you can identify these statements by forward-looking words such as “anticipate,” “believe,” “may,” “will,” “expect,” or the negative or plural of these words or similar expressions. Forward-looking statements are not guarantees of future performance and are subject to risks and uncertainties that could cause actual results and events to differ materially from those anticipated, including, but not limited to, our ability to develop and commercialize our product candidates, our ability to earn milestone payments under our collaboration agreements with Janssen and Takeda, our ability to use and expand our programs to build a pipeline of product candidates, our ability to obtain and maintain regulatory approval of our product candidates, our ability to operate in a competitive industry and compete successfully against competitors that have greater resources than we do, and our ability to obtain and adequately protect intellectual property rights for our product candidates. Additional information concerning these and other risk factors affecting our business can be found in our periodic filings with the Securities and Exchange Commission, including under the heading “Risk Factors” contained in our most recently filed periodic reports on Form 10-K and Form 10-Q filed with the Securities and Exchange Commission. Forward-looking statements are not guarantees of future performance, and our actual results of operations, financial condition, and liquidity, and the development of the industry in which we operate, may differ materially from the forward-looking statements contained in this press release. Any forward-looking statements that we make in this press release speak only as of the date of this press release. We assume no obligation to update our forward-looking statements, whether as a result of new information, future events, or otherwise, after the date of this press release.

Contacts

Takeda Media Contacts:

Japanese Media
Tsuyoshi Tada

tsuyoshi.tada@takeda.com

U.S. and International Media
Emy Gruppo

emy.gruppo@takeda.com

Protagonist Investor Relations Contact
Corey Davis, Ph.D.

LifeSci Advisors

cdavis@lifesciadvisors.com
+1 212 915 2577

Protagonist Media Relations Contact
Virginia Amann

ENTENTE Network of Companies

virginiaamann@ententeinc.com
+1 833 500 0061 ext 1
Genentech’s Fenebrutinib Confirms Its Potential as First and Only BTK Inhibitor for Relapsing and Primary Progressive MS in Third Positive Phase III Study (FENhance 1)

Genentech’s Fenebrutinib Confirms Its Potential as First and Only BTK Inhibitor for Relapsing and Primary Progressive MS in Third Positive Phase III Study (FENhance 1)


Genentech’s Fenebrutinib Confirms Its Potential as First and Only BTK Inhibitor for Relapsing and Primary Progressive MS in Third Positive Phase III Study (FENhance 1)

– FENhance 1 met its primary endpoint, showing investigational fenebrutinib significantly reduced relapses by 51% compared to teriflunomide in relapsing multiple sclerosis (RMS), consistent with FENhance 2 results showing 59% reduction –

– FENhance 1 is the final study readout of the fenebrutinib pivotal clinical development program in MS, following positive results for FENhance 2 in RMS and for FENtrepid in primary progressive multiple sclerosis (PPMS) –

– Fenebrutinib has the potential to become the first and only high-efficacy oral, brain-penetrant treatment for both RMS and PPMS, showing a profound benefit on relapsing and progressive disease biology –

– Totality of data from all three Phase III fenebrutinib studies will be submitted to regulatory authorities –

SOUTH SAN FRANCISCO, Calif.–(BUSINESS WIRE)–Genentech, a member of the Roche Group (SIX: RO, ROG; OTCQX: RHHBY), announced today that the pivotal Phase III study (FENhance 1) of fenebrutinib in RMS met its primary endpoint, showing clinically meaningful and statistically significant results. The study demonstrated that fenebrutinib, an investigational Bruton’s tyrosine kinase (BTK) inhibitor, markedly reduced the annualized relapse rate (ARR) by 51% compared to teriflunomide over a period of at least 96 weeks of treatment, consistent with FENhance 2 results showing a 59% reduction in ARR. Together, these results equate to approximately one relapse every 17 years. Secondary endpoints in both RMS studies show statistically significant and clinically meaningful reductions in brain lesions. Additionally, all progression endpoints show favorable trends for fenebrutinib.


Full data from the FENhance 1 and 2 studies will be shared at the American Academy of Neurology (AAN) Annual Meeting 2026 and submitted to regulatory authorities together with data from the FENtrepid study.

“These pivotal results, together with the earlier data, provide convincing evidence that fenebrutinib can become the first high-efficacy oral treatment for RMS and PPMS,” said Levi Garraway, M.D., Ph.D., chief medical officer and head of Global Product Development. “Building on a decade of transforming MS treatment, we are committed to advancing innovation to one day allow people with MS to live a life without disability.”

The positive FENhance 1 study follows positive results for FENhance 2 in RMS and for FENtrepid in PPMS, which were both announced in November. The collective positive results across all three pivotal studies demonstrate that fenebrutinib consistently shows a profound benefit on relapsing and progressive disease biology.

In both RMS studies, liver transaminase elevations were comparable with teriflunomide. In the FENhance 1 study, there was one Hy’s Law case in the fenebrutinib arm and one in the teriflunomide arm. Both cases were asymptomatic and resolved after study drug discontinuation. There were no additional Hy’s Law cases across all of the fenebrutinib clinical development program in MS or in other autoimmune diseases.

In the FENhance 1 and 2 studies in RMS, 1 fatal case was reported in the teriflunomide arm and 8 fatal cases with various causes and at different points in treatment in the fenebrutinib arms. Further analyses are ongoing to better understand these findings.

Fenebrutinib targets cells in the immune system known as B cells and microglia. Targeting B cells helps control the acute inflammation that causes relapses, while targeting microglia inside the brain addresses the chronic damage that is thought to drive long-term disability progression. Fenebrutinib, a non-covalent BTK inhibitor, is designed to have high potency, selectivity and reversibility. This design allows it to act throughout the body, and also to cross the blood-brain barrier into the central nervous system (CNS) targeting chronic inflammation.

About the FENhance 1 and 2 studies

FENhance 1 and 2 are two Phase III multicenter, randomized, double-blind, double-dummy, parallel-group studies to evaluate the efficacy and safety of investigational fenebrutinib compared with teriflunomide in a total of 1,497 adult patients with RMS. Eligible participants were randomized 1:1 to receive treatment with either oral fenebrutinib twice a day (and placebo matched to oral teriflunomide once a day) or oral teriflunomide once a day (and placebo matched to oral fenebrutinib twice a day) for at least 96 weeks.

The primary endpoint is annualized relapse rate (ARR). Secondary endpoints include total number of T1-gadolinium-enhancing MRI lesions, total number of new and/or enlarging T2-weighted MRI lesions, time to onset of 12-week composite confirmed disability progression (cCDP12) and 24-week cCDP (cCDP24). cCDP incorporates three measures of disability – total functional disability measured by confirmed disability progression (CDP) based on the Expanded Disability Status Scale (EDSS), walking speed measured by the timed 25-foot walk (T25FW) and upper limb function measured by the nine-hole peg test (9HPT).

Following the double-blind treatment period, patients have the option to enter an open-label extension (OLE) phase, in which all patients receive treatment with fenebrutinib.

About fenebrutinib

Fenebrutinib is an investigational oral, central nervous system (CNS)-penetrant, reversible and non-covalent Bruton’s tyrosine kinase (BTK) inhibitor with an optimized pharmacokinetics (PK) profile. Fenebrutinib can act throughout the body and also cross the blood-brain barrier into the CNS to target chronic inflammation. It is uniquely designed to target relapsing and progressive biology by inhibiting cells in the immune system known as B cells and microglia. Targeting B cells helps control the acute inflammation that causes relapses, while targeting microglia inside the brain addresses the chronic damage that is thought to drive long-term disability progression.

Fenebrutinib is designed to have high potency and reversibility, with a selectivity for BTK 130 times greater than other kinases. This high selectivity highlights fenebrutinib’s potential to bind to its intended target without interfering with other kinases. While most current BTK inhibitors are covalent and irreversible, meaning they form a permanent chemical bond with the enzyme, fenebrutinib is non-covalent and reversible, meaning it binds and then eventually releases the enzyme. These design features may help limit off-target effects.

The fenebrutinib Phase III program includes two similarly designed trials in relapsing multiple sclerosis (RMS) (FENhance 1 and 2) with active comparator teriflunomide and the only trial in primary progressive multiple sclerosis (PPMS) (FENtrepid) in which a BTK inhibitor is being evaluated against Ocrevus.

To date, more than 2,700 patients and healthy volunteers have been treated with fenebrutinib in Phase I, II and III clinical programs across multiple diseases, including multiple sclerosis and other autoimmune disorders.

About multiple sclerosis

Multiple sclerosis is a chronic disease that affects more than 2.9 million people worldwide. People with all forms of multiple sclerosis experience disease progression from the beginning of their disease. Therefore, an important goal of treating multiple sclerosis is to slow, stop and ideally prevent progression as early as possible.

Approximately 85% of people with multiple sclerosis are initially diagnosed with relapsing-remitting multiple sclerosis (RRMS). Relapsing forms of the disease (RMS) include RRMS and active secondary progressive MS, and people with RMS experience relapses and worsening disability over time. Primary progressive multiple sclerosis (PPMS) is a debilitating form of the disease marked by steadily worsening symptoms but typically without distinct relapses or periods of remission. Approximately 15% of people with multiple sclerosis are diagnosed with the primary progressive form of the disease. Until the FDA approval of Ocrevus®, there had been no FDA-approved treatments for PPMS and Ocrevus is still the only approved treatment for PPMS. Despite the availability of CD20s, 30% of patients remain on low-efficacy oral therapy today. Slowing or stopping progression while simultaneously stopping relapses remains a high unmet need in MS.

About Genentech in neuroscience

Neuroscience is a major focus of research and development at Genentech. Our goal is to pursue groundbreaking science to develop new treatments that help improve the lives of people with chronic and potentially devastating diseases.

Genentech and Roche are investigating more than a dozen medicines for neurological disorders, including multiple sclerosis, spinal muscular atrophy, neuromyelitis optica spectrum disorder, Alzheimer’s disease, Huntington’s disease, Parkinson’s disease and Duchenne muscular dystrophy. Together with our partners, we are committed to pushing the boundaries of scientific understanding to solve some of the most difficult challenges in neuroscience today.

About Genentech

Founded 50 years ago, Genentech is a leading biotechnology company that discovers, develops, manufactures and commercializes medicines to treat patients with serious and life-threatening medical conditions. The company, a member of the Roche Group, has headquarters in South San Francisco, California. For additional information about the company, please visit http://www.gene.com.

Indications and Important Safety Information

What is Ocrevus?

Ocrevus is a prescription medicine used to treat:

  • Relapsing forms of multiple sclerosis (MS), to include clinically isolated syndrome, relapsing-remitting disease, and active secondary progressive disease, in adults
  • Primary progressive MS, in adults.

It is not known if Ocrevus is safe and effective in children.

Who should not receive Ocrevus?

Do not receive Ocrevus if you have an active hepatitis B virus (HBV) infection.

Do not receive Ocrevus if you have had a life-threatening allergic reaction to Ocrevus. Tell your healthcare provider if you have had an allergic reaction to Ocrevus or any of its ingredients in the past.

What is the most important information I should know about Ocrevus?

Ocrevus can cause serious side effects, including:

  • Infusion reactions: Infusion reactions are a common side effect of Ocrevus, which can be serious and may require you to be hospitalized. You will be monitored during your infusion and for at least 1 hour after each infusion of Ocrevus for signs and symptoms of an infusion reaction. Tell your healthcare provider or nurse if you get any of these symptoms:

    • Itchy skin
    • Rash
    • Hives
    • Tiredness
    • Coughing or wheezing
    • Trouble breathing
    • Throat irritation or pain
    • Feeling faint
    • Fever
    • Redness on your face (flushing)
    • Nausea
    • Headache
    • Swelling of the throat
    • Dizziness
    • Shortness of breath
    • Fatigue
    • Fast heartbeat

These infusion reactions can happen for up to 24 hours after your infusion. It is important that you call your healthcare provider right away if you get any of the signs or symptoms listed above after each infusion.

If you get infusion reactions, your healthcare provider may need to stop or slow down the rate of your infusion.

  • Infection:

    • Infections are a common side effect. Ocrevus increases your risk of getting upper respiratory tract infections, lower respiratory tract infections, skin infections, and herpes infections. Serious infections can happen with Ocrevus, which can be life-threatening or cause death. Tell your healthcare provider if you have an infection or have any of the following signs of infection including fever, chills, a cough that does not go away, or painful urination. Signs of herpes infection include: cold sores, shingles, genital sores, skin rash, pain, and itching. Signs of more serious herpes infection include: changes in vision, eye redness or eye pain, severe or persistent headache, stiff neck, and confusion. Signs of infection can happen during treatment or after you have received your last dose of Ocrevus. Tell your healthcare provider right away if you have an infection. Your healthcare provider should delay your treatment with Ocrevus until your infection is gone.
    • Hepatitis B virus (HBV) reactivation: Before starting treatment with Ocrevus, your healthcare provider will do blood tests to check for hepatitis B viral infection. If you have ever had hepatitis B virus infection, the hepatitis B virus may become active again during or after treatment with Ocrevus. Hepatitis B virus becoming active again (called reactivation) may cause serious liver problems including liver failure or death. Your healthcare provider will monitor you if you are at risk for hepatitis B virus reactivation during treatment and after you stop receiving Ocrevus.
    • Weakened immune system: Ocrevus taken before or after other medicines that weaken the immune system could increase your risk of getting infections.
  • Progressive Multifocal Leukoencephalopathy (PML): PML is a rare brain infection that usually leads to death or severe disability and has been reported with Ocrevus. Symptoms of PML get worse over days to weeks. It is important that you call your healthcare provider right away if you have any new or worsening neurologic signs or symptoms that have lasted several days, including problems with:

    • Thinking
    • Eyesight
    • Strength
    • Balance
    • Weakness on 1 side of your body
    • Using your arms or legs
  • Decreased immunoglobulins: Ocrevus may cause a decrease in some types of antibodies. Your healthcare provider will do blood tests to check your blood immunoglobulin levels.

Before receiving Ocrevus, tell your healthcare provider about all of your medical conditions, including if you:

  • have ever taken, take, or plan to take medicines that affect your immune system, or other treatments for MS.
  • have ever had hepatitis B or are a carrier of the hepatitis B virus.
  • have a history of inflammatory bowel disease or colitis.
  • have a history of liver problems
  • have had a recent vaccination or are scheduled to receive any vaccinations.

    • You should receive any required ‘live’ or ‘live-attenuated’ vaccines at least 4 weeks before you start treatment with Ocrevus. You should not receive ‘live’ or ‘live-attenuated’ vaccines while you are being treated with Ocrevus and until your healthcare provider tells you that your immune system is no longer weakened.
    • When possible, you should receive any ‘non-live’ vaccines at least 2 weeks before you start treatment with Ocrevus. If you would like to receive any non-live (inactivated) vaccines, including the seasonal flu vaccine, while you are being treated with Ocrevus, talk to your healthcare provider.
    • If you have a baby and you received Ocrevus during your pregnancy, it is important to tell your baby’s healthcare provider about receiving Ocrevus so they can decide when your baby should be vaccinated.
  • are pregnant, think that you might be pregnant, or plan to become pregnant. It is not known if Ocrevus will harm your unborn baby. You should use birth control (contraception) during treatment with Ocrevus and for 6 months after your last infusion of Ocrevus. Talk with your healthcare provider about what birth control method is right for you during this time. Tell your healthcare provider if you become pregnant while receiving Ocrevus.
  • are breastfeeding or plan to breastfeed. It is not known if Ocrevus passes into your breast milk. Talk to your healthcare provider about the best way to feed your baby if you take Ocrevus.

Tell your healthcare provider about all the medicines you take, including prescription and over-the-counter medicines, vitamins, and herbal supplements.

What are the possible side effects of Ocrevus?

Ocrevus may cause serious side effects, including:

  • Risk of cancers (malignancies) including breast cancer. Follow your healthcare provider’s instructions about standard screening guidelines for breast cancer.
  • Inflammation of the colon, or colitis: Tell your healthcare provider if you have any symptoms of colitis, such as:

    • Diarrhea (loose stools) or more frequent bowel movements than usual
    • Stools that are black, tarry, sticky or have blood or mucus
    • Severe stomach-area (abdomen) pain or tenderness
  • Liver damage. Ocrevus may cause liver damage. Your healthcare provider will do blood tests to check your liver before you start Ocrevus and while you take Ocrevus if needed. Tell your healthcare provider right away if you have any symptoms of liver damage, such as:

    • yellowing of the skin and eyes (jaundice)
    • nausea
    • vomiting
    • unusual darkening of the urine
    • feeling tired or weak

These are not all the possible side effects of Ocrevus.

Call your doctor for medical advice about side effects. You may report side effects to the FDA at 1-800-FDA-1088. You may also report side effects to Genentech at (888) 835-2555.

For more information, go to https://www.Ocrevus.com or call 1-844-627-3887.

For additional safety information, please see the full Prescribing Information and Medication Guide.

Contacts

Media Contact:

Michelle McCourt, (650) 467-6800

Advocacy Contact:

Lily Rose Atherton, (202) 713-0083

Investor Contacts:

Loren Kalm, (650) 225-3217

Bruno Eschli, +41 61 687 5284

PHCbi Launches LiCellGrow™ Cell Expansion System to Support High-Quality and Efficient Production of Cell and Gene Therapies

PHCbi Launches LiCellGrow™ Cell Expansion System to Support High-Quality and Efficient Production of Cell and Gene Therapies


PHCbi Launches LiCellGrow™ Cell Expansion System to Support High-Quality and Efficient Production of Cell and Gene Therapies

— Proprietary In-Line Monitoring Technology Enables Seamless Scale-Up from Basic Research to Commercial Manufacturing —


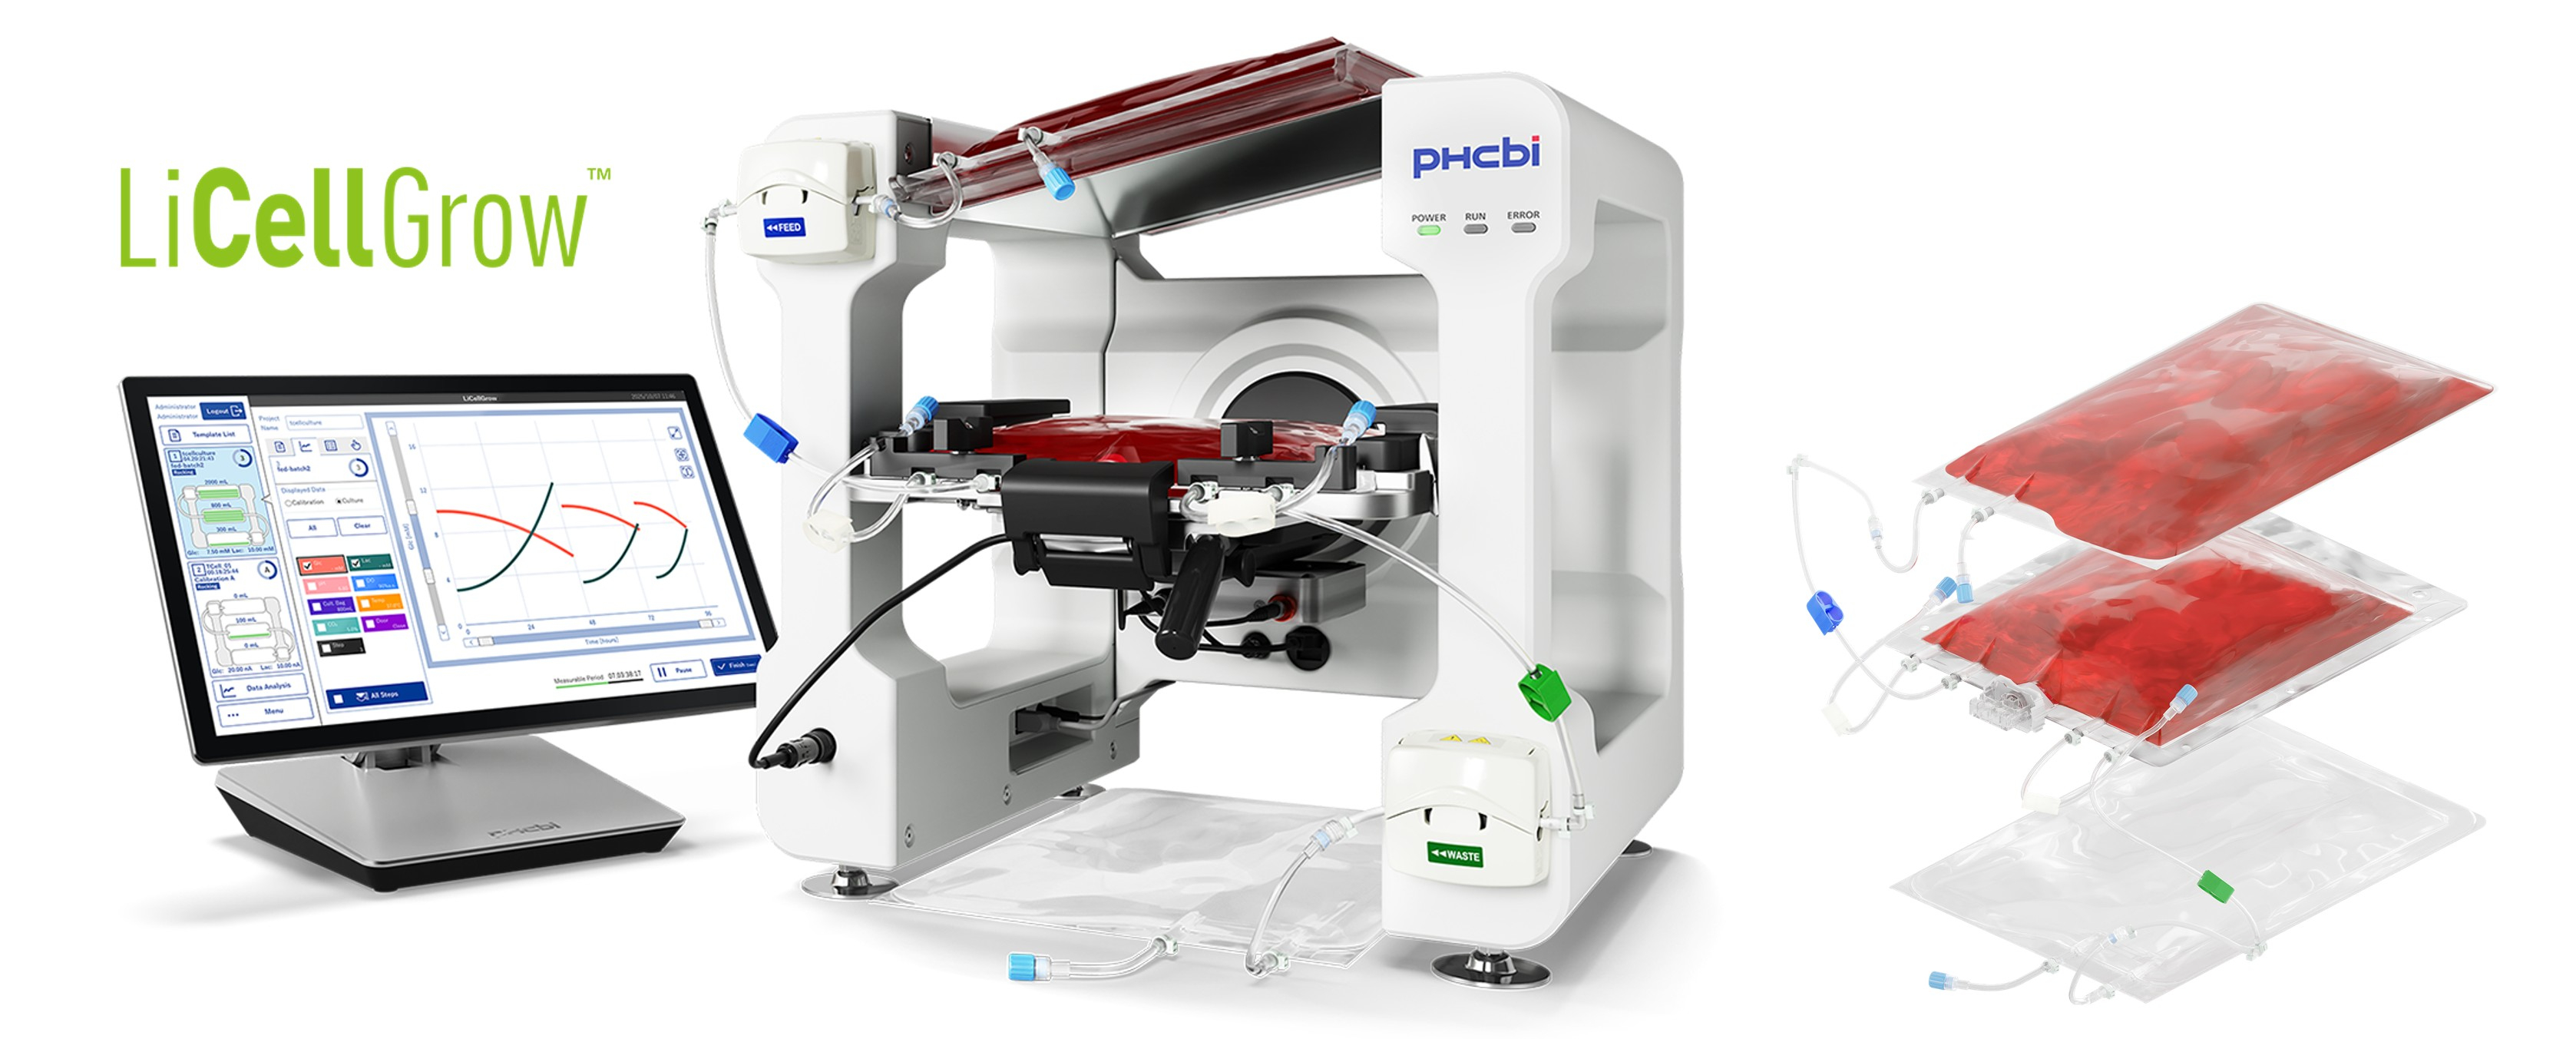
TOKYO–(BUSINESS WIRE)–PHC Corporation’s Biomedical Division (Head Office: Chiyoda-ku, Tokyo; President: Nobuaki Nakamura; hereinafter “PHCbi”), a global provider of laboratory sample storage and cell cultivation solutions and subsidiary of PHC Holdings Corporation (Head Office: Chiyoda-ku, Tokyo), today announced the launch of its new cell expansion system LiCellGrow™ (*1) for research use in Japan and other select countries worldwide(*2). The system is designed to improve quality and efficiency in the production of new advanced therapies by allowing therapy developers to visualize metabolic changes in cells in real time and automatically control culture conditions.

PHCbi will exhibit LiCellGrow™ at the 25th Congress of the Japanese Society for Regenerative Medicine, March 19-20 at the Kobe International Conference Center and Kobe International Exhibition Center in Kobe, Japan.

Cell and gene therapies (CGT), in which cells or genetic material are used to create personalized medical treatments, are rapidly advancing as promising treatment options for hard-to-treat diseases such as genetic disorders and cancer. In the manufacturing of CGT products, however, variations in cell characteristics and complexity of manufacturing are challenges to maintaining a consistent quality product. As a result, therapy developers are faced with reduced production efficiency, lower yields, and increased manufacturing costs compared to conventional therapies. To overcome these challenges, it is essential for therapy developers to be able to identify critical process parameters (CPPs) and critical quality attributes (CQAs) in CGT product manufacturing. The ability to monitor cell status and control culture conditions based on cell status are indispensable to developing quality CGT products at scale efficiently and at a lower cost.

PHCbi developed LiCellGrow™ to address these challenges and support CGT manufacturing at scale by helping to identify optimal culture conditions during the manufacturing process development. The system is equipped with proprietary In-Line monitoring technology that continuously measures, in real time, glucose and lactate concentrations, two key indicators of cell metabolism. It also incorporates culture control technology that adjusts cell culture medium at the optimal time based on these measurements. This technology is built on PHCbi’s proprietary electrochemical measurement platform, cultivated through more than 30 years of blood glucose sensor development. By visualizing the metabolic state of cultured cells, which is traditionally difficult to assess, and enabling precise control of culture conditions, LiCellGrow™ supports improved cell quality and enhanced efficiency as well as lower costs through reduced losses in the manufacturing of CGT products.

Under PHC Group’s Value Creation Plan 2027, which includes a focus on Diagnostics and Life Sciences, PHCbi is developing solutions to enhance efficiency and reduce costs in the manufacturing of CGT products. LiCellGrow™ builds on the In-Line monitoring technology introduced in LiCellMo™ (*3), a live cell metabolic analyzer launched in 2024 for research use only. From late 2026 through early 2028, PHCbi plans to expand its product lineup in phases, including culture bags with filtration functions that enhance cell recovery, pH/DO (dissolved oxygen) meter units, and cGMP-compliant dedicated consumables. These enhancements will support seamless scaling of CGT from basic research to commercial manufacturing in products such as CAR-T cell therapy.

[Key Features of the Product]

1. Automated Control and Optimization of Culture Conditions Using In-Line Monitoring Technology

By utilizing In-Line monitoring technology, LiCellGrow™ continuously tracks real-time changes in culture conditions and cell status, and automates medium exchange based on the acquired metabolic data. This enables the culture environment to be consistently maintained in an optimal state, contributing not only to improved cell quality and uniformity but also enhanced manufacturing efficiency. Furthermore, this process control approach aligns with the concept of Quality by Design (QbD) (*4), which embeds quality scientifically at the design stage rather than relying solely on final product testing. The system therefore supports the development of scientifically robust and highly reproducible manufacturing processes.

2. Single-Use Design Ensuring Aseptic Conditions and Highly Reproducible Cell Culture

LiCellGrow™ employs easy-to-attach, single-use dedicated culture bags. With automated measurement of culture conditions using In-Line sensors, the sampling tasks previously required to check glucose and lactate concentrations are no longer necessary. This reduces the risk of cross-contamination and enables closed-system cell culture that maintains aseptic conditions. In addition, the device can be placed inside a standard CO₂ incubator commonly used in laboratories, eliminating the need for special facility investments and allowing users to easily establish a reliable and highly reproducible culture environment.

Chikara Takauo, Director of PHC Corporation and Head of the Biomedical Division, commented:

“We are excited to launch LiCellGrow™ and offer therapy developers a new way to solve common challenges in quality, cost, and delivery of cell and gene therapies. Building on the success of our research-use system LiCellMo™, LiCellGrow™ represents an important step forward in making CGT manufacturing efficient, reliable, and scalable. We are committed to expanding our PHCbi offerings to continue to contribute to the adoption of CGT and other advanced therapy options for hard-to-treat conditions.”

****************************************************************

Overview of the 25th Congress of the Japanese Society for Regenerative Medicine

Dates: March 19 – 20, 2026

Venue: Kobe International Exhibition Hall, Building No. 2, 1st Floor (PHCbi Booth No.: T31)

Official Website: The 25th Congress of the Japanese Society for Regenerative Medicine
******************************************************************

(*1) https://www.phchd.com/jp/biomedical/incubation/cell-expansion-system-en
This product is intended for research use only and not for medical or clinical purposes.

(*2) Not available for purchase in the United States. Expected U.S. launch in the summer of 2026.

(*3) Live Cell Metabolic Analyzer | PHCbi
(*4) An approach to quality assurance defined in the ICH (International Council for Harmonisation of Technical Requirements for Pharmaceuticals for Human Use) guidelines (ICH Q8–Q11) and widely adopted in the pharmaceutical field.

About the Biomedical Division of PHC Corporation

Established in 1969, PHC Corporation is a Japanese subsidiary of PHC Holdings Corporation (TSE 6523), a global healthcare company that develops, manufactures, sells, and services solutions across diabetes management, healthcare solutions, life sciences and diagnostics. The Biomedical Division supports the life sciences industry helping researchers and healthcare providers in around 110 countries and regions through its laboratory and equipment and services including CO2 incubators and ultra-low temperature freezers.

www.phchd.com/global/phc

About PHC Holdings Corporation

PHC Holdings Corporation (TSE 6523) is a global healthcare company with a mission of contributing to the health of society through healthcare solutions that have a positive impact and improve the lives of people. Its subsidiaries include PHC Corporation, Ascensia Diabetes Care, Epredia, LSI Medience Corporation, Wemex Corporation, and Mediford Corporation. Together, these companies develop, manufacture, sell and service solutions across diabetes management, healthcare solutions, diagnostics and life sciences. The consolidated net sales in FY2024 were JPY 361.6 billion with global distribution of products and services in more than 125 countries and regions. PHC Group is a collective term referring to PHC Holdings Corporation and its subsidiaries.

www.phchd.com

Contacts

[Contact for product and service]

Masayo Okada

Marketing Department, Biomedical Division

PHC Corporation

Email: masayo.okada@phchd.com

[Contact for IR and media]

Investor Relations & Corporate Communications Department

PHC Holdings Corporation

Tel: +81-3-6778-5311 Email: phc-pr@gg.phchd.com

Otsuka Medical Devices/Otsuka Pharmaceutical: Paradise Ultrasound Renal Denervation System for the Treatment of Resistant Hypertension, Now Covered by Insurance and Commercially Available in Japan

Otsuka Medical Devices/Otsuka Pharmaceutical: Paradise Ultrasound Renal Denervation System for the Treatment of Resistant Hypertension, Now Covered by Insurance and Commercially Available in Japan


Otsuka Medical Devices/Otsuka Pharmaceutical: Paradise Ultrasound Renal Denervation System for the Treatment of Resistant Hypertension, Now Covered by Insurance and Commercially Available in Japan

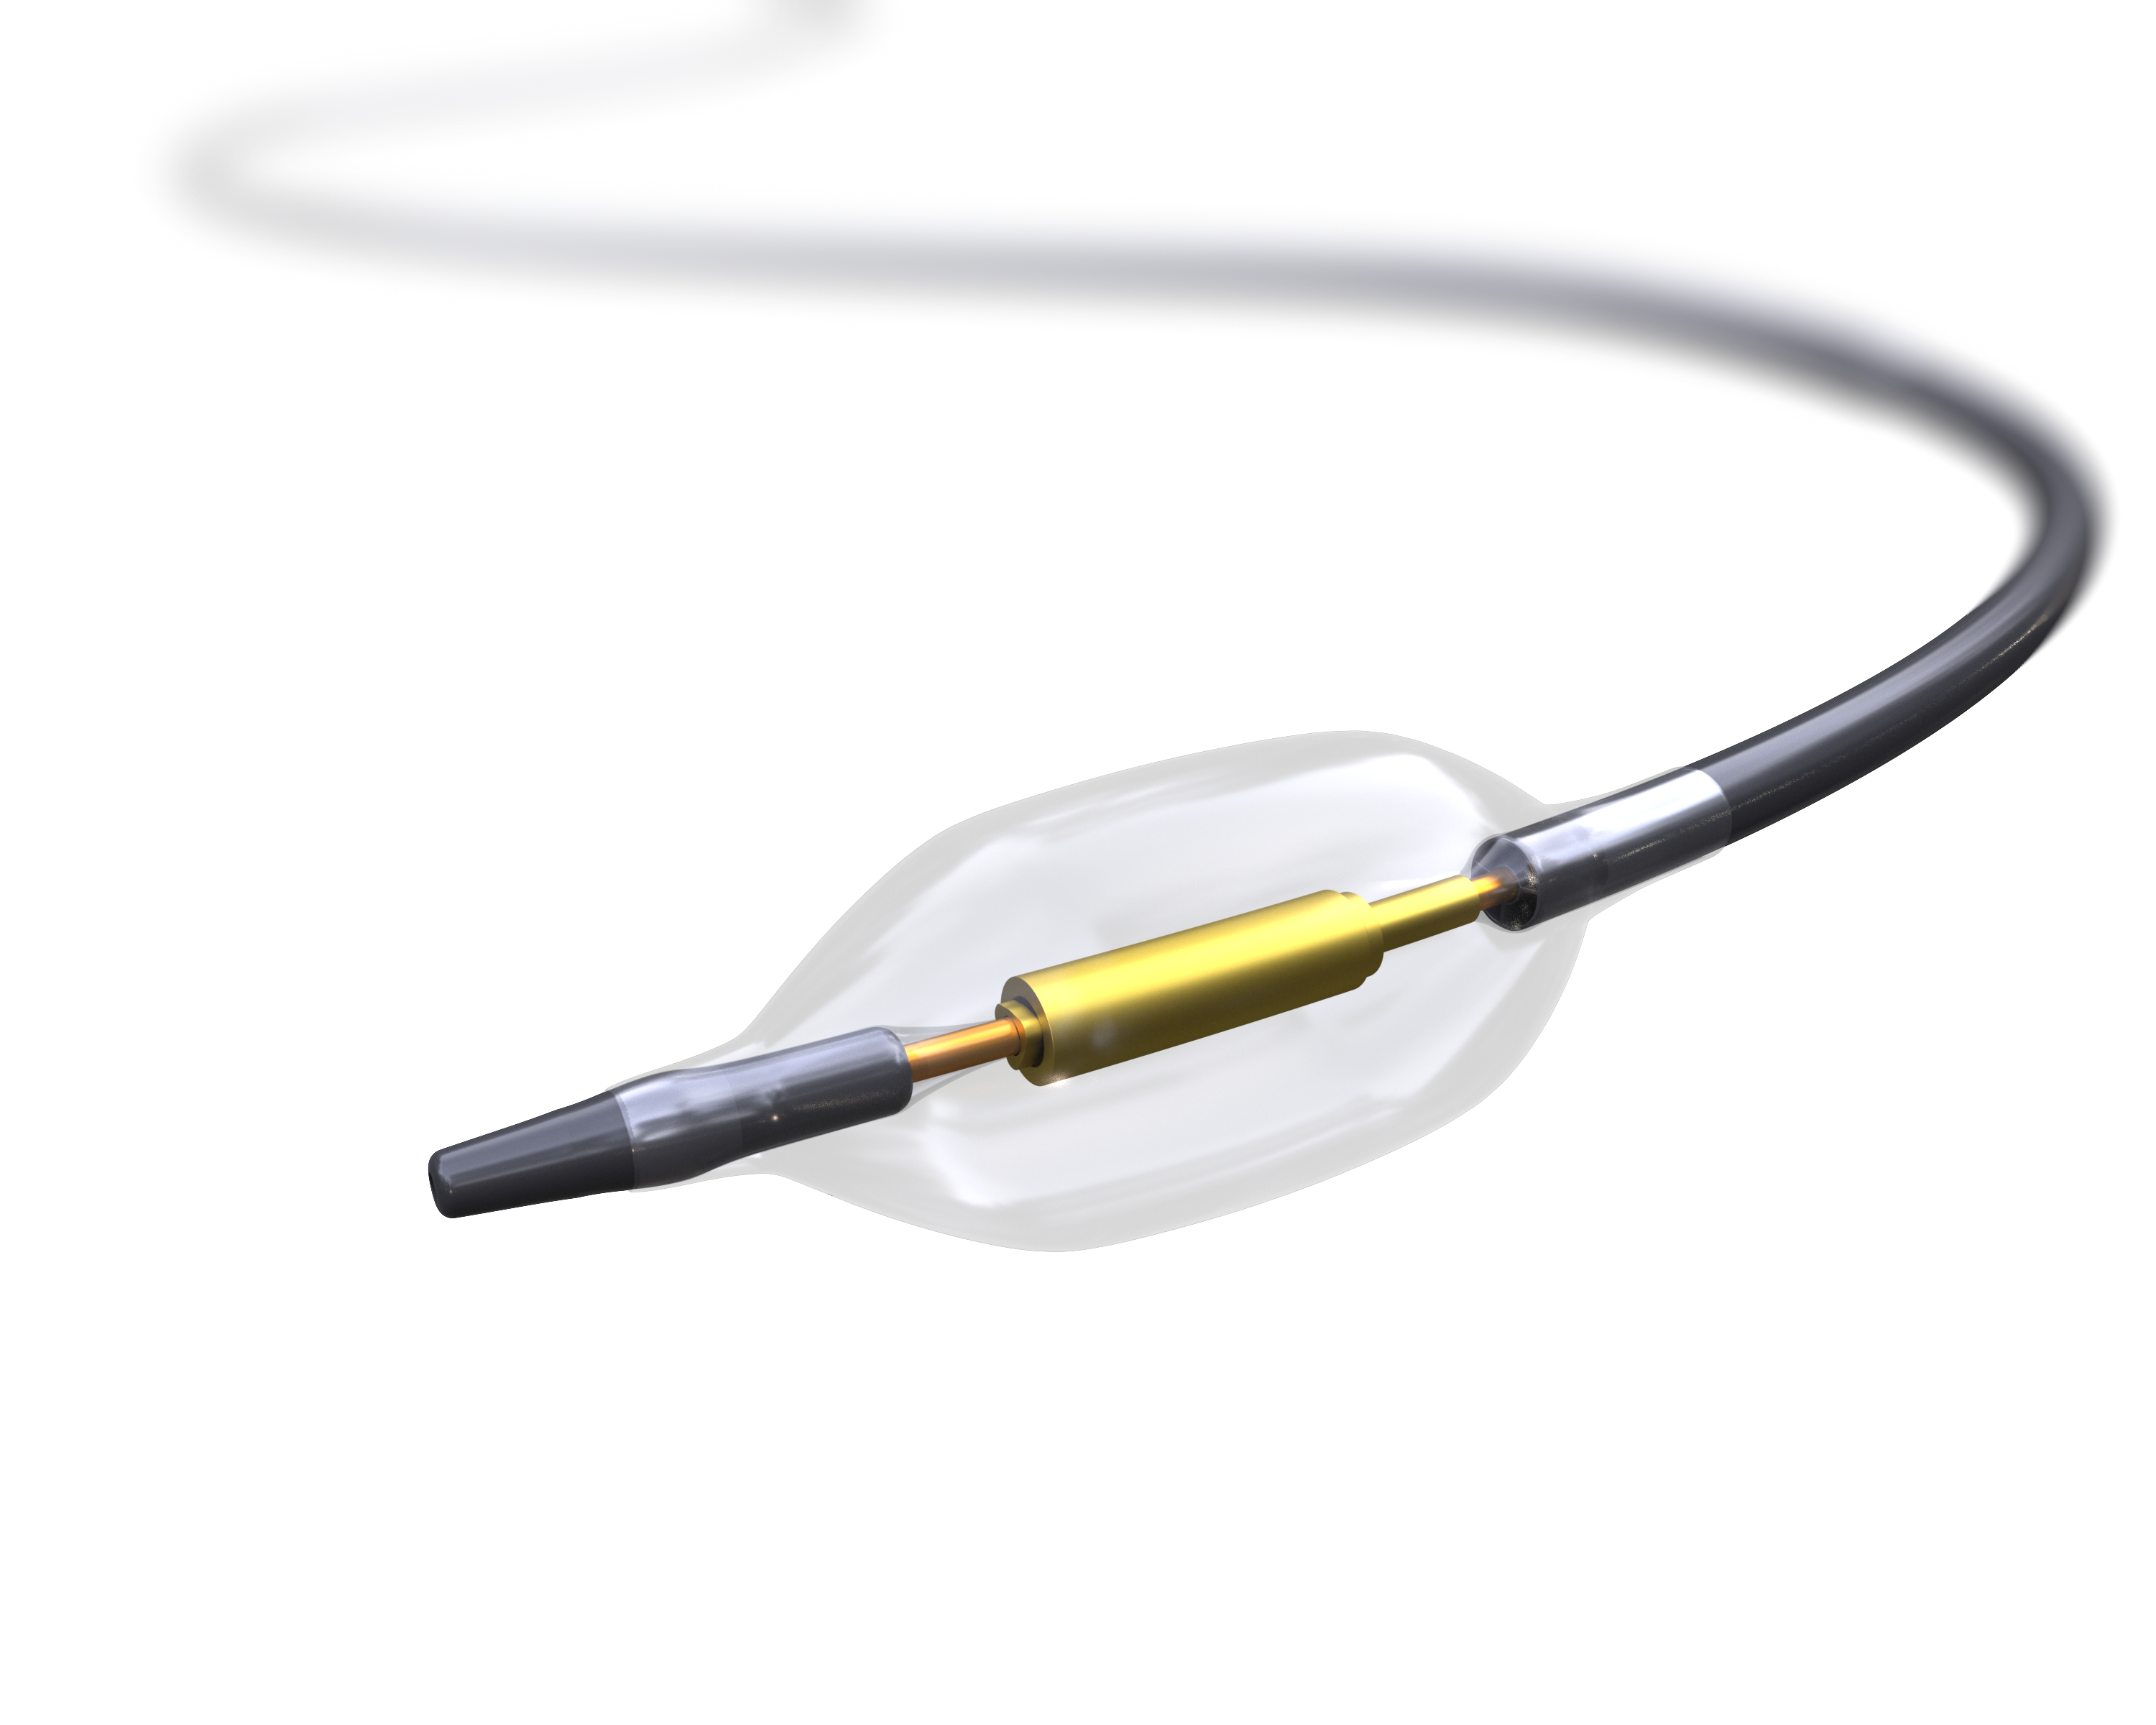
TOKYO–(BUSINESS WIRE)–Otsuka Medical Devices Co., Ltd. (Otsuka Medical Devices) and Otsuka Pharmaceutical Co., Ltd. (Otsuka Pharmaceutical) announce that the Paradise™ Ultrasound Renal Denervation (uRDN) system is covered by National Health Insurance system in Japan, effective March 1. Following the inception of insurance coverage, Otsuka Medical Devices has commenced sales of the system on March 2 and will conduct co-promotion with Otsuka Pharmaceutical.


The Paradise uRDN system was developed by Recor Medical, Inc. (Recor Medical), a U.S.-based subsidiary of Otsuka Medical Devices. This system is indicated for resistant hypertension patients whose blood pressure remains above target despite the use of three antihypertensive medications of different classes, including a diuretic.

Otsuka Medical Devices is introducing ultrasound renal denervation treatment to Japan for the first time. This novel treatment is designed to lower blood pressure by reducing overactivity of the sympathetic nerves surrounding the renal arteries.

With this insurance coverage, Paradise uRDN system becomes available as a new treatment option for resistant hypertension. Under a co-promotion framework, Otsuka Medical Devices and Otsuka Pharmaceutical will leverage Otsuka Pharmaceutical’s established expertise in the cardiovascular and renal fields to supply appropriate treatment options suited to patient conditions. The system will be prescribed by physicians to patients as defined in the guidelines for the proper use of renal denervation systems.

Kazuomi Kario, President of the Japanese Society of Hypertension and Professor of Cardiovascular Medicine at Jichi Medical University, commented, “Resistant hypertension carries an extremely high risk of leading to serious complications, such as cerebrovascular and cardiovascular diseases and renal failure. Being able to provide appropriate treatment opportunities to patients in need through this insurance coverage marks a significant step forward. Furthermore, the Japanese Society of Hypertension, the Japanese Association of Cardiovascular Intervention and Therapeutics, and the Japanese Circulation Society will work together to ensure that proper and safe treatment is delivered based on the facility requirements and patient eligibility criteria outlined in the guidelines for the proper use of renal denervation systems, which were formulated by our three societies.”

Makoto Inoue, President and Representative Director, CEO of Otsuka Holdings Co., Ltd., and President and Representative Director of Otsuka Pharmaceutical stated, “I am delighted that we can provide a new option to address the medical challenge of resistant hypertension by leveraging the know-how cultivated through the Otsuka group’s pharmaceutical business. Moving forward, we will bring together the collective strength of the Otsuka group to deliver optimal solutions for patients and consumers, as we strive to contribute to the total healthcare of every individual.”

“Paradise uRDN system received approval from the U.S. Food and Drug Administration (FDA) in 2023.* We are extremely pleased that patients in Japan with resistant hypertension now have access to this new treatment option under insurance coverage. We have built a framework with Otsuka Pharmaceutical to ensure this treatment reliably reaches patients in need, and we will continue to contribute to the health of patients and the advancement of medical care.” said Noriko Tojo, President and Representative Director of Otsuka Medical Devices.

Guided by the corporate philosophy of Otsuka-people creating new products for better health worldwide, Otsuka Medical Devices and Otsuka Pharmaceutical will continue to make every effort to deliver new value to address unmet medical needs.

*In the U.S, Paradise uRDN system received FDA approval in November 2023 as an adjunctive treatment option when lifestyle changes and medications have not adequately controlled a patient’s blood pressure.

About Otsuka Medical Devices

https://www.omd.otsuka.com/en/

About Recor Medical

https://www.recormedical.com/

About Otsuka Pharmaceutical

https://www.otsuka.co.jp/en/

Contacts

Otsuka Medical Devices

Inquiry form
+81-3-6361-7459

InnoCare’s next-generation TRKi Zurletrectinib Receives Priority Review for the Treatment of Pediatric Patients with Solid Tumors in China

InnoCare’s next-generation TRKi Zurletrectinib Receives Priority Review for the Treatment of Pediatric Patients with Solid Tumors in China


InnoCare’s next-generation TRKi Zurletrectinib Receives Priority Review for the Treatment of Pediatric Patients with Solid Tumors in China

BEIJING, March 01, 2026 (GLOBE NEWSWIRE) — InnoCare Pharma (HKEX: 09969), a leading biopharmaceutical company for the treatment of cancer and autoimmune diseases, announced today that its next generation TRK inhibitor zurletrectinib (ICP-723) has been granted priority review by the Center for Drug Evaluation (CDE) of the China National Medical Products Administration (NMPA), for the treatment of pediatric patients (aged 2 to 12) with solid tumors harboring NTRK gene fusions. Priority review is one of the key policies introduced by the CDE to accelerate drug approval.

Zurletrectinib has also been included in the “SPARK Program” by the CDE, a pilot initiative to encourage the development of pediatric anti-tumor drugs.

In December 2025, zurletrectinib received approval for the treatment of adult and adolescent patients (aged 12 years and older) with solid tumors harboring NTRK gene fusions in China. In the registrational clinical trial for patients with NTRK fusion-positive solid tumors, zurletrectinib demonstrated outstanding efficacy and a favorable safety profile. The study results showed an objective response rate (ORR) of 89.1%, a disease control rate (DCR) of 96.4%, and 24-month progression-free survival (PFS) and overall survival (OS) rates of 77.4% and 90.8% respectively.

In October 2025, the data from the Phase I/II clinical trial of zurletrectinib for the treatment of pediatric and adolescent patients with advanced solid tumors were released at the Congress of International Society of Pediatric Oncology (SIOP) 2025 as an oral presentation. Zurletrectinib demonstrated a well-tolerated safety profile and promising antitumor activity in pediatric/adolescent patients with NTRK/ROS1-altered solid tumors. The results highlight zurletrectinib’s strong potential as a next-generation therapy for NTRK/ROS1-driven malignancies, with the ability to overcome resistance to first-generation TRK inhibitors.

NTRK fusion genes occur in various types of adult and pediatric tumors. In some rare tumors, such as salivary gland carcinoma, secretory breast cancer, and infantile fibrosarcoma, the incidence of NTRK gene fusion exceeds 90%1. It is estimated that there are about 6,500 new cases of NTRK fusion-positive solid tumors diagnosed in China each year. There are significant unmet clinical needs in this area due to the lack of effective treatment options.

About InnoCare

InnoCare is a commercial stage biopharmaceutical company committed to discovering, developing, and commercializing first-in-class and/or best-in-class drugs for the treatment of cancers and autoimmune diseases with unmet medical needs in China and worldwide. InnoCare has branches in Beijing, Nanjing, Shanghai, Guangzhou, Hong Kong, and the United States.

InnoCare Forward-looking Statements

This report contains the disclosure of some forward-looking statements. Except for statements of facts, all other statements can be regarded as forward-looking statements, that is, about our or our management’s intentions, plans, beliefs, or expectations that will or may occur in the future. Such statements are assumptions and estimates made by our management based on its experience and knowledge of historical trends, current conditions, expected future development and other related factors. This forward-looking statement does not guarantee future performance, and actual results, development and business decisions may not match the expectations of the forward-looking statement. Our forward-looking statements are also subject to a large number of risks and uncertainties, which may affect our short-term and long-term performance.

Contact
   
Media Investors
Chunhua Lu
86-10-66609879
chunhua.lu@innocarepharma.com
86-10-66609999
ir@innocarepharma.com


1
Cocco, E., Scaltriti, M., and Drilon, A. (2018). NTRK fusion-positive cancers and TRK inhibitor therapy. Nature Reviews Clinical Oncology 15, 731-747.

Investor Education Webinar

Investor Education Webinar


Investor Education Webinar

The Future of Prostate Imaging: What do physicians want?

MELBOURNE, Australia and INDIANAPOLIS, March 02, 2026 (GLOBE NEWSWIRE) — Telix Pharmaceuticals Limited (ASX: TLX, NASDAQ: TLX, “Telix”) invites investors to join a webinar with key opinion leaders to discuss a physician’s perspective on innovations in PSMA-PET/CT imaging for prostate cancer.

Speakers:

  • Rodney Hicks, MD, Professor of Medicine at the University of Melbourne and Monash University; Founder, Executive Chairman and Chief Medical Officer at the Melbourne Theranostic Innovation Centre (MTIC);
  • Paul Yonover, MD, FACS, Chief Data Officer and Director of Clinical Navigation at UroPartners, a division of The Specialty Alliance; Chief of Urology at Ascension Saint Joseph Hospital, Chicago; and
  • Darren Patti, PharmD, Telix Group Chief Operating Officer.

The webinar will be moderated by David N. Cade, MD, Telix Group Chief Medical Officer.

The webinar will be held on:

EST: Wednesday March 4, 2026, 5:30 p.m.
AEDT: Thursday March 5, 2026, 9:30 a.m.
The event will run for approximately one hour.

Participants can register for the webcast at the following link: https://edge.media-server.com/mmc/p/n87mam6r

About Telix Pharmaceuticals Limited

Telix is a biopharmaceutical company focused on the development and commercialization of therapeutic and diagnostic radiopharmaceuticals and associated medical technologies. Telix is headquartered in Melbourne, Australia, with international operations in the United States, United Kingdom, Brazil, Canada, Europe (Belgium and Switzerland), and Japan. Telix is developing a portfolio of clinical and commercial stage products that aims to address significant unmet medical needs in oncology and rare diseases. Telix is listed on the Australian Securities Exchange (ASX: TLX) and the Nasdaq Global Select Market (NASDAQ: TLX).

Visit www.telixpharma.com for further information about Telix, including details of the latest share price, ASX and U.S. Securities and Exchange Commission (SEC) filings, investor and analyst presentations, news releases, event details and other publications that may be of interest. You can also follow Telix on LinkedIn, X and Facebook.

Telix Investor Relations (Global)

Ms. Kyahn Williamson
SVP Investor Relations and Corporate Communications

kyahn.williamson@telixpharma.com

Telix Investor Relations (Australia)

Ms. Charlene Jaw
Associate Director Investor Relations

charlene.jaw@telixpharma.com

Telix Investor Relations (U.S.) 

Ms. Annie Kasparian 
Director Investor Relations and Corporate Communications 

annie.kasparian@telixpharma.com

Legal Notices

Cautionary Statement Regarding Forward-Looking Statements. 

You should read this announcement together with our risk factors, as disclosed in our most recently filed reports with the Australian Securities Exchange (ASX), U.S. Securities and Exchange Commission (SEC), including our Annual Report on Form 20-F filed with the SEC, or on our website.

The information contained in this announcement is not intended to be an offer for subscription, invitation or recommendation with respect to securities of Telix Pharmaceuticals Limited (Telix) in any jurisdiction, including the United States. The information and opinions contained in this announcement are subject to change without notification.  To the maximum extent permitted by law, Telix disclaims any obligation or undertaking to update or revise any information or opinions contained in this announcement, including any forward-looking statements (as referred to below), whether as a result of new information, future developments, a change in expectations or assumptions, or otherwise. No representation or warranty, express or implied, is made in relation to the accuracy or completeness of the information contained or opinions expressed in the course of this announcement.

This announcement may contain forward-looking statements, including within the meaning of the U.S. Private Securities Litigation Reform Act of 1995, that relate to anticipated future events, financial performance, plans, strategies or business developments. Forward-looking statements can generally be identified by the use of words such as “may”, “expect”, “intend”, “plan”, “estimate”, “anticipate”, “believe”, “outlook”, “forecast” and “guidance”, or the negative of these words or other similar terms or expressions. Forward-looking statements involve known and unknown risks, uncertainties and other factors that may cause our actual results, levels of activity, performance or achievements to differ materially from any future results, levels of activity, performance or achievements expressed or implied by these forward-looking statements. Forward-looking statements are based on Telix’s good-faith assumptions as to the financial, market, regulatory and other risks and considerations that exist and affect Telix’s business and operations in the future and there can be no assurance that any of the assumptions will prove to be correct. In the context of Telix’s business, forward-looking statements may include, but are not limited to, statements about: the initiation, timing, progress, completion and results of Telix’s preclinical and clinical trials, and Telix’s research and development programs; Telix’s ability to advance product candidates into, enroll and successfully complete, clinical studies, including multi-national clinical trials; the timing or likelihood of regulatory filings and approvals for Telix’s product candidates, including the planned NDA resubmission for TLX101-Px and the planned BLA resubmission for TLX250-Px, manufacturing activities and product marketing activities; Telix’s sales, marketing and distribution and manufacturing capabilities and strategies; the commercialization of Telix’s product candidates, if or when they have been approved; Telix’s ability to obtain an adequate supply of raw materials at reasonable costs for its products and product candidates; estimates of Telix’s expenses, future revenues and capital requirements; Telix’s financial performance; developments relating to Telix’s competitors and industry; the anticipated impact of U.S. and foreign tariffs and other macroeconomic conditions on Telix’s business; and the pricing and reimbursement of Telix’s product candidates, if and after they have been approved. Telix’s actual results, performance or achievements may be materially different from those which may be expressed or implied by such statements, and the differences may be adverse. Accordingly, you should not place undue reliance on these forward-looking statements.

Trademarks and Trade Names. All trademarks and trade names referenced in this press release are the property of Telix Pharmaceuticals Limited (Telix) or, where applicable, the property of their respective owners. For convenience, trademarks and trade names may appear without the ® or ™ symbols. Such omissions are not intended to indicate any waiver of rights by Telix or the respective owners. Trademark registration status may vary from country to country. Telix does not intend the use or display of any third-party trademarks or trade names to imply any affiliation with, endorsement by, or sponsorship from those third parties.

©2026 Telix Pharmaceuticals Limited. All rights reserved.